
TVTX 10-Q
Travere Therapeutics, Inc. (TVTX)
10-Q
2026-08-04
For: 2026-06-30
View Original
Added on
August 04, 2026
UNITED STATES
SECURITIES AND EXCHANGE COMMISSION
WASHINGTON, D.C. 20549
_________________________________
FORM 10-Q
_________________________________
| QUARTERLY REPORT PURSUANT TO SECTION 13 OR 15(d) OF THE SECURITIES EXCHANGE ACT OF 1934 | ||||||||||||||
For the quarterly period ended June 30, 2026
or
| TRANSITION REPORT PURSUANT TO SECTION 13 OR 15(d) OF THE SECURITIES EXCHANGE ACT OF 1934 | ||||||||||||||
For the transition period from ___________ to ___________
Commission File Number: 001-36257
(Exact name of registrant as specified in its charter)
| (State or other jurisdiction of incorporation or organization) | (I.R.S. Employer Identification No.) | |||||||||||||
(Address of Principal Executive Offices)
(888 ) 969-7879
(Registrant's Telephone number including area code)
N/A | ||||||||
Former name, former address and former fiscal year, if changed since last report | ||||||||
Securities registered pursuant to Section 12(b) of the Act:
| Title of each class | Trading Symbol(s) | Name of each exchange on which registered | ||||||||||||
Indicate by check mark whether the registrant: (1) has filed all reports required to be filed by Section 13 or 15(d) of the Securities Exchange Act of 1934 during the preceding 12 months (or for such shorter period that the registrant was required to file such reports), and (2) has been subject to such filing requirements for the past 90 days. Yes ☑ No ☐
Indicate by check mark whether the registrant has submitted electronically every Interactive Data File required to be submitted pursuant to Rule 405 of Regulation S-T (§232.405 of this chapter) during the preceding 12 months (or for such shorter period that the registrant was required to submit such files). Yes ☑ No ☐
Indicate by check mark whether the registrant is a large accelerated filer, an accelerated filer, a non-accelerated filer, a smaller reporting company, or an emerging growth company. See the definitions of “large accelerated filer,” “accelerated filer,” “smaller reporting company,” and "emerging growth company" in Rule 12b-2 of the Exchange Act.
☑ | Accelerated filer | ☐ | |||||||||
| Non-accelerated filer | ☐ | Smaller reporting company | |||||||||
| Emerging growth company | |||||||||||
If an emerging growth company, indicate by check mark if the registrant has elected not to use the extended transition period for complying with any new or revised financial accounting standards provided pursuant to Section 13(a) of the Exchange Act. ☐
Indicate by check mark whether the registrant is a shell company (as defined in Rule 12b-2 of the Exchange Act). Yes ☐ No ☑
The number of shares of outstanding common stock, par value $0.0001 per share, of the Registrant as of July 30, 2026 was 94,261,082 .
TRAVERE THERAPEUTICS, INC.
Form 10-Q
For the Fiscal Quarter Ended June 30, 2026
TABLE OF CONTENTS
| Page No. | |||||||||||
1
FORWARD-LOOKING STATEMENTS
This report contains forward-looking statements regarding our business, financial condition, results of operations and prospects. Words such as “expects,” “anticipates,” “intends,” “plans,” “believes,” “seeks,” “estimates” and similar expressions or variations of such words are intended to identify forward-looking statements, but are not deemed to represent an all-inclusive means of identifying forward-looking statements as denoted in this report. Additionally, statements concerning future matters are forward-looking statements.
Although forward-looking statements in this report reflect the good faith judgment of our management, such statements can only be based on facts and factors currently known by us. Consequently, forward-looking statements are inherently subject to risks and uncertainties and actual results and outcomes may differ materially from the results and outcomes discussed in or anticipated by the forward-looking statements. Factors that could cause or contribute to such differences in results and outcomes include, without limitation, those specifically addressed under the headings “Risk Factors” and “Management’s Discussion and Analysis of Financial Condition and Results of Operations” in our Annual Report on Form 10-K for the fiscal year ended December 31, 2025, and in this Quarterly Report on Form 10-Q. You are urged not to place undue reliance on these forward-looking statements, which speak only as of the date of this report.
In addition, statements that "we believe" and similar statements reflect our beliefs and opinions on the relevant subject. These statements are based upon information available to us as of the date of this Quarterly Report on Form 10-Q, and while we believe such information forms a reasonable basis for such statements, such information may be limited or incomplete, and our statements should not be read to indicate that we have conducted an exhaustive inquiry into, or review of, all potentially available relevant information. These statements are inherently uncertain and you are cautioned to not unduly rely upon these statements.
We file reports with the Securities and Exchange Commission ("SEC"). The SEC maintains a website (www.sec.gov) that contains reports, proxy and information statements, and other information regarding issuers that file electronically with the SEC, including us.
We undertake no obligation to revise or update any forward-looking statements in order to reflect any event or circumstance that may arise after the date of this report, except as required by law. Readers are urged to carefully review and consider the various disclosures made throughout the entirety of this quarterly report, which are designed to advise interested parties of the risks and factors that may affect our business, financial condition, results of operations and prospects.
Risk Factor Summary
Below is a summary of material factors that make an investment in our common stock speculative or risky. This summary does not address all of the risks that we face. Additional discussion of the risks summarized in this risk factor summary, and other risks that we face, can be found under the heading “Risk Factors” in Item 1A of Part II of this Quarterly Report on Form 10-Q and should be carefully considered, together with other information in this Quarterly Report on Form 10-Q and our other filings with the SEC before making investment decisions regarding our common stock.
•Our future prospects are highly dependent upon our ability to successfully develop and execute commercialization strategies for our products, including FILSPARI, and to attain market acceptance among physicians, patients and healthcare payers.
•In order to operate our business and increase adoption and sales of our products, we need to continue to develop our commercial organization, including maintaining a highly experienced and skilled workforce with qualified sales representatives.
•We face substantial generic and other competition, and our operating results will suffer if we fail to compete effectively.
•Healthcare reform initiatives, unfavorable pricing regulations, and changes in reimbursement practices of third-party payers or patients' access to insurance coverage could affect the pricing of and demand for our products.
•We are dependent on third parties to manufacture and distribute our products.
•Our clinical trials are expensive and time-consuming and may fail to demonstrate the safety and efficacy of our product candidates.
•Success in nonclinical testing and early clinical trials does not ensure that later clinical trials will be successful.
•Communications and/or feedback from regulatory authorities related to our current or planned future clinical trials does not guarantee any particular outcome from or timeline for regulatory review, and expedited regulatory review pathways may not actually lead to faster development or approval.
•Interim, topline and preliminary data from our clinical trials that we announce or publish may change materially as more patient data become available and audit and verification procedures are complete.
•The market opportunities for our products and product candidates may be smaller than we believe they are.
•Our product candidates may cause undesirable side effects or have other properties that could delay or prevent their regulatory approval or commercialization.
•We do not currently have patent protection for certain of our commercial products. If we are unable to obtain and maintain protection for the intellectual property relating to our technology and products, their value will be adversely affected.
•We expect to rely on orphan drug status to develop and commercialize certain of our products and product candidates, but our orphan drug designations may not confer marketing exclusivity or other expected commercial benefits.
2
•If we are unable to obtain and maintain coverage and adequate reimbursement from governments or third-party payers for any products that we may develop or if we are unable to obtain acceptable prices for those products, our prospects for generating revenue and achieving profitability will suffer.
•International trade policies, including tariffs, sanctions and trade barriers may adversely affect our business, financial condition, results of operations and prospects.
•We will likely experience fluctuations in operating results and could incur substantial losses.
•Negative publicity regarding any of our products could impair our ability to market any such product and may require us to spend time and money to address these issues.
•We may need substantial funding and may be unable to raise capital when needed.
•We may not receive some or all of the potential milestone and/or royalty payments from our corporate and licensing transactions.
•We may be unable to successfully integrate new products or businesses we may acquire.
•We may become involved in litigation matters, which could result in substantial costs, divert management's attention and otherwise have a material adverse effect on our business, operating results or financial condition.
•We are subject to significant ongoing regulatory obligations and oversight, which may result in significant additional expense and may limit our commercial success.
3
PART I - FINANCIAL INFORMATION
Item 1. Financial Statements
TRAVERE THERAPEUTICS, INC. AND SUBSIDIARIES
CONSOLIDATED BALANCE SHEETS
(in thousands, except par value and share amounts)
| June 30, 2026 | December 31, 2025 | ||||||||||
| Assets | (unaudited) | ||||||||||
| Current assets: | |||||||||||
| Cash and cash equivalents | $ | $ | |||||||||
| Marketable debt securities, at fair value | |||||||||||
| Accounts receivable, net | |||||||||||
| Inventory | |||||||||||
| Prepaid expenses and other current assets | |||||||||||
| Total current assets | |||||||||||
| Long-term inventory | |||||||||||
| Property and equipment, net | |||||||||||
| Operating lease right of use assets | |||||||||||
| Intangible assets, net | |||||||||||
| Other assets | |||||||||||
| Total assets | $ | $ | |||||||||
Liabilities and Stockholders' Equity | |||||||||||
| Current liabilities: | |||||||||||
| Accounts payable | $ | $ | |||||||||
| Accrued expenses | |||||||||||
| Operating lease liabilities, current portion | |||||||||||
| Other current liabilities | |||||||||||
| Total current liabilities | |||||||||||
| Convertible debt | |||||||||||
| Operating lease liabilities, less current portion | |||||||||||
| Other non-current liabilities | |||||||||||
| Total liabilities | |||||||||||
| Commitments and Contingencies (See Note 13) | |||||||||||
Stockholders' Equity: | |||||||||||
Preferred stock $ | |||||||||||
Common stock $ | |||||||||||
| Additional paid-in capital | |||||||||||
| Accumulated deficit | ( | ( | |||||||||
| Accumulated other comprehensive loss | ( | ( | |||||||||
Total stockholders' equity | |||||||||||
Total liabilities and stockholders' equity | $ | $ | |||||||||
The accompanying notes are an integral part of these consolidated financial statements.
4
TRAVERE THERAPEUTICS, INC. AND SUBSIDIARIES
CONSOLIDATED STATEMENTS OF OPERATIONS AND COMPREHENSIVE LOSS
(unaudited, in thousands, except share and per share amounts)
| Three Months Ended June 30, | Six Months Ended June 30, | ||||||||||||||||||||||
| 2026 | 2025 | 2026 | 2025 | ||||||||||||||||||||
| Net product sales | $ | $ | $ | $ | |||||||||||||||||||
| License and collaboration revenue | |||||||||||||||||||||||
| Total revenue | |||||||||||||||||||||||
| Operating expenses: | |||||||||||||||||||||||
| Cost of goods sold | |||||||||||||||||||||||
| Research and development | |||||||||||||||||||||||
| Selling, general and administrative | |||||||||||||||||||||||
| Royalty expense | |||||||||||||||||||||||
| Total operating expenses | |||||||||||||||||||||||
Operating income (loss) | ( | ( | ( | ||||||||||||||||||||
Other (expense) income, net: | |||||||||||||||||||||||
| Interest income | |||||||||||||||||||||||
| Interest expense | ( | ( | ( | ( | |||||||||||||||||||
Other (expense) income, net | ( | ( | ( | ||||||||||||||||||||
Inducement expense | ( | ( | |||||||||||||||||||||
Total other (expense) income, net | ( | ( | ( | ||||||||||||||||||||
Loss from continuing operations before income tax | ( | ( | ( | ( | |||||||||||||||||||
Income tax benefit (provision) on continuing operations | ( | ( | |||||||||||||||||||||
Loss from continuing operations, net of tax | ( | ( | ( | ( | |||||||||||||||||||
| Loss from discontinued operations, net of tax | ( | ||||||||||||||||||||||
Net loss | $ | ( | $ | ( | $ | ( | $ | ( | |||||||||||||||
| Per share data | |||||||||||||||||||||||
| Basic and diluted: | |||||||||||||||||||||||
Net loss per common share | $ | ( | $ | ( | $ | ( | $ | ( | |||||||||||||||
Weighted average common shares outstanding | |||||||||||||||||||||||
| Comprehensive loss: | |||||||||||||||||||||||
Net loss | $ | ( | $ | ( | $ | ( | $ | ( | |||||||||||||||
Foreign currency translation gain (loss) | ( | ||||||||||||||||||||||
Unrealized loss on marketable debt securities | ( | ( | ( | ( | |||||||||||||||||||
| Comprehensive loss | $ | ( | $ | ( | $ | ( | $ | ( | |||||||||||||||
The accompanying notes are an integral part of these consolidated financial statements.
5
TRAVERE THERAPEUTICS, INC. AND SUBSIDIARIES
CONSOLIDATED STATEMENTS OF STOCKHOLDERS' EQUITY
(unaudited, in thousands, except share amounts)
| Three Months Ended June 30, 2026 | Three Months Ended June 30, 2025 | ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Common Stock | Additional Paid in Capital | Accumulated Other Comprehensive Loss | Accumulated Deficit | Total Stockholders' Equity | Common Stock | Additional Paid in Capital | Accumulated Other Comprehensive Loss | Accumulated Deficit | Total Stockholders' Equity | ||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Shares | Amount | Shares | Amount | ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Balance - March 31 | $ | $ | $ | ( | $ | ( | $ | $ | $ | $ | ( | $ | ( | $ | |||||||||||||||||||||||||||||||||||||||||||||||||||
| Share based compensation | — | — | — | — | — | — | — | — | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Issuance of common stock under the equity incentive plan and proceeds from exercise | — | — | — | — | — | — | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Employee stock purchase program purchase and expense | — | — | — | — | — | — | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
Induced conversion on 2029 Notes | — | — | ( | — | — | ( | — | — | — | — | — | ||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Foreign currency translation adjustments | — | — | — | — | — | — | — | — | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Unrealized loss on marketable debt securities | — | — | — | ( | — | ( | — | — | — | ( | — | ( | |||||||||||||||||||||||||||||||||||||||||||||||||||||
| Net loss | — | — | — | — | ( | ( | — | — | — | — | ( | ( | |||||||||||||||||||||||||||||||||||||||||||||||||||||
| Balance - June 30 | $ | $ | $ | ( | $ | ( | $ | $ | $ | $ | ( | $ | ( | $ | |||||||||||||||||||||||||||||||||||||||||||||||||||
The accompanying notes are an integral part of these consolidated financial statements.
6
TRAVERE THERAPEUTICS, INC. AND SUBSIDIARIES
CONSOLIDATED STATEMENTS OF STOCKHOLDERS' EQUITY
(unaudited, in thousands, except share amounts)
| Six Months Ended June 30, 2026 | Six Months Ended June 30, 2025 | ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Common Stock | Additional Paid in Capital | Accumulated Other Comprehensive Loss | Accumulated Deficit | Total Stockholders' Equity | Common Stock | Additional Paid in Capital | Accumulated Other Comprehensive Loss | Accumulated Deficit | Total Stockholders' Equity | ||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Shares | Amount | Shares | Amount | ||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Balance - December 31 | $ | $ | $ | ( | $ | ( | $ | $ | $ | $ | ( | $ | ( | $ | |||||||||||||||||||||||||||||||||||||||||||||||||||
| Share based compensation | — | — | — | — | — | — | — | — | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Issuance of common stock under the equity incentive plan and proceeds from exercise | — | — | — | — | — | — | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Employee stock purchase program purchase and expense | — | — | — | — | — | — | |||||||||||||||||||||||||||||||||||||||||||||||||||||||||||
Induced conversion on 2029 Notes | — | — | ( | — | ( | — | — | — | — | — | |||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Foreign currency translation adjustments | — | — | — | — | — | — | — | ( | — | ( | |||||||||||||||||||||||||||||||||||||||||||||||||||||||
| Unrealized loss on marketable debt securities | — | — | — | ( | — | ( | — | — | — | ( | — | ( | |||||||||||||||||||||||||||||||||||||||||||||||||||||
| Net loss | — | — | — | — | ( | ( | — | — | — | — | ( | ( | |||||||||||||||||||||||||||||||||||||||||||||||||||||
| Balance - June 30 | $ | $ | $ | ( | $ | ( | $ | $ | $ | $ | ( | $ | ( | $ | |||||||||||||||||||||||||||||||||||||||||||||||||||
7
TRAVERE THERAPEUTICS, INC. AND SUBSIDIARIES
CONSOLIDATED STATEMENTS OF CASH FLOWS
(unaudited, in thousands)
| For the Six Months Ended June 30, | |||||||||||
| 2026 | 2025 | ||||||||||
| Cash Flows From Operating Activities: | |||||||||||
| Net loss | $ | ( | $ | ( | |||||||
| Net loss from discontinued operations | ( | ||||||||||
| Net loss from continuing operations | ( | ( | |||||||||
| Adjustments to reconcile net loss to net cash provided by (used in) operating activities: | |||||||||||
| Depreciation and amortization | |||||||||||
| Share based compensation | |||||||||||
| Foreign currency remeasurement impact | ( | ||||||||||
Inducement expense | |||||||||||
| Other | ( | ( | |||||||||
| Changes in operating assets and liabilities: | |||||||||||
| Accounts receivable | ( | ( | |||||||||
| Inventory | ( | ||||||||||
| Prepaid expenses and other current and non-current assets | ( | ( | |||||||||
| Accounts payable | ( | ( | |||||||||
| Accrued expenses | ( | ||||||||||
| Deferred revenue | ( | ||||||||||
| Other current and non-current liabilities | |||||||||||
| Net cash provided by (used in) operating activities | ( | ||||||||||
| Cash Flows From Investing Activities: | |||||||||||
| Proceeds from the sale and maturity of marketable debt securities | |||||||||||
| Purchase of marketable debt securities | ( | ( | |||||||||
| Purchase of intangible assets | ( | ( | |||||||||
| Other | ( | ||||||||||
| Net cash (used in) provided by investing activities | ( | ||||||||||
| Cash Flows From Financing Activities: | |||||||||||
| Payment of guaranteed minimum royalty | ( | ( | |||||||||
Proceeds from issuance of 2032 convertible senior notes | |||||||||||
| Payment of debt issuance costs | ( | ||||||||||
Repurchase of 2029 convertible senior notes including premium | ( | ||||||||||
| Proceeds from exercise of stock options | |||||||||||
| Proceeds from issuances under the employee stock purchase plan | |||||||||||
| Net cash provided by financing activities | |||||||||||
| Effect of exchange rate changes on cash | |||||||||||
| Net increase in cash and cash equivalents | |||||||||||
| Cash and cash equivalents, beginning of year | |||||||||||
| Cash and cash equivalents, end of period | $ | $ | |||||||||
The accompanying notes are an integral part of these consolidated financial statements.
8
TRAVERE THERAPEUTICS, INC. AND SUBSIDIARIES
NOTES TO THE UNAUDITED CONSOLIDATED FINANCIAL STATEMENTS
NOTE 1. DESCRIPTION OF BUSINESS
Organization and Description of Business
Travere Therapeutics, Inc. (“we”, “our”, “us”, “Travere” and the “Company”) refers to Travere Therapeutics, Inc., a Delaware corporation, as well as its subsidiaries. Travere is a fully integrated biopharmaceutical company headquartered in San Diego, California focused on identifying, developing and delivering life-changing therapies to people living with rare kidney and metabolic diseases. The Company regularly evaluates and, where appropriate, acts on opportunities to expand its product pipeline and approved products through licenses and acquisitions of products in areas that will serve rare disease patients with serious unmet medical need and that the Company believes offer attractive growth characteristics.
Approved Products:
FILSPARI® (sparsentan)
Treatment of IgAN
In September 2024, the FDA granted full approval of FILSPARI® (sparsentan) to slow kidney function decline in adults with primary Immunoglobulin A nephropathy ("IgAN") who are at risk of disease progression. FILSPARI had previously been granted accelerated approval in February 2023 based on the surrogate marker of proteinuria. Full approval was based on positive long-term confirmatory results from the PROTECT Study demonstrating that FILSPARI significantly slowed kidney function decline over two years compared to irbesartan.
FILSPARI is the only oral, once-daily, non-immunosuppressive medication that directly targets glomerular injury in the kidney by blocking two critical pathways of IgAN disease progression (endothelin-1 and angiotensin II).
Treatment of FSGS
On April 13, 2026, the FDA granted full (traditional) approval of FILSPARI (sparsentan) to reduce proteinuria in adult and pediatric patients aged 8 years and older with focal segmental glomerulosclerosis (“FSGS”) without nephrotic syndrome. In the Phase 3 DUPLEX Study patients treated with FILSPARI in the overall study population experienced a significant reduction in proteinuria compared to those treated with maximum labeled dose irbesartan.
FILSPARI is the first and only medicine approved by the FDA for the treatment of FSGS.
License and Collaboration Agreements
In September 2021, the Company entered into a license and collaboration agreement (the "CSL Vifor License Agreement") with Vifor (International) Ltd. ("CSL Vifor"). In April 2024, the Company and CSL Vifor announced that the European Commission had granted conditional marketing authorization (“CMA”) for FILSPARI (sparsentan) for the treatment of adults with primary IgAN with a urine protein excretion ≥1.0 g/day (or urine protein-to-creatinine ratio ("UPCR") ≥0.75 g/g), and in April 2025, the Company and CSL Vifor announced that the European Commission had converted the CMA into a standard marketing authorization (“MA”) for FILSPARI for the treatment of adults with primary IgAN with a urine protein excretion ≥1.0 g/day (or UPCR ≥0.75 g/g). The MA is granted for all member states of the European Union, as well as in Iceland, Liechtenstein and Norway.
In January 2024, the Company entered into an exclusive licensing agreement with Renalys Pharma, Inc. ("Renalys"), to bring sparsentan for the treatment of IgAN to patients in Japan and other countries in Asia. In December 2024, Renalys announced that sparsentan received Orphan Drug Designation from the Japanese Ministry of Health, Labour and Welfare for the indication of primary IgA nephropathy in November 2024. In the fourth quarter of 2025, Renalys announced positive topline results from its Phase 3 study of sparsentan in Japanese patients with IgAN. Renalys has also announced that it has reached an agreement with the Pharmaceuticals and Medical Devices Agency ("PMDA") regarding development plans for two other Phase 3 clinical trials of sparsentan, one investigating the use of sparsentan in focal segmental glomerulosclerosis ("FSGS") and the other in Alport syndrome, in Japan. In the fourth quarter of 2025, Renalys was acquired by and merged into Chugai Pharmaceutical Co., Ltd. (“Chugai”). Through the acquisition, Chugai gained exclusive rights to develop and commercialize sparsentan in Japan, South Korea, and Taiwan. As a minority shareholder in Renalys, Travere received $10.2 million at the closing of the transaction in the fourth quarter of 2025, and Travere is also eligible to receive multiple milestones according to the progress of sparsentan regulatory approval, and consideration linked to sparsentan's net sales in the applicable territory. Under the terms of the licensing agreement, Chugai is responsible for development, regulatory matters, and commercialization in the licensed territories.
Thiola® and Thiola EC® (tiopronin tablets)
Thiola® and Thiola EC® (tiopronin tablets) are approved in the United States for the prevention of cystine (kidney) stone formation in patients with severe homozygous cystinuria.
Clinical-Stage Programs:
Pegtibatinase
Pegtibatinase is a novel investigational human enzyme replacement candidate being evaluated for the treatment of classical homocystinuria ("HCU"). Pegtibatinase has been granted Rare Pediatric Disease, Fast Track and Breakthrough Therapy designations by the FDA, as well as orphan drug designation in the United States and European Union. In May 2023, the Company announced positive topline results from cohort 6 in the Phase 1/2 COMPOSE Study. In December 2023, the Company initiated the pivotal Phase 3 HARMONY Study to support the potential approval of pegtibatinase for the treatment of classical HCU. The HARMONY Study is a global, randomized, multi-center, double-blind, placebo-controlled Phase 3 clinical trial designed to evaluate the efficacy
9
and safety of pegtibatinase as a novel treatment to reduce total homocysteine ("tHcy") levels. In the beginning of 2024, the first patients were dosed in the HARMONY Study.
In September 2024, the Company announced a voluntary pause of enrollment in the Phase 3 HARMONY Study. The voluntary enrollment pause was enacted following the Company's determination that the desired drug substance profile was not achieved in the initial scale-up process, and it enabled the Company to address necessary process improvements in manufacturing scale-up to support initial commercial scale manufacturing as well as full enrollment in the HARMONY Study. Following process improvements, the Company restarted enrollment activities in the first quarter of 2026 and dosed the first new patient following study restart in April 2026.
The Company acquired pegtibatinase as part of the November 2020 acquisition of Orphan Technologies Limited.
NOTE 2. BASIS OF PRESENTATION AND SIGNIFICANT ACCOUNTING POLICIES
The accompanying unaudited consolidated financial statements should be read in conjunction with the audited consolidated financial statements and notes thereto included in our Annual Report on Form 10-K for the year ended December 31, 2025, filed with the SEC on February 19, 2026. The accompanying consolidated financial statements have been prepared in accordance with accounting principles generally accepted in the United States (“GAAP”) for interim financial information, the instructions for Form 10-Q and the rules and regulations of the SEC. Accordingly, since they are interim statements, the accompanying consolidated financial statements do not include all of the information and notes required by GAAP for annual financial statements, but reflect all adjustments consisting of normal, recurring adjustments, that are necessary for a fair statement of the financial position, results of operations and cash flows for the interim periods presented. Interim results are not necessarily indicative of the results that may be expected for any future periods. The December 31, 2025 balance sheet information was derived from the audited financial statements as of that date.
Certain reclassifications have been made to the prior period consolidated financial statements to conform to the current period presentation. The Company revised the presentation of amortization expense of certain royalty and milestone payments associated with license agreements acquired in prior asset acquisitions that were accounted for under the asset accumulation method. Historically, the royalty amortization expense was included within selling, general and administrative expenses in the consolidated statements of operations. Beginning in the current year, the Company presents these amounts separately as royalty expense to better reflect the nature of these costs as consideration payable to licensors based on sales of licensed products and regulatory milestone payments. As a result of this change in presentation, $13.6 million and $26.1 million of amortization expense previously included within selling, general and administrative expenses has been reclassified to royalty expense for the three and six months ended June 30, 2025, respectively. This reclassification had no impact on total operating expenses, loss from operations, net loss, net loss per share, or cash flows for any period presented.
A summary of the significant accounting policies applied in the preparation of the accompanying consolidated financial statements follows:
Principles of Consolidation
The unaudited consolidated financial statements represent the consolidation of the accounts of the Company, its subsidiaries and variable interest entities for which the Company has been determined to be the primary beneficiary, in conformity with GAAP. All intercompany accounts and transactions have been eliminated in consolidation. See Note 6 for further discussion of variable interest entities (“VIE”) that the Company consolidates.
Revenue Recognition
Payments received under collaboration and licensing agreements may include non-refundable fees at the inception of the arrangements, milestone payments for specific achievements and royalties on the sale of products. At the inception of arrangements that include milestone payments, the Company uses judgment to evaluate whether the milestones are probable of being achieved and estimates the amount to include in the transaction price utilizing the most likely amount method. If it is probable that a significant revenue reversal will not occur, the estimated amount is included in the transaction price. Milestone payments that are not within the Company or the licensee’s control, such as regulatory approvals, are considered to be constrained due to a high degree of uncertainty and are not included in the transaction price until such uncertainty is resolved. At the end of each reporting period, the Company re-evaluates the probability of achievement of development milestones and any related constraint and adjusts the estimate of the overall transaction price, if necessary. The Company recognizes aggregate sales-based milestones and royalty payments from product sales of which the license is deemed to be the predominant item to which the royalties relate, at the later of when the related sales occur or when the performance obligation has been satisfied. Revenue from collaboration and licensing agreements may also include sales of inventory, at cost plus a margin, which is recorded in license and collaboration revenue.
The Company utilizes significant judgment to develop estimates of the stand-alone selling price for each distinct performance obligation based upon the relative stand-alone selling price. Variable consideration that relates specifically to the Company’s efforts to satisfy specific performance obligations is allocated entirely to those performance obligations. The stand-alone selling price for license-related performance obligations requires judgment in developing assumptions to project probability-weighted cash flows based upon estimates of forecasted revenues, clinical and regulatory timelines and discount rates. The stand-alone selling price for clinical development performance obligations is based on forecasted expected costs of satisfying a performance obligation plus an appropriate margin.
10
If the licenses to intellectual property are determined to be distinct from the other performance obligations identified in the arrangement and have stand-alone functionality, the Company recognizes revenues from non-refundable, upfront fees allocated to the license when the license is transferred to the licensee and the licensee is able to benefit from the license. For licenses that are not distinct from other promises, the Company applies judgment to assess the nature of the combined performance obligation to determine whether the combined performance obligation is satisfied over time or a point in time and, if over time, the appropriate method of measuring progress for purposes of recognizing revenue from non-refundable, upfront fees. The Company evaluates the measure of progress each reporting period and, if necessary, adjusts the related revenue recognition accordingly.
The selection of the method to measure progress towards completion requires judgment and is based on the nature of the products or services to be provided. Revenue is recorded proportionally as costs are incurred. The Company generally utilizes the cost-to-cost method of progress because it best measures the transfer of control to the customer which occurs as the Company incurs costs. Under the cost-to-cost measure of progress, the extent of progress towards completion is measured based on the ratio of costs incurred to date to the total estimated costs at completion of the performance obligation. The Company uses judgment to estimate the total costs expected to complete the clinical development performance obligations, which include subcontractor costs, labor, materials, other direct costs and an allocation of indirect costs. The Company evaluates these cost estimates and the progress each reporting period and adjusts the measure of progress, if necessary.
Cost of goods sold
Cost of goods sold includes the cost of inventory sold, third party manufacturing and supply chain costs, product shipping and handling costs, and provisions for excess and obsolete inventory. Cost of goods sold also includes the cost of goods sold under the Company's license and collaboration agreements, which currently consists of the sale of active pharmaceutical ingredients ("APIs") to the Company's collaboration partners, at cost or at cost plus a margin.
The following table summarizes cost of goods sold for the three and six months ended June 30, 2026 and 2025 (in thousands):
| Three Months Ended June 30, | Six Months Ended June 30, | ||||||||||||||||||||||
| 2026 | 2025 | 2026 | 2025 | ||||||||||||||||||||
| Cost of goods sold - product sales | $ | $ | $ | $ | |||||||||||||||||||
| Cost of goods sold - license and collaboration | |||||||||||||||||||||||
| Total cost of goods sold | $ | $ | $ | $ | |||||||||||||||||||
Capitalization of Inventory Costs
Prior to the regulatory approval of the Company's drug candidates, the Company incurs expenses for the manufacture of drug product supplies to support clinical development that could potentially be available to support the commercial launch of those drugs. The Company capitalizes inventory costs associated with its products after regulatory approval, when, based on management’s judgment, future commercialization is considered probable and the future economic benefit is expected to be realized. Until the date at which regulatory approval has been received, costs related to the production of inventory are recorded as research and development expenses as incurred. Any eventual sale of previously expensed ("zero-cost") inventories may impact future margins, for any periods in which those inventories are sold.
Prior to the February 2023 FDA accelerated approval of FILSPARI (sparsentan) in IgAN, the Company expensed the production of APIs purchased to support the commercial launch of FILSPARI, in research and development expenses. For the three and six months ended June 30, 2025 sales of FILSPARI primarily consisted of zero-cost inventories. The Company began capitalizing inventory costs associated with FILSPARI following the February 2023 accelerated approval.
Research and Development Expenses
Research and development includes expenses related to sparsentan, pegtibatinase, and the Company's other pipeline programs. The Company expenses all research and development costs as they are incurred. The Company's research and development costs are composed of salaries and bonuses, benefits, non-cash share-based compensation, license fees, milestones under license agreements, costs paid to third-party contractors to perform research, conduct clinical trials and develop drug materials and delivery methods, costs to manufacture drug product supplies to support clinical development, and associated overhead expenses and facilities costs. The Company charges direct internal and external program costs to the respective development programs. The Company also incurs indirect costs that are not allocated to specific programs because such costs benefit multiple development programs and allow us to increase our pharmaceutical development capabilities. These consist of internal shared resources related to the development and maintenance of systems and processes applicable to all of our programs.
Clinical Trial Expenses
The Company records expenses in connection with its clinical trials under contracts with contract research organizations ("CROs") that support conducting and managing clinical trials, as well as contract manufacturing organizations ("CMOs") for the manufacture of drug product supplies to support clinical development. The financial terms and activities of these agreements vary from contract to contract and may result in uneven expense levels. Generally, these agreements set forth activities that drive the recording of expenses such as start-up, initiation activities, enrollment, treatment of patients, or the completion of other clinical trial activities, and in the case of CMOs, costs associated with the production of drug product supplied and the procurement of raw materials to be consumed in the manufacturing process.
Expenses related to clinical trials are accrued based on our estimates of the progress of services performed, including actual level of patient enrollment, completion of patient studies and progress of the clinical trials or the delivery of goods. Other incidental costs related to patient enrollment or treatment are
11
accrued when reasonably certain. If the amounts we are obligated to pay under our clinical trial agreements are modified (for instance, as a result of changes in the clinical trial protocol or scope of work to be performed), the Company adjusts its estimates accordingly on a prospective basis. Revisions to the Company's contractual payment obligations are charged to expense in the period in which the facts that give rise to the revision become reasonably certain.
The Company currently has three Phase 3 clinical trials in process that are in varying stages of activity, with ongoing non-clinical support trials. As such, clinical trial expenses will vary depending on all the factors set forth above and may fluctuate significantly from quarter to quarter and year to year.
Earnings (Loss) Per Share
The Company calculates basic earnings per share by dividing net income/(loss) by the weighted average number of shares outstanding during the period.
The Company's diluted earnings/(loss) per share computation includes the effect, if any, of shares that would be issuable upon the exercise of outstanding stock options, convertible debt and RSUs using the treasury stock method. The treasury stock method reduces the number of shares issuable by the shares assumed to be purchased by the Company from the resulting proceeds at the average market price during the year, when such amounts are dilutive to the earnings/(loss) per share calculation. As the Company has reported net losses for all periods presented, all potentially dilutive securities are antidilutive and, accordingly, basic net loss per share equals diluted net loss per share. In accordance with ASC 260, Earnings per Share, if a company had a discontinued operation, the company uses income from continuing operations, adjusted for preferred dividend and similar adjustments, as its control number to determine whether potential common shares are dilutive.
Intangible Assets with Cost Accumulation Model
In 2014, the Company entered into a license agreement with Mission Pharmacal ("Mission") in which the Company obtained the exclusive right to license the trademark of Thiola ("Mission License Agreement"). The acquisition of the Thiola license qualified as an asset acquisition under the principles of ASC 805, Business Combinations ("ASC 805") in effect at the time of acquisition. The license agreement requires the Company to make royalty payments based on net sales of Thiola. The liability for royalties in excess of the annual contractual minimum is recognized in the period in which the royalties become probable and estimable, which is typically in the period corresponding with the respective sales. The Company records an offsetting increase to the cost basis of the intangible asset under the cost accumulation model ("Thiola Intangible"). The additional cost basis is subsequently amortized over the remaining estimated useful life of the license agreement.
Consistent with all prior periods since Thiola was acquired, the Company has not accrued any liability for future royalties in excess of the annual contractual minimum at June 30, 2026 as such royalties are not yet probable and estimable.
In 2012, the Company entered into an agreement with Ligand Pharmaceuticals, Inc. (“Ligand”) for a worldwide sublicense to develop, manufacture and commercialize sparsentan (the “Ligand License Agreement”). The acquisition of the Ligand License Agreement qualified as an asset acquisition under the principles of ASC 805 in effect at the time of acquisition. The license agreement requires the Company to make royalty payments based on net sales of FILSPARI (sparsentan) and milestone payments. The liabilities for royalties and milestone payments are recognized in the period in which they become probable and estimable, which is typically in the period corresponding with the respective sales or achievement of the milestone. The Company records an offsetting increase to the cost basis of the intangible asset under the cost accumulation model following the approval of FILSPARI. The additional cost basis is subsequently amortized over the remaining estimated useful life.
Royalty expense includes amortization of royalties and milestones paid that are capitalized to the intangible asset under the asset accumulation model as well as royalties expensed after the estimated useful life of the license agreement has been reached.
The following table summarizes royalty expense for the three and six months ended June 30, 2026 and 2025 (in thousands):
| Three Months Ended June 30, | Six Months Ended June 30, | ||||||||||||||||||||||
| 2026 | 2025 | 2026 | 2025 | ||||||||||||||||||||
Amortization of intangibles - royalties and milestones | $ | $ | $ | $ | |||||||||||||||||||
Royalties expensed | |||||||||||||||||||||||
Total royalty expense | $ | $ | $ | $ | |||||||||||||||||||
Royalties and milestones capitalized | $ | $ | $ | $ | |||||||||||||||||||
Variable Interest Entity
The Company reviews each investment and collaboration agreement to determine if it has a variable interest in the entity. In assessing whether the Company has a variable interest in the entity as a whole, the Company considers and makes judgments regarding the purpose and design of the entity, the value of the licensed assets to the entity, the value of the entity’s total assets and the significant activities of the entity. If the Company has a variable interest in the entity as a whole, the Company assesses whether or not the Company is a primary beneficiary of that VIE, based on a number of factors, including: (i) which party has the power to direct the activities that most significantly affect the VIE’s economic performance, (ii) the parties’ contractual rights and responsibilities pursuant to the collaboration agreement, and (iii) which party has the obligation to absorb losses of or the right to receive benefits from the VIE that could be significant to the VIE. If the Company determines that it is the primary beneficiary of a VIE at the onset of the collaboration, the collaboration is treated as a business combination and the Company consolidates the financial statements of the VIE into the Company’s consolidated financial statements. On a quarterly basis, the Company evaluates whether it continues to be the primary beneficiary of the consolidated VIE. If the Company determines that it is no longer the primary beneficiary of a consolidated VIE, it deconsolidates the VIE in the period in which the determination is made.
Assets and liabilities recorded as a result of consolidating the financial results of the VIE into the Company’s Consolidated Balance Sheets do not represent additional assets that could be used to satisfy claims against the Company’s general assets or liabilities for which creditors have recourse to the Company’s general assets.
12
Discontinued Operations
Recently Adopted Accounting Pronouncements
In July 2025, the FASB issued ASU No. 2025-05—Financial Instruments—Credit Losses (Topic 326): Measurement of Credit Losses for Accounts Receivable and Contract Assets. This ASU added a practical expedient that assumes that current conditions as of the balance sheet date do not change for the remaining life of the asset when estimating expected credit losses for current accounts receivable and current contract assets. The guidance is effective for annual periods beginning after December 15, 2025, including interim periods within those years. The Company adopted this standard effective January 1, 2026. The adoption did not have a material impact on the Company’s consolidated financial statements.
In November 2024, the FASB issued ASU No. 2024-04, Debt with Conversion and Other Options (Subtopic 470-20): Induced Conversions of Convertible Debts Instruments. This ASU clarifies the requirements for determining whether to account for certain early settlements of convertible debt instruments as induced conversions or extinguishments. The guidance is effective for fiscal years beginning after December 15, 2025, including interim periods within those years. The Company adopted this standard effective January 1, 2026 on a prospective basis. See Note 10 for further discussion of accounting for convertible debt transactions during 2026.
Recently Issued Accounting Pronouncements
From time to time, new accounting pronouncements are issued by the Financial Accounting Standards Board ("FASB") or other standard setting bodies. Unless otherwise discussed, the Company believes that the impact of recently issued standards that are not yet effective will not have a material impact on its consolidated financial position or results of operations upon adoption.
In September 2025, the FASB issued Accounting Standards Update ("ASU") No. 2025-06—Intangibles—Goodwill and Other—Internal-Use Software (Subtopic 350-40): Targeted Improvements to the Accounting for Internal-Use Software. This ASU removed the language around project stages that was used to assess when costs could be capitalized for an internal-use software. The update also requires internal-use software to be disclosed under the ASC 360 Property, Plant, and Equipment guidance. The guidance is effective for annual periods beginning after December 15, 2027. The Company is currently assessing the impact of this standard on the Company’s accounting policies and the financial statements.
In November 2024, the FASB issued ASU No. 2024-03, Income Statement - Reporting Comprehensive Income - Expense Recognition Disclosures. This ASU will require entities to provide enhanced disclosures related to certain expense categories included in income statement captions. The ASU aims to increase transparency and provide investors with more detailed information about the nature of expenses reported on the face of the income statement. The new standard does not change the requirements for the presentation of expenses on the face of the income statement. Under this ASU, entities are required to disaggregate, in a tabular format, expense captions presented on the face of the income statement — excluding earnings or losses from equity method investments — if they include any of the following expense categories: purchases of inventory, employee compensation, depreciation, intangible asset amortization, and depreciation or depletion. For any remaining items within each relevant expense caption, entities must provide a qualitative description of the nature of those expenses. The new ASU is effective, as clarified by ASU No. 2025-01, for annual reporting periods beginning after December 15, 2026 and interim reporting periods within annual reporting periods beginning after December 15, 2027. Early adoption is permitted. The Company is currently evaluating the impact of the adoption of this standard on the related disclosures.
NOTE 3. REVENUE RECOGNITION
Product Sales, Net
Product sales consist of FILSPARI and tiopronin products (Thiola and Thiola EC). The Company sells its products to specialty pharmacies and through direct-to-patient distributors worldwide, with the United States representing over 99 % of the Company's net product sales.
The Company sells FILSPARI to two direct-to-patient specialty pharmacies in the United States. The Company sells its tiopronin products to patients and pharmacies, with distribution facilitated through a single direct-to-patient distributor. Revenues from product sales are recognized in satisfaction of a single performance obligation when the customer obtains control of the Company’s product. For FILSPARI, sales are recognized upon delivery of the product to the specialty pharmacies. The Company receives payments from its FILSPARI sales based on terms that are generally 30 days from shipment of the product to the specialty pharmacy. For the Company's tiopronin products, product sales are recognized upon delivery to the patient. The Company receives payments from sales of its tiopronin products, primarily through third party payers, based on terms that generally are within 30 days of delivery of product to the patient. Contracts do not contain significant financing components based on the typical period of time between performance of services and collection of consideration.
Deductions from Revenue
Revenues from product sales are recorded at the net sales price, which includes provisions resulting from discounts, rebates and co-pay assistance that are offered to customers, payers and other indirect customers relating to the Company’s sales of its products. These provisions are based on the estimates of the amounts earned or to be claimed on the related sales. These amounts are treated as variable consideration, estimated and recognized as a reduction of the transaction price at the time of the sale, using the most likely amount method, and are classified as a reduction of accounts receivable (if the amount is payable to a customer) or as a current liability (if the amount is payable to a party other than a customer). The Company includes these estimated amounts in the transaction price to the extent it is probable that a significant reversal of cumulative revenue recognized for such transactions will not occur. Where
13
appropriate, these reserves take into consideration the Company’s historical experience, current contractual and statutory requirements and specific known market events and trends. Overall, these reserves reflect the Company’s best estimates of the amount of consideration to which it is entitled based on the terms of the contract. If actual results in the future vary from the Company’s provisions, the Company will adjust the estimate, which would affect net product revenue and earnings in the period such variances become known. For the six months ended June 30, 2026 and 2025, adjustments to net product revenue related to performance obligations satisfied in previous periods were $1.0 million and $1.3 million, respectively.
Government Rebates: The Company calculates the rebates that it will be obligated to provide to government programs and deducts these estimated amounts from its gross product sales at the time the revenues are recognized. Allowances for government rebates and discounts are established based on an estimated allocation of payers and the government-mandated discounts applicable to government-funded programs. Rebate discounts are included in accrued expenses in the accompanying Consolidated Balance Sheets.
Commercial Rebates: The Company calculates the rebates it incurs according to any contracts with certain commercial payers and deducts these amounts from its gross product sales at the time the revenues are recognized. Allowances for commercial rebates are established based on actual payer information, which is reasonably estimated at the time of delivery for applicable products. Rebate discounts are included in accrued expenses in the accompanying Consolidated Balance Sheets.
Prompt Pay Discounts: The Company offers discounts to certain customers for prompt payments. The Company accrues for the calculated prompt pay discount based on the gross amount of each invoice for those customers at the time of sale.
Other Fees: The Company pays service fees to certain customers based on a contractually fixed percentage of the wholesale acquisition cost ("WAC") and fees for data. Other fees are recorded as an offset to revenue based on contractual terms at the time revenue from the sale is recognized.
Product Returns: Consistent with industry practice, the Company offers its customers a limited right to return product purchased directly from the Company, which is principally based upon the product’s expiration date. Historically, returns have been immaterial.
Co-pay Assistance: The Company offers a co-pay assistance program, which is intended to provide financial assistance to qualified commercially insured patients with prescription drug co-payments required by payers. The calculation of the accrual for co-pay assistance is based on an estimate of claims and the estimated cost per claim associated with product that has been recognized as revenue.
The following table summarizes net product sales for the three and six months ended June 30, 2026 and 2025 (in thousands):
| Three Months Ended June 30, | Six Months Ended June 30, | ||||||||||||||||||||||
| 2026 | 2025 | 2026 | 2025 | ||||||||||||||||||||
| FILSPARI | $ | $ | $ | $ | |||||||||||||||||||
| Tiopronin products | |||||||||||||||||||||||
| Total net product sales | $ | $ | $ | $ | |||||||||||||||||||
NOTE 4. COLLABORATION AND LICENSE AGREEMENTS
License Agreement with CSL Vifor
In September 2021, the Company entered into the CSL Vifor License Agreement, pursuant to which the Company granted an exclusive license to CSL Vifor for the commercialization of FILSPARI in Europe, Australia and New Zealand. In June 2025, the CSL Vifor License Agreement was amended in order to expand the license to cover additional countries the ("CSL Vifor Licensed Territories") and to provide that the license rights to each additional country will revert to the Company if CSL Vifor does not take certain specified actions within specified timelines with respect to such country. CSL Vifor also has first right of negotiation to expand the licensed territories into Canada and/or Mexico. Under the terms of the CSL Vifor License Agreement, the Company received an upfront payment of $55.0 million and will be eligible for up to $135.0 million in aggregate regulatory and market access related milestone payments and up to $655.0 million in aggregate sales-based milestone payments for a total potential value of up to $845.0 million. The Company is also entitled to receive tiered double-digit royalties of up to 40 percent of annual net sales of FILSPARI in the CSL Vifor Licensed Territories.
Under the License Agreement, CSL Vifor is responsible for all commercialization activities in the CSL Vifor Licensed Territories. The Company remains responsible for the worldwide clinical development of sparsentan through regulatory approval as defined. Development costs for any post regulatory approval development activities, subject to approval by both parties, will be borne by the Company and CSL Vifor as defined, respectively. The CSL Vifor License Agreement will remain in effect, unless terminated earlier, until the expiration of all royalty terms for FILSPARI in the CSL Vifor Licensed Territories. Each party has the right to terminate the CSL Vifor License Agreement for the other party’s uncured material breach, insolvency or if the time required for performance under the CSL Vifor License Agreement by the other party is extended due to a force majeure event that continues for six months or more.
The Company assessed the CSL Vifor License Agreement and determined that it meets both criteria to be considered a collaborative agreement within the Scope of ASC 808, Collaborative Arrangements which are active participation by both parties and exposures to significant risks and rewards dependent on the commercial success of the activities. Both parties participate on joint steering and other committees overseeing the collaboration activities. Also, both parties are exposed to significant risks and rewards based on the economic outcomes of regulatory approvals and commercialization of sparsentan.
The Company determined the initial transaction price under the CSL Vifor License Agreement totaled $55.0 million, consisting of the fixed non-refundable upfront payment. The variable regulatory and access related milestones were considered variable consideration and excluded from the transaction price given the substantial uncertainty at inception related to their achievement, based on the most likely amount method. Sales-based milestone payments and royalties on net sales were excluded from the transaction price and are recognized at the later of when the related sales occur or when the performance
14
obligation to which the sales-based milestone or royalty has been allocated has been satisfied. The Company re-evaluates the transaction price for each reporting period and as uncertainties are resolved.
The Company concluded that CSL Vifor represented a customer and applied relevant guidance from ASC 606 to evaluate the accounting under the CSL Vifor License Agreement. In accordance with this guidance, the Company concluded at inception that the promise to grant the license is distinct from the promise to provide clinical development services resulting in two performance obligations. As a result, the Company allocated $12.0 million of the transaction price, based on the performance obligations' relative standalone selling prices, to the license, which was recognized in full in 2021. The remaining $43.0 million of the transaction price was allocated to the clinical development activities and recorded as deferred revenue, which was recognized over the development period based upon the ratio of costs incurred to date to the total estimated costs through June 30, 2025.
For the three and six months ended June 30, 2025, the Company achieved a regulatory milestone in the amount of $17.5
For the three months ended June 30, 2026 and 2025, the Company recognized $3.2 million and $19.6 million, respectively, in license and collaboration revenue associated with the CSL Vifor License Agreement. For the six months ended June 30, 2026 and 2025, the Company recognized $5.9 million and $25.5 million, respectively, in license and collaboration revenue associated with the CSL Vifor License Agreement.
Licensing Agreement with Chugai
In January 2024, the Company entered into an exclusive licensing agreement ("Chugai License Agreement") with Renalys, pursuant to which, the Company granted an exclusive license to Renalys (which has since been merged with and into Chugai) for the development and commercialization of sparsentan in Japan and other specified Asian countries. Under the terms of the Chugai License Agreement, the Company received a non-refundable upfront payment and will be eligible to receive up to $120.0 million in aggregate regulatory, development and sales-based milestones. The Company is also entitled to receive tiered double-digit to mid-20 percent royalties of annual net sales of sparsentan in the licensed territories. In addition, the Company received an option to purchase shares of common stock of Renalys (“Option Agreement”), which it exercised in January 2024. The Company also had the option to purchase all equity securities of Renalys at any time prior to the top-line results of the Phase 3 trial in Japan (“Buyout Right”).
Under the Chugai License Agreement, Chugai is responsible for all development and commercialization activities in the licensed territories. The Company will retain all rights to sparsentan in the United States and rest of world outside of the licensed territories and the CSL Vifor Licensed Territories, provided that CSL Vifor has a right of negotiation to expand the licensed territories into Canada and/or Mexico. The Chugai License Agreement will remain in effect, unless terminated earlier, until the expiration of all royalty terms for sparsentan in the licensed territories. Each party has the right to terminate the Chugai License Agreement for the other party’s uncured material breach or insolvency, or if the time required for performance under the Chugai License Agreement by the other party is extended due to a force majeure event that continues for nine months or more. Chugai may terminate the Chugai License Agreement for any reason upon prior written notice to the Company. The Company may terminate the Chugai License Agreement, in whole or with respect to Japan or South Korea if Chugai abandons development of all licensed products in the applicable country (or both countries) prior to the first commercial sale.
The Company assessed the Chugai License Agreement and determined that it meets both criteria to be considered a collaborative agreement within the Scope of ASC 808, Collaborative Arrangements which are active participation by both parties and exposures to significant risks and rewards dependent on the commercial success of the activities. Both parties participate on a joint steering committee overseeing the development and commercial activities. Also, both parties are exposed to significant risks and rewards based on the economic outcomes of regulatory approvals and commercialization of sparsentan.
The Company determined the transaction price under the Chugai License Agreement totaled $8.3 million, consisting of the fixed non-refundable upfront payment, milestone payment and estimated fair value of the Option Agreement. The variable development-related milestones were excluded from the transaction price given the substantial uncertainty related to their achievement under the most likely amount method. Sales-based milestone payments and royalties on net sales were excluded from the initial transaction price and will be recognized at the later of when the related sales occur or when the performance obligation to which the sales-based milestone or royalty has been allocated has been satisfied.
The Company concluded that Chugai represents a customer and applied relevant guidance from ASC 606 to evaluate the accounting under the Chugai License Agreement. In accordance with this guidance, the Company concluded that the promise to grant the license is distinct, resulting in one performance obligation as the license has stand-alone functionally at contract inception. The Buyout Right precluded transferring control of the license to Renalys under ASC 606 and the Company’s option to repurchase the common stock at a price greater than the original license premium results in accounting for the Chugai License Agreement as a financing arrangement. The transaction price was recorded in other current liabilities as a result of the Buyout Right. During the year ended December 31, 2025, the Buyout Right was relinquished, and the Company recognized the Renalys deferred revenue balance of $9.3 million in license and collaboration revenue.
See Note 6 for further discussion of VIEs.
For the three and six months ended June 30, 2026, the Company recognized a $5.0
NOTE 5. MARKETABLE DEBT SECURITIES
The Company's marketable debt securities as of June 30, 2026 and December 31, 2025 were composed of available-for-sale commercial paper and corporate and government debt securities and municipal bonds. The primary objective of the Company’s investment portfolio is to preserve capital and liquidity while enhancing overall returns. The Company’s investment policy limits interest-bearing security investments to certain types of instruments issued by institutions with primarily investment grade credit ratings and places restrictions on maturities and concentration by asset class and issuer.
15
Marketable debt securities consisted of the following (in thousands):
| June 30, 2026 | December 31, 2025 | ||||||||||
| Marketable debt securities: | |||||||||||
| Commercial paper | $ | $ | |||||||||
| Corporate debt securities | |||||||||||
| Securities of government sponsored entities | |||||||||||
| Municipal bonds | |||||||||||
| Total available-for-sale marketable debt securities | $ | $ | |||||||||
The following is a summary of short-term marketable debt securities classified as available-for-sale as of June 30, 2026 (in thousands):
| Remaining Contractual Maturity (in years) | Amortized Cost | Unrealized Gains | Unrealized Losses | Aggregate Estimated Fair Value | |||||||||||||||||||||||||
| Marketable debt securities: | |||||||||||||||||||||||||||||
| Commercial paper | Less than 1 | $ | $ | $ | ( | $ | |||||||||||||||||||||||
| Corporate debt securities | Less than 1 | ( | |||||||||||||||||||||||||||
| Securities of government-sponsored entities | Less than 1 | ( | |||||||||||||||||||||||||||
| Total maturity less than 1 year | ( | ||||||||||||||||||||||||||||
| Corporate debt securities | 1 to 2 | ( | |||||||||||||||||||||||||||
| Securities of government-sponsored entities | 1 to 2 | ( | |||||||||||||||||||||||||||
| Municipal bonds | 1 to 2 | ( | |||||||||||||||||||||||||||
| Total maturity 1 to 2 years | ( | ||||||||||||||||||||||||||||
| Total available-for-sale marketable debt securities | $ | $ | $ | ( | $ | ||||||||||||||||||||||||
The following is a summary of short-term marketable debt securities classified as available-for-sale as of December 31, 2025 (in thousands):
| Remaining Contractual Maturity (in years) | Amortized Cost | Unrealized Gains | Unrealized Losses | Aggregate Estimated Fair Value | |||||||||||||||||||||||||
| Marketable debt securities: | |||||||||||||||||||||||||||||
| Commercial paper | Less than 1 | $ | $ | $ | ( | $ | |||||||||||||||||||||||
| Corporate debt securities | Less than 1 | ( | |||||||||||||||||||||||||||
| Municipal bonds | Less than 1 | ||||||||||||||||||||||||||||
| Total maturity less than 1 year | ( | ||||||||||||||||||||||||||||
| Corporate debt securities | 1 to 2 | ( | |||||||||||||||||||||||||||
| Securities of government-sponsored entities | 1 to 2 | ( | |||||||||||||||||||||||||||
| Total maturity 1 to 2 years | ( | ||||||||||||||||||||||||||||
| Total available-for-sale marketable debt securities | $ | $ | $ | ( | $ | ||||||||||||||||||||||||
For the three and six months ended June 30, 2026 and 2025, realized gains and losses on marketable debt securities were immaterial. As of June 30, 2026 and December 31, 2025, the accrued interest receivable related to the Company's marketable debt securities was $2.3 million and $1.3 million, respectively, and was recorded in prepaid expenses and other current assets on the Consolidated Balance Sheets.
The Company reviews the available-for-sale marketable debt securities for declines in fair value below the cost basis each quarter. For any security whose fair value is below its amortized cost basis, the Company first evaluates whether it intends to sell the impaired security, or will otherwise be more likely than not required to sell the security before recovery. If either are true, the amortized cost basis of the security is written down to its fair value at the reporting date. If neither circumstance holds true, the Company assesses whether any portion of the unrealized loss is a result of a credit loss. Any amount deemed to be attributable to credit loss is recognized in the income statement, with the amount of the loss limited to the difference between fair value and amortized cost and recorded as an allowance for credit losses. The portion of the unrealized loss related to factors other than credit losses is recognized in other comprehensive income (loss).
16
The following is a summary of available-for-sale marketable debt securities in an unrealized loss position with no credit losses aggregated by investment category and length of time those individual securities have been in a continuous unrealized loss position reported as of June 30, 2026 (in thousands):
| Less Than 12 Months | 12 Months or Greater | Total | ||||||||||||||||||||||||||||||||||||
| Description of Securities | Fair Value | Unrealized Losses | Fair Value | Unrealized Losses | Fair Value | Unrealized Losses | ||||||||||||||||||||||||||||||||
| Commercial paper | $ | $ | $ | $ | $ | $ | ||||||||||||||||||||||||||||||||
| Corporate debt securities | ||||||||||||||||||||||||||||||||||||||
| Securities of government-sponsored entities | ||||||||||||||||||||||||||||||||||||||
| Municipal bonds | ||||||||||||||||||||||||||||||||||||||
| Total | $ | $ | $ | $ | $ | $ | ||||||||||||||||||||||||||||||||
The following is a summary of available-for-sale marketable debt securities in an unrealized loss position with no credit losses aggregated by investment category and length of time those individual securities have been in a continuous unrealized loss position reported as of December 31, 2025 (in thousands):
| Less Than 12 Months | 12 Months or Greater | Total | ||||||||||||||||||||||||||||||||||||
| Description of Securities | Fair Value | Unrealized Losses | Fair Value | Unrealized Losses | Fair Value | Unrealized Losses | ||||||||||||||||||||||||||||||||
| Commercial paper | $ | $ | $ | $ | $ | $ | ||||||||||||||||||||||||||||||||
| Corporate debt securities | ||||||||||||||||||||||||||||||||||||||
| Securities of government-sponsored entities | ||||||||||||||||||||||||||||||||||||||
| Total | $ | $ | $ | $ | $ | $ | ||||||||||||||||||||||||||||||||
As of June 30, 2026 and December 31, 2025, the amortized cost of the available-for-sale marketable debt securities in an unrealized loss position was $363.2 million and $140.9 million, respectively.
As of June 30, 2026 and December 31, 2025, the Company does not intend to sell these investments and it is not more likely than not that the Company will be required to sell the investments before recovery of their amortized cost basis. The increase in unrealized losses for the six months ended June 30, 2026 was primarily due to fluctuations in short-term interest rates. The Company does not believe the unrealized losses incurred during the period are due to credit-related factors. The credit ratings of the securities held remain of the highest quality. Moreover, the Company continues to receive payments of interest and principal as they become due, and our expectation is that those payments will continue to be received timely. Factors unknown to us at this time may cause actual results to differ and require adjustments to the Company’s estimates and assumptions in the future.
NOTE 6. VARIABLE INTEREST ENTITIES
Licensing Agreement with Chugai
In January 2024, the Chugai License Agreement between the Company and Renalys (which has since been acquired by and merged into Chugai) came into effect and the Company exercised its option to purchase shares of common stock of Renalys. The Company determined that Renalys was a VIE as they could require additional funding to support development and commercial activities. The Company had variable interests in Renalys, including an equity interest, Buyout Right and performance-related payments under the Chugai License Agreement that absorb variability from the performance of Renalys.
In order to determine the primary beneficiary of Renalys, the Company evaluated its variable interest to identify if the Company had the power to direct the activities that most significantly impact the economic performance. Based upon the capital structure, governing documents and overall business operations, the Company determined that it was not the primary beneficiary as it did not have the power to direct the activities that most significantly impact the economic performance of Renalys and did not have an obligation to absorb losses.
NOTE 7. LEASES
As of June 30, 2026, the Company had an operating lease with Kilroy Realty, L.P. (the "Landlord") for office space located in San Diego, California, which was entered into in April 2019 and subsequently amended in May 2020. Coinciding with the Company's ability to direct the use of the office space, which occurred in phases over 2020, and utilizing a discount rate equal to the Company's estimated incremental borrowing rate, the Company established right-of-use assets totaling $34.6 million and lease liabilities totaling $34.5 million. The total right-of-use asset and lease liability at measurement were each offset by lease incentives associated with tenant improvement allowances totaling $7.9 million.
The initial term of the office lease ends in August 2028, and the Landlord has granted the Company an option to extend the term of the lease by a period of five years . At lease inception, it was not reasonably certain that the Company will extend the term of the lease and therefore the renewal period has been
17
excluded from the aforementioned right-of-use asset and lease liability measurements. The measurement of the lease term occurs from the February 2021 occupancy date of the office space.
In November 2024, the Company entered into a sublease for a portion of the premises. The term of the sublease runs from January 2025 through August 2028. The Company’s sublease arrangement has been classified as an operating lease with sublease income recognized on a straight-line basis over the term of the sublease arrangement. To measure the Company’s periodic sublease income, the Company elected to use a practical expedient under ASC 842 to aggregate non-lease components with the related lease components because (i) the timing and pattern of transfer for the non-lease components and the related lease components are the same and (ii) the lease components, if accounted for separately, would be classified as an operating lease.
The following is a schedule of the future minimum rental commitments for the Company's operating lease reconciled to the lease liability and right-of-use asset as of June 30, 2026 (in thousands):
| June 30, 2026 | |||||
| 2026 (remaining) | $ | ||||
| 2027 | |||||
| 2028 | |||||
| 2029 and thereafter | |||||
| Total undiscounted future minimum payments | |||||
| Present value discount | ( | ||||
| Total lease liability | |||||
| Unamortized lease incentives | ( | ||||
| Cash payments in excess of straight-line lease expense | ( | ||||
| Total right-of-use asset | $ | ||||
The weighted-average remaining lease term and weighted-average discount rate of the Company's operating leases are as follows:
| June 30, 2026 | December 31, 2025 | ||||||||||
| Weighted-average remaining lease term in years | |||||||||||
| Weighted-average discount rate | % | % | |||||||||
NOTE 8. FAIR VALUE MEASUREMENTS
The Company utilizes a fair value hierarchy that prioritizes the inputs to valuation techniques used to measure fair value. The hierarchy gives the highest priority to unadjusted quoted prices in active markets for identical assets or liabilities (Level 1 measurements) and the lowest priority to unobservable inputs (Level 3 measurements). The three levels of the fair value hierarchy are described below:
Level 1 – Unadjusted quoted prices in active markets that are accessible at the measurement date for identical, unrestricted assets or liabilities;
Level 2 – Quoted prices in markets that are not active or financial instruments for which all significant inputs are observable, either directly or indirectly; and
Level 3 – Prices or valuations that require inputs that are both significant to the fair value measurement and unobservable.
Financial instruments with carrying values approximating fair value include cash and cash equivalents, accounts receivable, and accounts payable, due to their short-term nature. As of June 30, 2026 and December 31, 2025, the fair value of the Company's 2.25 180.5 million and $448.3 million, respectively. As of June 30, 2026 the fair value of the Company's 0.50 % Convertible Senior Notes due 2032 was $618.9 million. The fair values were estimated utilizing market quotations and are considered Level 2.
18
The following table presents the Company’s assets, measured and recognized at fair value on a recurring basis, classified under the appropriate level of the fair value hierarchy as of June 30, 2026 (in thousands):
| As of June 30, 2026 | |||||||||||||||||||||||
| Total carrying and estimated fair value | Quoted prices in active markets (Level 1) | Significant other observable inputs (Level 2) | Significant unobservable inputs (Level 3) | ||||||||||||||||||||
| Assets: | |||||||||||||||||||||||
| Cash and cash equivalents | $ | $ | $ | $ | |||||||||||||||||||
| Marketable debt securities, available-for-sale | |||||||||||||||||||||||
| Total | $ | $ | $ | $ | |||||||||||||||||||
The following table presents the Company’s assets, measured and recognized at fair value on a recurring basis, classified under the appropriate level of the fair value hierarchy as of December 31, 2025 (in thousands):
| As of December 31, 2025 | |||||||||||||||||||||||
| Total carrying and estimated fair value | Quoted prices in active markets (Level 1) | Significant other observable inputs (Level 2) | Significant unobservable inputs (Level 3) | ||||||||||||||||||||
| Assets: | |||||||||||||||||||||||
| Cash and cash equivalents | $ | $ | $ | $ | |||||||||||||||||||
| Marketable debt securities, available-for-sale | |||||||||||||||||||||||
| Total | $ | $ | $ | $ | |||||||||||||||||||
NOTE 9. INTANGIBLE ASSETS
Ligand License Agreement
In 2012, the Company entered into the Ligand License Agreement for a worldwide sublicense to develop, manufacture and commercialize FILSPARI. As consideration for the license, the Company is required to make substantial payments upon the achievement of certain milestones, totaling up to $114.1 million. Through June 30, 2026 the Company has capitalized $48.7 million for contractual milestones achieved under the Ligand License Agreement. Pursuant to the Ligand License Agreement, the Company is obligated to pay to Ligand (and Bristol-Myers Squibb Company) an escalating royalty between 15 % and 17 % of net sales of FILSPARI and any other products containing FILSPARI or related compounds, with payments due quarterly. The Company began incurring costs associated with such royalties following the February 2023 approval of FILSPARI for IgAN. For the three and six months ended June 30, 2026, the Company capitalized royalties owed on net sales of FILSPARI to intangible assets of $25.4 million and $43.4 million, respectively and $11.4 million and $20.3 million for the three and six months ended June 30, 2025, respectively. The cost of historical milestones paid and royalty payments are being amortized to royalty expense on a straight-line basis through the estimated useful life.
Mission License Agreement
In 2014, the Company entered into the Mission License Agreement, pursuant to which it obtained an exclusive, royalty-bearing license to market, sell and commercialize Thiola (tiopronin) in the United States and Canada, and a non-exclusive license to use know-how relating to Thiola to the extent necessary to market Thiola. Under the terms of the Mission License Agreement, as subsequently amended, which runs through May 2029, the Company is obligated to pay Mission the greater of $2.1 million, representing the guaranteed minimum royalty, or 20 % of its Thiola net sales generated globally during each calendar year. The Company capitalized a total of $188.4 million related to the Thiola intangible asset. For the three and six months ended June 30, 2025, the Company capitalized royalties owed on net sales of Thiola to intangible assets of $3.7 million and $6.9 million, respectively. The cost of the milestone payments and royalty payments were fully amortized to royalty expense on a straight-line basis through March 31, 2026. During the second quarter of 2026, the Company began expensing royalty payments during the period incurred.
The following table sets forth amortizable intangible assets as of June 30, 2026 and December 31, 2025 (in thousands):
| June 30, 2026 | December 31, 2025 | ||||||||||
| Finite-lived intangible assets | $ | $ | |||||||||
| Less: accumulated amortization | ( | ( | |||||||||
| Net carrying value | $ | $ | |||||||||
19
NOTE 10. CONVERTIBLE DEBT
The composition of the Company’s convertible senior notes are as follows (in thousands):
| June 30, 2026 | December 31, 2025 | ||||||||||
| $ | $ | ||||||||||
Unamortized debt issuance costs - | ( | ||||||||||
Unamortized debt issuance costs - | ( | ( | |||||||||
| Total convertible senior notes, net of unamortized debt issuance costs | $ | $ | |||||||||
Convertible Senior Notes Due 2032
On May 11, 2026, the Company completed a registered underwritten public offering of $525.0 million aggregate principal amount of 0.50 % Convertible Senior Notes due 2032 (“2032 Notes”), which includes $50.0 million aggregate principal amount of 2032 Notes sold pursuant to the full exercise of the underwriters’ over-allotment option. The Company issued the 2032 Notes under an indenture, dated as of May 11, 2026, as supplemented by the first supplemental indenture, dated as of May 11, 2026 (collectively, the “2032 Indenture”). The Company received net proceeds from the issuance of the 2032 Notes of $509.3 million, after deducting commissions and offering expenses of $16.3 million. The 2032 Notes will mature on May 15, 2032, unless earlier repurchased, redeemed, or converted. The 2032 Notes are senior unsecured obligations of the Company and bear interest at an annual rate of 0.50 %, payable semi-annually in arrears on May 15 and November 15 of each year, beginning on November 15, 2026.
At June 30, 2026, accrued interest on the 2032 Notes of $0.4 million is included in accrued expenses in the accompanying Consolidated Balance Sheets. The 2032 Notes comprise the Company’s senior, unsecured obligations and are (i) equal in right of payment with the Company’s existing and future senior, unsecured indebtedness, including the 2029 Notes; (ii) senior in right of payment to the Company’s existing and future indebtedness that is expressly subordinated to the 2032 Notes; (iii) effectively subordinated to the Company’s existing and future secured indebtedness, to the extent of the value of the collateral securing that indebtedness; and (iv) structurally subordinated to all existing and future indebtedness and other liabilities, including trade payables, of the Company's subsidiaries.
Holders may convert their 2032 Notes at their option only in the following circumstances: (1) during any calendar quarter commencing after the calendar quarter ending on September 30, 2026 (and only during such calendar quarter), if the last reported sale price per share of the Company’s common stock for each of at least 20 trading days, whether or not consecutive, during the period of 30 consecutive trading days ending on, and including, the last trading day of the immediately preceding calendar quarter exceeds 130 % of the conversion price on the applicable trading day; (2) during the five consecutive business days immediately after any 10 consecutive trading day period (such 10 consecutive trading day period, the “measurement period”) if the trading price per $1,000 principal amount of 2032 Notes for each trading day of the measurement period was less than 98 % of the product of the last reported sale price per share of the Company’s common stock on such trading day and the conversion rate on such trading day; (3) upon the occurrence of certain corporate events or distributions of the Company’s common stock; (4) if the Company calls the 2032 Notes for redemption; and (5) at any time from, and including, February 17, 2032 until the close of business on the second scheduled trading day immediately before the maturity date. The Company will settle conversions by paying or delivering, as applicable, cash, shares of the Company’s common stock, or a combination of cash and shares of the Company’s common stock, at the Company’s election, based on the applicable conversion rate. The initial conversion rate for the 2032 Notes is 15.4078 shares of the Company’s common stock per $1,000 principal amount of 2032 Notes, which represents an initial conversion price of approximately $64.90 per share. If a “make-whole fundamental change” (as defined in the 2032 Indenture) occurs, then the Company will, in certain circumstances, increase the conversion rate for a specified period of time.
The 2032 Notes will be redeemable, in whole or in part at the Company’s option at any time, and from time to time, on or after May 21, 2029 and, in the case of any partial redemption, on or before the 30 th scheduled trading day before the maturity date, at a cash redemption price equal to the principal amount of the 2032 Notes to be redeemed, plus accrued and unpaid interest, if any, to, but excluding, the redemption date but only if the last reported sale price per share of the Company’s common stock exceeds 130 % of the conversion price on (1) each of at least 20 trading days, whether or not consecutive, during the 30 consecutive trading days ending on, and including, the trading day immediately before the date the Company sends the related redemption notice; and (2) the trading day immediately before the date the Company sends such notice. However, the Company may not redeem less than all of the outstanding 2032 Notes unless at least $75.0 million aggregate principal amount of 2032 Notes are outstanding and not called for redemption as of the time the Company sends the related redemption notice. In addition, calling any 2032 Note for redemption will constitute a make-whole fundamental change with respect to that 2032 Note, in which case the conversion rate applicable to the conversion of that 2032 Note will be increased in certain circumstances if it is converted after it is called for redemption. If a fundamental change (as defined in the 2032 Indenture) occurs, then, except as described in the 2032 Indenture, holders may require the Company to repurchase their 2032 Notes at a cash repurchase price equal to the principal amount of the 2032 Notes to be repurchased, plus accrued and unpaid interest, if any, to, but excluding, the fundamental change repurchase date.
The Company incurred approximately $16.3 million of debt issuance costs relating to the issuance of the 2032 Notes, which were recorded as a reduction to the 2032 Notes on the Consolidated Balance Sheets. The debt issuance costs are being amortized and recognized as additional interest expense over the expected life of the 2032 Notes using the effective interest method. The Company determined the expected life of the debt is equal to the six-year term of the 2032 Notes. The effective interest rate on the 2032 Notes is 1.0 %.
Convertible Senior Notes Due 2029
On March 11, 2022, the Company completed a registered underwritten public offering of $316.3 million aggregate principal amount of 2.25 % Convertible Senior Notes due 2029 (“2029 Notes”), which includes $41.3 million aggregate principal amount of 2029 Notes sold pursuant to the full exercise of the underwriters’ option to purchase additional 2029 Notes. The Company issued the 2029 Notes under an indenture, dated as of September 10, 2018, as
20
supplemented by the second supplemental indenture, dated as of March 11, 2022 (collectively, the “2029 Indenture”). The Company received net proceeds from the issuance of the 2029 Notes of $306.4 million, after deducting commissions and offering expenses of $9.9 million. The 2029 Notes will mature on March 1, 2029, unless earlier repurchased, redeemed, or converted. The 2029 Notes are senior unsecured obligations of the Company and bear interest at an annual rate of 2.25 %, payable semi-annually in arrears on March 1 and September 1 of each year, beginning on September 1, 2022.
At June 30, 2026, accrued interest on the 2029 Notes of $0.7 million is included in accrued expenses in the accompanying Consolidated Balance Sheets. The 2029 Notes comprise the Company’s senior, unsecured obligations and are (i) equal in right of payment with the Company’s existing and future senior, unsecured indebtedness, including the 2032 Notes; (ii) senior in right of payment to the Company’s existing and future indebtedness that is expressly subordinated to the 2029 Notes; (iii) effectively subordinated to the Company’s existing and future secured indebtedness, to the extent of the value of the collateral securing that indebtedness; and (iv) structurally subordinated to all existing and future indebtedness and other liabilities, including trade payables, of the Company's subsidiaries.
Holders may convert their 2029 Notes at their option only in the following circumstances: (1) during any calendar quarter commencing after the calendar quarter ending on June 30, 2022 (and only during such calendar quarter), if the last reported sale price per share of the Company’s common stock for each of at least 20 trading days, whether or not consecutive, during the period of 30 consecutive trading days ending on, and including, the last trading day of the immediately preceding calendar quarter exceeds 130 % of the conversion price on the applicable trading day; (2) during the five consecutive business days immediately after any 10 consecutive trading day period (such 10 consecutive trading day period, the “measurement period”) if the trading price per $1,000 principal amount of 2029 Notes for each trading day of the measurement period was less than 98 % of the product of the last reported sale price per share of the Company’s common stock on such trading day and the conversion rate on such trading day; (3) upon the occurrence of certain corporate events or distributions of the Company’s common stock; (4) if the Company calls the 2029 Notes for redemption; and (5) at any time from, and including, December 1, 2028 until the close of business on the scheduled trading day immediately before the maturity date. The Company will settle conversions by paying or delivering, as applicable, cash, shares of the Company’s common stock, or a combination of cash and shares of the Company’s common stock, at the Company’s election, based on the applicable conversion rate. The initial conversion rate for the 2029 Notes is 31.3740 shares of the Company’s common stock per $1,000 principal amount of 2029 Notes, which represents an initial conversion price of approximately $31.87 per share. If a “make-whole fundamental change” (as defined in the 2029 Indenture) occurs, then the Company will, in certain circumstances, increase the conversion rate for a specified period of time.
The 2029 Notes will be redeemable, in whole or in part at the Company’s option at any time, and from time to time, on or after March 2, 2026 and, in the case of any partial redemption, on or before the 40 th scheduled trading day before the maturity date, at a cash redemption price equal to the principal amount of the 2029 Notes to be redeemed, plus accrued and unpaid interest, if any, to, but excluding, the redemption date but only if the last reported sale price per share of the Company’s common stock exceeds 130 % of the conversion price on (1) each of at least 20 trading days, whether or not consecutive, during the 30 consecutive trading days ending on, and including, the trading day immediately before the date the Company sends the related redemption notice; and (2) the trading day immediately before the date the Company sends such notice. However, the Company may not redeem less than all of the outstanding 2029 Notes unless at least $100.0 million aggregate principal amount of 2029 Notes are outstanding and not called for redemption as of the time the Company sends the related redemption notice. In addition, calling any 2029 Note for redemption will constitute a make-whole fundamental change with respect to that 2029 Note, in which case the conversion rate applicable to the conversion of that 2029 Note will be increased in certain circumstances if it is converted after it is called for redemption. If a fundamental change (as defined in the 2029 Indenture) occurs, then, except as described in the 2029 Indenture, holders may require the Company to repurchase their 2029 Notes at a cash repurchase price equal to the principal amount of the 2029 Notes to be repurchased, plus accrued and unpaid interest, if any, to, but excluding, the fundamental change repurchase date.
The Company incurred approximately $9.9 million of debt issuance costs relating to the issuance of the 2029 Notes, which were recorded as a reduction to the 2029 Notes on the Consolidated Balance Sheets. The debt issuance costs are being amortized and recognized as additional interest expense over the expected life of the 2029 Notes using the effective interest method. The Company determined the expected life of the debt is equal to the seven-year term of the 2029 Notes. The effective interest rate on the 2029 Notes is 2.74 %.
On May 11, 2026, the Company completed its repurchase of $221.4 million aggregate principal amount of 2029 Notes for cash, including accrued and unpaid interest, for a total of $350.9 million. The Company evaluated the transaction in accordance with amended accounting guidance under ASU 2024-04 and accounted for the repurchase as an induced conversion. As a result of the repurchase, the Company recognized inducement expense of $40.0 91.3 million of the premium, which represents the difference between the if-converted value and carrying value of the repurchased 2029 Notes and $2.8 million unamortized debt issuance costs related to the repurchased 2029 Notes were accounted for as reductions to additional paid-in capital in the Consolidated Balance Sheet. After giving effect to the repurchase, the total remaining principal amount outstanding under the 2029 Notes as of June 30, 2026 was $94.9 million.
The 2029 Notes and 2032 Notes are accounted for in accordance with ASC 470-20, Debt with conversion and Other Options (“ASC 470-20”) and ASC 815-40, Contracts in Entity’s Own Equity (“ASC 815-40”). Under ASC 815-40, to qualify for equity classification (or nonbifurcation, if embedded) the instrument (or embedded feature) must be both (1) indexed to the issuer’s stock and (2) meet the requirements of equity classification guidance. Based upon the Company’s analysis, it was determined that the 2029 Notes and 2032 Notes do not contain embedded features requiring recognition as derivatives and bifurcation, and therefore are measured at amortized cost and recorded as liabilities on the Consolidated Balance Sheets.
The 2029 Notes and 2032 Notes do not contain any financial or operating covenants or any restrictions on the payment of dividends, the issuance of other indebtedness or the issuance or repurchase of securities by the Company. There were no events of default for the 2029 Notes or 2032 Notes at June 30, 2026.
The 2029 Notes and 2032 Notes are classified as long-term convertible debt on the Company's Consolidated Balance Sheets at June 30, 2026.
21
Convertible Senior Notes Due 2025
On September 10, 2018, the Company completed a registered underwritten public offering of $276.0 million aggregate principal amount of 2.50 % Convertible Senior Notes due 2025 ("2025 Notes"), and entered into a base indenture and supplemental indenture agreement (collectively, the "2025 Indenture") with respect to the 2025 Notes.
The net proceeds from the issuance of the 2025 Notes were approximately $267.2 million, after deducting commissions and the offering expenses of $8.8 million payable by the Company. On March 11, 2022, the Company completed its repurchase of $207.1 million aggregate principal amount of 2025 Notes for cash, including accrued and unpaid interest, for a total of $213.8 million. After giving effect to the repurchase, the total remaining principal amount outstanding under the 2025 Notes was $68.9 million. On September 15, 2025, the 2025 Notes matured and the Company repaid the remaining principal amount outstanding of $68.9 million plus accrued interest.
Interest Expense
The following table sets forth total interest expense recognized related to the 2025, 2029 and 2032 Notes (in thousands):
| Three Months Ended June 30, | Six Months Ended June 30, | ||||||||||||||||||||||
| 2026 | 2025 | 2026 | 2025 | ||||||||||||||||||||
| Contractual interest expense | $ | $ | $ | $ | |||||||||||||||||||
| Amortization of debt issuance costs | |||||||||||||||||||||||
| Total interest expense | $ | $ | $ | $ | |||||||||||||||||||
Total interest expense recognized for the three and six months ended June 30, 2026 was $2.1 million and $4.4 million, respectively, and $2.8 million and $5.7 million for the three and six months ended June 30, 2025, respectively.
NOTE 11. ACCRUED EXPENSES
Accrued expenses at June 30, 2026 and December 31, 2025 consisted of the following (in thousands):
| June 30, 2026 | December 31, 2025 | ||||||||||
| Sales discounts, rebates, and allowances | $ | $ | |||||||||
| Accrued royalties | |||||||||||
| Compensation related costs | |||||||||||
| Research and development | |||||||||||
| Selling, general and administrative | |||||||||||
| Miscellaneous accrued expenses | |||||||||||
| Total accrued expenses | $ | $ | |||||||||
NOTE 12. NET INCOME (LOSS) PER COMMON SHARE
Basic and diluted net income (loss) per common share is calculated by dividing net loss applicable to common stockholders by the weighted-average number of common shares outstanding during the period. In accordance with ASC 260, Earnings per Share, if a company had a discontinued operation, the company uses income from continuing operations, adjusted for preferred dividend and similar adjustments, as its control number to determine whether potential common shares are dilutive.
The Company’s potentially dilutive shares, which include outstanding stock options, restricted stock units, and shares issuable upon conversion of the 2025 Notes, repaid in September 2025, 2029 Notes and 2032 Notes, are considered to be common stock equivalents. For the periods that the Company has reported net losses, all potentially dilutive securities are anti-dilutive and, accordingly, basic net loss per share equals diluted net loss per share.
22
Basic and diluted net loss per share is calculated as follows (in thousands, except share and per share amounts):
| Three Months Ended June 30, | |||||||||||||||||||||||||||||||||||
| 2026 | 2025 | ||||||||||||||||||||||||||||||||||
| Shares | Net Loss | EPS | Shares | Net Loss | EPS | ||||||||||||||||||||||||||||||
| Continuing operations | $ | ( | $ | ( | $ | ( | $ | ( | |||||||||||||||||||||||||||
| Discontinued operations | |||||||||||||||||||||||||||||||||||
| Basic and diluted loss per share | $ | ( | $ | ( | $ | ( | $ | ( | |||||||||||||||||||||||||||
| Six Months Ended June 30, | |||||||||||||||||||||||||||||||||||
| 2026 | 2025 | ||||||||||||||||||||||||||||||||||
| Shares | Net Loss | EPS | Shares | Net Loss | EPS | ||||||||||||||||||||||||||||||
| Continuing operations | $ | ( | $ | ( | $ | ( | $ | ( | |||||||||||||||||||||||||||
| Discontinued operations | ( | ( | |||||||||||||||||||||||||||||||||
| Basic and diluted loss per share | $ | ( | $ | ( | $ | ( | $ | ( | |||||||||||||||||||||||||||
The following common stock equivalents have been excluded because they were anti-dilutive:
| Three Months Ended June 30, | Six Months Ended June 30, | ||||||||||||||||||||||
| 2026 | 2025 | 2026 | 2025 | ||||||||||||||||||||
| Convertible debt | |||||||||||||||||||||||
| Options | |||||||||||||||||||||||
| Restricted stock units | |||||||||||||||||||||||
| Total anti-dilutive shares | |||||||||||||||||||||||
NOTE 13. COMMITMENTS AND CONTINGENCIES
Commitments
Certain of the Company's contractual arrangements with CMOs require binding forecasts or commitments to purchase minimum amounts for the manufacture of drug product supply, which may be material to the Company's financial statements.
Contingencies
In November 2020, the Company completed the acquisition of Orphan Technologies Limited (“Orphan”), including Orphan’s rare metabolic disorder drug pegtibatinase. The Company acquired Orphan by purchasing all of the outstanding shares. Under the stock purchase agreement, the Company has also agreed to make contingent cash payments up to an aggregate of $427.0 million based on the achievement of certain development, regulatory and commercialization events as set forth in the Agreement, as well as additional tiered mid-single digit royalty payments based upon future net sales of any pegtibatinase products in the U.S. and Europe, subject to certain reductions as set forth in the Agreement, and a contingent payment in the event a pediatric rare disease voucher for any pegtibatinase product is granted. Through June 30, 2026, the Company has paid $65.2 million for contractual milestones achieved under the stock purchase agreement.
In accordance with ASC 450, Contingencies, contingent cash payments will be accrued for when it is probable that a liability has been incurred and the amount can be reasonably estimated. As of June 30, 2026, no contingent cash payments have been accrued.
Legal Proceedings
From time to time in the normal course of business, the Company is subject to various legal matters such as threatened or pending claims or litigation. Although the results of claims and litigation cannot be predicted with certainty, the Company does not believe it is a party to any claim or litigation in which the outcome, if determined adversely to it, would individually or in the aggregate be reasonably expected to have a material adverse effect on its results of operations or financial condition.
23
NOTE 14. SHARE-BASED COMPENSATION
Stock Options
The following table summarizes stock option activity during the six months ended June 30, 2026:
Weighted Average | |||||||||||||||||||||||
| Shares Underlying Options | Exercise Price | Remaining Contractual Term (years) | Aggregate Intrinsic Value (in thousands) | ||||||||||||||||||||
| Outstanding at December 31, 2025 | $ | $ | |||||||||||||||||||||
| Granted | $ | ||||||||||||||||||||||
| Exercised | ( | $ | $ | ||||||||||||||||||||
| Forfeited/canceled | ( | $ | |||||||||||||||||||||
| Outstanding at June 30, 2026 | $ | $ | |||||||||||||||||||||
| Vested and expected to vest at June 30, 2026 | $ | $ | |||||||||||||||||||||
| Exercisable at June 30, 2026 | $ | $ | |||||||||||||||||||||
At June 30, 2026, unamortized stock compensation for stock options was $40.2 million, with a remaining weighted-average recognition period of 2.7 years.
Restricted Stock Units
Service Based Restricted Stock Units
The following table summarizes the Company’s service based restricted stock unit activity during the six months ended June 30, 2026:
| Number of Restricted Stock Units | Weighted Average Grant Date Fair Value | ||||||||||
| Unvested at December 31, 2025 | $ | ||||||||||
| Granted | $ | ||||||||||
| Vested | ( | $ | |||||||||
| Forfeited/canceled | ( | $ | |||||||||
| Unvested at June 30, 2026 | $ | ||||||||||
At June 30, 2026, unamortized stock compensation for service based restricted stock units was $108.0 million, with a remaining weighted-average recognition period of 2.5 years.
Performance Based Restricted Stock Units
The following table summarizes the Company’s performance based restricted stock unit activity during the six months ended June 30, 2026:
| Number of Restricted Stock Units | Weighted Average Grant Date Fair Value | ||||||||||
| Unvested at December 31, 2025 | $ | ||||||||||
| Granted | $ | ||||||||||
| Vested | ( | ||||||||||
| Forfeited/canceled | |||||||||||
| Unvested at June 30, 2026 | $ | ||||||||||
At June 30, 2026, unamortized stock compensation for performance based restricted stock units was $1.6 million, with a remaining weighted-average recognition period of 1.3 years.
Share-Based Compensation
The following table sets forth share-based compensation for continuing operations for the three and six months ended June 30, 2026 and 2025 (in thousands):
| Three Months Ended June 30, | Six Months Ended June 30, | ||||||||||||||||||||||
| 2026 | 2025 | 2026 | 2025 | ||||||||||||||||||||
| Research and development | $ | $ | $ | $ | |||||||||||||||||||
| Selling, general and administrative | |||||||||||||||||||||||
| Total share-based compensation | $ | $ | $ | $ | |||||||||||||||||||
24
NOTE 15. INVENTORY
Inventory consisted of the following at June 30, 2026 and December 31, 2025 (in thousands):
| June 30, 2026 | December 31, 2025 | ||||||||||
| Raw materials | $ | $ | |||||||||
| Work in process | |||||||||||
| Finished goods | |||||||||||
| Total inventory | $ | $ | |||||||||
| Classified as: | |||||||||||
| Current inventory | $ | $ | |||||||||
| Long-term inventory | |||||||||||
| Total inventory | $ | $ | |||||||||
NOTE 16. ACCOUNTS RECEIVABLE
Accounts receivable, net of reserves for prompt pay discounts and expected credit losses, was $80.6 million and $80.1 million at June 30, 2026 and December 31, 2025, respectively. The accounts receivable balance as of December 31, 2025 includes the $25.0 million milestone payment due from Mirum, received in April 2026. See Note 18 for further discussion. The total reserves for both periods were immaterial.
NOTE 17. EQUITY OFFERINGS
At-the-Market Equity Offering
NOTE 18. DIVESTITURES
Discontinued Operations
Sale of Bile Acid Product Portfolio
25
NOTE 19. SEGMENT INFORMATION
The Company operates in one
Long-lived assets located outside the U.S. were immaterial as of both June 30, 2026 and December 31, 2025. The measure of segment assets is reported on the Consolidated Balance Sheets as total assets. The accounting policies of the segment are the same as those described in Note 2, Summary of Significant Accounting Policies.
See Note 3 for further discussion of net product sales and Note 4 for discussion of license and collaboration revenues.
The following table presents reportable segment loss, including significant expenses regularly provided to the CODM, attributable to the Company’s reportable segment for the three and six months ended June 30, 2026 and 2025 (in thousands):
| Three Months Ended June 30, | Six Months Ended June 30, | ||||||||||||||||||||||
| 2026 | 2025 | 2026 | 2025 | ||||||||||||||||||||
| Revenue | $ | $ | $ | $ | |||||||||||||||||||
| Less: | |||||||||||||||||||||||
| Cost of goods sold | |||||||||||||||||||||||
| Research and development: | |||||||||||||||||||||||
| External research and development | |||||||||||||||||||||||
| Internal personnel costs | |||||||||||||||||||||||
Other research and development | |||||||||||||||||||||||
| Total research and development | |||||||||||||||||||||||
| Selling, general and administrative | |||||||||||||||||||||||
| Royalty expense | |||||||||||||||||||||||
| Total other (expense) income, net | ( | ( | ( | ||||||||||||||||||||
| Income tax benefit (provision) | ( | ( | |||||||||||||||||||||
| Loss from discontinued operations, net of tax | ( | ||||||||||||||||||||||
| Net loss | $ | ( | $ | ( | $ | ( | $ | ( | |||||||||||||||
NOTE 20. SUBSEQUENT EVENTS
In June 2026, the Company entered into a license and collaboration agreement with Everest Medicines (Singapore) Pte. Ltd. ("Everest"), granting the Company an exclusive license to develop, manufacture and commercialize civorebrutinib, an investigational oral, covalent reversible Bruton's tyrosine kinase ("BTK") inhibitor, and products containing civorebrutinib (“Products”) for any and all prophylactic, diagnostic and therapeutic uses and treatment of diseases and disorders in all countries outside of China and certain countries in East and Southeast Asia (the "Territory"). The agreement became effective in July 2026 following the satisfaction of customary closing conditions.
Under the terms of the agreement, the Company paid Everest an upfront payment of $112.5 million in July 2026. Pursuant to the agreement, Everest is eligible to receive up to $1.03 billion in additional development, regulatory and commercial milestone payments, as well as tiered royalties on net sales of Products in the Territory. The Company and Everest have agreed to collaborate on global clinical development, including sharing development costs for global clinical trials. Except for certain global development activities, the Company is responsible for development and commercialization of civorebrutinib in the Territory, while Everest retains responsibility for its territory.
26
Item 2. Management’s Discussion and Analysis of Financial Condition and Results of Operations
The following discussion and analysis of our financial condition and results of operations should be read in conjunction with our unaudited consolidated financial statements and related notes included in this Quarterly Report on Form 10-Q and the audited financial statements and notes thereto as of and for the year ended December 31, 2025 and the related Management’s Discussion and Analysis of Financial Condition and Results of Operations, both of which are contained in our Annual Report on Form 10-K for the year ended December 31, 2025, filed with the Securities and Exchange Commission ("SEC") on February 19, 2026. Past operating results are not necessarily indicative of results that may occur in future periods. In addition, see the discussion under the heading “Forward-Looking Statements” immediately preceding the consolidated financial statements included under Part I of this Quarterly Report on Form 10-Q.
Overview
We are a biopharmaceutical company headquartered in San Diego, California, focused on identifying, developing and delivering life-changing therapies to people living with rare kidney and metabolic diseases. Our approach centers on advancing our innovative pipeline with multiple late-stage clinical programs targeting rare diseases with significant unmet medical needs. Upon approval of any of our late-stage programs, we intend to leverage the skills of our talented commercial organization which has successfully identified, supported and treated patients prescribed our approved products for over ten years.
Our Pipeline and Approved Products
We have a diversified pipeline designed to address areas of high unmet need in rare kidney and metabolic diseases. We invest revenues from our commercial portfolio into our pipeline with the goal of delivering new treatments for diseases with limited or no approved therapies.
The following table summarizes the status of our clinical programs, preclinical programs and approved products, each of which is described in further detail below.
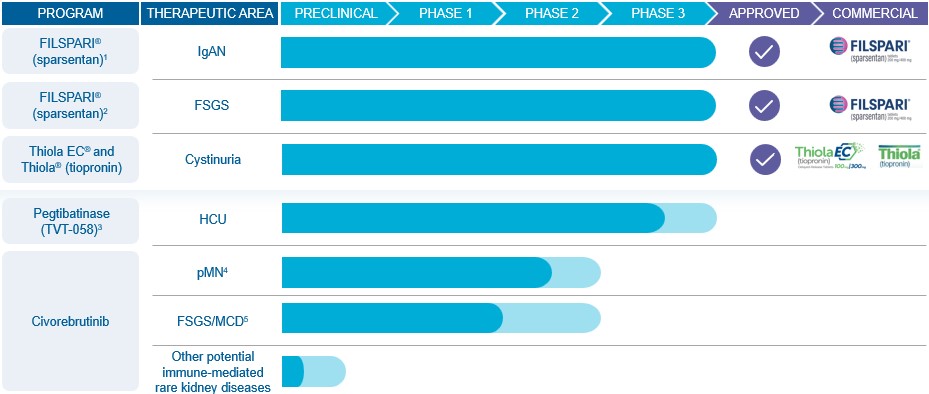
| 1 | On September 5, 2024, the FDA granted full approval of FILSPARI® (sparsentan) to slow kidney function decline in adults with primary Immunoglobulin A nephropathy ("IgAN”) who are at risk of disease progression. FILSPARI had previously been granted accelerated approval in February 2023. | ||||
| 2 | On April 13, 2026, the FDA granted full (traditional) approval of FILSPARI to reduce proteinuria in adult and pediatric patients aged 8 years and older with focal segmental glomerulosclerosis (“FSGS”) without nephrotic syndrome. | ||||
| 3 | Following a voluntarily pause in enrollment, we restarted enrollment activities for the pivotal Phase 3 HARMONY Study in the first quarter of 2026. | ||||
| 4 | Current Everest Medicines studies. | ||||
| 5 | As part of the ongoing basket study conducted by Everest Medicines. | ||||
FILSPARI® (sparsentan) for the treatment of IgAN
On September 5, 2024, the FDA granted full approval of FILSPARI® (sparsentan) to slow kidney function decline in adults with primary IgAN who are at risk of disease progression. FILSPARI had previously been granted accelerated approval in February 2023 based on the surrogate marker of proteinuria. Full approval was based on positive long-term confirmatory results from the PROTECT Study demonstrating that FILSPARI significantly slowed kidney function decline over two years compared to irbesartan.
FILSPARI is the only oral, once-daily, non-immunosuppressive medication that directly targets glomerular injury in the kidney by blocking two critical pathways of IgAN disease progression (endothelin-1 and angiotensin II).
27
The two-year efficacy data contained in the FDA-approved label is a modified intention to treat ("ITT") analysis and evaluates data from all patients regardless of treatment discontinuation. In the final analysis of the 404 randomized patients, FILSPARI significantly reduced the rate of decline in kidney function from baseline to Week 110 compared to irbesartan. In the ITT analysis included in the label, the mean eGFR slope from baseline to Week 110 was -3.0 mL/min/1.73 m2/year for FILSPARI and -4.2 mL/min/1.73 m2/year for irbesartan, corresponding to a statistically significant treatment effect of 1.2 mL/ min/1.73 m2/year (p=0.0168). The positive treatment effects on proteinuria compared to the active control irbesartan that were observed at Week 36 were durable out to the two-year measurement period. Additional results from the PROTECT Study demonstrated the benefit of FILSPARI on absolute eGFR accrued over time and by Week 110 resulted in a 3.8 mL/min/1.73 m2 difference in the mean change from baseline between FILSPARI and irbesartan.
Results from the PROTECT Study showed that FILSPARI was well tolerated with a clearly defined safety profile that has been consistent across all clinical trials conducted to date.
FILSPARI is a dual endothelin angiotensin receptor antagonist. Pre-clinical data have shown that blockade of both endothelin type A and angiotensin II type 1 pathways in forms of rare chronic kidney disease, reduces proteinuria, protects podocytes and prevents glomerulosclerosis and mesangial cell proliferation. FILSPARI has been granted seven years of Orphan Drug Exclusivity in the U.S. (running from the date of accelerated approval) for the reduction of proteinuria in adults with primary IgAN at risk of rapid disease progression, and has been granted a separate seven years of Orphan Drug Exclusivity in the U.S. (running from the date of full approval) to slow kidney function decline in adults with primary IgAN who are at risk for disease progression, excluding the use provided for in the aforementioned Orphan Drug Exclusivity granted in connection with the accelerated approval.
IgAN is characterized by hematuria, proteinuria, and variable rates of progressive renal failure. With an estimated prevalence of up to 150,000 people in the United States and greater numbers in Europe and Asia, IgAN is the most common primary glomerular disease. Most patients are diagnosed between the ages of 16 and 35, with up to 40% progressing to kidney failure within 15 years. FILSPARI is the first non-immunosuppressive therapy approved for IgAN and is the only oral, once-daily, non-immunosuppressive therapy approved for this condition that directly targets glomerular injury in the kidney by blocking two critical pathways of IgAN disease progression (endothelin-1 and angiotensin II). We estimate more than 70,000 patients in the United States to be addressable under FILSPARI's full approval indication.
Data to support the approval of FILSPARI was generated from the Phase 3 PROTECT Study, the largest head-to-head interventional study to date in IgAN. It is a global, randomized, multicenter, double-blind, parallel-arm, active-controlled clinical trial that evaluated the safety and efficacy of 400mg of sparsentan, compared to 300mg of irbesartan, in 404 patients ages 18 years and up with IgAN and persistent proteinuria despite available angiotensin converting enzyme ("ACE") inhibitor or angiotensin receptor blockers ("ARB") therapy, and is currently ongoing in the open label extension phase of the study.
FILSPARI is available only through a risk evaluation and mitigation strategy ("REMS") approved by the FDA for liver monitoring regarding potential risk of hepatotoxicity, as has been required for certain other approved endothelin antagonists. Initially, as part of the liver monitoring REMS, monthly monitoring of each patient was required for the first year a patient was on treatment, and quarterly thereafter. In August 2025, the FDA approved updated REMS labeling, reducing the frequency of liver monitoring to every three months from the onset of treatment and also removing the embryo-fetal toxicity monitoring requirement from the REMS.
In April 2024, we and our partner CSL Vifor announced that the European Commission had granted conditional marketing authorization (“CMA”) for FILSPARI (sparsentan) for the treatment of adults with primary IgAN with a urine protein excretion ≥1.0 g/day (or urine protein-to-creatinine ratio ≥0.75 g/g), and in April 2025, we and CSL Vifor announced that the European Commission had converted the CMA into a standard marketing authorization (“MA”) for FILSPARI for the treatment of adults with primary IgAN with a urine protein excretion ≥1.0 g/day (or urine protein-to-creatinine ratio ≥0.75 g/g). The MA was granted for all member states of the European Union, as well as in Iceland, Liechtenstein and Norway. As a result of the standard MA approval, we received a regulatory milestone payment of $17.5 million in May 2025 under the terms of the License Agreement. FILSPARI became commercially available in Europe under the CMA in August 2024, with an initial launch in Germany and Austria. In October 2024, we and CSL Vifor announced that Swissmedic has granted temporary marketing authorization for FILSPARI for the treatment of adults with primary IgAN with a urine protein excretion ≥1.0 g/day (or urine protein-to-creatinine ratio ≥0.75 g/g). In April 2025, the Medicines and Healthcare products Regulatory Agency ("MHRA") in the UK converted its conditional approval of FILSPARI in IgAN to standard approval. In the fourth quarter of 2025, we received a $40.0 million market access milestone payment from CSL Vifor.
In January 2024, we announced our entry into an exclusive licensing agreement with Renalys Pharma, Inc. ("Renalys"), to bring sparsentan for the treatment of IgAN to patients in Japan and other countries in Asia. In December 2024, Renalys announced that sparsentan received Orphan Drug Designation from the Japanese Ministry of Health, Labour and Welfare for the indication of primary IgA nephropathy as of November 27, 2024. In the fourth quarter of 2025, Renalys announced positive topline results from its Phase 3 study of sparsentan in Japanese patients with IgAN. Renalys has also announced that it has reached an agreement with the Pharmaceuticals and Medical Devices Agency ("PMDA") regarding development plans for two other Phase 3 clinical trials of sparsentan, one investigating the use of sparsentan in FSGS and the other in Alport syndrome, in Japan. In the fourth quarter of 2025, Renalys was acquired by and merged into Chugai Pharmaceutical Co., Ltd. (“Chugai”). Through the acquisition, Chugai gained exclusive rights to develop and commercialize sparsentan in Japan, South Korea, and Taiwan. As a minority shareholder in Renalys, we received $10.2 million at the closing of the transaction and we are also eligible to receive multiple milestones according to the progress of sparsentan regulatory approval, and consideration linked to sparsentan's net sales in the applicable territory. Under the terms of the licensing agreement, Chugai is responsible for development, regulatory matters, and commercialization in the licensed territories.
FILSPARI (sparsentan) for the treatment of FSGS
On April 13, 2026, the FDA granted full (traditional) approval of FILSPARI (sparsentan) to reduce proteinuria in adult and pediatric patients aged 8 years and older with FSGS without nephrotic syndrome.
FILSPARI is the first and only medicine approved by the FDA for the treatment of FSGS.
FSGS is a leading cause of kidney failure and nephrotic syndrome. Prior to the approval of FILSPARI in April 2026, there were no FDA-approved pharmacologic treatments for FSGS, and there remains a high unmet need for patients living with FSGS as off-label treatments such as ACE/ARBs, steroids, and immunosuppressant agents are effective in only a subset of patients and use of some of these off-label treatments may be further inhibited by their safety profiles.
28
People with FSGS who do not have nephrotic syndrome span across types of FSGS and represent a population aligned with the KDIGO guidelines for treating glomerular diseases. Nephrotic syndrome is commonly defined as the presence of three concurrent criteria: proteinuria greater than 3.5 g/24h, edema, and albumin less than 3.0 g/dL. We estimate that the addressable population in the U.S. is more than 30,000 individuals with FSGS who do not have nephrotic syndrome.
In the Phase 3 DUPLEX Study, the largest head-to-head interventional study in FSGS to date, patients treated with FILSPARI in the overall study population experienced a statistically significant 46% reduction in proteinuria from baseline to Week 108 compared to 30% for those treated with maximum labeled dose irbesartan (nominal p-value, 0.0299). In patients without nephrotic syndrome, FILSPARI demonstrated even greater improvements compared to maximum labeled dose irbesartan across proteinuria and eGFR. Those without nephrotic syndrome who were treated with FILSPARI experienced a 48% reduction in proteinuria from baseline to Week 108 compared to 27% for those treated with irbesartan, which was statistically significant (nominal p-value, 0.0075). FILSPARI-treated patients without nephrotic syndrome also demonstrated a benefit in eGFR with a treatment difference of 1.1 mL/min/1.73 m2 based on mean change from baseline to Week 108 (-11.3 mL/min/1.73 m2 for FILSPARI compared to -12.4 mL/min/1.73 m2 for maximum labeled dose irbesartan). Across both adult and pediatric patients, FILSPARI was generally well tolerated, with a safety profile comparable to irbesartan and consistent across clinical programs.
The Phase 3 DUPLEX Study was a global, randomized, multicenter, double-blind, parallel-arm, active-controlled Phase 3 clinical trial that assessed the efficacy and safety of FILSPARI in 371 patients ages 8 to 75 years with biopsy-proven or genetic FSGS. After a two-week washout period, patients were randomized 1:1 to receive either FILSPARI or irbesartan, the active control, and subsequently dose titrated to the maximum dose of 800 mg of sparsentan or 300 mg of irbesartan, as tolerated. In the study, nephrotic syndrome was defined as (a) documentation of nephrotic syndrome in the medical history, or (b) the concurrent presence of proteinuria >3.5 g/24 hours (adults) or UPCR >2.0 g/g (pediatric patients <18 years of age), serum albumin <3.0 g/dL, and edema at baseline. The primary efficacy endpoint at the final analysis was the rate of change in eGFR from baseline to Week 108. The two-year results from the study were published in the New England Journal of Medicine. Patients who completed the DUPLEX double-blind portion of the study on treatment were eligible to participate in the open-label extension of the trial.
Sparsentan has been granted Orphan Drug Designation for the treatment of FSGS in the U.S. and the EEA.
As noted above in more detail, FILSPARI is available only through a REMS approved by the FDA for liver monitoring regarding potential risk of hepatotoxicity, as has been required for certain other approved endothelin antagonists.
Together with CSL Vifor and Chugai, we continue to evaluate the potential for a regulatory pathway forward for sparsentan in FSGS in Europe and Japan.
Under the terms of our exclusive license to CSL Vifor, CSL Vifor is responsible for all commercialization activities in its licensed territories. We remain responsible for the clinical development of sparsentan in the applicable territories. If sparsentan receives marketing authorization in any of the territories covered by the exclusive license to Chugai, Chugai will be responsible for all development, regulatory matters, and commercialization activities in such licensed territories. We will retain all rights to sparsentan in the United States and rest of world outside of the territories licensed to CSL Vifor and Chugai, provided that CSL Vifor has a right of negotiation to expand the licensed territories into Canada and/or Mexico.
Clinical-Stage Programs:
Pegtibatinase
Pegtibatinase is a novel investigational human enzyme replacement candidate being evaluated for the treatment of classical homocystinuria ("HCU"). Classical HCU is a rare metabolic disorder characterized by elevated levels of plasma homocysteine that can lead to vision, skeletal, circulatory and central nervous system complications. We estimate that there are approximately 7,000 to 10,000 addressable HCU patients globally. Pegtibatinase has been granted Rare Pediatric Disease, Fast Track and Breakthrough Therapy designations by the FDA, as well as orphan drug designation in the United States and European Union.
In December 2021, we announced positive topline results from the Phase 1/2 COMPOSE Study, a double blind, randomized, placebo-controlled dose escalation study to assess its safety, tolerability, pharmacokinetics, pharmacodynamics and clinical effects in patients with classical HCU. Pegtibatinase demonstrated dose-dependent reductions in total homocysteine ("tHcy") during the 12 weeks of treatment, and in the highest dose cohort to date evaluating 1.5 mg/kg of pegtibatinase twice weekly ("BIW"), treatment with pegtibatinase resulted in rapid and sustained reductions in tHcy through 12 weeks of treatment, including a 55.1% mean relative reduction in tHcy from baseline as well as maintenance of tHcy below a clinically meaningful threshold of 100 μmol. Additionally, in a dose-dependent manner in the study to date, methionine levels were substantially reduced and cystathionine levels were substantially elevated following treatment with pegtibatinase, suggesting that pegtibatinase acts in a manner similar to the native CBS enzyme.
In May 2023, we announced positive topline results from the sixth cohort of the Phase 1/2 COMPOSE Study, which was initiated to inform and refine formulation work for future development and commercial purposes and to further evaluate the dose response curve for pegtibatinase, and to further inform our pivotal development program to ultimately support potential approval of pegtibatinase for the treatment of HCU. In this cohort, five patients were randomized in a blinded fashion to receive 2.5 mg/kg of lyophilized pegtibatinase or placebo BIW, with four patients assigned to the treatment group. In this highest dose cohort to date, treatment with pegtibatinase resulted in rapid and sustained reductions in tHcy, with a 67.1% mean relative reduction in tHcy from baseline, as well as maintenance of mean tHcy below the clinically meaningful threshold of 100 μmol, over weeks 6 to 12. In the double-blind period, pegtibatinase was generally well-tolerated, with no discontinuations due to treatment-related adverse events.
In December 2023, we initiated the pivotal Phase 3 HARMONY Study to support the potential approval of pegtibatinase for the treatment of classical HCU. The HARMONY Study is a global, randomized, multi-center, double-blind, placebo-controlled Phase 3 clinical trial designed to evaluate the efficacy and safety of pegtibatinase as a novel treatment to reduce tHcy levels. In the beginning of 2024, the first patients were dosed in the HARMONY Study.
In September 2024, we announced a voluntary pause of enrollment in the Phase 3 HARMONY Study. The voluntary enrollment pause was enacted following our determination that the desired drug substance profile was not achieved in the initial scale-up process, and it enabled us to address necessary process improvements in manufacturing scale-up to support initial commercial scale manufacturing as well as full enrollment in the HARMONY Study. Following
29
successful process improvements, the Company restarted enrollment activities in the first quarter of 2026 and dosed the first new patient following study restart in April 2026, with topline data anticipated in the second half of 2027.
We acquired pegtibatinase as part of the November 2020 acquisition of Orphan Technologies Limited.
Civorebrutinib
Civorebrutinib is an investigational oral, covalent reversible Bruton’s tyrosine kinase (“BTK”) inhibitor designed to provide differentiated efficacy, safety and convenience for patients with rare, immune-mediated kidney diseases, including primary membranous nephropathy (“PMN”), with planned evaluation in FSGS, minimal change disease (“MCD”) and potentially additional indications. BTK is a key mediator of B-cell receptor signaling and plays an important role in B-cell activation, maturation, proliferation, and differentiation into antibody-producing cells.
In immune-mediated kidney diseases, B-cell activation and autoantibody production are believed to contribute directly to kidney injury. Civorebrutinib has demonstrated proof of concept in a Phase 1/2 clinical trial of patients with PMN. The previously reported Phase 1/2 data demonstrated rapid and sustained reductions in anti-PLA2R autoantibodies and proteinuria, with high rates of immunologic and clinical remission and stable kidney function through 52 weeks of follow-up. Civorebrutinib has been generally well tolerated throughout the development program to date.
In June 2026, we entered into a license and collaboration agreement (the “Everest Agreement”) with Everest Medicines (Singapore) Pte. Ltd. (“Everest”), pursuant to which we obtained an exclusive license to develop, manufacture and commercialize civorebrutinib in the United States and global markets excluding Greater China and certain countries in East and Southeast Asia (the “Territory”). The Everest Agreement became effective in July 2026 following the satisfaction of customary closing conditions.
We and Everest will collaborate on global clinical development, including sharing development costs for global clinical trials. Except for certain global development activities, we are responsible for development and commercialization of civorebrutinib in the Territory, while Everest retains responsibility for its territory.
Under the Everest Agreement, we paid Everest an upfront payment of $112.5 million in July 2026 and Everest is eligible to receive up to approximately $1.03 billion in additional development, regulatory and commercial milestone payments, as well as tiered royalties on net sales of civorebrutinib products in the Territory.
Other Commercial Products:
Thiola and Thiola EC (tiopronin)
Thiola and Thiola EC are approved by the FDA for the treatment of cystinuria, a rare genetic cystine transport disorder that causes high cystine levels in the urine and the formation of recurring kidney stones. Due to the larger stone size, cystine stones may be more difficult to pass, often requiring surgical procedures to remove. More than 80 percent of people with cystinuria develop their first stone by the age of 20. More than 25 percent will develop cystine stones by the age of 10. Recurring stone formation can cause loss of kidney function in addition to substantial pain and loss of productivity associated with renal colic and stone passage. While a portion of people living with the disease are able to manage symptoms through diet and fluid intake, the prevalence of cystinuria in the U.S. is estimated to be 10,000 to 12,000, indicating that there may be as many as 4,000 to 5,000 affected individuals with cystinuria in the U.S. that would be candidates for Thiola or Thiola EC.
In June 2019 we announced that the FDA approved 100mg and 300mg tablets of Thiola EC, an enteric-coated formulation of Thiola, to be used for the treatment of cystinuria. Thiola EC offers the potential for administration with or without food, and the ability to reduce the number of tablets necessary to manage cystinuria. Thiola EC became available to patients in July 2019.
In May 2021, a generic option for the 100mg version of the original formulation of Thiola (tiopronin tablets) became available and in June 2022, a second option for the 100mg version of the original formulation of Thiola (tiopronin tablets) was approved. These generic versions of the original formulation of Thiola have impacted our sales, and these or additional generic versions of either formulation could have a material adverse impact on sales. To date, several generic options for the 100mg and 300mg versions of Thiola EC have been approved by the FDA and become available. Accordingly, Thiola EC is subject to generic competition.
30
Results of Operations
Results of operations for the three and six months ended June 30, 2026 compared to the three and six months ended June 30, 2025
Revenue
The following table provides information regarding revenue, including net product sales and license and collaboration revenue (in thousands):
| Three Months Ended June 30, | Six Months Ended June 30, | ||||||||||||||||||||||||||||||||||
| 2026 | 2025 | Change | 2026 | 2025 | Change | ||||||||||||||||||||||||||||||
| FILSPARI | $ | 141,078 | $ | 71,887 | $ | 69,191 | $ | 246,230 | $ | 127,768 | $ | 118,462 | |||||||||||||||||||||||
| Tiopronin products | 20,274 | 22,955 | (2,681) | 39,614 | 42,934 | (3,320) | |||||||||||||||||||||||||||||
| Total net product sales | 161,352 | 94,842 | 66,510 | 285,844 | 170,702 | 115,142 | |||||||||||||||||||||||||||||
| License and collaboration revenue | 8,232 | 19,607 | (11,375) | 10,938 | 25,478 | (14,540) | |||||||||||||||||||||||||||||
| Total revenue | $ | 169,584 | $ | 114,449 | $ | 55,135 | $ | 296,782 | $ | 196,180 | $ | 100,602 | |||||||||||||||||||||||
Net product sales
The increase in total net product sales for the three and six months ended June 30, 2026 compared to the three and six months ended June 30, 2025 was primarily due to growth in sales of FILSPARI.
License and collaboration revenue
The decrease in license and collaboration revenue for the three months ended June 30, 2026 compared to the three months ended June 30, 2025 was primarily due to recognition of a $17.5 million regulatory milestone associated with the CSL Vifor License Agreement during the three months ended June 30, 2025, offset by recognition of a $5.0 million regulatory milestone associated with the Chugai License Agreement during the three months ended June 30, 2026. The decrease in license and collaboration revenue for the six months ended June 30, 2026 compared to the six months ended June 30, 2025 was primarily due to recognition of a $17.5 million regulatory milestone associated with the CSL Vifor License Agreement and the sale of $3.8 million in active pharmaceutical ingredients ("API") during the three months ended June 30, 2025, offset by recognition of a $5.0 million regulatory milestone associated with the Chugai License Agreement during the six months ended June 30, 2026.
Operating Expenses
We revised the presentation of amortization expense of certain royalty and milestone payments associated with license agreements. Historically, these royalties were included within selling, general and administrative expenses in the consolidated statements of operations. Beginning in the current year, we present these amounts separately as royalty expense to better reflect the nature of these costs as consideration payable to licensors based on sales of licensed products and regulatory milestone payments.
The following table provides information regarding operating expenses (in thousands):
| Three Months Ended June 30, | Six Months Ended June 30, | ||||||||||||||||||||||||||||||||||
| 2026 | 2025 | Change | 2026 | 2025 | Change | ||||||||||||||||||||||||||||||
| Cost of goods sold - product sales | $ | 2,174 | $ | 1,521 | $ | 653 | $ | 4,123 | $ | 2,775 | $ | 1,348 | |||||||||||||||||||||||
| Cost of goods sold - license and collaboration | — | — | — | — | 3,426 | (3,426) | |||||||||||||||||||||||||||||
| Total cost of goods sold | 2,174 | 1,521 | 653 | 4,123 | 6,201 | (2,078) | |||||||||||||||||||||||||||||
| Research and development | 60,283 | 49,362 | 10,921 | 117,377 | 96,251 | 21,126 | |||||||||||||||||||||||||||||
| Selling, general and administrative | 96,112 | 62,581 | 33,531 | 176,366 | 122,995 | 53,371 | |||||||||||||||||||||||||||||
| Royalty expense | 7,061 | 13,635 | (6,574) | 31,877 | 26,060 | 5,817 | |||||||||||||||||||||||||||||
| Total operating expenses | $ | 165,630 | $ | 127,099 | $ | 38,531 | $ | 329,743 | $ | 251,507 | $ | 78,236 | |||||||||||||||||||||||
Cost of goods sold
Cost of goods sold includes the cost of inventory sold, third party manufacturing and supply chain costs, product shipping and handling costs, and provisions for excess and obsolete inventory. Cost of goods sold also includes the cost of goods sold under our license and collaboration agreements, which currently consists of the sale of APIs to our collaboration partners, at cost or at cost plus a margin.
Prior to the February 2023 FDA accelerated approval of FILSPARI (sparsentan) in IgAN, we expensed the production of APIs purchased to support the commercial launch of FILSPARI, in research and development expenses. For the three and six months ended June 30, 2025 sales of FILSPARI primarily consisted of zero-cost inventories, and therefore cost of goods sold did not increase proportionally to the increase in product sales. We began capitalizing inventory costs associated with FILSPARI following the February 2023 accelerated approval.
For the six months ended June 30, 2026 compared to the six months ended June 30, 2025, our cost of goods sold - license and collaboration decreased by $3.4 million, due to the sale of API in 2025.
31
Research and development expenses
Research and development costs include expenses related to sparsentan, pegtibatinase and our other pipeline programs. We expense all research and development costs as they are incurred. Our research and development costs are comprised of salaries and bonuses, benefits, non-cash share-based compensation, license fees, milestones under license agreements, costs paid to third-party contractors to perform research, conduct clinical trials, and develop drug materials and delivery methods, costs to manufacture drug product supplies to support clinical development, and associated overhead expenses and facilities costs. We charge direct internal and external program costs to the respective development programs. We also incur indirect costs that are not allocated to specific programs because such costs benefit multiple development programs and allow us to increase our pharmaceutical development capabilities. These consist of internal shared resources related to the development and maintenance of systems and processes applicable to all of our programs.
We currently have three Phase 3 clinical trials in process that are in various stages of activity, with ongoing non-clinical support trials. As such, clinical trial expenses will vary depending on all the factors set forth above and may fluctuate significantly from quarter to quarter and year to year.
We routinely engage vendors and service providers for scientific research, clinical trial, regulatory compliance, manufacturing and other consulting services. We also make grants to research and non-profit organizations to conduct research which may lead to new intellectual properties that we may subsequently license under separately negotiated license agreements. Such grants may be funded in lump sums or installments.
The following table provides information regarding research and development expenses (in thousands):
| Three Months Ended June 30, | Six Months Ended June 30, | ||||||||||||||||||||||||||||||||||
| 2026 | 2025 | Change | 2026 | 2025 | Change | ||||||||||||||||||||||||||||||
| External service provider costs: | |||||||||||||||||||||||||||||||||||
| Sparsentan | $ | 6,971 | $ | 13,737 | $ | (6,766) | $ | 16,470 | $ | 25,056 | $ | (8,586) | |||||||||||||||||||||||
| Pegtibatinase | 24,244 | 13,283 | 10,961 | 42,473 | 24,319 | 18,154 | |||||||||||||||||||||||||||||
| General and other product candidates | 5,106 | 4,127 | 979 | 9,947 | 8,559 | 1,388 | |||||||||||||||||||||||||||||
| Total external service provider costs | 36,321 | 31,147 | 5,174 | 68,890 | 57,934 | 10,956 | |||||||||||||||||||||||||||||
| Internal personnel costs | 23,962 | 18,215 | 5,747 | 48,487 | 38,317 | 10,170 | |||||||||||||||||||||||||||||
| Total research and development | $ | 60,283 | $ | 49,362 | $ | 10,921 | $ | 117,377 | $ | 96,251 | $ | 21,126 | |||||||||||||||||||||||
For the three and six months ended June 30, 2026 compared to 2025, our research and development expenses increased by $10.9 million and $21.1 million, respectively. External service provider costs increased by $5.2 million and $11.0 million, respectively, for the three and six months ended June 30, 2026 compared to 2025, largely driven by an increase in costs associated with pegtibatinase due to restarting enrollment activities for the HARMONY Study in 2026, as well as additional manufacturing costs, offset by a decrease in costs associated with the development of sparsentan as our Phase 3 programs advanced towards completion. The increase in internal personnel costs of $5.7 million and $10.2 million, respectively, for the three and six months ended June 30, 2026 compared to 2025, is due to an increase in headcount.
Selling, general and administrative expenses
Selling, general and administrative expenses consist of salaries and bonuses, benefits, non-cash share-based compensation, legal and other professional fees, rent, depreciation and amortization, travel, insurance, business development, sales and marketing programs, and other operating expenses.
For the three and six months ended June 30, 2026 compared to 2025, our selling, general and administrative expenses increased by $33.5 million and $53.4 million, respectively, primarily as a result of an increase in commercial investments to support FILSPARI in IgAN and commercial investments for the launch of FSGS, including an expansion of our field force to support both indications.
Royalty expense
Royalty expense includes amortization of royalties and milestones paid that are capitalized to the intangible asset under the asset accumulation model as well as royalties expensed after the estimated useful life of the license agreement has been reached.
For the three months ended June 30, 2026 compared to 2025, our royalty expense decreased by $6.6 million due to the Thiola intangible asset reaching the end of its useful life on March 31, 2026. For the six months ended June 30, 2026 compared to 2025, royalty expense increased by $5.8 million, primarily due to elevated amortization of the Thiola intangible asset in the first quarter of 2026 as it reached the end of its useful life on March 31, 2026.
Other Income/Expenses
The following table provides information regarding other (expense) income, net (in thousands):
| Three Months Ended June 30, | Six Months Ended June 30, | ||||||||||||||||||||||||||||||||||
| 2026 | 2025 | Change | 2026 | 2025 | Change | ||||||||||||||||||||||||||||||
| Interest income | $ | 3,356 | $ | 3,287 | $ | 69 | $ | 5,939 | $ | 7,083 | $ | (1,144) | |||||||||||||||||||||||
| Interest expense | (2,134) | (2,846) | 712 | (4,434) | (5,702) | 1,268 | |||||||||||||||||||||||||||||
Other (expense) income, net | (169) | (526) | 357 | (259) | 24 | (283) | |||||||||||||||||||||||||||||
Inducement expense | (40,008) | — | (40,008) | (40,008) | — | (40,008) | |||||||||||||||||||||||||||||
Total other (expense) income, net | $ | (38,955) | $ | (85) | $ | (38,870) | $ | (38,762) | $ | 1,405 | $ | (40,167) | |||||||||||||||||||||||
32
The change in our total other (expense) income, net for the three and six months ended June 30, 2026 as compared to the three and six months ended June 30, 2025 is primarily due to the $40.0 million inducement expense related to the partial repurchase of the 2.25% Convertible Senior Notes due 2029 (“2029 Notes”).
Liquidity and Capital Resources
We have financed our operations through a combination of borrowings, sales of our equity securities, and revenues generated from our commercialized products, along with proceeds from license and collaboration agreements and the divestiture of our bile acid business. We experienced significant growth in recent years in the number of our employees and the scope of our operations. We also expanded our sales and marketing, compliance and legal functions in addition to expansion of all functions to support a commercial organization, including by adding additional members to our sales force in connection with the commercial launch of FILSPARI in the United States for IgAN and for the commercial launch of FILSPARI in the United States for FSGS.
We believe that our available cash and short-term investments as of the date of this filing, together with anticipated cash generated from operations, will be sufficient to fund our anticipated level of operations beyond the next 12 months from the date of this filing. We expect that our operating results will vary from quarter-to-quarter and year-to-year depending upon various factors including revenues, selling, general and administrative expenses, and research and development expenses, particularly with respect to our clinical and preclinical development activities. Our ability to fund our operations in subsequent years will depend upon certain factors which are beyond our control and may require us to obtain additional debt or equity capital or refinance all or a portion of our debt, including the 2029 Notes and 0.50% Convertible Senior Notes due 2032 (“2032 Notes”), on or before maturity. Though we generate revenues from product sales, we may incur significant operating losses over the next several years. Our ability to achieve profitable operations in the future will depend in large part upon completing development of products in our pipeline, obtaining regulatory approvals for these products and bringing these products to market, along with potential in-licensing of additional products approved by the FDA and manufacturing and selling these products.
We had the following balances and financial performance at June 30, 2026 and December 31, 2025 (in thousands):
| June 30, 2026 | December 31, 2025 | ||||||||||
| Cash and cash equivalents | $ | 117,735 | $ | 93,035 | |||||||
| Marketable debt securities, at fair value | $ | 371,442 | $ | 229,761 | |||||||
| Convertible debt | $ | 602,781 | $ | 311,724 | |||||||
| Accumulated deficit | $ | (1,544,612) | $ | (1,472,713) | |||||||
Stockholders' equity | $ | 24,522 | $ | 114,828 | |||||||
| Net working capital* | $ | 463,116 | $ | 277,661 | |||||||
| Net working capital ratio** | 4.05 | 2.74 | |||||||||
| * Current assets less current liabilities. **Current assets divided by current liabilities. | |||||||||||
As of June 30, 2026, we had cash and cash equivalents of $117.7 million and available-for-sale marketable debt securities of $371.4 million. In July 2026, we paid Everest an upfront payment of $112.5 million pursuant to the terms of the Everest Agreement. Substantial sources of funds over the past year, as summarized further below, include net proceeds of $158.4 million from the issuance of 2032 Notes after giving effect to the partial repurchase of 2029 Notes, milestone payments from CSL Vifor totaling $40.0 million, a milestone payment of $25.0 million from Mirum and $10.2 million from the sale of Renalys stock to Chugai.
Over the next 12 months, our expected financial obligations include, but are not limited to, funding our operations, operating lease payments, interest payments on our outstanding debt, anticipated milestone payments, royalties on sales of our existing commercialized products, research and development expenses pertaining to clinical and preclinical development activities across our pipeline, expenses associated with the ongoing commercialization of FILSPARI in IgAN and expenses associated with the commercial launch of FILSPARI in FSGS. Sources of cash over this period include net revenues from sales of our products, the sale or maturity of investments in our portfolio of marketable debt securities, FILSPARI royalties and certain earned and potential milestone payments.
Beyond the next 12 months and over the foreseeable future, our known commitments and potential financial obligations will likely include ongoing operations funding, operating lease payments, interest payments on our outstanding debt, royalties on sales of our existing commercialized products, research and development expenses pertaining to clinical and preclinical development activities across our pipeline, milestone and royalty payments associated with FILSPARI, pegtibatinase, and other developmental programs based upon the achievement of certain agreement-specific criteria, along with sales-based royalties and the repayment of principal on the outstanding 2029 Notes, which mature on March 1, 2029, and 2032 Notes, which mature on May 15, 2032. Potential sources of cash over this time horizon may include net revenues from sales of our existing products and, if commercialized, our pipeline products, licensing revenue, the sale or maturity of marketable debt securities in our investment portfolio, the refinancing of all or a portion of our debt, on or before maturity, or the issuance of additional debt or equity. In addition, depending on prevailing market conditions, our liquidity requirements, contractual restrictions, and other factors, we may also from time to time seek to retire or purchase our outstanding debt through cash purchases and/or exchanges for equity securities, in open market purchases, privately negotiated transactions or otherwise, and the amounts involved in such purchases and/or exchanges, individually or in the aggregate, may be material. We may not be able to successfully conduct financing or refinancing activity on favorable terms or at all.
33
Purchase Agreement Proceeds
Sale of Bile Acid Product Portfolio
In July 2023, we entered into the Purchase Agreement with Mirum, pursuant to which Mirum agreed to purchase substantially all of the assets primarily related to our business of development, manufacture and commercialization of the Products, which comprised our bile acid business. Upon the Closing of the transaction in August 2023, we received an upfront cash payment of $210.0 million. Pursuant to the Purchase Agreement, we are eligible to receive up to $235.0 million upon the achievement of certain milestones based on specified amounts of annual net sales (tiered from $125.0 million to $500.0 million) of the Products. Mirum achieved the first such milestone based on its annual net sales in 2025, and we received a milestone payment of $25.0 million in April 2026, as a result of such achievement.
Collaboration and License Proceeds
License and Collaboration Agreement with CSL Vifor
In September 2021, we entered into a license agreement with CSL Vifor, pursuant to which we granted an exclusive license to CSL Vifor for the commercialization of FILSPARI in the licensed territories. Under the terms of the license agreement, we will be eligible for up to $135.0 million in aggregate regulatory and market access related milestone payments and up to $655.0 million in aggregate sales-based milestone payments for a total potential value of up to $845.0 million. Through June 30, 2026, we have received milestone payments totaling $57.5 million associated with the license agreement. We are also entitled to receive tiered double-digit royalties of up to 40 percent of annual net sales of sparsentan in the licensed territories.
See Note 4 to our unaudited Consolidated Financial Statements for further discussion.
Licensing Agreement with Chugai
In January 2024, our license agreement with Renalys Pharma, Inc. came into effect. Under the terms of the agreement, we granted an exclusive license to Renalys for the development and commercialization of sparsentan in Japan and other specified countries in Asia. Pursuant to the terms of the agreement, we are eligible to receive up to $120.0 million in aggregate regulatory, development and sales-based milestone payments. We are also entitled to receive tiered double-digit to mid-20 percent royalties of annual net sales of sparsentan in the licensed territories. In addition, we received an option to purchase shares of common stock of Renalys, which we exercised in January 2024. In the fourth quarter of 2025, Renalys was acquired by and merged into Chugai. Through the acquisition, Chugai gained exclusive rights to develop and commercialize sparsentan in Japan, South Korea, and Taiwan. As a minority shareholder in Renalys, we received $10.2 million at the closing of the transaction and we are also eligible to receive multiple milestones according to the progress of sparsentan regulatory approval, and consideration linked to sparsentan's net sales in the applicable territory. Under the terms of the licensing agreement, Chugai is responsible for development, regulatory matters, and commercialization in the licensed territories.
See Note 4 to our unaudited Consolidated Financial Statements for further discussion.
Equity Offerings
At-the-Market Equity Offering
In October 2024, we filed a prospectus supplement to the prospectus included in our registration statement on Form S-3 (File No. 333-281194), pursuant to which we may offer and sell, from time to time through Jefferies LLC, as agent (“Jefferies”), up to $100.0 million of our common stock pursuant to an Amended and Restated Open Market Sale Agreement ("ATM Agreement") with Jefferies. We have not sold any shares under the ATM Agreement.
Operating Leases
Future Minimum Rental Commitments
As of June 30, 2026, we have future minimum rental commitments totaling $15.2 million arising from our operating lease and sublease income totaling $2.8 million. These commitments represent the aggregate base rent through August 2028.
See Note 7 to our unaudited Consolidated Financial Statements for further discussion.
Purchase Commitments
Manufactured Product
Certain of our contractual arrangements with contract manufacturing organizations ("CMOs") require binding forecasts or commitments to purchase minimum amounts for the manufacture of drug product supply, which may be material to our financial statements.
Royalties and Contingent Cash Payments
Ligand License Agreement
In 2012, we entered into an agreement with Ligand Pharmaceuticals, Inc. ("Ligand") for a worldwide sublicense to develop, manufacture and commercialize FILSPARI (the “Ligand License Agreement”). As consideration for the license, we are required to make substantial payments upon the achievement of certain milestones, totaling up to $114.1 million. Through June 30, 2026, we have paid $48.7 million for contractual milestones achieved under the Ligand License Agreement. Pursuant to the terms of the Ligand License Agreement, we are obligated to pay to Ligand an escalating royalty between 15% and 17% of net sales of FILSPARI and any other products containing FILSPARI or related compounds, with payments due quarterly. We began incurring costs associated with such royalties following the February 2023 approval of FILSPARI.
34
The Ligand License Agreement will continue until neither party has any further payment obligations under the agreement and is expected to continue for up to 20 years from the effective date. Ligand may terminate the Ligand License Agreement due to (i) our insolvency, (ii) our material uncured breach of the agreement, (iii) our failure to use commercially reasonable efforts to develop and commercialize FILSPARI as described above or (iv) certain other conditions. We may terminate the Ligand License Agreement due to a material uncured breach of the agreement by Ligand.
See Note 9 to our unaudited Consolidated Financial Statements for further discussion.
Mission License Agreement
In 2014, we entered into a license agreement with Mission Pharmacal ("Mission"), pursuant to which we obtained an exclusive, royalty-bearing license to market, sell and commercialize Thiola (tiopronin) in the United States and Canada, and a non-exclusive license to use know-how relating to Thiola to the extent necessary to market Thiola ("Mission License Agreement"). Under the terms of the Mission License Agreement, as subsequently amended, which runs through May 2029, we are obligated to pay to Mission the greater of $2.1 million, representing the guaranteed minimum royalty, or 20% of our Thiola net sales generated globally during each calendar year.
See Note 9 to our unaudited Consolidated Financial Statements for further discussion.
Acquisition of Orphan Technologies Limited
In November 2020, we completed the acquisition of Orphan Technologies Limited (“Orphan”), including Orphan’s rare metabolic disorder drug pegtibatinase. We acquired Orphan by purchasing all of its outstanding shares. Under the Stock Purchase Agreement (the "Agreement"), we agreed to make contingent cash payments up to an aggregate of $427.0 million based on the achievement of certain development, regulatory and commercialization events as set forth in the Agreement, as well as additional tiered mid-single digit royalty payments based upon future net sales of any pegtibatinase products in the U.S. and Europe, subject to certain reductions as set forth in the Agreement, and a contingent payment in the event a pediatric rare disease voucher for any pegtibatinase product is granted. Through June 30, 2026, we have paid $65.2 million for contractual milestones achieved under the stock purchase agreement.
See Note 13 to our unaudited Consolidated Financial Statements for further discussion.
French Rebate Accrual
In October 2021, our distributor in France for our previously marketed product Kolbam informed us that they had received a notice that the price previously paid for Kolbam during its period on the market in France had been recalculated by the agency responsible for pharmaceutical pricing in France. In October 2024, we received an invoice from the government authority in the amount of approximately $6.2 million for reimbursement of amounts previously paid for Kolbam, which we paid in November 2024. We have appealed the pricing decision and will pursue an appeal of the amount owed with the Competent Administrative Court.
Borrowings
Convertible Senior Notes Due 2032
On May 11, 2026, we completed a registered underwritten public offering of $525.0 million aggregate principal amount of our 2032 Notes. We issued the 2032 Notes under an indenture dated as of May 11, 2026, as supplemented by the first supplemental indenture, dated as of May 11, 2026 (collectively, the “2032 Indenture”). The 2032 Notes will mature on May 15, 2032, unless earlier repurchased, redeemed, or converted. The 2032 Notes are senior unsecured obligations of ours and bear interest at an annual rate of 0.50%, payable semi-annually in arrears on May 15 and November 15 of each year, beginning on November 15, 2026. The 2032 Notes do not contain any financial or operating covenants or any restrictions on the payment of dividends, the issuance of other indebtedness or the issuance or repurchase of securities by us.
Convertible Senior Notes Due 2029
On March 11, 2022, we completed a registered underwritten public offering of $316.3 million aggregate principal amount of our 2029 Notes. We issued the 2029 Notes under an indenture, dated as of September 10, 2018, as supplemented by the second supplemental indenture, dated as of March 11, 2022 (collectively, the “2029 Indenture”). The 2029 Notes will mature on March 1, 2029, unless earlier repurchased, redeemed, or converted. The 2029 Notes are senior unsecured obligations of ours and bear interest at an annual rate of 2.25%, payable semi-annually in arrears on March 1 and September 1 of each year, beginning on September 1, 2022. On May 11, 2026, coinciding with the issuance of the 2032 Notes, we completed our repurchase of $221.4 million aggregate principal amount of 2029 Notes for cash. After giving effect to the repurchase, the total remaining principal amount outstanding under the 2029 Notes as of June 30, 2026 was $94.9 million. The 2029 Notes do not contain any financial or operating covenants or any restrictions on the payment of dividends, the issuance of other indebtedness or the issuance or repurchase of securities by us.
See Note 10 to our unaudited Consolidated Financial Statements for further discussion.
Funding Requirements
We believe that our available cash and short-term investments as of the date of this filing will be sufficient to fund our anticipated level of operations beyond the next 12 months from the date of this filing. We expect to use cash flows from operations and, when necessary, outside financings, to meet our current and future financial obligations, including funding our operations, debt service and capital expenditures. Our ability to make these payments depends on our future performance, which will be affected by financial, business, economic, regulatory and other factors, many of which we cannot control. Factors that may affect financing requirements include, but are not limited to:
•the timing, progress, cost and results of our clinical trials, preclinical studies and other discovery and research and development activities;
35
•the timing and outcome of, and costs involved in, seeking and obtaining marketing approvals for our products, and in maintaining quality systems standards for our products;
•the timing of, and costs involved in, commercial activities, including product marketing, sales and distribution;
•our ability to successfully commercialize FILSPARI for the treatment of IgAN and FSGS, and to obtain regulatory approval for, and successfully commercialize our other or future product candidates;
•increases or decreases in revenue from our marketed products, including decreases in revenue resulting from generic entrants, changes in reimbursement or rebates, and/or health epidemics or pandemics;
•payment obligations related to the 2029 Notes and 2032 Notes;
•the number and development requirements of other product candidates that we pursue;
•our ability to manufacture sufficient quantities of our products to meet expected demand;
•the costs of preparing, filing, prosecuting, maintaining and enforcing any patent claims and other intellectual property rights, litigation costs and the results of litigation;
•our ability to enter into collaboration, licensing or distribution arrangements and the terms and timing of these arrangements;
•the potential need to expand our business, resulting in additional payroll and other overhead expenses;
•the potential in-licensing of other products or technologies;
•the emergence of competing technologies or other adverse market or technological developments;
•the potential impacts of actions taken by the current administration, including but not limited to tariffs and changes at the FDA and other government agencies; and
•the impacts of inflation and resulting cost increases.
Future capital requirements will also depend on the extent to which we acquire or invest in additional complementary businesses, products and technologies.
Cash Flows from Continuing Operations
Cash Flows from Operating Activities
Cash provided by operating activities for the six months ended June 30, 2026 was $17.7 million compared to cash used of $37.2 million for the six months ended June 30, 2025. The change was due to a $115.1 million increase in total net product sales, offset by an increase in operational expenses and the timing of payments from normal operations.
Cash Flows from Investing Activities
Cash used in investing activities for the six months ended June 30, 2026 was $184.7 million compared to cash provided by of $46.1 million for the six months ended June 30, 2025. The change resulted primarily from the investment of the net proceeds of the 2032 Notes and partial repurchase of the 2029 Notes, as well as timing differences in investment purchases, sales and maturities, and the fluctuation of our portfolio mix between cash equivalents and short-term investment holdings and an increase in intangible asset purchases.
Cash Flows from Financing Activities
Cash provided by financing activities from continuing operations for the six months ended June 30, 2026 was $191.7 million compared to cash provided by of $5.2 million for the six months ended June 30, 2025. The change was primarily due to net proceeds of $158.4 million from the issuance of the 2032 Notes and the partial repurchase of the 2029 Notes and an increase in proceeds from the exercise of stock options during the six months ended June 30, 2026.
Other Matters
Adoption of New Accounting Standards
See Note 2 to our unaudited Consolidated Financial Statements in this report for a discussion of adoption of new accounting standards.
Recently Issued Accounting Pronouncements
See Note 2 to our unaudited Consolidated Financial Statements in this report for a discussion of recently issued accounting pronouncements.
Critical Accounting Estimates
Our consolidated financial statements are prepared in accordance with accounting principles generally accepted in the United States of America. The preparation of our consolidated financial statements and related disclosures requires us to make estimates and assumptions that affect the reported amount of assets, liabilities, revenue, costs and expenses, and related disclosures. We evaluate our estimates and assumptions on an ongoing basis. Our actual results may differ from these estimates under different assumptions and conditions. See our Annual Report on Form 10-K for the fiscal year ended December 31, 2025 for information about critical accounting estimates as well as a description of our other significant accounting policies.
36
Item 3. Quantitative and Qualitative Disclosures About Market Risk
Our primary exposure to market risk is related to changes in interest rates. As of June 30, 2026, we had cash equivalents and marketable debt securities of approximately $489.2 million, consisting of money market funds, U.S. government agency debt, municipal bonds, corporate debt and commercial paper. This exposure to market risk is interest rate sensitivity, which is affected by changes in the general level of U.S. interest rates, particularly because our investments are in short-term debt securities. Our marketable debt securities are subject to interest rate risk and will fall in value if market interest rates continue to increase. A change in interest rates of 100 basis points as of June 30, 2026 would have had approximately a $3.1 million impact on the fair value of our investments at June 30, 2026.
The marketable debt securities held in our investment portfolio may subject us to credit risk, though our investment policy limits interest-bearing security investments to certain types of instruments issued by institutions with primarily investment grade credit ratings and places restrictions on maturities and concentration by asset class and issuer. Given these policy restrictions and our emphasis on preserving capital and liquidity while enhancing overall returns, we have not experienced material credit-related losses with our securities holdings.
We are also exposed to market risk related to changes in foreign currency exchange rates. From time to time, we enter into contracts with vendors that are located outside of the United States, which contracts are denominated in foreign currencies. We are subject to fluctuations in foreign currency rated in connection with these agreements. We do not currently hedge our foreign currency exchange rate risk.
Inflation generally affects us by increasing our salaries and fees paid to third-party contract service providers. Recent inflationary pressures have primarily impacted our operations through increased labor costs. While we continue to monitor the effects of macroeconomic factors, inflationary pressures have not affected our current outlook or business objectives.
Item 4. Controls and Procedures
Evaluation of Disclosure Controls and Procedures
We maintain disclosure controls and procedures that are designed to ensure that information required to be disclosed in our reports required by the Securities Exchange Act of 1934, as amended ("Exchange Act"), is recorded, processed, summarized and reported within the timelines specified in the SEC’s rules and forms, and that such information is accumulated and communicated to our management, including our Chief Executive Officer and Chief Financial Officer, as appropriate, to allow timely decisions regarding required disclosure. In designing and evaluating the disclosure controls and procedures, management recognized that any controls and procedures, no matter how well designed and operated, can only provide reasonable assurance of achieving the desired control objectives, and in reaching a reasonable level of assurance, management necessarily was required to apply its judgment in evaluating the cost-benefit relationship of possible controls and procedures.
As required by Rule 13a-15(b) of the Exchange Act, we carried out an evaluation, under the supervision and with the participation of our management, including our Chief Executive Officer and Chief Financial Officer, of the effectiveness of the design and operation of our disclosure controls and procedures (as defined in Exchange Act Rule 13a–15(e) and 15d-15(e)), as of the end of the quarter covered by this report. Based on the foregoing, our Chief Executive Officer and Chief Financial Officer concluded that our disclosure controls and procedures were effective at the reasonable assurance level.
Changes in Internal Control over Financial Reporting
An evaluation was also performed under the supervision and with the participation of our management, including our Chief Executive Officer and Chief Financial Officer, of any change to our internal control over financial reporting that occurred during the quarter covered by this report and that has materially affected, or is reasonably likely to materially affect, our internal control over financial reporting. Our evaluation did not identify any change in our internal control over financial reporting (as defined in Rules 13a-15(f) and 15d-15(f) under the Securities Exchange Act of 1934) that occurred during the quarter ended June 30, 2026 that have materially affected, or are reasonably likely to materially affect, our internal control over financial reporting.
PART II - OTHER INFORMATION
Item 1. Legal Proceedings
The information required by this Item is incorporated herein by reference to the Notes to the Unaudited Consolidated Financial Statements--Note 13 Commitments and Contingencies: Legal Proceedings in Part I, Item 1, of this Quarterly Report on Form 10-Q.
37
Item 1A. Risk Factors
Our business, as well as an investment in our common stock, is highly speculative in nature and involves a high degree of risk. Our securities should be purchased only by persons who can afford to lose their entire investment. Carefully consider the risks and uncertainties described below together with all of the other information included herein, including the financial statements and related notes, before deciding to invest in our common stock. If any of the following risks actually occur, they could adversely affect our business, prospects, financial condition and results of operations. In such event(s), the market price of our common stock could decline and result in a loss of part or all of your investment. Accordingly, prospective investors should carefully consider, along with other matters referred to herein, the following risk factors in evaluating our business before purchasing any shares of our common stock. We have marked with an asterisk (*) those risk factors that were not included as separate risk factors in, or reflect changes to the similarly titled risk factors included in, Item 1A of our Annual Report on Form 10-K for the fiscal year ended December 31, 2025 as filed with the Securities and Exchange Commission ("SEC") on February 19, 2026.
Risks Related to the Commercialization of Our Products
Our future prospects are highly dependent upon our ability to successfully develop and execute commercialization strategies for our products, including FILSPARI, and to attain market acceptance among physicians, patients and healthcare payers.*
Our ability to generate significant product revenues and to achieve commercial success in the near-term will depend almost entirely on our ability to successfully commercialize our products in the United States, including FILSPARI (sparsentan) to slow kidney function decline in adults with primary IgAN who are at risk of disease progression, which was granted full approval by the FDA in September 2024, and FILSPARI (sparsentan) to reduce proteinuria in adult and pediatric patients aged 8 years and older with focal segmental glomerulosclerosis (“FSGS”) without nephrotic syndrome, which was granted full (traditional) approval by the FDA in April 2026. FILSPARI had previously been granted accelerated approval for IgAN in February 2023 based on the surrogate marker of proteinuria. The successful launch and commercialization of FILSPARI in IgAN and FSGS is subject to many risks. There are numerous examples of unsuccessful product launches and failures to meet high expectations of market potential, including by pharmaceutical companies with more experience and resources than we have. While we have established our commercial team and U.S. sales force, we will need to continue to train and further develop the team in order to successfully coordinate the ongoing launch and commercialization of FILSPARI in the United States, in particular as it relates to the launch and commercialization in FSGS. There are many factors that could cause the launch and commercialization of FILSPARI to be unsuccessful, including a number of factors that are outside our control. Because no non-immunosuppressive product had previously been approved by the FDA for the treatment of IgAN, and because FILSPARI is the first and only medicine approved by the FDA for the treatment of FSGS, it is difficult to accurately estimate FILSPARI’s market potential or the time it will take to increase patient and physician awareness of FILSPARI and change current treatment paradigms.
The commercial success of FILSPARI depends on the extent to which patients and physicians accept and adopt FILSPARI for IgAN and FSGS patients. For example, if the addressable patient population suffering from primary IgAN or FSGS is smaller than we estimate, if it proves difficult to educate physicians as to the availability and potential benefits of FILSPARI, or if physicians are unwilling to prescribe or patients are unwilling to take FILSPARI, the commercial potential of FILSPARI will be limited. Although we have gained experience in recent years regarding physician, patient and payer acceptance of FILSPARI, the degree of future adoption and utilization of FILSPARI may be affected by numerous factors, including its pricing, the recent approval in FSGS, the details of the approved labeling, the evolving clinical practice guidelines and treatment paradigms, payer coverage and reimbursement policies, developments related to existing and future competitive products, and any future publications in an evolving treatment landscape. Physicians may not prescribe FILSPARI and patients may be unwilling to use FILSPARI if coverage is not provided or reimbursement is inadequate to cover a significant portion of the cost. Thus, significant uncertainty remains regarding the commercial potential of FILSPARI. If the launch or commercialization of FILSPARI is unsuccessful or perceived as disappointing, the price of our common stock could decline significantly and long-term success of the product and our company could be harmed.
In order to operate our business and increase adoption and sales of our products, we need to continue to develop our commercial organization, including maintaining a highly experienced and skilled workforce with qualified sales representatives.*
In order to successfully commercialize our products in the United States, we have built a specialized sales force. In order to successfully commercialize any approved products, we must continue to build our sales, marketing, distribution, managerial and other non-technical capabilities. Factors that may hinder our ability to successfully market and commercially distribute our products include:
•inability of sales personnel to obtain access to or educate adequate numbers of physicians on the benefits and safety of prescribing our products;
•inability to recruit, retain and effectively manage adequate numbers of effective sales personnel;
•lack of complementary products to be offered by sales personnel, which may put us at a competitive disadvantage relative to companies that have more extensive product lines; and
•unforeseen delays, costs and expenses associated with maintaining our sales organization.
If we are unable to maintain an effective sales force for our products, including the recently expanded sales force for FILSPARI or any other potential future approved products, we may not be able to generate sufficient product revenue in the United States. In addition, until the commencement of our commercial launch in IgAN in February 2023, and in FSGS in April 2026, no one in our sales force had promoted FILSPARI for the treatment of IgAN or FSGS patients, respectively. We are required to expend significant time and resources to train our sales force to be credible in educating physicians and pharmacists on the benefits of our products. In addition, we must continually train our sales force to ensure that a consistent and appropriate message about our products is being delivered to our potential customers. We currently have limited resources compared to some of our competitors, and the continued development of our own commercial organization to market our products and any additional products we may develop or acquire will be expensive and time-consuming. We also cannot be certain that we will be able to continue to successfully develop this capability.
We have granted exclusive licenses to third parties for the commercialization of sparsentan in certain territories outside of the United States, including Europe, Australia, New Zealand, Japan, South Korea, Taiwan, Brazil, Chile, Israel and the Gulf Cooperation Council Countries. If these third parties do not
38
effectively engage or maintain their sales force for sparsentan if approved in the applicable territories, our ability to recognize milestone payments and royalties from the sales in such territories will be adversely affected.
We will need to continue to expend significant time and resources to train our sales forces to be credible in discussing our products with the specialists treating the patients indicated under the product’s label. In addition, if we are unable to effectively train our sales force and equip them with effective marketing materials our ability to successfully commercialize our products could be diminished, which would have a material adverse effect on our business, results of operations and financial condition.
We are dependent on third parties for the successful commercialization of sparsentan in certain key territories outside of the United States, if approved, and such third parties' commercialization efforts may fail to meet our expectations. We may not be able to establish additional collaborations or other arrangements for sparsentan in other territories, which may adversely impact our ability to generate product revenue in additional jurisdictions.
We have granted exclusive licenses to third parties for the commercialization of sparsentan in certain territories outside of the United States, including Europe, Australia, New Zealand, Japan, South Korea, Taiwan, Brazil, Chile, Israel and the Gulf Cooperation Council Countries. Consequently, the commercial success of sparsentan in these territories will depend in significant part on the efforts of such third parties, over which we will have limited control. In August 2022, Vifor Pharma Group was acquired by CSL Limited, parent company to CSL Behring and is now operating under the brand CSL Vifor. Similarly, in the fourth quarter of 2025, Renalys was acquired by and merged into Chugai Pharmaceutical Co., Ltd. ("Chugai"). While our agreements with these partners remain in place following the acquisitions, there is no guarantee that our collaboration with these partners will not be affected, adversely or otherwise, by the changes in ownership. Moreover, in connection with the acquisitions and related restructurings, substantially less resources could be devoted to the commercialization of sparsentan in the territories licensed to such parties, or such efforts could be discontinued entirely. If we are unable to establish sales and marketing capabilities or enter into agreements with third parties to market and sell sparsentan in territories outside of the United States, if approved, our ability to generate product revenue outside of the United States may be limited.
The commercial success of our products depends on them being considered to be effective drugs with advantages over other therapies.
The commercial success of our products depends on them being considered to be effective drugs with advantages over other therapies. A number of factors, as discussed in greater detail below, may adversely impact the degree of acceptance of these products, including their efficacy, safety, price and benefits over competing therapies, as well as the coverage and reimbursement policies of third-party payers, such as government and private insurance plans.
Pegtibatinase, if approved, is expected to be administered by patients or caregivers through self-administration using a delivery device. While we have not observed material issues related to self-administration in clinical studies to date, patient acceptance, proper use of the device, and adherence in broader or longer-term real-world use may differ from clinical trial experience, which could adversely affect utilization, outcomes, or commercial success.
If unexpected adverse events are reported in connection with the use of any of these products, physician and patient acceptance of the product could deteriorate and the commercial success of such product could be adversely affected. We are required to report to the FDA events associated with our products relating to death or injury. Adverse events could result in additional regulatory controls, such as a requirement for costly post-approval clinical studies or revisions to our approved labeling which could limit the indications or patient population for a product or could even lead to the withdrawal of a product from the market.
We face substantial generic and other competition, and our operating results will suffer if we fail to compete effectively.
Under the Hatch-Waxman Amendments of the Federal Food, Drug, and Cosmetic Act, a pharmaceutical manufacturer may file an ANDA seeking approval of a generic copy of an approved innovator product or an NDA under Section 505(b)(2) that relies on the FDA’s prior findings of safety and effectiveness in approving the innovator product. A Section 505(b)(2) NDA may be for a new or improved version of the original innovator product. Our product Thiola, and products from which we may receive milestone payments such as Cholbam, are subject to immediate competition from compounded and generic entrants, as the ANDA and/or NDA for these drug products have no remaining or current patent or non-patent exclusivity. In April 2021, a generic option for the 100mg version of the original formulation of Thiola (tiopronin tablets) was approved by the FDA and an additional generic option of the original formulation of Thiola (tiopronin tablets) was approved in June 2022 and during the year ended December 31, 2022, we experienced a decrease in total net product revenues compared to the year ended December 31, 2021, which was due in part to competition from generic tiopronin tablets (100mg version of the original formulation). Additional generic versions of Thiola may be approved in the future. Several generic options for the 100mg and 300mg versions of Thiola EC have been approved by the FDA and become available. Our future net product revenues from Thiola and/or Thiola EC may be materially impacted by competition from existing or additional generic versions of Thiola or Thiola EC.
In addition, there have been a number of recent regulatory and legislative initiatives designed to encourage generic competition for pharmaceutical products, including expedited review procedures for generic manufacturers, proposed "skinny label" legislation, and incentives designed to spur generic competition of branded drugs. In particular, the FDA and the U.S. Federal Trade Commission (“FTC”) have been focused on brand companies’ denial of drug supply to potential generic competitors for testing. In December 2019, the CREATES Act was enacted, which provides a legislatively defined private right of action under which generic companies can bring suit against companies who refuse access to product for the bioequivalence testing needed to support approval of a generic product.
In 2020, we completed our response to a civil investigative demand from the FTC related to the marketing, sale, distribution and pricing of our products, including Thiola. While the investigation remains open, at this time the FTC has not indicated that it has additional questions for us and has not initiated any claim or proceeding against us relating to these matters.
We cannot currently predict the specific outcome or impact on our business of such regulatory and legislative initiatives, litigation or investigation. However, it is our policy, which is in compliance with the CREATES Act, to evaluate requests for samples of our branded products, and to provide samples in response to bona fide requests from qualified third parties, including generic manufacturers, subject to specified conditions. We have provided samples to certain generic manufacturers.
39
If additional generic versions of Thiola or Thiola EC, any generic versions of FILSPARI following the expiration of patent or regulatory exclusivity for the product, or generic versions of any other current or future products are approved, sales of that product likely would be negatively impacted, which could have a material adverse impact on our revenue and profitability. If generic versions of products from which we may receive milestone payments, such as Cholbam, are approved, our potential to receive milestone payments may be negatively impacted. In addition, the defense of litigation and response to investigation requests could result in substantial costs, reputational impact, and the diversion of management attention and resources.
The Drug Price Competition and Patent Term Restoration Act (commonly referred to as the "Hatch-Waxman Act") requires an ANDA applicant seeking FDA approval of its proposed generic product prior to the expiration of an Orange Book-listed patent (as defined below) to certify that the applicant believes that the patent is invalid, unenforceable and/or will not be infringed by the manufacture, use or sale of the drug for which the application has been submitted (a paragraph IV certification) and notify the NDA and patent holder of such certification (a paragraph IV notice). Upon receipt of a paragraph IV notice, the Hatch-Waxman Act allows the patent holder, with proper basis, to bring an action for patent infringement against the ANDA filer, asking that the proposed generic product not be approved until after the patent expires. For ANDAs that are filed (“received”) after the listing of the patent in the Orange Book, if the patent holder commences a lawsuit within 45 days from receipt of the paragraph IV notice, the Hatch-Waxman Act provides a 30-month stay during which time the FDA cannot finally approve the generic’s application. If the litigation is resolved in favor of the ANDA applicant during the 30-month stay period, the stay is lifted and the FDA may finally approve the ANDA if it is otherwise ready for approval. For ANDAs that are filed (“received”) before the listing of the patent in the Orange Book, the 30-month stay provision of the Hatch-Waxman Act does not apply. It also may be possible, depending on the approved label, for an ANDA applicant to elect to submit a section viii statement certifying that its proposed ANDA label does not contain (or carves out) any language regarding the patented method-of-use rather than certify to a listed method-of-use patent.
Healthcare reform initiatives, unfavorable pricing regulations, and changes in reimbursement practices of third-party payers or patients' access to insurance coverage could affect the pricing of and demand for our products.*
The business and financial condition of healthcare-related businesses will continue to be affected by efforts of governments and third-party payers to contain or reduce the cost of healthcare through various means. In the United States and some foreign jurisdictions, there have been a number of legislative and regulatory changes and proposed changes regarding the healthcare system that could prevent or delay marketing approval for our current product candidates or any future product candidate that we develop, restrict or regulate post-approval activities and affect our ability to profitably sell sparsentan, pegtibatinase, civorebrutinib, or any other product candidate for which we obtain marketing approval.
Our products are sold to patients whose healthcare costs are met by third-party payers, such as government programs, private insurance plans and managed-care programs. These third-party payers are increasingly attempting to contain healthcare costs by limiting both coverage and the level of reimbursement for medical products and services. Levels of reimbursement, if any, may be decreased in the future, and future healthcare reform legislation, regulations or changes to reimbursement policies of third-party payers may otherwise adversely affect the demand for and price levels of our products, which could have a material adverse effect on our sales and profitability.
The current administration has indicated that it plans to pursue additional policies aimed at lowering prescription drug costs. For example, in May 2025, the administration published an executive order regarding most favored nation (“MFN”) drug pricing, which is sometimes referred to as international reference pricing. This executive order directed the Secretary of Health and Human Services to communicate MFN price targets to pharmaceutical manufacturers, and if significant progress towards MFN pricing is not delivered, to propose a rulemaking plan to impose MFN pricing. The scope, timing, and potential impact of current and future policy initiatives remain uncertain, and accordingly, we cannot predict how such legal and regulatory changes may affect our business, operations, or financial condition. If MFN pricing or other legal or regulatory changes are implemented in a way that is broadly applicable to our products, there could be a material negative impact.
Economic, social, and congressional pressure may result in individuals and government entities increasingly seeking to achieve cost savings through mechanisms that limit coverage or payment for our products. For example, state Medicaid programs are increasingly requesting manufacturers to pay supplemental rebates and are requiring prior authorization for use of drugs. Managed care organizations continue to seek price discounts and, in some cases, to impose restrictions on the coverage of particular drugs. Government efforts to reduce Medicaid expenses may lead to increased use of managed care organizations by Medicaid programs. This may result in managed care organizations influencing prescription decisions for a larger segment of the population and a corresponding constraint on prices and reimbursement for our products.
In addition, patients’ access to employer sponsored insurance coverage may be negatively impacted by economic factors that result in increased rates of unemployment. To the extent patients taking our approved therapies become unemployed and experience a reduction to, or increased costs associated with, their insurance coverage, demand for our products could decline, which could have a material adverse effect on our sales and profitability, either as a result of decreased sales of our products and/or increased provision by us of free product to uninsured or commercially insured patients. The extent and duration of this potential impact on our business is currently unknown.
We are dependent on third parties to manufacture and distribute our products.
We have no manufacturing capabilities and rely on third-party manufacturers who are currently sole source suppliers for manufacturing of FILSPARI and Thiola. The facilities used by our third-party manufacturers must be approved by the FDA and comparable foreign regulatory authorities. Our dependence on third parties for the manufacture of our products may harm our profit margin on the sale of products and our ability to deliver products on a timely and competitive basis. Because we are ultimately responsible for ensuring that our API and finished products are manufactured in accordance with cGMP regulations and similar regulatory requirements outside the United States, it is critical that we maintain effective management practices and oversight with respect to our third-party manufacturers, including routine auditing. If our third-party manufacturers are unable to manufacture to specifications or in compliance with applicable regulatory requirements, our ability to commercialize our products will be adversely impacted and could affect our ability to gain market acceptance for our products and negatively impact our revenues.
Based on the complex relationships between the United States and certain foreign countries, there is inherent risk that political, diplomatic and national security influences might lead to trade disputes and impacts and/or disruptions to our third-party manufacturers and product supply. There is currently significant uncertainty about the future relationship between the United States and Mexico, Canada, China and certain other countries, including potential
40
changes with respect to trade policies, treaties, tariffs, taxes, and other limitations on cross-border operations. We currently source products and various materials that are necessary for the manufacturing of our products from countries that are subject to tariffs, and any changes in tariffs, trade barriers, and other regulatory requirements could lead to higher cost of goods, which would have an adverse effect on our business, financial condition and operating results, the extent of which cannot be predicted with certainty at this time.
We currently have no in-house distribution channels for FILSPARI or Thiola and we are dependent on third-party distributors to distribute such products. The outsourcing of our distribution function is complex, and we may experience difficulties that could reduce, delay or stop shipments of such products. If we encounter such distribution problems, and we are unable to quickly enter into a similar agreement with another distributor on substantially similar terms, distribution of FILSPARI and/or Thiola could become disrupted, resulting in lost revenues, provider dissatisfaction, and/or patient dissatisfaction.
Governments outside the United States tend to impose strict price controls and reimbursement approval policies, which may adversely affect our prospects for generating revenue.
Outside the United States, reimbursement and healthcare payment systems vary significantly by country, and many countries have instituted price ceilings on specific products and therapies. In some countries, particularly EU Member States and EFTA countries, the pricing of prescription pharmaceuticals is subject to governmental control. In these countries, pricing negotiations with governmental authorities can take considerable time (6 to 12 months or longer) after the receipt of marketing approval for a product. The EU provides options for EU Member States to restrict the range of medicinal products for which their national health insurance systems provide reimbursement and to control the prices of medicinal products for human use. An EU Member State may approve a specific price for the medicinal product, it may refuse to reimburse a product at the price set by the manufacturer or it may instead adopt a system of direct or indirect controls on the profitability of the company placing the medicinal product on the market. Many EU Member States also periodically review their reimbursement procedures for medicinal products, which could have an adverse impact on reimbursement status.
Moreover, to obtain reimbursement or pricing approval in some countries, we may be required to conduct a clinical trial that compares the cost effectiveness of our product candidate to other available therapies. This Health Technology Assessment (“HTA”) of medicinal products is becoming an increasingly common part of the pricing and reimbursement procedures in some EU Member States, including those representing the larger markets. The HTA process is the procedure to assess therapeutic, economic and societal impact of a given medicinal product in the national healthcare systems of the individual country. The outcome of an HTA will often influence the pricing and reimbursement status granted to these medicinal products by the competent authorities of individual EU Member States. The extent to which pricing and reimbursement decisions are influenced by the HTA of the specific medicinal product currently varies between EU Member States.
In December 2021, Regulation No 2021/2282 on HTA amending Directive 2011/24/EU, was adopted in the EU. This Regulation entered into force in January 2022 and began to apply on January 12, 2025, through a phased implementation. The Regulation is intended to boost cooperation among EU Member States in assessing health technologies, including new medicinal products, and providing the basis for cooperation at EU level for joint clinical assessments in these areas. The Regulation permits EU Member States to use common HTA tools, methodologies, and procedures across the EU, working together in four main areas, including joint clinical assessment of the innovative health technologies with the most potential impact for patients, joint scientific consultations whereby developers can seek advice from HTA authorities, identification of emerging health technologies to identify promising technologies early, and continuing voluntary cooperation in other areas. Individual EU Member States will continue to be responsible for assessing non-clinical (e.g., economic, social, ethical) aspects of health technologies, and making decisions on pricing and reimbursement. If we or our partners are unable to maintain favorable pricing and reimbursement status in EU Member States for drug candidates that we or our partners may successfully develop and for which we or our partners may obtain regulatory approval, any anticipated revenue from and growth prospects for those products in the EU could be negatively affected. In light of the fact that the United Kingdom has left the EU, Regulation No 2021/2282 on HTA does not apply in the United Kingdom. However, the UK Medicines and Healthcare products Regulation Agency (“MHRA”) is working with UK HTA bodies and other national organizations, such as the Scottish Medicines Consortium (“SMC”), the National Institute for Health and Care Excellence (“NICE”), and the All-Wales Medicines Strategy Group, to introduce new pathways supporting innovative approaches to the safe, timely and efficient development of medicinal products, including, effective as of March 31, 2025, relaunching the Innovative Licensing and Access Pathway with more predicable timelines and closer involvement of the National Health Service.
In addition, certain governmental authorities may conduct reviews of reimbursement previously provided and assert for various reasons that amounts need to be repaid. For example, in October 2021 our distributor/exploitant in France for our previously marketed product Kolbam (which has since been divested) informed us that they had received a notice that the price previously paid for Kolbam during its period on the market in France had been recalculated by the agency responsible for pharmaceutical pricing in France. Such notice was confirmed by a decision in October 2023, asserting percentages of our turnover owed for repayment. In April 2024, we filed an appeal with the Competent Administrative Court regarding this matter. In October 2024, we received an invoice from the government authority for approximately $6.2 million, which we paid while we continue to pursue an appeal of the decision and the amount paid. While we cannot predict the amount that we may ultimately need to repay following ongoing review and future potential appeal proceedings, from 2015 through 2020, the period during which we had sales of Kolbam in France, our aggregate revenues from sales of Kolbam in France attributable to all purchasers/payers were approximately $8 million. If reimbursement of our products is unavailable or limited in scope or amount, or if pricing is set at unsatisfactory levels or subject to re-assessment and recoupment procedures, our prospects for generating revenue outside of the United States, if any, could be adversely affected and our business may suffer.
We may not be able to rely on orphan drug exclusivity for our products.*
Regulatory authorities in some jurisdictions, including the United States and the EU, may designate drugs for relatively small patient populations as orphan drugs, providing eligibility for orphan drug exclusivity upon regulatory approval if certain jurisdictional-specific conditions are met. For example, FILSPARI has been granted orphan drug designation for the treatment of IgAN and has been awarded seven years of orphan drug exclusivity in the United States (running from the date of accelerated approval) to reduce proteinuria in adults with primary IgAN at risk of rapid disease progression, and has been granted a separate seven years of Orphan Drug Exclusivity in the U.S. (running from the date of full approval) to slow kidney function decline in adults with primary IgAN who are at risk for disease progression, excluding the use provided for in the aforementioned Orphan Drug Exclusivity granted in connection with the accelerated approval. Generally, if a product with an orphan drug designation subsequently receives the first marketing approval for the indication for which it has such designation, that product is entitled to a period of marketing exclusivity, which precludes the applicable regulatory authority from approving another
41
marketing application for the same drug for the same indication for that time period. The applicable period is seven years in the United States and ten years in the EU or, in the case of orphan drugs for which a pediatric investigation plan has been completed, 12 years. Even though we have been awarded orphan drug designation in the United States and the EU for sparsentan for the treatment of IgAN and FSGS and for pegtibatinase for the treatment of classical homocystinuria ("HCU"), we may not be able to maintain it in the EU and the orphan drug designation may not result in orphan drug exclusivity for pegtibatinase if approved, or FSGS in the EU if approved. For example, if a competitive product that contains the same active moiety and treats the same disease as our product is shown to be clinically superior to our product, any orphan drug exclusivity we have obtained will not block the approval of such competitive product and we may effectively lose orphan drug exclusivity. Similarly, if a competitive product that contains the same active moiety and treats the same disease as our product candidate is approved for orphan drug exclusivity before our product candidate, we may not be able to obtain approval for our product candidate until the expiration of the competitive product’s orphan drug exclusivity unless our product candidate is shown to be clinically superior to the competitive product.
Guidelines and recommendations published by various organizations may impact the use of our products.
Government agencies promulgate regulations and guidelines directly applicable to us and to our products. However, professional societies, industry groups, practice management groups, insurance carriers, physicians, private foundations and other organizations involved in various diseases or conditions from time to time may also publish guidelines or recommendations to healthcare providers, administrators and payers, and patient communities. Recommendations by government agencies or those other groups/organizations may relate to such matters as clinical guidelines, usage and reimbursement of our products by government and private payers. Recommendations or guidelines that are followed by patients, healthcare providers and payers could impact the use of our products in positive or negative ways. In addition, recommendations or guidelines may not be followed by patients, healthcare providers or payers, and thus any such positive recommendations or guidelines may not have a positive impact on the use of our products. Any such recommendations or guidelines may be updated over time as the treatment landscape evolves, and future changes to guidelines or recommendations could have a material adverse impact on the use of our products. Any recommendations or guidelines, or changes thereto, that result in decreased use or reimbursement of our products could materially and adversely affect our product sales, business and operating results.
Risks Related to the Development of our Product Candidates
Our clinical trials are expensive and time-consuming and may fail to demonstrate the safety and efficacy of our product candidates.*
Before obtaining regulatory approval for the sale of any of our current or future product candidates, we must subject these product candidates to extensive nonclinical and clinical testing to demonstrate their safety and efficacy for humans. Clinical trials are expensive, time-consuming and may take years to complete.
We may experience numerous unforeseen events during, or as a result of, preclinical or nonclinical testing and the clinical trial process that could delay or prevent our ability to obtain, or impact our willingness to pursue, regulatory approval or commercialize our product candidates, including:
•our preclinical or nonclinical tests or clinical trials may produce negative or inconclusive results, and we may decide, or regulators may require us, to conduct additional nonclinical testing or clinical trials or we may abandon projects that we expect to be clinically promising in light of cost or strategic considerations;
•regulators may require us to conduct studies of the long-term effects associated with the use of our product candidates;
•regulators, institutional review boards or ethics committees may not authorize us to commence a clinical trial or conduct a clinical trial at a prospective trial site;
•the FDA or any non-United States regulatory authority may impose conditions on us regarding the scope or design of our clinical trials or may require us to resubmit our clinical trial protocols to institutional review boards or ethics committees for re-inspection due to changes in the regulatory environment;
•the number of patients required for our clinical trials may be larger than we anticipate or participants may drop out of our clinical trials at a higher rate than we anticipate;
•our third-party contractors or clinical investigators may fail to comply with regulatory requirements or fail to meet their contractual obligations to us in a timely manner;
•we might have to suspend, vary or terminate one or more of our clinical trials if we, regulators or institutional review boards or ethics committees determine that the participants are being exposed to unacceptable health risks;
•regulators, institutional review boards or ethics committees may require that we hold, suspend, vary or terminate clinical research for various reasons, including noncompliance with regulatory requirements;
•the cost of our clinical trials or the anticipated commercialization costs may be greater than we anticipate;
•the supply or quality of our product candidates or other materials necessary to conduct our clinical trials may be insufficient or inadequate, or more expensive than we originally anticipated, or we may not be able to reach agreements on acceptable terms with prospective suppliers or clinical research organizations; and
•the effects of our product candidates may not be the desired effects or may include undesirable side effects or the product candidates may have other unexpected characteristics.
42
We will only obtain regulatory approval to commercialize a product candidate if we can demonstrate to the satisfaction of the FDA, and in the case of foreign commercialization, to the applicable foreign regulatory authorities, in well-designed and conducted clinical trials, that our product candidates are safe and effective and otherwise meet the appropriate standards required for approval for a particular indication.
Conducting clinical trials effectively in pursuit of regulatory approval requires significant resources, and the costs of conducting clinical trials varies depending on a number of factors, including the dosage of the study therapy, trial size and duration. These costs may prove greater than we originally anticipated, which may result in us choosing to abandon or forgo clinical trials that we deem clinically promising as we actively strategize over time with respect to the allocation of our resources.
Our product development costs will also increase if we experience delays in testing or approvals. We do not know whether any nonclinical tests or clinical trials will be initiated as planned, will need to be restructured or will be completed on schedule, if at all. Significant nonclinical or clinical trial delays also could shorten the patent protection period during which we may have the exclusive right to commercialize our product candidates. In addition, such delays could allow our competitors to bring products to market before we do and impair our ability to commercialize our products or product candidates.
If we are required to conduct additional clinical trials or other testing of our product candidates beyond those that we currently contemplate, if we are unable to successfully complete our clinical trials or other testing, if the results of these trials or tests are not positive or are only modestly positive or if there are safety concerns, we may:
•be delayed in obtaining, or may not be able to obtain, marketing approval for one or more of our product candidates;
•obtain approval for indications that are not as broad as intended or entirely different than those indications for which we sought approval; and
•have the product removed from the market after obtaining marketing approval.
We may not be able to initiate or continue clinical trials in the rare diseases on which we are focused if we are unable to locate a sufficient number of eligible patients willing and able to participate in the clinical trials required by the FDA or foreign regulatory authorities. In addition, as other companies and researchers may be concurrently developing therapies for the same or similar indications that we are focused on, we could face competition for a limited number of patients, investigators and clinical trial sites willing to participate in clinical trials. Our inability to enroll and maintain a sufficient number of patients for any of our current or future clinical trials would result in significant delays or may require us to abandon one or more clinical trials altogether.
Success in nonclinical testing and early clinical trials does not ensure that later clinical trials will be successful.*
Success in nonclinical testing and early clinical trials does not ensure that later clinical trials will be successful. For example, the positive nonclinical data we have seen from pegtibatinase being tested in a mouse model of homocystinuria and the positive topline results we reported in December 2021 and May 2023 from the ongoing Phase 1/2 clinical trial of pegtibatinase may not be replicated in future studies. We cannot assure that any current or future clinical trials of sparsentan, pegtibatinase or civorebrutinib will ultimately be successful. Before obtaining regulatory approval to conduct clinical trials of our product candidates, we must conduct extensive nonclinical tests to demonstrate the safety of our product candidates in animals. Nonclinical testing is expensive, difficult to design and implement, and can take many years to complete. In addition, during the clinical development process, additional nonclinical toxicology studies are routinely conducted concurrently with the clinical development of a product candidate. If any of our product candidates show unexpected findings in concurrent toxicology studies, we could experience potentially significant delays in, or be required to abandon, development of that product candidate. A failure of one or more of our nonclinical studies can occur at any stage of testing.
Communications and/or feedback from regulatory authorities related to our current or planned future clinical trials does not guarantee any particular outcome from or timeline for regulatory review, and expedited regulatory review pathways may not actually lead to faster development or approval.*
Communications and/or feedback from regulatory authorities, including the FDA or EMA, related to our current or future clinical trials does not guarantee any particular outcome from or timeline for regulatory review for such clinical trials, and expedited regulatory review pathways may not actually lead to faster development or approval.
There is no guarantee that our collaborator CSL Vifor will be able to establish a pathway to a potential submission of sparsentan for FSGS in Europe based on the results from the DUPLEX Study, that the EMA will support an application for sparsentan in FSGS, or that sparsentan will be approved for FSGS in Europe.
In December 2023, we initiated the pivotal Phase 3 HARMONY Study to support the potential approval of pegtibatinase for the treatment of classical HCU. The HARMONY Study is a global, randomized, multi-center, double-blind, placebo-controlled Phase 3 clinical trial designed to evaluate the efficacy and safety of pegtibatinase as a novel treatment to reduce total homocysteine ("tHcy") levels. In September 2024, we announced a voluntary pause of enrollment in the HARMONY Study. The voluntary enrollment pause was enacted following our determination that the desired drug substance profile was not achieved in the initial scale-up process. While we have made process improvements and have manufactured material at the larger scale, and while we have restarted enrollment activities for the Phase 3 HARMONY Study, and while the FDA has granted Fast Track and Breakthrough Therapy designations to pegtibatinase for the treatment of HCU, there is no guarantee that our pivotal Phase 3 HARMONY Study will be conducted or completed on the anticipated timeline or be successful, or that pegtibatinase will be approved for HCU in the future.
Obtaining access to an expedited program (such as Fast Track and Breakthrough Therapy designations) may not in fact lead to faster development timelines or achieve faster review or approval than conventional FDA procedures. We may experience delays in approval timelines attributable to, among other things, acquiring sufficient supply of our product to conduct clinical trials, identifying and resolving issues relating to chemistry, manufacturing and controls, or conducting additional nonclinical or clinical studies. In addition, the FDA may withdraw access to an expedited program if it believes the access or designation is no longer supported by the data from our program.
43
Interim, topline and preliminary data from our clinical trials that we announce or publish may change materially as more patient data become available and audit and verification procedures are complete.
From time to time, we may publicly disclose preliminary or topline or interim data from our clinical studies, which is based on a preliminary analysis of then-available data, and the results and related findings and conclusions are subject to change following a more comprehensive review of the data related to the particular study. We also make assumptions, estimations, calculations and conclusions as part of our analyses of data, and we may not have received or had the opportunity to fully and carefully evaluate all data. As a result, the topline results that we report may differ from future results of the same studies, or different conclusions or considerations may qualify such results, once additional data have been received and fully evaluated. Topline data also remain subject to audit and verification procedures that may result in the final data being materially different from the preliminary data we previously published. As a result, topline data should be viewed with caution. From time to time, we may also disclose interim data from our clinical trials. Interim data from clinical trials that we may complete are subject to the risk that one or more of the clinical outcomes may materially change as patient enrollment and dosing continues and more patient data become available. Adverse differences between preliminary or interim data and final or confirmatory data could significantly harm our business prospects.
Further, others, including regulatory authorities, may not accept or agree with our assumptions, estimates, calculations, conclusions or analyses or may interpret or weigh the importance of data differently, which could impact the value of the particular program, the approvability or commercialization of the particular product candidate or product and our company in general. In addition, the information we choose to publicly disclose regarding a particular study or clinical trial is based on what is typically extensive information, and you or others may not agree with what we determine is the material or otherwise appropriate information to include in our disclosure, and any information we determine not to disclose may ultimately be deemed significant with respect to future decisions, conclusions, views, activities or otherwise regarding a particular therapy, therapeutic candidate or our business. If the topline data that we report differ from actual results, or if others, including regulatory authorities, disagree with the conclusions reached, our ability to obtain approval for, and commercialize, our product candidates may be harmed, which could harm our business, operating results, prospects or financial condition.
We and/or a collaborative partner are or will be subject to ongoing regulatory obligations and continued regulatory review for our approved products and any product candidates that receive regulatory approval.*
In September 2024, the FDA granted full approval of FILSPARI to slow kidney function decline in adults with primary IgAN who are at risk of disease progression, and in April 2026, the FDA granted full (traditional) approval of FILSPARI to reduce proteinuria in adult and pediatric patients aged 8 years and older with FSGS without nephrotic syndrome. Any future regulatory approvals that sparsentan or any of our other product candidates receives may be subject to significant limitations on the approved indicated uses for which the product may be marketed or to the conditions of approval, or contain requirements for potentially costly post-marketing testing, including Phase 4 clinical trials, and surveillance to monitor the safety and efficacy of the product candidate.
In addition, our products, including FILSPARI, and any of our product candidates that are approved by the FDA or a comparable foreign regulatory authority, are or will be subject to extensive and ongoing regulatory requirements, including for the manufacturing, labeling, packaging, distribution, adverse event reporting, storage, advertising, promotion, import, export, recordkeeping, conduct of potential post-marketing studies and post-market submission requirements. These requirements include submissions of safety and other post-marketing information and reports, registration, as well as continued compliance with current good manufacturing practices and good clinical practices, for any clinical trials that we conduct post-approval. Later discovery of previously unknown problems with a product, including adverse events of unanticipated severity or frequency, undesirable side effects caused by the product, problems encountered by our third-party manufacturers or manufacturing processes, or failure to comply with regulatory requirements, either before or after product approval, may result in, among other things:
•restrictions on the marketing, manufacturing, or distribution of the product;
•requirements to include additional warnings on the label;
•requirements to create or enhance a medication guide outlining the risks to patients;
•withdrawal of the product from the market;
•voluntary or mandatory product recalls;
•requirements to change the way the product is administered or for us to conduct additional clinical trials;
•fines, warning or untitled letters or holds on clinical trials;
•refusal by the FDA or comparable foreign regulatory authorities to approve pending applications or supplements to approved applications filed by us or our strategic partners, or suspension, variation or revocation of product license approvals;
•product seizure or detention, or refusal to permit the import or export of products;
•injunctions or the imposition of civil or criminal penalties; and
•harm to our reputation.
For example, we have certain post-marketing requirements and commitments associated with FILSPARI. Further, we face risks relating to those post-marketing obligations, as well as the commercial acceptance of FILSPARI. If the regulatory approval for FILSPARI and/or Thiola are withdrawn for any reason, it would have a material adverse impact on our sales and profitability. Furthermore, if the regulatory approval for Chenodal and/or Cholbam are withdrawn for any reason, it would reduce the chance that we will receive certain milestone payments from the sale of our bile acid product portfolio in August 2023. On July 28, 2026, we received an untitled letter from FDA's Office of Prescription Drug Promotion ("OPDP") regarding claims made in a professional visual aid for FILSPARI. The letter asserts that the professional visual aid includes certain false or misleading claims about the safety and efficacy of FILSPARI. We intend to respond to FDA within the requested timeframe.
44
The third-party clinical investigators and contract research organizations that we rely upon to conduct our clinical trials may not be diligent, careful or timely, and may make mistakes, in the conduct of our trials.
We depend on third-party clinical investigators and contract research organizations (“CROs”) to conduct our clinical trials under agreements with us. The CROs play a significant role in the conduct of our clinical trials. Failure of the CROs to meet their obligations could adversely affect clinical development of our product candidates. The third-party clinical investigators are not our employees and we cannot control the timing or amount of resources they devote to our studies. If their performance is substandard, it could delay or prevent approval of our FDA applications. Moreover, these third-party investigators and CROs may also have relationships with other commercial entities, some of which may compete with us. If third-party investigators and CROs allocate their resources to assist our competitors at our expense, it could harm our competitive position. The introduction of new third parties into our ongoing clinical trials increases the risks associated with our dependence on third parties, including the risk that substandard performance by, or competing interests of, such third parties could have a negative impact on our clinical trials.
Risks Related to our Products and Product Candidates
Our products may not achieve or maintain expected levels of market acceptance or commercial success.
The success of our products is dependent upon achieving and maintaining market acceptance. Commercializing products is time consuming, expensive and unpredictable. There can be no assurance that we will be able to, either by ourselves or in collaboration with our partners or through our licensees, successfully commercialize new products or current products or gain market acceptance for such products. New product candidates that appear promising in development may fail to reach the market or may have only limited or no commercial success.
Further, the discovery of significant problems with a product similar to one of our products that implicate (or are perceived to implicate) an entire class of products could have an adverse effect on sales of the affected products. Accordingly, new data about our products, or products similar to our products, could negatively impact demand for our products due to real or perceived side effects or uncertainty regarding efficacy and, in some cases, could result in product withdrawal.
Our current products, including FILSPARI, and any product candidates that receive marketing approval, that we or a collaboration partner bring to the market may not gain market acceptance by physicians, patients, third-party payers, and others in the medical community. If these products do not achieve an adequate level of acceptance, we may not generate significant product revenue and we may not become profitable. The degree of market acceptance of our current products and product candidates, if approved for commercial sale, will depend on a number of factors, including:
•the prevalence and severity of any side effects, including any limitations or warnings contained in a product’s approved labeling;
•the efficacy and potential advantages over alternative treatments;
•the pricing of our product candidates;
•the relative convenience and ease of administration;
•the willingness of the target patient population to try new therapies and of physicians to prescribe these therapies;
•the strength of marketing and distribution support and timing of market introduction of competitive products;
•publicity concerning our products or competing products and treatments; and
•sufficient third-party insurance coverage and reimbursement.
As part of the NDA review process for sparsentan for IgAN, the FDA initially required us to include a REMS and a boxed warning on the label regarding mandatory birth control for patients of child-bearing potential regarding risk of embryo-fetal toxicity, as has been required for certain other approved endothelin antagonists, and a REMS and boxed warning on the label for liver monitoring regarding potential risk of hepatotoxicity, as has been required for certain other approved endothelin antagonists. Initially, as part of the liver monitoring REMS, monthly monitoring of each patient was required for the first year a patient was on treatment, and quarterly thereafter. In August 2025, we announced that the FDA approved updated REMS labeling for FILSPARI, reducing the frequency of liver function monitoring to every three months from the onset of treatment with FILSPARI, and removing the embryo-fetal toxicity monitoring requirement from the REMS. While we intend to utilize our continued clinical trial experience with FILSPARI and post-marketing data gathering commitment to potentially support lifting of the liver monitoring REMS in the future following sufficient experience with FILSPARI and if supported by the data, there is no guarantee that such efforts will be successful.
Even if a potential or current product displays a favorable efficacy and safety profile in nonclinical and clinical trials, market acceptance of the product will not be known until after it is launched. The efforts by us or any applicable collaboration partner to educate patients, the medical community, and third-party payers on the benefits of our products may require significant resources and may never be successful. Such efforts to educate the marketplace may require more resources than are required by the conventional marketing technologies employed by our competitors.
The market opportunities for our products and product candidates may be smaller than we believe they are.*
Certain of the diseases that our current and future product candidates are being developed to address, including but not limited to IgAN, FSGS, PMN and HCU, are relatively rare. Our estimates and projections of both the number of people who have a particular disease or condition, as well as the subset of people who have the potential to benefit from treatment with our product candidates, may not be accurate.
Currently, most reported estimates of the prevalence of such diseases or conditions are based on studies of small subsets of the population of specific geographic areas, which are then extrapolated to estimate the prevalence in the broader world population. As new studies are performed, the estimated prevalence of a particular disease or condition may change. There can be no assurance that the prevalence of any such disease or condition in study populations accurately reflect the prevalence in the broader world population.
45
If our estimates of the prevalence of a particular disease or condition, or of the number of patients who may benefit from treatment with sparsentan, pegtibatinase or civorebrutinib prove to be incorrect or if regulatory approval is conditioned on label restrictions that limit the approved patient population, the market opportunities for our product candidates may be smaller than we believe they are, our prospects for generating revenue may be adversely affected and our business may suffer.
Our product candidates may cause undesirable side effects or have other properties that could delay or prevent their regulatory approval or commercialization.
Undesirable side effects caused by our product candidates could interrupt, delay or halt clinical trials and could result in the denial of regulatory approval by the FDA or other regulatory authorities for any or all targeted indications, and in turn prevent us from commercializing our product candidates and generating revenues from their sale.
In addition, if any of our product candidates receive marketing approval and we or others later identify undesirable side effects caused by the product:
•regulatory authorities may require the addition of restrictive labeling statements;
•regulatory authorities may withdraw, suspend or vary their approval of the product; and
•we may be required to change the way the product is administered or conduct additional clinical trials.
Any of these events could prevent us from achieving or maintaining market acceptance of the affected product or could substantially increase the costs and expenses of commercializing the product candidate, which in turn could delay or prevent us from generating significant revenues from its sale or adversely affect our reputation.
We do not currently have patent protection for certain of our commercial products. If we are unable to obtain and maintain protection for the intellectual property relating to our technology and products, their value will be adversely affected.*
Our success will depend in large part on our ability to obtain and maintain protection in the United States and other countries for the intellectual property covering, or incorporated into, our technology and products. The patent situation in the field of biotechnology and pharmaceuticals generally is highly uncertain and involves complex legal, technical, scientific and factual questions. We do not have, and do not expect to obtain, patent protection for the original formulation of Thiola. Additionally, although we have a license to a granted U.S. patent covering the treatment of cystinuria by administering Thiola EC with food (U.S. Patent No. 11,458,104, "the '104 patent”), certain generic manufacturers have been able to obtain “skinny-label” approvals of generic versions of tiopronin EC as described below. More generally, we may not be able to obtain additional issued patents relating to our technology or products. Even if issued, patents issued to us or our licensors may be challenged, narrowed, invalidated, held to be unenforceable or circumvented, which could limit our ability to stop competitors from marketing similar products or reduce the term of patent protection we may have for our products. In addition, in certain circumstances with respect to method of use patents, an ANDA applicant may certify that its proposed ANDA label does not contain (or carves out) any language regarding the patented method-of-use rather than certify to a listed method-of-use patent. To date, several generic options for the 100mg and 300mg versions of Thiola EC have been approved by the FDA and become available. Accordingly, Thiola EC is subject to generic competition. Changes in either patent laws or in interpretations of patent laws in the United States and other countries may diminish the value of our intellectual property or narrow the scope of our patent protection.
Patent laws vary by country. Some countries have compulsory licensing laws under which a patent owner may be required to grant licenses to third parties. Some countries do not grant or enforce patents related to medical treatments, or limit enforceability in the case of a public emergency. In addition, many countries limit the enforceability of patents against government agencies or government contractors. If we are unable to obtain or enforce patents related to medical treatments in certain countries, or we or any of our licensors is forced to grant a license to third parties with respect to any patents relevant to our business, our business may be adversely affected.
The intellectual property systems in other countries can be destabilized as a result of political events, during which the ability to obtain, maintain and enforce intellectual property protection in the affected country may be uncertain and evolving. For example, as a result of the ongoing war between Ukraine and Russia, Russian officials have suggested that they may treat patents or patent applications owned by parties from certain countries, including the United States, as unenforceable and/or provide for zero compensation compulsory licenses to such patents or patent applications. Recent court decisions in Russia have raised questions about the strength of trademark protections in Russia. The U.S. government’s response to political events may also negatively affect our ability to obtain, maintain and enforce intellectual property protection in the affected country. For example, the U.S. government has issued sanctions against Russia related to the ongoing war in Ukraine, and as a result of these sanctions, it may not be possible to pay fees necessary for prosecution and maintenance of Russian patent applications and patents in the absence of licenses or exclusions set forth by the U.S. government authorizing transactions in connection with intellectual property. Payments for trademark protection may be similarly impacted. The U.S. Department of the Treasury has issued General License No. 31, authorizing such transactions to allow filing, prosecution and maintenance of Russian patents and trademarks. Uncertainties regarding political events, including the ongoing war between Ukraine and Russia, as well as any resulting losses of intellectual property protection, could harm our business.
Our product FILSPARI was covered by U.S. Patent No. 6,638,937, which expired in 2019 and to which we had an exclusive license. In addition, U.S. Patent No. 9,662,312, to which we also have an exclusive license and which was granted on May 30, 2017 and expires in 2030, covers the use of sparsentan for treating glomerulosclerosis, including FSGS. U.S. Patent No. 9,993,461, to which we also have an exclusive license and which was granted on June 12, 2018 and expires in 2030, covers the use of sparsentan for treating IgAN as well as glomerulosclerosis, including FSGS. While we have additional pending U.S. and foreign patent applications directed to sparsentan and its uses, including an allowed U.S. patent application directed to certain methods of using sparsentan in IgAN, there is no guarantee that any pending, allowed or future patent applications will result in issued patents, issue on a timeline that provides material protection, or, if issued, be listed in the Orange Book or contain claims of commercially meaningful scope.
For products we develop based on a new chemical entity not previously approved by the FDA, we expect that in addition to the protection afforded by our patent filings that we will be able to obtain five years regulatory exclusivity via the provisions of the Food, Drug, and Cosmetic Act ("FDC Act") and possibly
46
seven years regulatory exclusivity via the orphan drug provisions of the FDC Act. The periods of regulatory exclusivity may, if certain conditions are satisfied, be extended by six months on the basis of pediatric exclusivity, thereby resulting in exclusivity periods of 5.5 years and 7.5 years, respectively. In addition, companies may be able to obtain up to five years patent term extension (to compensate for regulatory approval delay) for one patent covering such a product for its FDA-approved use. Such a patent, like the periods of regulatory exclusivity, also may be extended by a further six months on the basis of pediatric exclusivity if certain conditions are satisfied. While we have filed an application for patent term extension of U.S. Patent No. 9,993,461, and subsequently filed a supplement to this application to reflect the FDA -determined regulatory review period for sparsentan, which, if granted could extend the term of U.S. Patent No. 9,993,461 to January 2033, there is no guarantee that such patent term extension will be granted to such date, or at all. In addition, while we intend to seek pediatric exclusivity for FILSPARI based on our ongoing development efforts, which, if granted, could extend the term of such patent by an additional six months, the granting of pediatric exclusivity requires a series of regulatory interactions to reach agreement with the FDA, and there is no guarantee that our pediatric development efforts will support a path to pediatric exclusivity or that pediatric exclusivity will be granted by the FDA on a timeline that confers benefit on the term of patent coverage or regulatory exclusivity for FILSPARI, or at all.
The degree of future protection for our proprietary rights is uncertain, and we cannot ensure that:
•we or our licensors were the first to make the inventions covered by each of our pending patent applications;
•we or our licensors were the first to file patent applications for these inventions;
•others will not independently develop similar or alternative technologies or duplicate any of our technologies;
•any patents issued to us or our licensors that provide a basis for commercially viable products will provide us with any competitive advantages or will not be challenged by third parties;
•we will develop additional proprietary technologies that are patentable;
•we will file patent applications for new proprietary technologies promptly or at all;
•the claims we make in our patents will be upheld by patent offices in the United States and elsewhere;
•our patents will not expire prior to or shortly after commencing commercialization of a product; and
•the patents of others will not have a negative effect on our ability to do business.
We have a license agreement with Ligand Pharmaceuticals for the rights to sparsentan for which we have obtained FDA approval in the areas of IgAN and FSGS. This license subjects us to various commercialization, reporting and other obligations. If we were to default on our obligations, we and our licensees (including CSL Vifor and Chugai) could lose our rights to sparsentan. We have obtained a U.S. patent and European patent each covering the use of sparsentan for treating glomerulosclerosis, including FSGS, as well as a second U.S. patent and a second European patent each covering both the use of sparsentan for treating IgAN and the use of sparsentan for treating glomerulosclerosis, including FSGS. In November 2020, a third party filed an opposition to our second European patent (European Patent No. EP3222277, “the ‘277 EP Patent”), in the EPO. In March 2026, following an appeal hearing at the Technical Boards of Appeal of the EPO, the opposing party prevailed and the ’277 EP Patent was revoked. Accordingly, previously granted supplementary protection certificates, or SPCs, based on the ’277 EP Patent have been or will be invalidated. As a result of this outcome with respect to the ‘277 EP Patent and associated SPCs, we expect to rely on the data and/or marketing exclusivity that may be available in the EU. In addition, future changes to EU legislation may affect the availability, scope, or duration of rights granted after such changes take effect.
We have a license and collaboration agreement with Everest Medicines for certain rights to develop and commercialize civorebrutinib, including in the United States, and rely on licenses and sublicenses to certain patents and know-how owned or controlled by Everest Medicines and third parties. If we were to default on our obligations under the agreement or applicable upstream third-party agreements, or if the agreement were terminated, we could lose our rights to civorebrutinib and our ability to develop, manufacture or commercialize civorebrutinib could be adversely affected.
Our patents also may not afford us protection against competitors with similar technology. Because patent applications in the United States and many other jurisdictions are typically not published until 18 months after filing, or in some cases not at all, and because publications of discoveries in the scientific literature often lag behind the actual discoveries, neither we nor our licensors can be certain that we or they were the first to make the inventions claimed in our or their issued patents or pending patent applications, or that we or they were the first to file for protection of the inventions set forth in these patent applications. If a third party has also filed a United States patent application prior to the effective date of the relevant provisions of the America Invents Act (i.e. before March 16, 2013) covering our product candidates or a similar invention, we may have to participate in an adversarial proceeding, known as an interference, declared by the USPTO to determine priority of invention in the United States. The costs of these proceedings could be substantial, and it is possible that our efforts could be unsuccessful, resulting in a loss of our United States patent position.
We cannot assure you that third parties will not assert patent or other intellectual property infringement claims against us with respect to technologies used in our products. If patent infringement suits were brought against us, we may be unable to commercialize some of our products which could severely harm our business. Litigation proceedings, even if not successful, could result in substantial costs and harm our business.
We expect to rely on orphan drug status to develop and commercialize certain of our products and product candidates, but our orphan drug designations may not confer marketing exclusivity or other expected commercial benefits.
We expect to rely on orphan drug exclusivity for sparsentan and potential future product candidates that we may develop. Orphan drug status currently confers seven years of marketing exclusivity in the United States under the FDC Act, and up to ten years of marketing exclusivity in the EU for a particular product in a specified indication or, in the case of orphan drugs for which a pediatric investigation plan has been completed, 12 years. The FDA and European Commission have granted orphan designation for sparsentan for the treatment of IgAN and FSGS, and pegtibatinase for the treatment of homocystinuria. While we have been granted these orphan designations, we will not be able to rely on these designations to exclude other companies from manufacturing or selling these molecules for the same indication beyond these time frames. Furthermore, in the EU, orphan drug status is re-evaluated in connection with the marketing authorization review process and a product candidate must re-qualify as of such time in order to maintain orphan drug status
47
and benefit from the potential regulatory exclusivity periods related to marketing authorizations granted to orphan products. The period of market exclusivity in the EU may, however, be reduced to six years if, at the end of the fifth year, it is established that the product no longer meets the criteria on the basis of which it received orphan medicinal product destination, including where it can be demonstrated on the basis of available evidence that the original orphan medicinal product is sufficiently profitable not to justify maintenance of market exclusivity or where the prevalence of the condition has increased above the threshold. Additionally, a marketing authorization may be granted to a similar medicinal product with the same orphan indication during the 10 year period if: (i) the applicant consents to a second original orphan medicinal product application, (ii) the manufacturer of the original orphan medicinal product is unable to supply sufficient quantities; or (iii) the second applicant can establish that its product, although similar, is safer, more effective or otherwise clinically superior to the original orphan medicinal product. A company may voluntarily remove a product from the register of orphan products.
For any product candidate for which we have been granted orphan drug designation in a particular indication, it is possible that another company also holding orphan drug designation for the same product candidate will receive marketing approval for the same indication before we do. If that were to happen, our applications for that indication may not be approved until the competing company's period of exclusivity expires. Even if we are the first to obtain marketing authorization for an orphan drug indication in the United States, there are circumstances under which a competing product may be approved for the same indication during the seven-year period of marketing exclusivity, such as if the later product is shown to be clinically superior to our orphan product, or if the later product is deemed a different product than ours. Further, the seven-year marketing exclusivity would not prevent competitors from obtaining approval of the same product candidate as ours for indications other than those in which we have been granted orphan drug designation, or for the use of other types of products in the same indications as our orphan product.
Any therapies we develop may become subject to unfavorable pricing regulations, third-party reimbursement practices or healthcare reform initiatives, thereby harming our business.
In March 2010, the Patient Protection and Affordable Care Act, as amended by the Health Care and Education Reconciliation Act of 2010 (collectively, the "PPACA"), was signed into law, which intended to, among other things, broaden access to health insurance, reduce or constrain the growth of healthcare spending, enhance remedies against fraud and abuse, add new transparency requirements for healthcare and health insurance industries, impose new taxes and fees on the health industry and impose additional health policy reforms. There have been executive, judicial, Congressional, and political challenges and amendments to certain aspects of the PPACA. For example, on July 4, 2025, the One Big Beautiful Bill Act, or OBBBA, was signed into law, which narrowed access to PPACA marketplace exchange enrollment and declined to extend the PPACA enhanced advanced premium tax credits that expired at the end of 2025, which, among other provisions in the law, are anticipated to reduce the number of Americans with health insurance. The OBBBA also is expected to reduce Medicaid spending and enrollment by implementing work requirements for some beneficiaries, capping state-directed payments, reducing federal funding, and limiting provider taxes used to fund the program. Congress is considering proposed legislation intended to further reduce healthcare costs with alternatives to replace the expired PPACA subsidies. It is possible that the PPACA will be subject to judicial or Congressional challenges in the future. It is unclear how any potential future healthcare reform measures of the current administration will impact the PPACA and our business.
Additionally, in August 2011, President Obama signed into law the Budget Control Act of 2011, which, among other things, includes aggregate reductions to Medicare payments to providers of up to 2% per fiscal year, which went into effect beginning on April 1, 2013 and, due to subsequent legislative amendments, will stay in effect until 2032 unless additional Congressional action is taken. Additionally, in January 2013, the American Taxpayer Relief Act of 2012 was signed into law, which, among other things, reduced Medicare payments to several providers, including hospitals and imaging centers.
In addition, in the EU, in April 2023, the European Commission adopted a proposal for a new Directive and Regulation to revise the existing pharmaceutical legislation. In April 2024, the Parliament adopted its related position and on 4 June 2025 the European Council agreed on its position. The Council, the Parliament and the European Commission have begun trilogue negotiations with a view to reaching an agreement on the package. A decrease in data and market exclusivity opportunities for our product candidates in the EU could make them open to generic or biosimilar competition earlier than is currently the case with a related reduction in reimbursement status.
If we are unable to obtain and maintain coverage and adequate reimbursement from governments or third-party payers for any products that we may develop or if we are unable to obtain acceptable prices for those products, our prospects for generating revenue and achieving profitability will suffer.*
Our prospects for generating revenue and achieving profitability will depend heavily upon the availability of coverage and adequate reimbursement for the use of our approved product candidates from governmental and other third-party payers, both in the United States and in other markets. Reimbursement by a third-party payer may depend upon a number of factors, including the third-party payer’s determination that use of a product is:
•a covered benefit under its health plan;
•safe, effective and medically necessary;
•appropriate for the specific patient;
•cost-effective; and
•neither experimental nor investigational.
Obtaining reimbursement approval for a product from each government or other third-party payer is a time consuming and costly process that could require us to provide supporting scientific, clinical and cost effectiveness data for the use of our products to each payer. We may not be able to provide data sufficient to gain acceptance with respect to reimbursement. Additionally, we might need to conduct post-marketing studies in order to demonstrate the cost-effectiveness of any future products to such payers’ satisfaction. Such studies might require us to commit a significant amount of management time and financial and other resources. Even when a payer determines that a product is eligible for reimbursement, the payer may impose coverage limitations that preclude payment for some uses that are approved by the FDA or non-United States regulatory authorities. Also, prior authorization for a product may be required. In addition, there is a risk that full reimbursement may not be available for high-priced products. Moreover, eligibility for coverage does not imply that any product will be reimbursed in all cases or at a rate that allows us to make a profit or even cover our costs. Interim payments for new products, if
48
applicable, may also not be sufficient to cover our costs and may not be made permanent. Further, coverage policies and third-party payer reimbursement rates may change at any time. Even if favorable coverage and reimbursement status is attained, less favorable coverage policies and reimbursement rates may be implemented in the future.
A primary trend in the United States healthcare industry and elsewhere is toward cost containment. We expect the changes made by PPACA, other legislation impacting the Medicare program and the 340B program, and the increasing emphasis on managed care to continue to put pressure on pharmaceutical product pricing. As these concerns continue to grow over the need for tighter oversight, there remains the possibility that the Heath Resources and Services Administration or another agency under the U.S. Department of Health and Human Services ("HHS") will propose regulations or that Congress will explore changes to the 340B program through legislation. There have also been a number of initiatives pending at the state and federal level that could negatively impact the reimbursement for products approved under the accelerated approval pathway in the United States by restricting patient access or establishing differential payment models. Certain states are also in the process of establishing Patient Drug Affordability Boards with the authority in some cases to set upper payment limits.
Further, there has been increasing legislative and enforcement interest in the United States with respect to drug pricing practices, including several recent U.S. congressional inquiries and federal and state legislation designed to, among other things, increase drug pricing transparency, expedite generic competition, review relationships between pricing and manufacturer patient assistance programs, and reform government program drug reimbursement methodologies. For example, HHS imposes rebates on many Medicare Part B and Medicare Part D products to penalize price increases that outpace inflation. In addition, HHS has been empowered to negotiate the price of certain single-source biologics that have been on the market for at least 11 years covered under Medicare as part of the Medicare Drug Price Negotiation Program. Each year up to twenty (20) products will be selected by HHS for the Medicare Drug Price Negotiation Program. Products subject to the Medicare Drug Price Negotiation Program are expected to experience a significant reduction in reimbursement from the Medicare program on a per unit basis. If coverage and adequate reimbursement are not available, or are available only to limited levels, we may not be able to successfully commercialize our current and any future product candidates that we develop, which could have an adverse effect on our operating results and our overall financial condition.
At the state level, legislatures are increasingly passing legislation and implementing regulations designed to control pharmaceutical and biological product pricing, including price or patient reimbursement constraints, discounts, restrictions on certain product access and marketing cost disclosure and transparency measures, and, in some cases, designed to encourage importation from other countries and bulk purchasing. For example, on June 15, 2026, the FDA approved Colorado’s Section 804 Importation Program (“SIP”) proposal to import certain drugs from Canada for specific state healthcare programs. It is unclear how this and Florida’s similar program, approved by the FDA in 2024, will be implemented and whether they will overcome potential legal, regulatory, or industry challenges in the United States and/or Canada. Any such approved importation plans, when implemented, may result in lower drug prices for products covered by those programs.
In addition, the current administration is pursuing policies to reduce regulations and expenditures across government including at the HHS, the FDA, Centers for Medicare & Medicaid Services (“CMS”) and related agencies and has made significant staff reductions at the FDA and other agencies. These actions, presently directed by executive orders or memoranda from the Office of Management and Budget, may propose policy changes that create additional uncertainty for our business. For example, the current administration has announced agreements with certain pharmaceutical companies that require the drug manufacturers to offer, through a direct-to-consumer platform, U.S. patients and Medicaid programs prescription drug Most-Favored Nation pricing equal to or lower than those paid in other developed nations, with additional mandates for direct-to-patient discounts and repatriation of foreign revenues. Other recent actions and proposals include for example, (1) directing agencies to reduce agency workforce and cut programs; (2) directing HHS and other agencies to lower prescription drug costs for Medicare through a variety of initiatives, including by improving upon the Medicare Drug Price Negotiation Program and establishing MFN pricing for pharmaceutical products; (3) imposing tariffs on certain imported pharmaceutical products; and (4) as part of the Make America Healthy Again Commission’s recent Strategy Report, working across government agencies to increase enforcement on direct-to-consumer pharmaceutical advertising. Additionally, the current administration recently called on Congress to enact “The Great Healthcare Plan,” to codify and expand Most-Favored Nation pricing, lower government subsidies to private insurance companies, increase healthcare price transparency, expand pharmaceutical drugs available for over-the-counter purchase, and enact restrictions on pharmacy benefit manager payment methodologies, among other things. These actions and policies may significantly reduce U.S. drug prices, potentially impacting manufacturers’ global pricing strategies and profitability, while increasing their operational costs and compliance risks. In June 2024, the U.S. Supreme Court’s Loper Bright decision greatly reduced judicial deference to regulatory agencies, which could increase successful legal challenges to federal regulations affecting our operations. Finally, Congress may introduce and ultimately pass health care related legislation that could impact the drug approval process and make changes to the Medicare Drug Price Negotiation Program. We cannot predict which additional measures may be adopted or the impact of current and additional measures on the marketing, pricing and demand for our products, which could have a material adverse effect on our business, financial condition and results of operations.
Any reduction in reimbursement from Medicare, Medicaid or other government-funded programs may result in a similar reduction in payments from private payers. The implementation of cost containment measures or other healthcare reforms may prevent us from being able to generate revenue, attain profitability or commercialize our therapies. Additionally, we are currently unable to predict what additional legislation or regulation, if any, relating to the healthcare industry may be enacted in the future or what effect recently enacted federal legislation or any such additional legislation or regulation would have on our business.
We face potential product liability exposure far in excess of our limited insurance coverage.
The use of any of our potential products in clinical trials, and the sale of any approved products, may expose us to liability claims. These claims might be made directly by consumers, health care providers, pharmaceutical companies or others selling our products. We have obtained limited product liability insurance coverage for our clinical trials in the amount of $10 million per occurrence and $35 million in the aggregate. However, our insurance may not reimburse us or may not be sufficient to reimburse us for any expenses or losses we may suffer. Moreover, insurance coverage is becoming increasingly expensive, and we may not be able to maintain insurance coverage at a reasonable cost or in sufficient amounts to protect us against losses due to liability. We intend to expand our insurance coverage as we obtain marketing approval for additional product candidates in development, but we may be unable to obtain commercially reasonable product liability insurance. On occasion, juries have awarded large judgments in class action lawsuits based on drugs that
49
had unanticipated side effects. A successful product liability claim or series of claims brought against us would decrease our cash reserves and could cause our stock price to fall.
We face substantial competition, which may result in others discovering, developing or commercializing products before or more successfully than we do. Our operating results will suffer if we fail to compete effectively.*
Several of our competitors have substantially greater financial, research and development, distribution, manufacturing and marketing experience and resources than we do and represent substantial long-term competition for us. Other companies may succeed in developing and marketing products that are more effective and/or less costly than any products that may be developed and marketed by us, or that are commercially accepted before or perceived as preferred relative to any of our products, or that obtain preferential formulary and reimbursement status. Factors affecting competition in the pharmaceutical and therapeutic industries vary, depending on the extent to which a competitor is able to achieve a competitive advantage based on its proprietary technology and ability to market and sell therapeutics. The industry in which we compete is characterized by extensive research and development efforts and rapid technological progress. In particular, the competitive landscape for IgAN is rapidly evolving and is expected to continue to evolve as multiple new modalities advance in development and begin to gain approval. While the competitive landscape for FSGS is limited at this point in time, we expect that it will evolve as other companies advance their programs toward potential approvals in the future. Furthermore, although we believe that our orphan drug status and proprietary position with respect to sparsentan may give us a competitive advantage, new developments are expected to continue and there can be no assurance that discoveries by others will not render our products and product candidates noncompetitive.
Furthermore, competitors could enter the market with generic versions of our products. For example, a generic option for the 100mg version of the original formulation of Thiola (tiopronin tablets) was approved by the FDA in May 2021 and a second 100mg version of the original formulation of Thiola (tiopronin tablets) was approved by the FDA in June 2022. In addition, several generic options for the 100mg and 300mg versions of Thiola EC have been approved by the FDA and become available.
Our competitive position also depends on our ability to enter into strategic alliances with one or more large pharmaceutical and contract manufacturing companies, attract and retain qualified personnel, develop effective proprietary products, implement development and marketing plans, obtain patent protection, secure adequate capital resources and successfully sell and market our approved products. There can be no assurance that we will be able to successfully achieve all of the foregoing objectives.
Use of third parties to manufacture our products and product candidates may increase the risk that we will not have sufficient quantities of our product and product candidates or such quantities at an acceptable cost, and clinical development and commercialization of our product and product candidates could be delayed, prevented or impaired.
We do not own or operate manufacturing facilities for clinical or commercial production of our products or product candidates. We have limited personnel with experience in drug manufacturing and we lack the resources and the capabilities to manufacture any of our product candidates on a clinical or commercial scale. We outsource all manufacturing and packaging of our nonclinical, clinical, and commercial products to third parties. The manufacture of pharmaceutical products in general, and biologics in particular, requires significant expertise and capital investment, including the development of advanced manufacturing techniques and process controls. Manufacturers of pharmaceutical products often encounter difficulties in production, particularly in scaling up initial production and in maintaining required quality control. These problems include difficulties with production costs and yields and quality control, including stability of the product candidate.
In September 2024, we announced a voluntary pause of enrollment in the Phase 3 HARMONY Study evaluating pegtibatinase for the treatment of classical HCU. The voluntary enrollment pause enabled us to address necessary process improvements in manufacturing scale-up to support the initial commercial scale manufacturing as well as full enrollment in the HARMONY Study. The voluntary enrollment pause was enacted following our determination that the desired drug substance profile was not achieved in the initial scale-up process. While we have made process improvements and have manufactured material at the larger scale, and while we restarted enrollment activities for the pivotal Phase 3 HARMONY Study in the first quarter of 2026, there is no guarantee that the Phase 3 HARMONY Study will be conducted or completed on the anticipated timeline or be successful. In addition, external factors including supply chain risks, geopolitical factors, and matters related to the staffing, resources and prioritization at the FDA and other government agencies, among others, could have impacts on our anticipated timeline.
We intend to rely on third-party manufacturers for the long-term commercial supply of FILSPARI and for our development stage product candidates. We expect the manufacturers of each product or product candidate to, at least initially and potentially for a significant period of time, be single source suppliers to us. Reliance on third-party manufacturers entails risks to which we may not be subject if we manufactured our product candidates or products ourselves, including:
•reliance on the third party for regulatory compliance and quality assurance;
•limitations on supply availability resulting from capacity and scheduling constraints of the third parties;
•less control over cost increases resulting from inflationary pressures affecting raw materials and other supply chain components;
•impact on our reputation in the marketplace if manufacturers of our products fail to meet the demands of our customers;
•the possible breach of the manufacturing agreement by the third party because of factors beyond our control; and
•the possible termination or nonrenewal of the agreement by the third party, based on its own business priorities, at a time that is costly or inconvenient for us.
The failure of any of our contract manufacturers to maintain high manufacturing standards could result in injury or death of clinical trial participants or patients using our products. Such failure could also result in product liability claims, product recalls, product seizures or withdrawals, delays or failures in testing or delivery, cost overruns or other problems that could seriously harm our business or profitability.
50
Our contract manufacturers are required to adhere to FDA regulations setting forth cGMP and comparable foreign regulatory authority requirements. These regulations cover all aspects of the manufacturing, testing, quality control and recordkeeping relating to our product candidates and any products that we may commercialize. Our manufacturers may not be able to comply with cGMP regulations or similar regulatory requirements outside the United States. Our manufacturers are subject to unannounced inspections by the FDA, state regulators and similar regulators outside the United States to monitor and ensure compliance with cGMP. We are ultimately responsible for ensuring that our API and finished products are manufactured in accordance with cGMP regulations and similar regulatory requirements outside the United States, and it is therefore critical that we maintain effective management practices and oversight with respect to our third-party manufacturers, including routine auditing. Our failure, or the failure of our third-party manufacturers, to comply with applicable regulations could result in sanctions being imposed on us, including shutdown of the third-party vendor or invalidation of drug product lots or processes, fines, injunctions, civil penalties, failure of regulatory authorities to grant marketing approval of our product candidates, delays, suspension, variation or withdrawal of approvals, license revocation, seizures or recalls of product candidates or products, operating restrictions and criminal prosecutions, any of which could significantly and adversely affect regulatory approval and supplies of our product candidates.
Our product and any products that we may develop may compete with other product candidates and products for access to manufacturing facilities. There are a limited number of manufacturers that operate under cGMP regulations and that are both capable of manufacturing for us and willing to do so. A health epidemic or pandemic and associated vaccine or treatment development and manufacturing efforts may increase demand for the services supplied by many third-party manufacturers, including some of those that we utilize for our products and product candidates, which may result in decreased availability of manufacturing slots at many such facilities. If the third parties that we engage to manufacture products for our developmental or commercial products should halt or cease to continue to do so for any reason, we likely would experience interruptions in cash flows and/or delays in advancing our clinical trials while we identify and qualify replacement suppliers, and we may be unable to obtain replacement supplies on terms that are favorable to us. Later relocation to another manufacturer will also require notification, review and other regulatory approvals from the FDA and other regulators and will subject our production to further cost and instability in the availability of our product candidates. In addition, if we are not able to obtain adequate supplies of our products and product candidates, or the drug substances used to manufacture them, it will be more difficult for us to sell our products and to develop our product candidates. This could greatly reduce our competitiveness and negatively affect our results of operations.
Our current and anticipated future dependence upon others for the manufacture of our products and product candidates may adversely affect our future profit margins and our ability to develop product candidates and commercialize our marketed products and any other products that may obtain regulatory approval on a timely and competitive basis.
Materials necessary to manufacture our products and product candidates may not be available on commercially reasonable terms, or at all, which may delay the development and commercialization of our products and product candidates.*
We rely on the manufacturers of our products and product candidates to purchase from third-party suppliers the materials necessary to produce the compounds or biologic substances used in our nonclinical and clinical studies and for the commercial supply of our products. Suppliers may not sell these materials to our manufacturers at the time we need them or on commercially reasonable terms and all such prices are susceptible to fluctuations in price and availability due to transportation costs, government regulations, price controls, and changes in economic climate or other foreseen circumstances. We do not have any control over the process or timing of the acquisition of these materials by our manufacturers. In addition, inflation and global supply chain disruptions, as well as past disruptions related to COVID-19 and potential future disruptions related to a future health epidemic or pandemic, wars, armed conflicts, tariffs and global geopolitical tension, including between the U.S. and China, have had and may continue to have a negative impact on our manufacturers’ ability to acquire the materials necessary for our business. If our manufacturers are unable to obtain these materials, our nonclinical and clinical studies, product testing, regulatory activities, manufacturing and commercial supply of our products and product candidates could be delayed or interrupted. If our manufacturers or we are unable to purchase these materials for our commercial products or after regulatory approval has been obtained for any of our product candidates, commercial supply could be interrupted, commercial launches could be delayed, or shortages could occur, which could materially affect our ability to meet clinical trial timelines, and/or generate revenues from the sale of our products . For example, in 2021 a membrane used in pegtibatinase drug substance manufacturing became more difficult to acquire due to the same or similar membranes being used in certain of the COVID-19 vaccine manufacturing processes. We also may encounter manufacturing process or supply challenges. For example, in September 2024, we announced a voluntary pause of enrollment in the Phase 3 HARMONY Study to enable us to address necessary process improvements in manufacturing scale-up; these process improvements have been achieved and enrollment has restarted. From time to time we continue to, and may in the future, face supply challenges or shortages of other materials necessary to manufacture pegtibatinase or our other products and product candidates. If our risk mitigation plans are not successful in overcoming these challenges, our pegtibatinase program or other products and product candidates, could be delayed.
Changes in legislation could also potentially impact our ability to secure the materials we need for our products and product candidates. For example, the United States passed legislation in December 2025, namely the BIOSECURE Act (the “BIOSECURE Act”), to prohibit U.S. federal executive agencies from procuring or obtaining any biotechnology equipment or service produced or provided by a “biotechnology company of concern” or entering into or renewing a contract, loan, or grant with an entity that uses such biotechnology equipment or services. The BIOSECURE Act prohibits the U.S. government from procuring or obtaining biotechnology equipment or services produced or provided by a “biotechnology company of concern” (“BCC”); entering into, extending, or renewing government contracts with an entity that directly or indirectly uses biotechnology equipment or services from a BCC in performance of that federal contract; and/or issuing grants or loans to purchase, obtain, or use biotechnology equipment or services produced by a BCC. The BIOSECURE Act also prohibits U.S. government loan and grant recipients from using federal loan or grant money to enter into contracts with entities that use equipment from BCCs in the performance of any federal prime contract or subcontract. Companies designated as a BCC include those that are identified on the U.S. Department of Defense’s annual List of Chinese Military Companies, also known as the 1260H List, and the U.S. government also has the ability to designate entities as BCCs through a separate designation process. There is a “safe harbor” provision providing that the restrictions do not apply to equipment or services that were formerly but are no longer provided by a BCC, as well as a “grandfathering” provision providing that the prohibitions shall not apply for a five-year period to biotechnology equipment or services produced or provided under a contract or agreement entered into before the applicable effective date. Given the BIOSECURE Act, we may be restricted in our ability to work with certain Chinese biotechnology companies to the extent we would contract with, or otherwise receive funding from, the U.S. government. In June 2026, the Section 1260H list was expanded to include our current primary API supplier for sparsentan, and, if the required implementing agency actions are completed as currently anticipated, our supplier would become subject to the BIOSECURE Act's restrictions. As part of our regular supply continuity planning, we have taken steps over the past several years to identify, secure and
51
qualify alternative sources of sparsentan API; however, there can be no assurance that any alternative source will be successfully qualified on a timely basis, obtain any necessary regulatory approvals, manufacture API at commercial scale, or supply API in sufficient quantities or on commercially reasonable terms. We may ultimately be required, or choose to transition away from our current primary API supply relationship for sparsentan if and when the BIOSECURE Act's restrictions become applicable following completion of the required implementing agency actions and prior to or upon the expiration of the expected five-year grandfathering period. Any such transition could increase costs or adversely affect the commercial supply of sparsentan if we are unable to complete it successfully and on a timely basis. In addition, BCC designations, implementing regulations or other changes under the BIOSECURE Act could affect supply arrangements for certain of our other products or product candidates. As a result, we or our licensors, manufacturers or other supply chain partners may be required to identify and qualify alternative suppliers, which could increase costs or delay development, manufacturing or commercialization activities. For certain products and product candidates, there may be a limited number of suppliers and manufacturers with the required technical capabilities, manufacturing capacity or regulatory qualifications.
Risks Related to Our Business
International trade policies, including tariffs, sanctions and trade barriers may adversely affect our business, financial condition, results of operations and prospects.*
We operate in a global economy, and our business depends on a global supply chain for the development, manufacturing, and distribution of our pharmaceutical products, and for the advancement of our pre/nonclinical and clinical development programs. There is inherent risk, based on the complex relationships among the U.S. and the countries in which we conduct our business, that political, diplomatic, and national security factors can lead to global trade restrictions and changes in trade policies and export regulations that may adversely affect our business and operations. The current international trade and regulatory environment is subject to significant ongoing uncertainty.
We source quantities of active pharmaceutical ingredients ("APIs"), precursor chemicals, and specialized equipment from international suppliers, including from manufacturers in China, consistent with broader industry practices. While the impact of tariff policies on our business has been minimal to date, current or future tariff policies, particularly those affecting China and pharmaceutical products, could further increase our costs, and may affect profitability, particularly in formulary-based markets where pricing flexibility may be limited. Recent and potential future changes in international trade policies, particularly regarding U.S.-China trade relations and pharmaceutical-specific tariffs, present potential risks to our future operations and financial performance.
The ongoing trade tensions between the United States and other countries including China have resulted in multiple rounds of tariffs affecting pharmaceutical ingredients, manufacturing equipment, and related supplies. The evolving tariff and trade landscape contributes to planning challenges for global pharmaceutical operations. Changes in tariff classifications, country-of-origin requirements, or customs procedures can occur with limited notice. This uncertainty complicates our long-term investment decisions regarding manufacturing facilities, supply chain optimization, and research and development locations.
Recent policy discussions have included potential targeted tariffs or other trade measures specifically aimed at pharmaceutical products and ingredients as part of broader healthcare cost control or national security initiatives. The Bureau of Industry and Security, U.S. Department of Commerce, initiated an investigation to determine whether pharmaceutical ingredients, including finished drug product, manufactured outside the United States pose a national security risk and should be subject to additional tariffs. Following that investigation, the President announced a proclamation which will impose 100% tariffs on certain patented pharmaceutical products and associated pharmaceutical ingredients. Although we expect FILSPARI to be eligible for the 0% tariff rate for orphan drugs under the proclamation, the application of that rate is subject to the terms of the proclamation and future agency determinations, and we will continue to monitor and assess the evolving tariff landscape on our business.
Pharmaceuticals and biologics face regulatory and technical constraints that make rapid supply chain adjustments challenging, complex and costly. Identifying and qualifying a new alternative supplier with available capacity and capabilities— whether in the U.S. or in another country with a more favorable tariff regime—requires a substantial monetary investment and investment of personnel and other resources, including those related to contracting, qualification, technology transfer, and regulatory approvals, and the process may take an extended period of time to complete.
While the impact of tariff policies on our business has been minimal to date, current or future tariffs may result in increased research and development expenses, including with respect to raw materials, APIs, laboratory equipment and research materials and components. Trade restrictions affecting the import of materials necessary for clinical trials could result in delays to development timelines. Increased costs and extended development timelines could affect competitiveness relative to companies operating in regions with more favorable trade relationships and could impact investor confidence.
The complexity of announced or future tariffs may also increase the risk of enforcement actions related to trade compliance. Foreign governments may adopt non-tariff measures, such as procurement preferences or informal disincentives to engage with, purchase from or invest in U.S. entities. These developments could affect our ability to compete internationally or engage with global suppliers, customers and partners. Retaliatory actions, such as changes to intellectual property protection, increased enforcement, or delays in regulatory approvals, could result in legal and operational risks.
In addition, the United States and other governments have imposed and may continue to impose additional sanctions, such as trade restrictions or trade barriers, which could restrict us from doing business directly or indirectly in or with certain countries or parties and may impose additional costs and complexity to our business.
Trade disputes, tariffs, restrictions and other political tensions between the United States and other countries may also exacerbate unfavorable macroeconomic conditions including inflationary pressures, foreign exchange volatility, financial market instability, and economic recessions or downturns. The ultimate impact of current or future tariffs and trade restrictions remains uncertain. While we actively monitor these risks and manage our supply chain accordingly, prolonged economic or geopolitical disruptions could adversely affect our business, ability to access the capital markets or other financing sources, results of operations, financial condition and prospects. In addition, tariffs and other trade developments have and may continue to heighten the risks related to the other risk factors described elsewhere in this report.
52
Many of the trade, tariff, pricing, and related policy actions described herein remain subject to significant uncertainty, including with respect to their scope, duration, implementation, and enforceability. Such measures may be modified, delayed, suspended, repealed, or invalidated through administrative action, changes in policy priorities, or judicial challenges, and it is uncertain whether, when, or to what extent any such measures will ultimately be implemented or apply to our products, suppliers, or operations. Although we have taken and may continue to take certain actions intended to reduce potential exposure to supply chain or cost impacts, including efforts to diversify suppliers, there can be no assurance that such actions will be effective, timely, or sufficient to mitigate any adverse effects arising from these policies.
Our limited operating history makes it difficult to evaluate our future prospects, and our profitability in the future is uncertain.
We face the problems, expenses, difficulties, complications and delays, many of which are beyond our control, associated with any business in its early stages and have a limited operating history on which an evaluation of our prospects can be made. Such prospects should be considered in light of the risks, expenses and difficulties frequently encountered in the establishment of a business in a new industry, characterized by a number of market entrants and intense competition, and in the shift from development to commercialization of new products based on innovative technologies.
We have experienced significant growth over the past five years in the number of our employees and the scope of our operations. We have expanded our sales and marketing, compliance and legal functions in addition to expansion of all functions to support a commercial organization. We have also expanded our operations in connection with the commercial launch of FILSPARI in the United States, including by adding additional members to our sales force. To appropriately manage for our future, we must continue to implement and improve our managerial, operational and financial systems, continue to recruit, train and retain qualified personnel as needed, and successfully integrate any changes into our existing business. To succeed, we must recruit, retain, manage and motivate qualified clinical, scientific, technical, commercial and management personnel, and we face significant competition for experienced personnel.
Due to our limited resources, we may not be able to effectively manage the expansion of our operations or recruit, train and retain qualified personnel, including in connection with the ongoing commercial launch of FILSPARI in the United States. The management of changes to our operations may lead to significant costs and may divert our management and business development resources. Any inability on the part of our management to manage growth or other changes in our organization could delay the execution of our business plans or disrupt our operations.
Factors that may inhibit our efforts to commercialize our products without strategic partners or licensees include:
•our inability to recruit and retain adequate numbers of effective sales and marketing personnel;
•the inability of sales personnel to obtain access to or educate adequate numbers of physicians to prescribe our products;
•the lack of complementary products to be offered by our sales personnel, which may put us at a competitive disadvantage against companies with broader product lines;
•unforeseen costs associated with expanding our own sales and marketing team for new products or with entering into a partnering agreement with an independent sales and marketing organization; and
•efforts by our competitors to commercialize competitive products.
Moreover, though we generate revenues from product sales arrangements, we may incur significant operating losses over the next several years. Our ability to achieve profitable operations in the future will depend in large part upon successful in-licensing of products approved by the FDA, selling and manufacturing these products, completing development of our products, obtaining regulatory approvals for these products, and bringing these products to market. The likelihood of the long-term success of our company must be considered in light of the expenses, difficulties and delays frequently encountered in the development and commercialization of new therapeutics, competitive factors in the marketplace, as well as the regulatory environment in which we operate.
In addition, we may encounter unforeseen expenses, difficulties, complications, delays and other known and unknown factors.
We depend on a highly experienced and skilled workforce to grow and operate our business. If we are unable to attract, retain and engage our employees, we may not be able to grow effectively.
The execution of our strategic objectives and future success will depend upon our continued ability to identify, hire, develop, motivate and retain a highly qualified workforce. We depend on contributions from our employees, and, in particular, our senior management team, to execute efficiently and effectively. Our success further depends on our ability to attract, retain and motivate highly skilled mid-level and senior managers as well as team members at various levels in the scientific, development, medical and commercial areas of the business, particularly in connection with our ongoing commercial launch of FILSPARI in the United States.
Our headquarters are based in San Diego, California. This region is home to many other biopharmaceutical companies and many academic and research institutions. Competition for qualified key talent in our market is intense and may limit our ability to hire and retain employees on acceptable terms, or at all. As a result, we may not be able to retain our existing employees or hire new employees quickly enough to meet our needs.
To induce valuable employees to remain at our company, in addition to salary, cash incentives and other employee benefits, we have provided stock options and restricted stock unit ("RSU") awards that vest over time. The value to employees of stock options and RSU awards that vest over time may be significantly affected by movements in our stock price that are beyond our control and may at any time be insufficient to counteract more lucrative offers from other companies. Current market conditions and the potential for extreme stock price volatility exacerbates this risk. Despite our efforts to retain valuable employees, members of our management, scientific, development and commercial teams may terminate their employment with us on short notice. All of our employees have at-will employment, which means that they could leave our employment at any time, with or without notice. We do not maintain “key person” insurance policies on the lives of any of our employees.
If we fail to effectively manage our hiring and retention needs, our ability to meet our strategic objectives and our business and operating results may be adversely impacted.
53
Health epidemics or pandemics could materially adversely affect our business, results of operations and financial condition.
A health epidemic or pandemic poses the risk that we or our clinical trial subjects, employees, contractors, collaborators, suppliers and vendors may be prevented from conducting certain clinical trials or other business activities for an indefinite period of time, including due to travel restrictions, quarantines, “stay-at-home” and "shelter-in-place" orders or shutdowns that have been or may be requested or mandated by governmental authorities, or that our or their ability to conduct operations will be negatively impacted by staffing shortages while employees quarantine as a result of exposure to or transmission of the virus. In addition, a health epidemic or pandemic could impact personnel at third-party manufacturing facilities in the United States and other countries, including China, or the availability or cost of materials, which could potentially disrupt the supply chain for our commercial products, our product candidates or the comparator products in our ongoing clinical trials.
The timelines and conduct of our ongoing clinical trials previously have been affected by COVID-19 and we may experience similar delays or interruptions due to other health epidemics or pandemics in the future. For example, in 2020 we experienced a reduction in the rates of patient enrollment in our ongoing clinical trials as a result of the COVID-19 pandemic. New health epidemics or pandemics may emerge that result in similar or more severe disruptions to our business, which could adversely impact our business and operating results.
We will likely experience fluctuations in operating results and could incur substantial losses.*
We expect that our operating results will vary significantly from quarter-to-quarter and year-to-year as a result of investments in research and development, specifically our clinical and nonclinical development activities. We anticipate that certain of our expenses will continue to increase, depending on factors including but not limited to: the continuation and cost of our clinical trials and the research and development of additional product candidates; the costs involved in seeking and obtaining marketing approvals for our products, and in maintaining quality systems standards for our products; the timing of, and costs involved in, commercial activities, including product marketing, sales and distribution, costs related to our operational, financial, and management information systems and personnel, including personnel to support product development efforts and our obligations as a public company.
To attain and sustain profitability, we must succeed in developing and commercializing therapies with significant market potential. This will require us to be successful in a range of challenging activities, including the discovery of product candidates, successful completion of nonclinical testing and clinical trials of our product candidates, obtaining regulatory approval for these product candidates and manufacturing, marketing and selling those products for which we may obtain regulatory approval. We may not be successful enough in these activities to generate revenues that are substantial enough to recoup the expenses we have expended in conducting these activities to achieve profitability. Pursuant to the Ligand License Agreement, we are obligated to pay to Ligand an escalating annual royalty between 15% and 17% of net sales of FILSPARI and any other products containing sparsentan or related compounds, which will impact our potential future profit from the commercialization of FILSPARI in the United States and other parts of the world where it is approved or may be approved in the future. Even if we do achieve profitability, we may not be able to sustain or increase profitability on a quarterly or annual basis. Our failure to become or remain profitable could depress the market price of our common stock and could impair our ability to raise capital, expand our business, diversify our product offerings or continue our operations. A decline in the market price of our common stock may also cause a loss of a part or all of your investment.
Negative publicity regarding any of our products could impair our ability to market any such product and may require us to spend time and money to address these issues.
If any of our products or any similar products distributed by other companies prove to be, or are asserted to be, harmful to consumers and/or subject to FDA or comparable foreign regulatory authority enforcement action, our ability to successfully market and sell our products could be impaired. Because of our dependence on patient and physician perceptions, any adverse publicity associated with illness or other adverse effects resulting from the use or misuse of our products or any similar products distributed by other companies could limit the commercial potential of our products and expose us to potential liabilities.
We may not have sufficient insurance to cover our liability in any current or future litigation claims either due to coverage limits or as a result of insurance carriers seeking to deny coverage of such claims.
We face a variety of litigation-related liability risks. Our certificate of incorporation, bylaws, other applicable agreements, and/or Delaware law require us to indemnify (and advance expenses to) our current and past directors and officers and employees from reasonable expenses related to the defense of any action arising from their service to us, including circumstances under which indemnification is otherwise discretionary. While our directors and officers are included in a director and officer liability insurance policy ("D&O insurance"), which covers all our directors and officers in some circumstances, our insurance coverage does not cover all of our indemnification obligations and may not be adequate to cover any indemnification or other claims against us. In addition, the underwriters of our present coverage may seek to avoid coverage in certain circumstances based upon the terms of the respective policies. If we incur liabilities that exceed our coverage under our D&O insurance or incur liabilities not covered by our insurance, we would have to self-fund any indemnification amounts owed to our directors and officers and employees in which case our results of operations and financial condition could be materially adversely affected. Further, if D&O insurance becomes prohibitively expensive to maintain in the future, we may be unable to renew such insurance on economic terms or unable to renew such insurance at all. The potential lack of D&O insurance may make it difficult for us to retain and attract talented and skilled directors and officers to serve our company, which could adversely affect our business.
We may need substantial funding and may be unable to raise capital when needed.*
We expect our general and research and development expenses to increase in connection with our ongoing and planned activities, particularly as we conduct later-stage clinical trials of our product candidates. In addition, in connection with the commercial launch of FILSPARI in the United States, we have begun to incur significant commercialization expenses and expect to continue to incur significant commercialization expenses for FILSPARI and any other future approved products, including for product sales and marketing, securing commercial quantities of product from our manufacturers, and product distribution. Our expenses have and may continue to increase as a result of inflation in the United States and abroad. We currently have no additional commitments or arrangements for any additional financing to fund the research and development and commercial launch of our product candidates. General market conditions, including high interest rates and stock price volatility, actual or anticipated bank failures, new or increased tariffs, and ongoing issues
54
arising global geopolitical tensions, including the wars and other armed conflicts, as well as market conditions affecting companies in the life sciences industry in general, may make it difficult for us to seek financing from the capital markets on attractive terms, or at all.
Management believes our ability to continue our operations depends on our ability to sustain and grow revenue, results of operations and our ability to access capital markets when necessary to accomplish our strategic objectives. Management believes that we may incur losses in the immediate future. We expect that our operating results will vary significantly from quarter-to-quarter and year-to-year as a result of investments in research and development, specifically our clinical and nonclinical development activities. We expect to finance our cash needs from cash on hand and results of operations, and depending on results of operations we may either need additional equity or debt financing, or need to enter into strategic alliances on products in development to continue our operations until we can achieve sustained profitability and positive cash flows from operating activities. Additional funds may not be available to us when we need them on terms that are acceptable to us, or at all. If adequate funds are not available to us on a timely basis, we may be required to reduce or eliminate research development programs or commercial efforts.
Our future capital requirements will depend on many factors, including:
•the timing, progress, cost and results of our clinical trials, preclinical studies and other discovery and research and development activities;
•the timing of, and costs involved in, seeking and obtaining marketing approvals for our products, and in maintaining quality systems standards for our products;
•the timing of, and costs involved in, commercial activities, including product marketing, sales and distribution;
•our ability to successfully commercialize FILSPARI, and to obtain regulatory approval for, and successfully commercialize, our other or future product candidates;
•increases or decreases in revenue from our marketed products, including decreases resulting from generic entrants or health epidemics or pandemics;
•debt service obligations on the 2029 Notes and 2032 Notes;
•the number and development requirements of other product candidates that we pursue;
•our ability to manufacture sufficient quantities of our products to meet expected demand;
•the costs of preparing, filing and prosecuting patent applications and maintaining, enforcing and defending intellectual property related claims;
•our ability to enter into collaboration, licensing or distribution arrangements and the terms and timing of these arrangements;
•the potential need to expand our business, resulting in additional payroll and other overhead expenses;
•the potential in-licensing of other products or technologies;
•the emergence of competing products and technologies and other adverse market developments;
•the extent to which we acquire or invest in businesses, products and technologies; and
•the potential impacts of inflation and resulting cost increases.
The market price for shares of our common stock may be volatile and purchasers of our common stock could incur substantial losses.
The price of our stock is likely to be volatile. The stock market in general, and the market for biotechnology companies in particular, have experienced extreme volatility that has often been unrelated to the operating performance of particular companies. The market price for our common stock has been in the past, and may be in the future, influenced by many factors, including:
•results of clinical trials of our product candidates or those of our competitors;
•our entry into or the loss of a significant collaboration;
•regulatory or legal developments in the United States and other countries, including changes in the health care payment systems;
•our ability to obtain and maintain marketing approvals from the FDA or similar regulatory authorities outside the United States;
•variations in our financial results or those of companies that are perceived to be similar to us;
•changes in the structure of healthcare payment systems;
•market conditions in the pharmaceutical and biotechnology sectors and issuance of new or changed securities analysts’ reports or recommendations;
•general economic, industry and market conditions, including the impacts thereon of inflation and high interest rates, actual or anticipated bank failures, new or increased tariffs, wars, armed conflicts and global geopolitical tensions;
•results of clinical trials conducted by others on therapies that would compete with our product candidates;
•developments or disputes concerning patents or other proprietary rights;
•public concern over our product candidates or any products approved in the future;
•litigation;
55
•communications from government officials regarding health care costs or pharmaceutical pricing;
•future sales or anticipated sales of our common stock by us or our stockholders; and
•the other factors described in this “Risk Factors” section.
In addition, the stock markets, and in particular, the Nasdaq Stock Market, have experienced extreme price and volume fluctuations that have affected and continue to affect the market prices of equity securities of many pharmaceutical companies. The realization of any of the above risks or any of a broad range of other risks, including those described in these “Risk Factors” could have a dramatic and material adverse impact on the market price of our common stock.
We may not receive some or all of the potential milestone and/or royalty payments from our corporate and licensing transactions.*
From time to time, we engage in corporate transactions and licensing transactions that include potential milestone payments and/or royalties. For example, on July 16, 2023, we entered into a definitive asset purchase agreement (the "Purchase Agreement") with Mirum Pharmaceuticals, Inc. ("Mirum"), pursuant to which we agreed to sell to Mirum, subject to the terms of the Purchase Agreement, our bile acid product portfolio including Chenodal and Cholbam (also known as Kolbam) (the "Products"). The closing of the transaction occurred on August 31, 2023. A portion of the consideration for the sale is in the form of milestone payments that will only be payable upon the achievement of certain milestones based on specified amounts of annual net sales of the Products. We are also party to license agreements with CSL Vifor and Chugai pursuant to which we are entitled to receive certain payments contingent on the future achievement of specified milestones, and royalty payments based on potential future sales in specified licensed territories. There is a risk that any or all of the milestone events under these various agreements might not be achieved, that our licensees may not achieve sales that would entitle us to royalty payments, and that any or all of the consideration tied to the achievement of the milestone events and/or royalties might not be received.
In the fourth quarter of 2025, our partner Renalys was acquired by and merged into Chugai, and as a result of the acquisition, Chugai gained exclusive rights to develop and commercialize sparsentan in Japan, South Korea, and Taiwan. Travere received a portion of the upfront payment due to the fact that it was a minority shareholder in Renalys prior to the acquisition, and Travere is also eligible to receive future payments upon the achievement of specified regulatory milestones for sparsentan and royalties on net sales in Japan, South Korea, and Taiwan. There is no guarantee that we will receive any or all of the consideration that is tied to the regulatory milestones or net sales.
We may be unable to successfully integrate new products or businesses we may acquire.*
We may in the future expand our product pipeline by pursuing acquisition of pharmaceutical products. For example, in June 2026, we signed a license and collaboration agreement (the “Everest Agreement”) with Everest Medicines (Singapore) Pte. Ltd. (“Everest”), pursuant to which Everest granted us an exclusive license for the development and commercialization of civorebrutinib and products containing civorebrutinib for any and all prophylactic, diagnostic and therapeutic uses and treatment of diseases and disorders in the U.S. and global markets excluding Greater China and certain countries in East and Southeast Asia. The Everest Agreement became effective in July 2026.
If an acquisition is consummated, the integration of the acquired business, product or other assets into our company may also be complex and time-consuming and, if such businesses, products and assets are not successfully integrated, we may not achieve the anticipated benefits, cost-savings or growth opportunities. Potential difficulties that may be encountered in the integration process include the following:
•integrating personnel, operations and systems, while maintaining focus on producing and delivering consistent, high quality products;
•coordinating geographically dispersed organizations;
•distracting employees from operations;
•retaining existing customers and attracting new customers; and
•managing inefficiencies associated with integrating the operations of the acquired company or product into our own operations.
Furthermore, these acquisitions and other arrangements, even if successfully integrated, may fail to further our business strategy as anticipated, expose us to increased competition or challenges with respect to our products or geographic markets, and expose us to additional liabilities associated with an acquired business, product, technology or other asset or arrangement. Any one of these challenges or risks could impair our ability to realize any benefit from our acquisitions or arrangements after we have expended resources on them.
Our ability to realize the anticipated benefits of in-licensed products and collaboration arrangements depends on several factors, many of which are outside of our control.*
We have entered into, and may in the future enter into, in-license, collaboration and other strategic agreements to acquire rights to develop and commercialize product candidates. For example, in June 2026 we entered into an exclusive license and collaboration agreement with Everest Medicines for the development and commercialization of civorebrutinib. Our ability to realize the anticipated benefits of these types of arrangements depend on a number of factors, many of which are outside of our control.
Under the terms of these agreements, we may rely on our licensors, collaborators or other counterparties to perform obligations that are important to the successful development and commercialization of licensed product candidates, including technology transfer activities, manufacturing support, the conduct of certain development activities, regulatory support and other responsibilities allocated to such parties. These agreements are often complex and require ongoing coordination among the parties. As a result, disagreements may arise regarding the interpretation of contractual terms, the parties' respective rights and obligations, development strategy or execution, intellectual property matters or differing business priorities. Delays or failures by these counterparties in performing their obligations, disagreements regarding development strategy or execution, differing business priorities, changes in a counterparty's financial condition or strategic focus, or other disputes between the parties could delay development, regulatory submissions or commercialization of licensed product candidates, increase our costs or otherwise adversely affect the value of the applicable license or collaboration. For example, under our agreement with Everest, we rely on Everest to perform certain technology transfer, development and other obligations with respect to civorebrutinib.
56
In addition, our rights to in-licensed products and product candidates are subject to the terms and conditions of the applicable license or collaboration agreements and may be subject to additional upstream license agreements. Accordingly, our rights may depend, in part, on agreements to which we are not a party and over which we have limited or no control. Disputes involving our licensors and their upstream licensors, or differing interpretations of applicable agreements, could adversely affect the scope or availability of rights that are important to our business. Furthermore, our ability to fully exploit these products and product candidates may be subject to contractual restrictions, diligence obligations, milestone and royalty requirements and other contractual provisions, including developmental cost-sharing obligations and reimbursements, and our failure, or our counterparty's failure, to comply with applicable contractual obligations could adversely affect our rights under these arrangements. If any such agreement is modified or terminated, whether as a result of a material breach, insolvency, failure to satisfy contractual obligations or otherwise, we could lose some or all of our rights to the applicable product or product candidate, be required to transfer or provide rights to certain regulatory filings or other assets developed under the license or collaboration, lose the benefit of investments we have made in the program, or incur additional costs in pursuing alternative development or licensing arrangements
Product liability lawsuits against us could cause us to incur substantial liabilities and to limit commercialization of any products that we may develop.
Our business exposes us to potential liability risks inherent in the research, development, manufacturing and marketing of pharmaceutical products. If any of our product candidates in clinical trials or commercialized products harm people, we may be subject to costly and damaging product liability claims. We have clinical trial insurance and commercial product liability coverage. However, this insurance may not be adequate to cover all claims. We may be exposed to product liability claims and product recalls, including those which may arise from misuse or malfunction of, or design flaws in, such products, whether or not such problems directly relate to the products we have provided. If we cannot successfully defend ourselves against claims that our product candidates or products caused injuries, we will incur substantial liabilities. Regardless of merit or eventual outcome, liability claims may result in:
•decreased demand for any product candidates or products that we may develop;
•damage to our reputation;
•regulatory investigations that could require costly recalls or product modifications;
•withdrawal of clinical trial participants;
•costs to defend the related litigation;
•substantial monetary awards to trial participants or patients, including awards that substantially exceed our product liability insurance, which we would then be required to pay from other sources, if available, and would damage our ability to obtain liability insurance at reasonable costs, or at all, in the future;
•loss of revenue;
•the diversion of management’s attention from managing our business; and
•the inability to commercialize any products that we may develop.
A successful product liability claim or a series of claims brought against us could cause our stock price to fall and, if judgments exceed our insurance coverage, could decrease our available cash and adversely affect our business.
We may become involved in litigation matters, which could result in substantial costs, divert management's attention and otherwise have a material adverse effect on our business, operating results or financial condition.
From time to time we may become involved in certain litigation matters, including those described in Note 13 of the Consolidated Financial Statements included in this report. Although we intend to vigorously defend our interests in each matter, there is no guarantee that we will be successful and we may have to pay damages awards or otherwise may enter into settlement arrangements in connection with such matters. Any such payments or settlement arrangements could have material adverse effects on our business, operating results or financial condition. Even if we are successful in defending our interests in each matter, litigation with respect to such matters could result in substantial costs and significant adverse impact on our reputation and divert management's attention and resources, which could have a material adverse effect on our business, operating results or financial condition.
We are subject to significant ongoing regulatory obligations and oversight, which may result in significant additional expense and may limit our commercial success.*
We are subject to significant ongoing regulatory obligations, such as safety reporting requirements and additional post-marketing obligations, including regulatory oversight of the promotion and marketing of our products. In addition, the manufacture, quality control, labeling, packaging, safety surveillance, adverse event reporting, storage and recordkeeping for our products are subject to extensive and ongoing regulatory requirements. If we become aware of previously unknown problems with any of our products, a regulatory authority may impose restrictions on our products, our contract manufacturers or us. If we, our products and product candidates, or the manufacturing facilities for our products and product candidates fail to comply with applicable regulatory requirements, a regulatory authority, including the FDA, may send enforcement letters, mandate labeling changes, suspend, vary or withdraw regulatory approval, suspend, vary or terminate any ongoing clinical trials, refuse to approve pending applications or supplements filed by us, suspend or impose restrictions on manufacturing operations, request a recall of, seize or detain a product, seek criminal prosecution or an injunction, or impose civil or criminal penalties or monetary fines. In such instances, we could experience a significant drop in the sales of the affected products, our product revenues and reputation in the marketplace may suffer, and we could become the target of lawsuits.
We are also subject to regulation by supranational, national, regional, state and local agencies and regulatory authorities, including but not limited to the FDA, CMS, Department of Justice, the Federal Trade Commission, the HHS Office of Inspector General and other regulatory bodies. The FDC Act, Social Security Act, Public Health Service Act and other federal and state statutes and regulations, and comparable foreign regulatory acts, govern to varying degrees the research, development, manufacturing and commercial activities relating to prescription pharmaceutical products, including nonclinical testing, clinical
57
research, approval, production, labeling, sale, distribution, post-market surveillance, advertising, dissemination of information, promotion, marketing, and pricing to government purchasers and government health care programs. Our manufacturing partners are subject to many of the same requirements.
Companies may not promote drugs for “off-label” uses—that is, uses that are not described in the product’s labeling and that differ from those approved by the FDA or other applicable regulatory authorities. However, a company may share truthful and not misleading information that is otherwise consistent with the product’s labeling. A company that is found to have improperly promoted off-label uses may be subject to significant liability, including civil and administrative remedies as well as criminal sanctions. In addition, management’s attention could be diverted from our business operations and our reputation could be damaged.
The federal health care program Anti-Kickback Statute prohibits, among other things, knowingly and willfully offering, paying, soliciting, or receiving remuneration to induce or in return for purchasing, leasing, ordering or arranging for the purchase, lease or order of any health care item or service reimbursable under Medicare, Medicaid or other federally financed healthcare programs. This statute has been interpreted broadly to apply to arrangements that pharmaceutical companies have with prescribers, purchasers and formulary managers, among others. Further, the PPACA, among other things, amends the intent requirement of the federal Anti-Kickback Statute so that a person or entity no longer needs to have actual knowledge of this statute or specific intent to violate it. In addition, the PPACA provides that the government may assert that a claim including items or services resulting from a violation of the federal Anti-Kickback Statute constitutes a false or fraudulent claim for purposes of the civil False Claims Act. Although there are a number of statutory exceptions and regulatory safe harbors under the federal Anti-Kickback Statute protecting certain common manufacturer business arrangements and activities from prosecution, the exceptions and safe harbors are drawn narrowly and an arrangement must meet all of the conditions specified in order to be fully protected from scrutiny under the federal Anti-Kickback Statute. We seek to comply with the exceptions and safe harbors whenever possible, but our practices, such as our patient assistance programs and discounts with certain customers, may not in all cases meet all of the criteria for protection from Anti-Kickback liability and may be subject to scrutiny.
The federal false claims laws, including the federal False Claims Act, prohibit any person or entity from knowingly presenting, or causing to be presented, a false claim for payment to the federal government, or knowingly making, or causing to be made, a false statement to get a false claim paid. Additionally, the civil monetary penalties statute imposes penalties against any person or entity that, among other things, is determined to have presented or caused to be presented a claim to a federal health program that the person knows or should know is for an item or service that was not provided as claimed or is false or fraudulent. Many pharmaceutical and other health care companies have been investigated and have reached substantial financial settlements with the federal government under the federal False Claims Act for a variety of alleged marketing activities, including providing free product to customers with the expectation that the customers would bill federal programs for the product; providing consulting fees, grants, free travel, and other benefits to physicians to induce them to prescribe the company’s products; and inflating prices reported to private price publication services, which may be used by states to set drug payment rates under government health care programs. Companies have been prosecuted for causing false claims to be submitted because of the marketing of their products for unapproved uses. Pharmaceutical and other health care companies have also been prosecuted on other legal theories of Medicare and Medicaid fraud.
Legislative and regulatory proposals have been made to expand post-approval requirements and restrict sales and promotional activities for pharmaceutical products. It is not clear whether additional legislative changes will be enacted, or whether the FDA regulations, guidance or interpretations will be changed, or what the impact of such changes on the marketing approvals of any Travere products, if any, may be. In addition, increased scrutiny by the U.S. Congress of the FDA’s approval process may significantly delay or prevent marketing approval, as well as subject us to more stringent product labeling and post-marketing testing and other requirements.
We also could become subject to government investigations and related subpoenas. Such subpoenas are often associated with previously filed qui tam actions, or lawsuits filed under seal under the federal False Claims Act. Qui tam actions are brought by private plaintiffs suing on behalf of the federal government for alleged violations of the federal False Claims Act. The time and expense associated with responding to such subpoenas, and any related qui tam or other actions, may be extensive, and we cannot predict the results of our review of the responsive documents and underlying facts or the results of such actions. Responding to government investigations, defending any claims raised, and any resulting fines, restitution, damages and penalties, settlement payments or administrative actions, as well as any related actions brought by stockholders or other third parties, could have a material impact on our reputation, business and financial condition and divert the attention of our management from operating our business.
The number and complexity of both federal and state laws continues to increase, and additional governmental resources are being added to enforce these laws and to prosecute companies and individuals who are believed to be violating them. In particular, the PPACA includes a number of provisions aimed at strengthening the government’s ability to pursue Anti-Kickback and False Claims Act cases against pharmaceutical manufacturers and other healthcare entities, including substantially increased funding for healthcare fraud enforcement activities, enhanced investigative powers, amendments to the federal False Claims Act that make it easier for the government and whistleblowers to pursue cases for alleged kickback and false claim violations and public reporting of certain payments and transfers of value by certain pharmaceutical manufacturers to physicians and teaching hospitals nationwide. We anticipate that government scrutiny of pharmaceutical sales and marketing practices will continue for the foreseeable future and subject us to the risk of further government investigations and enforcement actions. Responding to a government investigation or enforcement action would be expensive and time-consuming, and could have a material adverse effect on our business, financial condition, results of operations and growth prospects.
The U.S. Foreign Corrupt Practices Act, and similar worldwide anti-bribery laws generally prohibit companies and their intermediaries from making improper payments to government officials for the purpose of obtaining or retaining business. Our policies mandate compliance with these anti-bribery laws. We operate in parts of the world that have experienced governmental corruption to some degree and in certain circumstances, strict compliance with anti-bribery laws may conflict with local customs and practices or may require us to interact with doctors and hospitals, some of which may be state controlled, in a manner that is different than in the United States. We cannot assure that our internal control policies and procedures will protect us from reckless or criminal acts committed by our employees or agents. Violations of these laws, or allegations of such violations, could disrupt our business and result in criminal or civil penalties or remedial measures, any of which could have a material adverse effect on our business, financial condition and results of operations and could cause the market value of our common stock to decline.
The federal Health Insurance Portability and Accountability Act of 1996 ("HIPAA"), created new federal criminal statutes that prohibit, among other actions, knowingly and willfully executing, or attempting to execute, a scheme to defraud any healthcare benefit program, including private third-party payers, and
58
knowingly and willfully falsifying, concealing or covering up a material fact or making any materially false, fictitious or fraudulent statement in connection with the delivery of or payment for healthcare benefits, items or services. Like the federal Anti-Kickback Statute, the PPACA amended the intent standard for certain healthcare fraud provisions under HIPAA such that a person or entity no longer needs to have actual knowledge of the statute or specific intent to violate it in order to have committed a violation.
Additionally, the federal Physician Payments Sunshine Act within the PPACA, and its implementing regulations, require that certain manufacturers of drugs, devices, biologicals and medical supplies to report annually information related to certain payments or other transfers of value made or distributed to physicians (defined to include doctors, dentists, optometrists, podiatrists, and chiropractors), other healthcare professionals (such as physician assistants and nurse practitioners), and teaching hospitals, or to entities or individuals at the request of, or designated on behalf of, physicians and teaching hospitals and to report annually certain ownership and investment interests held by physicians and their immediate family members.
Also, many states have similar fraud and abuse statutes or regulations, including state anti-kickback and false claims laws, that apply to items and services reimbursed under Medicaid and other state programs, or, in several states, apply regardless of the payer. Further, certain states require implementation of commercial compliance programs and marketing codes, compliance with the pharmaceutical industry’s voluntary compliance guidelines, and compliance with the applicable compliance guidance promulgated by the federal government. Other various state level requirements include restricting payments or the provision of other items of value that may be made to healthcare providers and other potential referral sources; restricting various marketing practices; requiring prescription drug companies to report expenses relating to the marketing and promotion of drug products; requiring the posting of information relating to clinical studies and their outcomes; requiring certain regulatory licenses to manufacture or distribute our products commercially and/or the registration of sales representatives; requiring the reporting of certain information related to drug pricing; and requiring drug manufacturers to track and report information related to payments, gifts, compensation, and other items of value to physicians and other healthcare providers.
If our operations are found to be in violation of any of the health regulatory laws described above or any other laws that apply to us, we may be subject to significant penalties, including imprisonment, criminal fines, civil monetary penalties, administrative penalties, disgorgement, exclusion from participation in federal healthcare programs, contractual damages, injunctions, recall or seizure of products, total or partial suspension of production, reputational harm, administrative burdens, additional oversight and reporting obligations if we become subject to a corporate integrity agreement or similar agreement to resolve allegation of non-compliance with these laws, diminished profits and future earnings, and the curtailment or restructuring of our operations, any of which could adversely affect our ability to operate our business and our results of operations.
We are also subject to foreign requirements comparable to those established above. Outside the United States, interactions between pharmaceutical companies and health care professionals are also governed by strict laws, such as national anti-bribery laws of European countries, national sunshine rules, regulations, industry self-regulation codes of conduct and physicians’ codes of professional conduct. Failure to comply with these requirements could result in reputational risk, public reprimands, administrative penalties, fines or imprisonment.
If we are not able to obtain and maintain required regulatory approvals, we will not be able to commercialize our products, and our ability to generate revenue will be materially impaired.*
Our product candidates, once approved, and the activities associated with their manufacture, marketing, distribution, and sales are subject to extensive regulation by the FDA and other regulatory agencies in the United States and by comparable authorities in other countries. Failure to adhere to regulations set out by these bodies for one or more of our commercial products could prevent us from commercializing the product candidate in the jurisdiction of the regulatory authority. We have only limited experience in meeting the regulatory requirements incumbent on the sale of drugs in the United States and elsewhere, and expect to rely on third parties to assist us in these processes. If these third parties fail to adequately adhere to the regulations governing drug distribution and promotion, we may be unable to sell our products, which could have a material effect on our ability to generate revenue.
Our product candidates and the activities associated with their development and commercialization, including testing, manufacture, safety, efficacy, recordkeeping, labeling, storage, approval, advertising, promotion, sale and distribution, are subject to comprehensive regulation by the FDA and other regulatory agencies in the United States and by comparable authorities in other countries. Failure to obtain regulatory approval for a product candidate will prevent us from commercializing the product candidate in the jurisdiction of the regulatory authority. We have only limited experience in filing and prosecuting the applications necessary to obtain regulatory approvals and expect to rely on third-party contract research organizations to assist us in this process.
Securing FDA approval requires the submission of extensive nonclinical and clinical data and supporting information to the FDA for each therapeutic indication to establish the product candidate’s safety and efficacy. Securing FDA approval also requires the submission of information about the product manufacturing process to, and successful inspection of manufacturing facilities by, the FDA. Our future products may not be effective, may be only moderately effective or may prove to have undesirable or unintended side effects, toxicities or other characteristics that may preclude our obtaining regulatory approval or prevent or limit commercial use. Comparable requirements are applicable outside the United States.
Our product candidates may fail to obtain regulatory approval for many reasons, including:
•our failure to demonstrate to the satisfaction of the FDA or comparable regulatory authorities that a product candidate is safe and effective for a particular indication;
•the results of clinical trials may not meet the level of statistical significance required by the FDA or comparable regulatory authorities for approval;
•our inability to demonstrate that a product candidate’s benefits outweigh its risks;
•our inability to demonstrate that the product candidate presents an advantage over existing therapies;
•the FDA’s or comparable regulatory authorities’ disagreement with the manner in which we interpret the data from nonclinical studies or clinical trials;
•the FDA or comparable regulatory authorities determine that data from clinical trials conducted outside the United States, including studies conducted solely in China, are not adequate to support regulatory approval or require additional nonclinical, clinical or other studies before such data may be relied upon;
59
•our inability to obtain or maintain authorization to conduct clinical trials, including FDA clearance of an IND, on a timely basis or at all;
•failure of the third-party manufacturers with which we contract for clinical or commercial supplies to satisfactorily complete an FDA or comparable foreign regulatory authority pre-approval inspection of the facility or facilities at which the product is manufactured to assess compliance with the FDA’s cGMP regulations or comparable foreign regulatory authority requirements to assure that the facilities, methods and controls are adequate to preserve the drug’s identity, strength, quality and purity; and
•a change in the approval policies or regulations of the FDA or comparable regulatory authorities or a change in the laws governing the approval process.
The process of obtaining regulatory approvals is expensive, often takes many years, if approval is obtained at all, and can vary substantially based upon a variety of factors, including the type, complexity and novelty of the product candidates involved. Changes in regulatory approval policies during the development period, changes in or the enactment of additional statutes or regulations, or changes in regulatory review for each submitted product application may cause delays in the approval or rejection of an application. The FDA and non-United States regulatory authorities have substantial discretion in the approval process and may refuse to accept any application or may decide that our data is insufficient for approval and require additional nonclinical, clinical or other studies. For certain of our product candidates, including civorebrutinib, clinical development to date has been conducted outside of the United States. Although the FDA may accept data from foreign clinical trials under applicable regulations, it may require additional studies, including bridging or U.S. clinical studies, or determine that such data are insufficient to support approval. In addition, the FDA's approach to the use of clinical data generated in China may continue to evolve. If the FDA requires us to generate additional data or repeat studies, our development timelines could be delayed, our costs could increase and our ability to obtain regulatory approval could be adversely affected.
In addition, varying interpretations of the data obtained from nonclinical and clinical testing could delay, limit or prevent regulatory approval of a product candidate. Any regulatory approval we ultimately obtain may be limited or subject to restrictions or post approval commitments that render the approved product not commercially viable. Any FDA or other regulatory approval of our product candidates, once obtained, may be suspended, varied or withdrawn, including for failure to comply with regulatory requirements or if clinical or manufacturing problems follow initial marketing.
We and the third parties with whom we work are subject to stringent and changing U.S. and foreign laws, regulations, and rules, contractual obligations, industry standards, policies and other obligations related to data privacy and security. Our (or the third parties with whom we work) actual or perceived failure to comply with such obligations could lead to regulatory investigations or actions; litigation (including class claims) and mass arbitration demands; fines and penalties; disruptions of our business operations; reputational harm; loss of revenue or profits; loss of customers or sales; and other adverse business consequences.
In the ordinary course of business, we collect, receive, store, process, generate, use, transfer, disclose, make accessible, protect, secure, dispose of, transmit, and share (collectively, "process") personal data and other sensitive information, including proprietary and confidential business data, trade secrets, intellectual property, data we collect about trial participants in connection with clinical trials, and sensitive third-party data (collectively, "sensitive information"). Our data processing activities subject us to numerous data privacy and security obligations, such as various laws, regulations, guidance, industry standards, external and internal privacy and security policies, contracts, and other obligations that govern the processing of personal data by us and on our behalf.
In the United States, federal, state, and local governments have enacted numerous data privacy and security laws, including data breach notification laws, personal data privacy laws, consumer health data laws, consumer protection laws (e.g., Section 5 of the Federal Trade Commission Act) and other similar laws (e.g., wiretapping laws). For example, HIPAA, as amended by the Health Information Technology for Economic and Clinical Health Act ("HITECH"), and their respective implementing regulations, imposes specific requirements relating to the privacy, security and transmission of individually identifiable health information. Among other things, HITECH, through its implementing regulations, makes certain of HIPAA’s privacy and security standards directly applicable to business associates, defined as a person or organization, other than a member of a covered entity’s workforce, that creates, receives, maintains or transmits protected health information for or on behalf of a covered entity for a function or activity regulated by HIPAA as well as their covered subcontractors.
Numerous U.S. states have enacted comprehensive privacy laws that impose certain obligations on covered businesses, including providing specific disclosures in privacy notices and affording residents with certain rights concerning their personal data. As applicable, such rights include the right to access, correct, or delete certain personal data, and to opt-out of certain data processing activities, such as targeted advertising, profiling, and automated decision-making. The exercise of these rights may impact our business and ability to provide our products and services. Certain states also impose stricter requirements for processing certain personal data, including sensitive personal information, such as conducting data privacy impact assessments or risk assessments. These state laws allow for statutory fines for noncompliance. For example, the California Consumer Privacy Act of 2018 (“CCPA”), as amended, applies to personal data of consumers, business representatives, and employees who are California residents, and requires businesses to provide specific disclosures in privacy notices, and affords California residents certain privacy rights related to their personal data, such as those noted herein. The CCPA allows for fines for certain noncompliance and allows private litigants affected by certain data breaches to recover significant statutory damages. The CCPA and other U.S. comprehensive privacy laws exempt some data processed in the context of clinical trials, but these laws increase compliance costs and potential liability with respect to certain other personal data we maintain about residents in certain states. Similar laws are being considered in several other states, as well as at the local level, and we expect more jurisdictions to pass similar laws in the future.
In addition, numerous U.S. states have enacted new laws governing the privacy of consumer health data. For example, Washington’s My Health My Data Act broadly defines consumer health data, places restrictions on processing consumer health data (including imposing stringent requirements for consents), provides consumers certain rights with respect to their health data, and creates a private right of action to allow individuals to sue for violations of the law. Other states have passed, are considering, and may adopt similar laws.
Additionally, under various privacy laws and other obligations, we are required to obtain certain consents to process personal data. For example, some of our data processing practices may be challenged under wiretapping laws, since we obtain consumer information from third parties through various methods,
60
including via cookies or third-party marketing pixels. These practices may be subject to increased challenges by class action plaintiffs. Our inability or failure to obtain consent for these practices could result in adverse consequences, including class action litigation and mass arbitration demands.
Outside the United States, an increasing number of laws, regulations, and industry standards apply to data privacy and security. For example, the European Union’s General Data Protection Regulation (“EU GDPR”), the United Kingdom’s GDPR (“UK GDPR”) (EU GDPR and UK GDPR, collectively "GDPR"), Brazil’s General Data Protection Law (Lei Geral de Proteção de Dados Pessoais, or “LGPD”) (Law No. 13,709/2018), and China’s Personal Information Protection Law (“PIPL”) impose strict requirements for processing personal data. For example, the GDPR imposes significant and complex burdens on processing personal data, which is relevant to our operations in the context of our conduct of clinical trials and is of interest to relevant regulators. Under the GDPR, government regulators can impose temporary or definitive bans on data processing, as well as fines of up to 20 million euros under the EU GDPR, 17.5 million pounds sterling under the UK GDPR or, in each case, or 4% of annual global revenue, whichever is greater. Further, under the GDPR, individuals may initiate litigation related to processing of their personal data, as well as consumer protection organizations authorized at law to represent data subjects' interests.
In addition, privacy advocates and industry groups around the world have proposed, and may propose, standards with which we are legally or contractually bound to comply, and may become subject to in the future. We are also bound by contractual obligations related to data privacy and security, and our efforts to comply with such obligations may not be successful. Additionally, we publish privacy policies, marketing materials and other statements, such as compliance with certain certifications, regarding data privacy and security. Regulators in the United States are increasingly scrutinizing these statements. If these policies, materials or statements are found to be deficient, lacking in transparency, deceptive, unfair, misleading, or misrepresentative of our practices, we may be subject to investigation, enforcement actions by regulators or other adverse consequences.
In the ordinary course of business, we transfer personal data from Europe and other jurisdictions to the United States or other countries. Europe and other jurisdictions have enacted laws requiring data to be localized or limiting the transfer of personal data to other countries. In particular, the European Economic Area ("EEA") and the United Kingdom ("UK") have significantly restricted the transfer of personal data to the United States and other countries whose privacy laws it generally believes are inadequate. Other jurisdictions may adopt or have already adopted similarly stringent localization and cross-border transfer laws, which could make it more difficult to transfer information across jurisdictions or prevent us from conducting business in certain countries. Although there are currently various mechanisms that are used to transfer personal data from the EEA and UK to the United States in compliance with these laws, such as the EU Standard Contractual Clauses ("EU SCCs"), the UK’s International Data Transfer Agreement / International Data Transfer Addendum to the EU SCCs, and the EU-U.S. Data Privacy Framework and the UK extension thereto (which allows for transfers to relevant U.S.-based organizations who self-certify compliance and participate in the applicable frameworks), these mechanisms may be subject to legal challenges, and there is no assurance that we can satisfy or rely on the Data Privacy Framework to lawfully transfer personal data to the United States.
If we are unable to implement a valid compliance mechanism for cross-border personal data transfers, or if the requirements for a legally-compliant transfer are too onerous, we may face significant adverse consequences, including increased exposure to regulatory actions, substantial fines and injunctions against processing or transferring personal data from Europe. Inability to import personal data from Europe to the United States may significantly and negatively impact our business operations, including by limiting our ability to conduct clinical trial activities in Europe and elsewhere; limiting our ability to collaborate with third parties with whom we work (such as CROs, service providers, contractors and other companies) that are subject to such cross-border data transfer or localization laws; the need to relocate part of or all of our business or data processing activities to other jurisdictions (such as Europe) at significant expense; or requiring us to increase our personal data processing capabilities and infrastructure in foreign jurisdictions at significant expense. Additionally, companies that transfer personal data out of the EEA and UK to other jurisdictions, particularly to the United States, are subject to increased scrutiny from regulators, individual litigants, and activist groups. Some European regulators have ordered certain companies to suspend or permanently cease certain transfers out of Europe for allegedly violating the GDPR’s cross-border data transfer limitations.
Additionally, the U.S. Department of Justice issued a rule entitled the Preventing Access to U.S. Sensitive Personal Data and Government-Related Data by Countries of Concern or Covered Persons, which places additional restriction on certain data transactions involving countries of concern (e.g., China, Russia, Iran) and covered persons that may impact certain business activities such as vendor engagements, sale or sharing of data, employment of certain individuals, and investor agreements. Violations of the rule could lead to significant civil and criminal fines and penalties. The rule applies regardless of whether data is anonymized, key-coded, pseudonymized, de-identified or encrypted, which presents particular challenges for companies like ours and may impact our ability to transfer data in connection with certain transactions or agreements.
In Europe, the Network and Information Security Directive (“NIS 2”) regulates the cyber resilience and incident response capabilities of entities operating in a number of sectors, including the health sector. Although NIS 2 has not yet been transposed into domestic law in Ireland, we may be required to comply with its provisions. Achieving compliance with NIS 2 may require significant investment of our time and resources.
Our obligations related to data privacy and security (and consumers’ data privacy expectations) are quickly changing in an increasingly stringent fashion, creating uncertainty. Additionally, these obligations are subject to differing applications and interpretations, which may be inconsistent or conflict among jurisdictions. Preparing for and complying with these obligations requires significant resources and may necessitate changes to our information technologies, systems, and practices and to those of any third parties with whom we work. Although we endeavor to comply with all applicable data privacy and security obligations, we may at times fail (or be perceived to have failed) to do so. Moreover, despite our efforts, our personnel or third parties with whom we work may fail to comply with such obligations, which could negatively impact our business operations and compliance posture. For example, any failure by a third party with whom we work to comply with applicable law, regulations, or contractual obligations could result in adverse effects, including proceedings against us by governmental entities or others. If we or any of the third parties with whom we work fail to comply or are perceived to have failed to comply with applicable obligations, we or they could be subject to a range of regulatory actions, litigation (including class actions), or mass arbitration demands that could affect our or our partners' ability to commercialize our products and conduct necessary research and development, and could harm or prevent sales of the affected products, or could substantially increase the costs and expenses of commercializing and marketing our products. In particular, plaintiffs have become increasingly more active in bringing privacy-related claims against companies, including class claims and mass arbitration demands. Some of these claims allow for the recovery of statutory damages on a per violation basis, and, if viable, carry the potential for monumental statutory damages, depending on the volume of data and the number of violations. Any threatened or actual government enforcement action or litigation could also generate adverse publicity and require that we devote substantial resources that could otherwise be used in other aspects of our business. Compliance with applicable federal,
61
state, and foreign laws is difficult and time consuming, and companies that violate them may face substantial penalties. The potential sanctions include significant criminal fines, civil monetary penalties, administrative penalties, disgorgement, exclusion from participation in federal health care programs, individual imprisonment, injunctions, recall or seizure of products, total or partial suspension of production, reputational harm, administrative burdens, interruption or cessation of clinical trials, additional oversight and reporting obligations if we become subject to a corporate integrity agreement or similar agreement to resolve allegations of non-compliance with these laws, diminished profits and future earnings, and the curtailment or restructuring of our operations, and other sanctions. Because of the breadth of these laws, it is possible that some of our business activities could be subject to challenge under one or more of these laws. Such a challenge, irrespective of the underlying merits of the challenge or the ultimate outcome of the matter, could have a material adverse effect on our business, financial condition, results of operations and growth prospects. Moreover, clinical trial subjects and other individuals about whom we or the third parties with whom we work obtain personal data, as well as the third parties with whom we work who share this information with us, may limit our ability to collect, use and disclose the information. Claims that we have violated individuals’ privacy rights, failed to comply with data protection laws, or breached our contractual obligations, even if we are not found liable, could be expensive and time-consuming to defend and could result in adverse publicity that could harm our business.
If our information technology systems or data, or those of third parties with whom we work, are or were compromised, we could experience adverse impacts resulting from such compromise, including, but not limited to, regulatory investigations or actions; litigation; fines and penalties; interruptions to our commercial operations, clinical trials or other operations; harm to our reputation; loss of revenue or profits; loss of sales; and other adverse consequences.
In the ordinary course of our business, we and the third parties with whom we work process sensitive information.
Cyberattacks, malicious internet-based activity, and online and offline fraud are prevalent and continue to increase. These threats are becoming increasingly difficult to detect. These threats come from a variety of sources, including traditional computer “hackers,” threat actors, personnel (such as through theft or misuse), "hacktivists", organized criminal threat actors, sophisticated nation-states, and nation-state-supported actors. Some actors now engage and are expected to continue to engage in cyberattacks, including without limitation nation-state actors for geopolitical reasons and in conjunction with military conflicts and defense activities. During times of war and other major conflicts, we and the third parties with whom we work may be vulnerable to a heightened risk of these attacks, including retaliatory cyberattacks that could materially disrupt our systems and operations, supply chain, and ability to produce, sell and distribute our products. For example, we work with third parties to support our business located in unstable regions and regions experiencing (or expected to experience) geopolitical or other conflicts, including in Israel, where businesses have experienced an increase in cyberattacks in relation to the Israel/Hamas conflict. We and the third parties with whom we work are subject to a variety of other evolving threats, including, but not limited to, social-engineering attacks (including through deep fakes, which are increasingly more difficult to identify as fake, and phishing attacks), malicious code (such as viruses and worms), malware (including as a result of advanced persistent threat intrusions), denial-of-service attacks, credential stuffing, credential harvesting, personnel misconduct or error, ransomware attacks, supply-chain attacks, software bugs, server malfunctions, software or hardware failures, loss of data or other information technology assets, adware, telecommunications failures, attacks enhanced or facilitated by artificial intelligence, and other similar threats. In particular, ransomware attacks, including those from organized criminal threat actors, nation-states and nation-state supported actors, are becoming increasingly prevalent and severe and can lead to significant interruptions, delays, or outages in our operations, ability to provide our products, disruption of clinical trials, loss of data (including data related to clinical trials), loss of income, significant extra expenses to restore data or systems, reputational loss and the diversion of funds. To alleviate the financial, operational and reputational impact of a ransomware attack, it may be preferable to make extortion payments, but we may be unwilling or unable to do so (including, for example, if applicable laws prohibit such payments). Additionally, hybrid and remote work has increased risks to our information technology systems and data, as our employees utilize network connections, computers, and devices outside our premises or network, including working at home, while in transit, and in public locations. Future or past business transactions (such as acquisitions or integrations) could also expose us to additional cybersecurity risks and vulnerabilities, as our systems could be negatively affected by vulnerabilities present in acquired or integrated entities’ systems and technologies. Furthermore, we may discover security issues that were not found during due diligence of such acquired or integrated entities, and it may be difficult to integrate companies into our information technology environment and security program.
We rely upon third parties and technologies to operate critical business systems to process sensitive information in a variety of contexts, including, without limitation, cloud-based infrastructure, encryption and authentication technology, employee email, and other functions. We also rely on third parties to provide certain products, including active pharmaceutical ingredients or API, to operate our business, including in China. Our ability to monitor these third parties’ information security practices is limited, and these third parties may not have adequate information security measures in place. While we may be entitled to damages if the third parties with whom we work fail to satisfy their privacy or security-related obligations to us, any award may be insufficient to cover our damages, or we may be unable to recover such award. In addition, supply-chain attacks have increased in frequency and severity, and we cannot guarantee that third parties’ infrastructure in our supply chain or that of the third parties with whom we work have not been compromised.
While we have implemented security measures designed to protect against security incidents, there can be no assurance that these measures will be effective. We take steps designed to detect, mitigate and remediate vulnerabilities in our information security systems (such as our hardware and/or software, including that of third parties with whom we work), but we have not in the past and may not in the future be able to detect, mitigate, and remediate all such vulnerabilities including on a timely basis. Despite our efforts, there can be no assurance that these vulnerability mitigation measures will be effective. It may also be difficult and/or costly to detect, investigate, mitigate, contain, and remediate a security incident. Further, we have experienced, and may in the future experience, delays in developing and deploying remedial measures and patches designed to address identified vulnerabilities. If exploited, certain vulnerabilities can result in a security incident. Actions taken by us or the third parties with whom we work to detect, investigate, mitigate, contain, and remediate a security incident could result in outages, data losses, and disruptions of our business. Threat actors may also gain access to other networks and systems after a compromise of our networks and systems. For example, threat actors may use an initial compromise of one part of our environment to gain access to other parts of our environment, or leverage a compromise of our networks or systems to gain access to the networks or systems of third parties with whom we work, such as through phishing or supply chain attacks.
Certain of the previously identified or similar threats have in the past, and may in the future, cause a security incident or other interruption that could result in unauthorized, unlawful, or accidental acquisition, modification, destruction, loss, alteration, encryption, disclosure of, or access to our sensitive information or our information technology systems, or those of the third parties with whom we work. We have experienced, and may in the future experience, security
62
incidents involving our systems and the systems of third parties with whom we conduct business. A security incident or other interruption could disrupt our ability (and that of third parties with whom we work) to provide our products. We expend resources and may have to modify our business activities (including our clinical trial activities) to try to protect against security incidents. Certain data privacy and security obligations require us to implement and maintain specific security measures, industry-standard or reasonable security measures to protect our information technology systems and sensitive information.
Applicable data security and public company disclosure obligations require us, or we may voluntarily choose, to notify relevant stakeholders of certain security incidents, including affected individuals, customers, regulators and investors, or to take other actions in certain circumstances, such as providing credit monitoring and identity theft protection services. Whether a cybersecurity incident is reportable to our investors may not be straightforward, may take considerable time to determine, and may be subject to change as the investigation of the incident progresses, including changes that may significantly alter any initial disclosure that we provide. Such disclosures and related actions can be costly, and the disclosures or the failure to comply with such applicable requirements, could lead to adverse consequences. Any incidents or perceived incidents involving our systems (or those of a third party with whom we work) may lead to adverse consequences, such as government enforcement actions (for example, investigations, fines, penalties, audits, and inspections); additional reporting requirements and/or oversight; restrictions on processing sensitive information (including personal data); litigation (including class claims); indemnification obligations; negative publicity; reputational harm; loss of customer, investor or partner confidence in the effectiveness of our cybersecurity measures; monetary fund diversions; the expenditure of significant capital and other resources; diversion of management attention; interruptions in our operations (including availability of data); financial loss and other similar harms. For example, the loss of clinical trial data from completed or ongoing clinical trials for any of our product candidates could result in delays in our regulatory approval efforts and significantly increase our costs to recover or reproduce the data.
Some of our contracts do not contain limitations of liability, and even where they do, there can be no assurance that limitations of liability in our contracts are sufficient to protect us from liabilities, damages, or claims related to our data privacy and security obligations. In addition, our insurance coverage may not be adequate or sufficient to protect us from or to mitigate liabilities arising out of our privacy and security practices or that such coverage will continue to be available on commercially reasonable terms or at all, or that such coverage will pay future claims.
In addition to experiencing a security incident, third parties may gather, collect, or infer sensitive information about us from public sources, data brokers, or other means that reveals competitively sensitive details about our organization and could be used to undermine our competitive advantage or market position. Sensitive information of us or our customers could also be leaked, disclosed, or revealed as a result of or in connection with our employee’s, personnel’s, or third parties with whom we work use of generative AI technologies.
Risks related to the use of artificial intelligence technologies could adversely affect our business, financial condition and/or operating results.
Our employees and personnel use generative artificial intelligence, machine learning and other artificial intelligence technologies (together, “AI/ML”) and/or automated decision-making technologies to perform their work, and the disclosure and use of personal data in AI/ML technologies is subject to various privacy laws and other privacy obligations. Governments have passed and are likely to pass additional laws and regulations regulating AI/ML and/or automated decision-making technologies. Our use of this technology could result in additional compliance costs, regulatory investigations and actions, and lawsuits. If we are unable to use AI/ML and/or automated decision-making technologies in the future, it could make our business less efficient and result in competitive disadvantages. Any sensitive information (including confidential, competitive, proprietary, or personal data) that we input into a third-party generative AI/ML platform could be leaked or disclosed to others, including if sensitive information is used to train the third parties’ AI/ML model. Additionally, where an AI/ML model ingests personal data and makes connections using such data, those technologies may reveal other personal or sensitive information generated by the model. Moreover, AI/ML models may create flawed, incomplete, or inaccurate outputs, some of which may appear correct. This may happen if the inputs that the model relied on were inaccurate, incomplete or flawed (including if a bad actor “poisons” the AI/ML with bad inputs or logic), or if the logic of the AI/ML is flawed (a so-called “hallucination”). We may use AI/ML outputs to make certain decisions. Due to these potential inaccuracies or flaws, the model could be biased and could lead us to make decisions that could bias certain individuals (or classes of individuals), and adversely impact their rights, employment, and ability to obtain certain pricing, products, services, or benefits.
Several jurisdictions around the globe, including Europe and certain U.S. states, have proposed, enacted, or are considering laws governing AI/ML, including the EU’s AI Act and the Colorado AI Act. For example, the EU AI Act sets out a risk-based framework, subjecting certain AI technologies to numerous compliance obligations, including transparency, conformity and risk assessment, monitoring and human oversight requirements. Under the EU AI Act, non-compliant companies may be subject to administrative fines of up to 35 million Euros or 7% of a company’s total worldwide annual turnover for the preceding financial year, whichever is the higher. Certain of our activities subject us to the EU AI Act and depending on how the EU AI Act is implemented and interpreted, we may have to adapt our business practices, contractual arrangements, and services to comply with such obligations. We expect other jurisdictions will adopt similar laws. Additionally, certain privacy laws extend rights to consumers (such as the right to delete certain personal data) and regulate automated decision making, which may be incompatible with our use of AI/ML. These obligations may make it harder for us and our employees and personnel to use AI/ML, lead to regulatory fines or penalties, require us to change our business practices or make changes to the AI/ML that we use, or prevent or limit our use of AI/ML. For example, the FTC has required other companies to turn over (or disgorge) valuable insights or trainings generated through the use of AI/ML where they allege the company has violated privacy and consumer protection laws. If we cannot use AI/ML or that use is restricted, our business may be less efficient, or we may be at a competitive disadvantage.
Additionally, sensitive information of the Company or our employees or other individuals could be leaked, disclosed, or revealed as a result of or in connection with our employee’s, personnel’s, or vendor’s use of AI/ML technologies.
If we fail to establish or maintain successful collaborations or licensing arrangements, including cross-border collaborations, our business could be adversely affected.*
Our current and future collaborations, partnerships and licensing arrangements may be affected by evolving U.S. and foreign laws, regulations and government policies governing cross-border investments, technology transfers, licensing arrangements and collaborations involving biotechnology and pharmaceutical companies. In particular, U.S. legislative and regulatory proposals have sought to expand governmental review of certain investments, licensing transactions and other business arrangements involving companies with operations or significant activities in China. Although these proposals
63
remain subject to legislative and regulatory developments, if enacted or implemented, they could delay, restrict, increase the cost of, or otherwise adversely affect our ability to enter into, maintain or perform collaborations, licensing arrangements or other strategic transactions involving existing or prospective partners. Any such developments could require modifications to transaction structures, delay development timelines, increase compliance costs or limit our ability to realize the anticipated benefits of our collaborations.
Uncertainties in the interpretation and application of existing, new and proposed tax laws and regulations could materially affect our tax obligations and effective tax rate.
The tax regimes to which we are subject or under which we operate are unsettled and may be subject to significant change. The issuance of additional guidance related to existing or future tax laws, or changes to tax laws or regulations proposed or implemented by the current or a future U.S. presidential administration, Congress, or taxing authorities in other jurisdictions, including jurisdictions outside of the United States, could materially affect our tax obligations and effective tax rate. To the extent that such changes have a negative impact on us, including as a result of related uncertainty, these changes may adversely impact our business, financial condition, results of operations, and cash flows.
The amount of taxes we pay in different jurisdictions depends on the application of the tax laws of various jurisdictions, including the United States, to our international business activities, tax rates, new or revised tax laws, or interpretations of tax laws and policies, and our ability to operate our business in a manner consistent with our corporate structure and intercompany arrangements. The taxing authorities of the jurisdictions in which we operate may challenge our methodologies for pricing intercompany transactions pursuant to our intercompany arrangements or disagree with our determinations as to the income and expenses attributable to specific jurisdictions. If such a challenge or disagreement were to occur, and our position was not sustained, we could be required to pay additional taxes, interest, and penalties, which could result in one-time tax charges, higher effective tax rates, reduced cash flows, and lower overall profitability of our operations. Our financial statements could fail to reflect adequate reserves to cover such a contingency. Similarly, a taxing authority could assert that we are subject to tax in a jurisdiction where we believe we have not established a taxable connection, often referred to as a “permanent establishment” under international tax treaties, and such an assertion, if successful, could increase our expected tax liability in one or more jurisdictions.
In July 2025, the OBBBA was signed into law, and it (along with other recent U.S. federal tax reform) has resulted in significant changes to the taxation of business entities including, among other changes, changes to the taxation of income derived from international operations, changes in the deduction and amortization of research and development expenditures, and limitations on the deductibility of business interest. Future guidance from the Internal Revenue Service and other tax authorities with respect to any legislation may affect us, and certain aspects of such legislation could be repealed or modified or sunset in future years.
Our ability to use net operating loss carryforwards and certain other tax attributes to offset future taxable income and taxes may be subject to limitations.
Our U.S. federal net operating losses ("NOLs") generated in tax years beginning after December 31, 2017, may be carried forward indefinitely, but the deductibility of such federal NOL carryforwards in a taxable year is limited to 80% of taxable income in such year. As of December 31, 2025, we had federal NOL carryforwards of $232.6 million. In addition, under Sections 382 and 383 of the Internal Revenue Code of 1986, as amended, and corresponding provisions of state law, if a corporation undergoes an “ownership change,” which is generally defined as a greater than 50% change, by value, in its equity ownership over a three-year period, the corporation’s ability to use its pre-change NOL carryforwards and other pre-change U.S. tax attributes (such as research tax credits) to offset its post-change income or taxes may be limited. We continue to evaluate potential historical ownership changes and we may experience ownership changes in the future as a result of subsequent shifts in our stock ownership, some of which may be outside of our control.
As a result, our federal NOL carryforwards may be subject to a percentage limitation if used to offset income in tax years following an ownership change. In addition, it is possible that we have in the past undergone, and in the future may undergo, additional ownership changes that could limit our ability to use all of our pre-change NOL carryforwards and other pre-change tax attributes (such as research tax credits) to offset our post-change income or taxes. Similar provisions of state tax law may also apply to limit our use of accumulated state tax attributes. In addition, at the state level, there may be periods during which the use of NOL carryforwards is suspended or otherwise limited, which could accelerate or permanently increase state taxes owed. For example, California imposed limits on the usability of California state net operating losses to offset taxable income in tax years beginning after 2023 and before 2027. As a result, we may be unable to use all or a material portion of our NOL carryforwards and other tax attributes, which would harm our future operating results by effectively increasing our future tax obligations.
Changes in funding for the FDA, the SEC and other government agencies or regulatory authorities could hinder their ability to hire and retain key leadership and other personnel, prevent new products and services from being developed or commercialized in a timely manner, or otherwise prevent those agencies from performing normal functions on which the operation of our business may rely, which could negatively impact our business.
The ability of the FDA to review and approve new products can be affected by a variety of factors, including government budget and funding levels, ability to hire and retain key personnel and accept payment of user fees, and statutory, regulatory, and policy changes. Average review times at the agency have fluctuated in recent years as a result. In addition, government funding of the SEC and other government agencies on which our operations may rely, including those that fund research and development activities is subject to the political process, which is inherently fluid and unpredictable.
Disruptions at the FDA and other agencies may also slow the time necessary for new therapies to be reviewed and/or approved by necessary government agencies, which would adversely affect our business. For example, over the last several years, the U.S. government has shut down several times, including in the fourth calendar quarter of 2025, and certain regulatory agencies, such as the FDA and the SEC, have had to furlough critical FDA, SEC and other government employees and stop critical activities. In addition, there have been significant staff reductions at the FDA and other agencies, some of which are or may be subject to legal challenges. If a prolonged government shutdown occurs, or if the FDA or EDA experience resource constraints, it could significantly impact the ability of the applicable regulatory agency to timely review and process our regulatory submissions, which could have a material adverse effect on our business. Further, future government shutdowns could impact our ability to access the public markets and obtain necessary capital in order to properly capitalize and continue our operations. Comparable considerations may be applicable in relation to foreign regulatory authorities.
64
Our business could be negatively impacted by environmental, social and corporate governance (ESG) matters or our reporting of such matters.
In recent years, there has been an increased focus from certain investors, employees, partners, and other stakeholders concerning ESG matters. While we have internal efforts directed at ESG matters and preparations for potential increased future disclosures, we may be perceived by certain stakeholders as not acting responsibly in connection with these matters, which could negatively impact us. Various policymakers within and outside of the U.S., including the State of California and the European Union, among others, have adopted (or are considering adopting) requirements for certain companies to undertake various actions, including disclosures, on climate-related or other ESG-related matters. These requirements are not uniform and may not be interpreted or applied uniformly, which may increase the cost and complexity of compliance and any associated risks. Some of these initiatives have been subject to legal challenges, delays, stays, or changes in enforcement posture, and their ultimate scope, applicability, and timing remain uncertain. The ESG landscape has been evolving rapidly, and it can be difficult to predict future developments. If other climate-related disclosure rules or other ESG rules become effective or become applicable to us, they may significantly increase our compliance and reporting costs and may also result in disclosures that certain investors or other stakeholders deem to negatively impact our reputation and/or that harm our stock price.
The withdrawal of the United Kingdom from the European Union, commonly referred to as “Brexit,” may adversely impact our ability to obtain regulatory approvals of our product candidates in the United Kingdom, result in restrictions or imposition of taxes and duties for importing our product candidates into the United Kingdom, and may require us to incur additional expenses in order to develop, manufacture and commercialize our product candidates in the United Kingdom.
The UK withdrawal from the EU on January 31, 2020, commonly referred to as Brexit, has changed the regulatory relationship between the UK and the EU. The MHRA is now the UK’s standalone regulator for medicinal products and medical devices. The United Kingdom is no longer subject to EU regulations. Northern Ireland continues to follow certain limited EU regulatory rules, but not in relation to medicinal products.
The UK regulatory framework in relation to clinical trials is governed by the Medicines for Human Use (Clinical Trials) Regulations 2004, as amended, which is derived from the CTD, as implemented into UK national law through secondary legislation. On January 17, 2022, the MHRA launched an eight-week consultation on reframing the UK legislation for clinical trials. The UK Government published its response to the consultation on March 21, 2023 confirming that it would bring forward changes to the legislation, and such changes were laid before Parliament on December 12, 2024. and signed into law on April 11, 2025. The changes include risk-proportionate regulation of clinical trials, with low-risk trials able to receive faster approval through automatic authorization, a streamlined approval process that integrates both regulatory and ethics committee approvals, leading to a single UK decision for clinical trials, and new legal obligations mandating the registration of clinical trials in public registries and the publication of trial results within 12 months of trial conclusion.
Marketing authorizations in the UK are governed by the Human Medicines Regulations (SI 2012/1916), as amended. Since January 1, 2021, an applicant for the EU centralized procedure marketing authorization can no longer be established in the UK. As a result, since this date, companies established in the UK cannot use the EU centralized procedure and instead must follow one of the UK national authorization procedures or one of the remaining post-Brexit international cooperation procedures to obtain a marketing authorization to market products in the UK. All existing EU marketing authorizations for centrally authorized products were automatically converted or grandfathered into UK marketing authorization, effective in Great Britain only, free of charge on January 1, 2021, unless the marketing authorization holder opted-out of this possibility. Northern Ireland remained within the scope of EU authorizations in relation to centrally authorized medicinal products until January 1, 2025. However, on January 1, 2025, a new arrangement as part of the so-called "Windsor Framework" came into effect and reintegrated Northern Ireland under the regulatory authority of the MHRA with respect to medicinal products. The Windsor Framework also removes EU licensing processes and EU labelling and serialization requirements in relation to Northern Ireland and introduces a UK-wide licensing process for medicines.
The MHRA has also introduced changes to national marketing authorization procedures. This includes introduction of procedures to prioritize access to new medicines that will benefit patients, including a 150-day assessment route, a rolling review procedure and the International Recognition Procedures which entered into application on January 1, 2024. Since January 1, 2024, the MHRA may rely on the International Recognition Procedure, or IRP, when reviewing certain types of marketing authorization applications. This procedure is available for applicants for marketing authorization who have already received an authorization for the same product from a reference regulator. These include the FDA, the EMA, and national competent authorities of individual EEA countries. A positive opinion from the EMA and CHMP, or a positive end of procedure outcome from the mutual recognition or decentralized procedures are considered to be authorizations for the purposes of the IRP.
There is no pre-marketing authorization orphan designation for medicinal products in the UK. Instead, the MHRA reviews applications for orphan designation in parallel to the corresponding marketing authorization application. The criteria are essentially the same as those in the EU, but have been tailored for the market. This includes the criterion that prevalence of the condition in the United Kingdom, rather than the EU, must not be more than five in 10,000. Upon the grant of a marketing authorization with orphan status, the medicinal product will benefit from up to 10 years of market exclusivity from similar products in the approved orphan indication. The start of this market exclusivity period will be set from the date of first approval of the product in the United Kingdom.
Business disruptions could seriously harm our future revenue and financial condition and increase our costs and expenses.
Our operations, and those of our third-party manufacturers, CROs and other contractors and consultants, could be subject to disruptions resulting from earthquakes, power shortages, telecommunications failures, water shortages, floods, hurricanes, typhoons, fires, extreme weather conditions, health epidemics or pandemics, wars and other geopolitical conflicts, and other natural or man-made disasters or business interruptions, for which we are predominantly self-insured. The occurrence of any of these business disruptions could seriously harm our operations and financial condition and increase our costs and expenses.
Our corporate headquarters are located in San Diego, California, an area prone to wildfires and earthquakes. These and other natural disasters could severely disrupt our operations, and have a material adverse effect on our business, results of operations, financial condition and prospects. If a natural disaster, power outage or other event occurred that prevented us from using all or a significant portion of our headquarters, that damaged critical infrastructure, such as the manufacturing facilities of our third-party contract manufacturers, or that otherwise disrupted operations, it may be difficult or, in certain cases, impossible for us to continue our business for a substantial period of time. Any disaster recovery and business continuity plans we have in
65
place may prove inadequate in the event of a serious disaster or similar event. We may incur substantial expenses as a result of the limited nature of our disaster recovery and business continuity plans, which could have a material adverse effect on our business.
In addition, we rely on third-party manufacturers, some of whom are located in China, to manufacture API for FILSPARI and certain of our product candidates. Any disruption in production or inability of our manufacturers in China to produce or ship adequate quantities to meet our needs, whether as a result of a natural disaster or other causes (such as staffing shortages, or a health epidemic or pandemic), could impair our ability to meet commercial demand for FILSPARI, to operate our business on a day-to-day basis and to continue our research and development of our product candidates. In addition, we are exposed to the possibility of product supply disruption and increased costs in the event of changes in the policies of the United States or Chinese governments (such as tariffs on products or materials that we use that are manufactured in China), political unrest or unstable economic conditions in China. Any recall of the manufacturing lots or similar action regarding our API used in clinical trials could delay the trials or detract from the integrity of the trial data and its potential use in future regulatory filings. In addition, manufacturing interruptions or failure to comply with regulatory requirements by any of these manufacturers could significantly delay clinical development of potential products and reduce third-party or clinical researcher interest and support of proposed trials. These interruptions or failures could also impede commercialization of our product candidates and impair our competitive position.
If material weaknesses in our internal control over financial reporting are discovered or occur in the future, our consolidated financial statements may contain material misstatements and we could be required to restate our financial results, which could adversely affect our stock price and result in an inability to maintain compliance with applicable stock exchange listing requirements.
A material weakness is a deficiency, or a combination of deficiencies, in internal control over financial reporting such that there is a reasonable possibility that a material misstatement of our annual or interim consolidated financial statements will not be prevented or detected on a timely basis. If material weaknesses in our internal control over financial reporting are discovered or occur in the future, or if we are unable to maintain effective internal control over financial reporting or disclosure controls and procedures for any reason, our ability to record, process and report financial information accurately, and to prepare financial statements within required time periods, could be adversely affected, which could subject us to litigation or investigations requiring management resources and payment of legal and other expenses and negatively impact the price of our common stock. In addition, we could be subject to sanctions or investigations by the SEC or other regulatory authorities, which would require additional financial and management resources.
Furthermore, investor perceptions of our company may suffer as a result of any material weakness in our internal controls, and this could cause a decline in the market price of our stock. Any failure of our internal control over financial reporting could have a material adverse effect on our stated operating results, result in an adverse opinion on our internal control over financial reporting from our independent registered public accounting firm, and harm our reputation.
Adverse developments affecting the financial services industry could adversely affect our current and projected business operations and our financial condition and results of operations.
Adverse developments that affect financial institutions, such as events involving liquidity that are rumored or actual, have in the past and may in the future lead to bank failures and market-wide liquidity problems. For example, on March 10, 2023, Silicon Valley Bank ("SVB") was closed by the California Department of Financial Protection and Innovation, which appointed the Federal Deposit Insurance Corporation ("FDIC") as receiver. Similarly, on March 12, 2023, Signature Bank and Silvergate Capital Corp. were each swept into receivership. In addition, on May 1, 2023, the FDIC seized First Republic Bank and sold its assets to JPMorgan Chase & Co. It is uncertain whether the U.S. Department of Treasury, FDIC and Federal Reserve Board will provide access to uninsured funds in the future in the event of the closure of other banks or financial institutions, or that they would do so in a timely fashion.
Although we assess our banking relationships as we believe necessary or appropriate, our access to cash in amounts adequate to finance or capitalize our current and projected future business operations could be significantly impaired by factors that affect the financial institutions with which we have banking relationships. These factors could include, among others, events such as liquidity constraints or failures, the ability to perform obligations under various types of financial, credit or liquidity agreements or arrangements, disruptions or instability in the financial services industry or financial markets, or concerns or negative expectations about the prospects for companies in the financial services industry. These factors could also include factors involving financial markets or the financial services industry generally. The results of events or concerns that involve one or more of these factors could include a variety of material and adverse impacts on our current and projected business operations and our financial condition and results of operations. These could include, but may not be limited to, delayed access to deposits or other financial assets or the uninsured loss of deposits or other financial assets; or termination of cash management arrangements and/or delays in accessing or actual loss of funds subject to cash management arrangements.
In addition, widespread investor concerns regarding the U.S. or international financial systems could result in less favorable commercial financing terms, including concerns or uncertainty relating to tariffs and their potential impact on the economy, levels of interest rates or costs and tighter financial and operating covenants, or systemic limitations on access to credit and liquidity sources, thereby making it more difficult for us to acquire financing on acceptable terms or at all. Any decline in available funding or access to our cash and liquidity resources could, among other risks, adversely impact our ability to meet our operating expenses, financial obligations or fulfill our other obligations, result in breaches of our financial and/or contractual obligations or result in violations of federal or state wage and hour laws. Any of these impacts, or any other impacts resulting from the factors described above or other related or similar factors not described above, could have material adverse impacts on our liquidity and our current and/or projected business operations and financial condition and results of operations.
We maintain our cash at financial institutions, often in balances that exceed federally insured limits.
We maintain the majority of our cash and cash equivalents in accounts at banking institutions in the United States that we believe are of high quality. Cash held in these accounts often exceed the FDIC insurance limits. If such banking institutions were to fail, we could lose all or a portion of amounts held in excess of such insurance limitations. In the event of failure of any of the financial institutions where we maintain our cash and cash equivalents, there can be no assurance that we would be able to access uninsured funds in a timely manner or at all. Any inability to access or delay in accessing these funds could adversely affect our business and financial position.
66
Risks Related to our Indebtedness and Financial Condition
Our indebtedness could adversely affect our financial condition.*
As of the end of the period covered by this report, we had approximately $620 million of total debt outstanding, all of which is classified as long term. As a result of our indebtedness, a portion of our cash flow will be required to pay interest and principal on the 2029 Notes and 2032 Notes if the notes are not converted into shares of common stock prior to maturity. We may not generate sufficient cash flow from operations or have future borrowings available to enable us to repay our indebtedness or to fund other liquidity needs.
Our indebtedness pursuant to the 2029 Notes and 2032 Notes could have important consequences. For example, it could:
•make it more difficult for us to satisfy our obligations with respect to any other debt we may incur in the future;
•increase our vulnerability to general adverse economic and industry conditions;
•require us to dedicate a substantial portion of our cash flow from operations to payments on our indebtedness and related interest, thereby reducing the availability of our cash flow to fund working capital, capital expenditures and other general corporate purposes;
•limit our flexibility in planning for, or reacting to, changes in our business and the industry in which we operate;
•increase our cost of borrowing;
•place us at a competitive disadvantage compared to our competitors that may have less debt; and
•limit our ability to obtain additional financing for working capital, capital expenditures, acquisitions, debt service requirements or general corporate purposes.
We expect to use cash flow from operations and outside financings to meet our current and future financial obligations, including funding our operations, debt service and capital expenditures. Our ability to make these payments depends on our future performance, which will be affected by financial, business, economic and other factors, many of which we cannot control. Our business may not generate sufficient cash flow from operations in the future, which could result in our being unable to repay indebtedness, or to fund other liquidity needs. If we do not generate sufficient cash from operations, we may be forced to reduce or delay our business activities and capital expenditures, sell assets, obtain additional debt or equity capital or restructure or refinance all or a portion of our debt, including the 2029 Notes and 2032 Notes, on or before maturity. We cannot make any assurances that we will be able to accomplish any of these alternatives on terms acceptable to us, or at all. In addition, the terms of existing or future indebtedness may limit our ability to pursue any of these alternatives. In addition, we may from time to time seek to retire or purchase our outstanding debt, including the 2029 Notes and 2032 Notes, through cash purchases and/or exchanges for equity securities, in open market purchases, privately negotiated transactions or otherwise. Such repurchases or exchanges, if any, will depend on prevailing market conditions, our liquidity requirements, contractual restrictions, and other factors. The amounts involved in any such transactions, individually or in the aggregate, may be material. Further, any such purchases or exchanges may result in us acquiring and retiring a substantial amount of such indebtedness, which could impact the trading liquidity of the 2029 Notes, the 2032 Notes or any other indebtedness.
We may be unable to raise the funds necessary to repurchase the 2029 Notes and 2032 Notes for cash following a fundamental change, or to pay any cash amounts due upon conversion, and our future indebtedness may limit our ability to repurchase the 2029 Notes and 2032 Notes or pay cash upon their conversion.*
Noteholders may require us to repurchase their 2029 Notes or 2032 Notes, as applicable, following a fundamental change at a cash repurchase price generally equal to the principal amount of the 2029 Notes and 2032 Notes to be repurchased, plus accrued and unpaid interest to, but excluding, the fundamental change repurchase date. In addition, upon conversion of the 2029 Notes or the 2032 Notes, we may be required or elect to satisfy all or a portion of our conversion obligation in cash, shares of our common stock or a combination thereof, depending on the terms of the applicable notes and any settlement election we make.
We may not have enough available cash or be able to obtain financing at the time we are required to repurchase the 2029 Notes or 2032 Notes, or pay the cash amounts due upon conversion thereof. In addition, applicable law, regulatory authorities and the agreements governing our future indebtedness may restrict our ability to repurchase the 2029 Notes or 2032 Notes or pay the cash amounts due upon conversion thereof. Our failure to repurchase the 2029 Notes or 2032 Notes or to pay the cash amounts due upon conversion thereof when required will constitute a default under the base and supplemental indentures that govern the 2029 Notes and 2032 Notes, which we refer to collectively as the “indenture.” We may not have sufficient funds to satisfy all amounts due under the 2029 Notes, the 2032 Notes and any other indebtedness.
A default under the 2029 Notes or 2032 Notes may have a material adverse effect on our financial condition.*
If an event of default under the 2029 Notes or 2032 Notes occurs, the principal amount of the 2029 Notes or 2032 Notes, as applicable, plus accrued and unpaid interest (including additional interest, if any) may be declared immediately due and payable, subject to certain conditions set forth in the indenture governing such notes. Events of default under the indentures governing the 2029 Notes and the 2032 Notes include, among other things:
•certain failures to make required payments of principal or interest;
•certain failures to comply with conversion, notice or other covenant obligations;
•specified defaults or accelerations with respect to other indebtedness; and
•certain bankruptcy, insolvency or reorganization events.
Accordingly, the occurrence of a default under the 2029 Notes or 2032 Notes, unless cured or waived, may have a material adverse effect on our results of operations.
67
Provisions of the 2029 Notes and 2032 Notes could discourage an acquisition of us by a third party.*
Certain provisions of the 2029 Notes and 2032 Notes could make it more difficult or more expensive for a third party to acquire us. Upon the occurrence of certain transactions constituting a fundamental change, under the applicable indenture governing the 2029 Notes and 2032 Notes, holders of the 2029 Notes or 2032 Notes, as the case may be, will have the right, at their option, to require us to repurchase all of their notes or any portion of the principal amount of such notes in integral multiples of $1,000. We may also be required to increase the conversion rate for conversions of the 2029 Notes and/or 2032 Notes, as applicable, in connection with certain make-whole fundamental changes.
Conversion of the 2029 Notes or 2032 Notes may dilute the ownership interest of existing stockholders, including holders who had previously converted their 2029 Notes or 2032 Notes.*
To the extent we issue shares of common stock upon conversion of the 2029 Notes or the 2032 Notes, the conversion of some or all of such notes may dilute the ownership interests of existing stockholders. Any sales in the public market of shares of the common stock issuable upon such conversion could adversely affect prevailing market prices of shares of our common stock. In addition, the existence of the 2029 Notes and 2032 Notes may encourage short selling by market participants because the conversion of the notes could be used to satisfy short positions, which could depress the price of shares of our common stock.
General Risk Factors
Unstable market, economic and geopolitical conditions may have serious adverse consequences on our business, financial condition and stock price.*
The global credit and financial markets have experienced extreme volatility and disruptions, including as a result of inflation and high interest rates, bank failures, tariffs, other restrictive trade policies, wars, armed conflicts and global geopolitical tension, and may experience disruptions in the future. These disruptions can result in severely diminished liquidity and credit availability, increased inflation, declines in consumer confidence, declines in economic growth, increases in unemployment rates, recessions and uncertainty about economic stability. There can be no assurance that further deterioration in credit and financial markets and confidence in economic conditions will not occur. Our general business strategy may be adversely affected by any such economic downturn, volatile business environment, higher inflation, or unpredictable and unstable market conditions. If the current equity and credit markets deteriorate, it may make any necessary debt or equity financing more difficult, more costly and more dilutive. Failure to secure any necessary financing in a timely manner and on favorable terms could have a material adverse effect on our operations, growth strategy, financial performance and stock price and could require us to delay or abandon clinical development plans.
Other international and geopolitical events could also have a serious adverse impact on our business. While we cannot predict the broader consequences, conflicts and retaliatory and counter-retaliatory actions could materially adversely affect global trade, currency exchange rates, inflation, regional economies, and the global economy, which in turn may increase our costs, disrupt our supply chain, impair our ability to raise or access additional capital when needed on acceptable terms, if at all, or otherwise adversely affect our business, financial condition, and results of operations.
Item 2. Unregistered Sales of Equity Securities and Use of Proceeds
None.
Item 3. Defaults Upon Senior Securities
None.
Item 4. Mine Safety Disclosures
Not applicable.
68
Item 5. Other Information
Trading Arrangements
During the fiscal quarter ended June 30, 2026, certain of our directors and officers (as defined in Rule 16a-1(f) under the Exchange Act) adopted or terminated a “Rule 10b5-1 trading arrangement” or “non-Rule 10b5-1 trading arrangement ,” as those terms are defined in Regulation S-K, Item 408, as set forth below. Each 10b5-1 plan listed below is part of such individual’s long-term asset diversification, tax and/or financial planning strategy, and has been entered into in accordance with the terms of our Insider Trading Policy. Under the terms of the plans, once the plans are in effect, the individuals listed below have no discretion or control over the timing or effectuation of any transactions in our securities pursuant to the plans, and sales will be made only if the stock meets the minimum price thresholds and/or other requirements specified in the applicable plan. Some of the planned sales relate to shares required to be sold by such individual to cover the tax withholding obligation in connection with the settlement of vested restricted stock units and/or performance restricted stock units. Each such director and/or officer continues to be subject to our stock ownership guidelines, and the execution of all potential sales under the plan would not by themselves cause any such individual to be out of compliance with the requirements of our stock ownership guidelines.
Trading Arrangements Adopted:
| Name & Title | Date Adopted | Character of Trading Arrangement (1) | Aggregate Number of Shares of Common Stock to be Sold Pursuant to Trading Arrangement | Expiration Date (2) | ||||||||||
Up to | ||||||||||||||
Up to | ||||||||||||||
Up to | ||||||||||||||
Up to | ||||||||||||||
Up to | ||||||||||||||
Up to | ||||||||||||||
Up to | ||||||||||||||
Up to | ||||||||||||||
Up to | ||||||||||||||
| 1 | Each trading arrangement marked as a “Rule 10b5-1 Trading Arrangement” is intended to satisfy the affirmative defense of Rule 10b5-1(c) under the Exchange Act (the “Rule”). | ||||
| 2 | Each trading arrangement permits transactions through and including the earlier to occur of (a) the completion of all sales and (b) the date listed in the table. Each trading arrangement marked as a “Rule 10b5-1 Trading Arrangement” only permits transactions upon expiration of the applicable mandatory cooling-off period under the Rule. | ||||
| 3 | Includes shares underlying stock options expiring in May 2027. | ||||
| 4 | Includes shares underlying stock options expiring in May 2028. | ||||
| 5 | Consists of shares underlying stock options expiring in May 2027. | ||||
| 6 | Consists of shares underlying stock options expiring in May 2027 and May 2028. | ||||
Trading Arrangements Terminated:
69
Item 6. Exhibits
(a) Exhibits
| 3.1 | |||||
| 3.2 | |||||
| 3.3 | |||||
| 3.4 | |||||
| 3.5 | |||||
| 3.6 | |||||
| 4.1 | |||||
| 4.2 | Supplemental Indenture, dated May 11, 2026, between the Company and U.S. Bank Trust Company, National Association, as Trustee (including the form of 0.50% Convertible Senior Note due 2032) (incorporated by reference to Exhibit 4.2 to the Company’s Current Report on Form 8-K, filed with the SEC on May 11, 2026). | ||||
| 10.1† | |||||
| 10.2↡* | |||||
| 31.1 | |||||
| 31.2 | |||||
| 32.1 | |||||
| 32.2 | |||||
| 101.INS | Inline XBRL Instance Document | ||||
| 101.SCH | Inline XBRL Taxonomy Extension Schema Document | ||||
| 101.CAL | Inline XBRL Taxonomy Extension Calculation Linkbase Document | ||||
| 101.DEF | Inline XBRL Taxonomy Extension Definition Linkbase Document | ||||
| 101.LAB | Inline XBRL Taxonomy Extension Label Linkbase Document | ||||
| 101.PRE | Taxonomy Extension Presentation Linkbase Document | ||||
| 104 | The cover page to this Quarterly Report on Form 10-Q has been formatted in Inline XBRL | ||||
| † | Indicates management contract or compensatory plan. | |||||||
| ↡ | Schedules have been omitted pursuant to Item 601(a)(5) of Regulation S-K. The Company undertakes to furnish supplemental copies of any of the omitted schedules upon request by the SEC. | |||||||
| * | Certain confidential information contained in this Exhibit, marked in brackets, has been omitted, because it is both not material and of the type of information that the registrant treats as private or confidential. | |||||||
70
SIGNATURES
Pursuant to the requirements of the Securities Exchange Act of 1934, as amended, the registrant caused this report to be signed on its behalf by the undersigned, thereunto duly authorized.
| Date: August 4, 2026 | TRAVERE THERAPEUTICS, INC. | ||||||||||
| By: | /s/ Eric Dube | ||||||||||
| Name: | Eric Dube | ||||||||||
| Title: | Chief Executive Officer | ||||||||||
| By: | /s/ Christopher Cline | ||||||||||
| Name: | Christopher Cline | ||||||||||
| Title: | Chief Financial Officer | ||||||||||
71
Exhibit 10.2
Execution Version
Confidential
CERTAIN CONFIDENTIAL INFORMATION CONTAINED IN THIS DOCUMENT, MARKED BY [*], HAS BEEN OMITTED BECAUSE IT IS BOTH (I) NOT MATERIAL AND (II) IS THE TYPE OF INFORMATION THAT THE REGISTRANT TREATS AS PRIVATE OR CONFIDENTIAL.
EXCLUSIVE LICENSE AND COLLABORATION AGREEMENT
by and between
Everest Medicines (Singapore) Pte. Ltd.,
Travere Therapeutics, Inc.
and
solely for the purpose of Section 16.15 (Parent Guarantee)
Everest Medicines Limited
dated as of June 1, 2026
TABLE OF CONTENTS
Page
ARTICLE 1 DEFINITIONS 1
ARTICLE 2 LICENSE 19
ARTICLE 3 GOVERNANCE 23
ARTICLE 4 DEVELOPMENT 27
ARTICLE 5 REGULATORY 32
ARTICLE 6 MANUFACTURE & SUPPLY 35
ARTICLE 7 COMMERCIALIZATION MATTERS 36
ARTICLE 8 FINANCIAL TERMS 38
ARTICLE 9 PAYMENT; RECORDS; AUDITS 44
ARTICLE 10 CONFIDENTIALITY 47
ARTICLE 11 REPRESENTATIONS AND WARRANTIES; LIMITATION OF LIABILITY 52
ARTICLE 12 INTELLECTUAL PROPERTY 58
ARTICLE 13 TERM; TERMINATION 64
ARTICLE 14 INDEMNIFICATION 72
ARTICLE 15 DISPUTE RESOLUTION 74
ARTICLE 16 MISCELLANEOUS 75
Schedules
Schedule 1.34 EVER001
Schedule 1.37 List of Everest Patents
Schedule 1.60 Everest Knowledge Parties
Schedule 1.83 Planned Global Trials
Schedule 4.4 List of Initial Technology Transfer
Schedule 10.4 Initial Press Release
i
EXCLUSIVE LICENSE AND COLLABORATION AGREEMENT
This Exclusive License and Collaboration Agreement (the “Agreement”) is entered into as of June 1, 2026 (the “Execution Date”), by and between Everest Medicines (Singapore) Pte. Ltd., a private company organized and existing under the laws of Singapore, having a registered office at 36 Robinson Road, #20-01 City House, Singapore 068877 (“Everest”), and Travere Therapeutics, Inc., a Delaware corporation, having a place of business at 3611 Valley Centre Drive, Suite 300, San Diego, California 92130, USA (“Licensee”) and, solely for the purpose of Section 16.15 (Parent Guarantee), Everest Medicines Limited, a Cayman Islands company with limited liability having an address at PO Box 309, Ugland House, Grand Cayman, KY1-1104, Cayman Islands (“Parent”).
Recitals
Whereas, Everest is a commercial stage biopharmaceutical company in the business of discovering, developing, manufacturing, and commercializing biologic and pharmaceutical products;
Whereas, Licensee is a biopharmaceutical company that is developing and commercializing rare disease therapies;
Whereas, Licensee desires to obtain from Everest, and Everest desires to grant to Licensee, an exclusive license under the Everest Licensed IP to Exploit the Licensed Compound and Licensed Products in the Field in the Territory (each as defined below), subject to the terms and conditions of this Agreement; and
Whereas, the Parties desire to collaborate in the Development of the Licensed Products in accordance with the terms and conditions of this Agreement.
Now, Therefore, in consideration of the foregoing premises and the mutual covenants herein contained, and for other good and valuable consideration, the receipt and sufficiency of which are hereby acknowledged, Everest and Licensee hereby agree as follows:
Article 1
DEFINITIONS
DEFINITIONS
Unless the context otherwise requires, the terms in this Agreement with initial letters capitalized (and their correlatives), shall have the meanings set forth below, or the meaning as designated in the indicated places throughout this Agreement. Cognates of such terms have correlative meanings.
1.1“Accounting Standards” means with respect to a Selling Party (as defined below), (a) International Financial Reporting Standards of the International Accounting Standards Board or (b) the U.S. generally accepted accounting principles, in each case, consistently applied.
1
1.2“Additional Active(s)” means any active pharmaceutical or biologic ingredient(s) that is not the Licensed Compound. For clarity, Additional Actives shall exclude formulation components such as coatings, stabilizers, excipients, solvents or controlled release technologies.
1.3“Affiliate” means any company or entity controlled by, controlling, or under common control with a Party, but only for so long as such control exists. For the purpose of this definition, an entity shall be deemed to “control” another entity, if it owns directly or indirectly, more than fifty percent (50%) of the outstanding voting securities, capital stock, or other comparable equity or ownership interest of such entity, or possession, direct or indirect, of the power to direct or cause the direction of the management or policies of an entity (whether through ownership of securities or other ownership interests, by contract or otherwise).
1.4“Annual Net Sales” means, with respect to any Calendar Year, the aggregate amount of Net Sales for such Calendar Year.
1.5“Applicable Laws” means collectively the applicable provisions of any and all national, supranational, regional, state and local laws, treaties, statutes, rules, regulations, administrative codes, guidance, ordinances, judgments, decrees, directives, injunctions, orders, and any license, franchise, permit or similar right granted under any of the foregoing (including Regulatory Approvals) and any policies and other requirements of or from any court, arbitrator, Regulatory Authority or governmental agency or authority having jurisdiction over or related to the subject item or subject person, including all applicable Data Protection Laws and anti-bribery and anti-corruption laws and applicable national and international guidelines such as ICH, GCP, GLP and GMP.
1.6“Basket Renal Trial” means the Basket Renal Trial described in Schedule 1.83 (Planned Global Trials).
1.7“Basket Renal Trial Development Costs” means any Development Cost incurred by or on behalf of a Party in the conduct of the Basket Renal Trial, including COGS, without mark-up, for the Manufacture of Licensed Products and placebo for the Basket Renal Trial.
1.8“BTK” means Bruton tyrosine kinase.
1.9“Business Day” means any day that is not a Saturday, a Sunday or other day on which banks are required or authorized by law to close in San Diego, California, Shanghai, China or Hong Kong Special Administrative Region.
1.10“Calendar Quarter” means each period of three (3) consecutive months commencing on January 1, April 1, July 1 or October 1 (or any portion thereof at the beginning or end of the Term or other relevant period).
1.11“Calendar Year” means each period of twelve (12) consecutive months commencing on January 1 and ending on December 31 (or any portion thereof at the beginning or end of the Term or other relevant period).
1.12“Change of Control” means, with respect to either Party: (a) a merger, acquisition, reorganization, or consolidation of such Party with a Third Party that results in the voting securities of such Party outstanding immediately prior thereto, or any securities into which such voting securities have been converted or exchanged, ceasing to represent more than fifty percent (50%) of the combined voting power of the surviving entity immediately after such merger, acquisition, reorganization, or consolidation; (b) a transaction or series of related transactions in which a Third Party, together with its Affiliates, becomes the beneficial owner,
2
directly or indirectly, of more than fifty percent (50%) of the combined voting power of the outstanding securities of such Party; (c) the sale or other transfer (in one transaction or a series of related transactions) to a Third Party of all or substantially all of such Party’s or its parent Affiliate’s assets relating to this Agreement; or (d) such other arrangement or agreement whereby a Third Party (or a group of Third Parties acting in concert) obtains the right to control the board of directors or equivalent governing body that has the ability to cause the direction of the management, policies or affairs of such Party.
1.13“Clinical Trial” means a clinical study (or a portion thereof) involving the administration of a pharmaceutical or biological product to a human.
1.14“CMC” means chemistry, manufacturing and controls with respect to the Licensed Compound or Licensed Products.
1.15“CMO” means any Third Party contract manufacturing organization.
1.16“COGS” means, the cost of Manufacturing the Licensed Product. COGS shall be a “standard cost” per unit (which standard cost shall reasonably approximate actual costs and be reviewed and adjusted periodically for material variances), comprised of the following elements calculated in accordance with U.S. generally accepted accounting principles (ASC 330) and International Financial Reporting Standards (IAS 2), as applicable: (a) direct labor (the actual cost of employees engaged in direct manufacturing activities and manufacturing-related quality control and quality assurance activities of the Licensed Compound and Licensed Products), (b) direct materials (the actual costs incurred in manufacturing or purchasing materials for manufacture, including freight-in costs, sales and excise taxes imposed thereon that are non-recoverable and customs duties, tariffs and charges levied by government authorities, and all costs of packaging components), (c) pro-rata facility costs (meaning facility occupancy costs, including depreciation of owned facilities and right-of-use assets, rent, depreciation of leaseholds, utilities, and maintenance costs) reasonably allocated to the manufacture of the Licensed Product but not including construction or capital improvement and without regard to idle space or abnormal unused capacity, (d) manufacturing-related quality assurance/quality control costs necessary to bring the Licensed Compound and Licensed Products to a saleable condition, and (e) overheads that are directly related to manufacturing and reasonably allocable to the provision of the Licensed Compound and Licensed Products using a systematic and rational allocation methodology, excluding selling, general and administrative costs, research and development costs, and other non-manufacturing period costs. To the extent that the Licensed Compound and Licensed Products are sourced from one or more CMOs, COGS shall be the actual price (e.g., net of any rebates, discounts, credits or similar deductions) paid to such CMO(s) for the manufacture, pre-sale storage, shipping and supply of the Licensed Compound and Licensed Products, including any associated tax and fees.
1.17“Combination Product” means a Licensed Product comprising the Licensed Compound and one or more Additional Active(s), where the Licensed Compound and the Additional Active(s) are (a) sold as a fixed dose/unit or otherwise co-formulated, or (b) sold as separate doses/units in a single package or otherwise co-packaged for a single invoiced price.
1.18“Commercialization” means, with respect to a product, all activities undertaken before and after obtaining Regulatory Approvals relating specifically to the pre-launch, launch, promotion, detailing, marketing, pricing, reimbursement, sale, and distribution of such product, including strategic marketing, sales force detailing, advertising, market product support, all customer support, product distribution, and invoicing and sales activities, but excluding Development and Manufacturing.
1.19“Commercially Reasonable Efforts” means [*].
3
1.20“Competing Product” means [*].
1.21“Complete” means, with respect to the Ongoing Clinical Trial for the Licensed Product, the finalization of the final clinical study report for such Ongoing Clinical Trial.
1.22“Confidential Information” means all Know-How and other confidential or proprietary scientific, technical, clinical, marketing, financial or commercial information or Data Controlled by a Party or its Affiliates, that is made available by or on behalf of such Party or any of its Affiliates to the other Party or its Affiliates, whether in oral, written, electronic or other form.
1.23“Confidentiality Agreement” means the Mutual Confidential Disclosure Agreement between the Parties dated October 24, 2024.
1.24“Control” (including any variations such as “Controlled” and “Controlling”) means, with respect to any Know-How, Patents or other intellectual property rights, possession by a Party or Third Party of the right, power and authority (whether by ownership, license or otherwise, other than by virtue of any rights granted under this Agreement) to grant access to, to grant use of, or to grant a license or a sublicense to such Know-How, Patents or intellectual property rights without violating the terms of any agreement or other arrangement with any Third Party. Notwithstanding anything in this Agreement to the contrary, a Party and its Affiliates shall be deemed to not Control any Know-How, Patents, or other intellectual property rights that are in-licensed or acquired by such Party or its Affiliates from a Third Party after the Effective Date, unless the other Party agrees to (a) comply with the terms and conditions of the agreement under which such Know-How, Patents, or other intellectual property rights were in-licensed or acquired by such Party; and (b) pay all amounts that such Party would be obligated to pay in connection with the grant, maintenance and exercise of a (sub)license as reasonably allocable to such other Party for use in its territory under such Know-How, Patents, or other intellectual property rights.
1.25“Cover” means, with respect to a Patent and a product, that a Valid Claim of such Patent would (absent a license or ownership thereof) be infringed (or, in the case of a claim of a Patent that has not yet issued, would be infringed if it were issued) by the Exploitation of such product.
1.26“Data” means all data, including nonclinical data and clinical data, generated by or on behalf of a Party or its Affiliates or their respective Sublicensees pursuant to activities conducted under this Agreement.
1.27“Data Protection Laws” means any Applicable Laws governing the Processing of the relevant Personal Data under the Agreement.
1.28“Development” means, with respect to a product, all non-clinical and clinical drug development activities and processes, including toxicology, pharmacology, project management and other non-clinical efforts, statistical analysis, delivery system development, the performance of Clinical Trials (excluding the Manufacturing of such product for use in Clinical Trials) and other activities, in each case, which are reasonably necessary to prepare submissions for, and obtain or maintain, Regulatory Approval of such product, including lifecycle management studies and other activities.
1.29“Development Budget” means the budget for Development Costs as agreed upon between the Parties through the JSC for conducting any Development activities pursuant to the Development Plan, which are subject to Development Cost Sharing and Reimbursement.
4
1.30“Development Cost Sharing and Reimbursement” means (i) the Development Cost sharing mechanism, pursuant to which the Parties shall share any Other Trial Development Costs pursuant to Section 4.3(a)(ii) (Other Trial Development Costs), or (ii) the Development Cost reimbursement mechanism, pursuant to which any Development Cost incurred by Everest or its Affiliates in connection with its or their performance of any Development activities under the Development Plan shall be reimbursed by Licensee pursuant to Section 4.3(a)(iii) (Other Development Costs).
1.31“Development Costs” means all Internal Costs and Out-of-Pocket Costs incurred by a Party or its Affiliates in connection with any Development activities pursuant to the Development Plan, including any Internal Costs or Out-of-Pocket Costs incurred by a Party following the Effective Date for any nonclinical or CMC Development activities for the Licensed Compound or Licensed Products, or Clinical Trial of the Licensed Products, provided that, to the extent incurred pursuant to the Development Plan, Development Costs shall not include any Internal Costs or Out-of-Pocket Costs incurred by a Party or its Affiliates solely in connection with any regulatory activities that are specifically and solely directed to preparing submissions for, and obtaining and maintaining, Regulatory Approvals and Pricing and Reimbursement Approvals of the Licensed Products in such Party’s territory, including communicating with Regulatory Authorities in such Party’s territory.
1.32“Development Plan” means a written plan that meets the requirements as set forth in Section 4.2 (Development Plan) and is approved by the JSC, describing (a) the global Development strategy for the Licensed Compound and Licensed Products in the Territory and, as applicable, outside the Territory, (b) all material (with respect to Licensee) or all (with respect to Everest) non-clinical studies and Clinical Trials (including any Global Trial), in each case, that the applicable Party or its Affiliates or Sublicensees plans to perform to obtain and maintain Regulatory Approval for the Licensed Products in the Field in the Territory or outside the Territory, as applicable, (c) all other material (with respect to Licensee) or all other (with respect to Everest) Development or regulatory activities, in each case, that the applicable Party or its Affiliates or Sublicensees plans to perform to obtain and maintain Regulatory Approval for the Licensed Products in the Field in the Territory or outside the Territory, as applicable, and (d) any corresponding Development Budget, as may be amended in accordance with this Agreement from time to time.
1.33“Divestiture” means [*]. “Divest” means to complete a Divestiture.
1.34“EVER001” means Everest’s proprietary chemical compound designated by Everest as EVER001, a reversible BTK inhibitor having the chemical structure as set forth on Schedule 1.34 (EVER001).
1.35“Everest Know-How” means any and all Know-How that (a) is Controlled by Everest or any of its Affiliates as of the Effective Date, during the Term, as of the Execution Date or at any time during the period between the Execution Date and the Effective Date, and (b) is necessary or reasonably useful for the Exploitation of the Licensed Compound or Licensed Products in the Field in the Territory, including the Know-How contained in Everest Solely Owned Inventions or in Everest’s interest in Know-How contained in the Jointly Owned Inventions. Notwithstanding the foregoing, Everest Know-How shall not include (i) any Know-How Controlled by any Third Party that becomes an Affiliate of Everest after the Effective Date as a result of a Change of Control transaction, or a merger, acquisition or other similar transaction, unless such Know-How is used by Everest in its performance of Development or Manufacture activities under this Agreement or any supply agreement between the Parties or their Affiliates, or (ii) any Know-How that is related to any Additional Active or other proprietary compound or product Controlled by Everest or any of its Affiliates to the extent that it does not relate to the Licensed Compound or Licensed Products.
5
1.36“Everest Licensed IP” means Everest Know-How and Everest Patents.
1.37“Everest Patents” means any and all Patents that (a) are Controlled by Everest or any of its Affiliates as of the Effective Date, during the Term, as of the Execution Date or at any time during the period between the Execution Date and the Effective Date and (b) are necessary or reasonably useful for the Exploitation of the Licensed Compound or Licensed Products in the Field in the Territory, including Everest Solely Owned Invention Patents and Everest’s interest in the Jointly Owned Invention Patents that satisfy clauses (a) and (b). Notwithstanding the foregoing, Everest Patents shall not include (i) any Patent Controlled by any Third Party that becomes an Affiliate of Everest after the Effective Date as a result of a Change of Control transaction or a merger, acquisition, consolidation or other similar transaction unless such Patent is practiced by Everest in its performance of Development or Manufacture activities under this Agreement or any supply agreement between the Parties or their Affiliates, or (ii) any Patents to the extent that claims in such Patents Cover any Additional Active or other proprietary compound or product Controlled by Everest or any of its Affiliates and do not Cover the Licensed Compound or Licensed Products. A list of Everest Patents as of the Effective Date is attached hereto on Schedule 1.37 (List of Everest Patents).
1.38“Exploit” means to make, have made, import, have imported, export, have exported, distribute, have distributed, use, have used, sell, have sold, offer for sale, or have offered for sale, including to Develop, Manufacture, and Commercialize.
1.39“FDA” means the United States Food and Drug Administration or any successor agency thereto performing similar functions.
1.40“Field” means any and all prophylactic, diagnostic and therapeutic uses and treatment of diseases and disorders.
1.41“First Commercial Sale” means, with respect to a Licensed Product in a country in the Territory, the first sale for monetary value by or on behalf of Licensee, its Affiliate or Sublicensee of the Licensed Product in such country after Regulatory Approval of such Licensed Product has been granted in such country. Sales of Licensed Product in a country prior to receipt of Regulatory Approval in such country, including “treatment IND sales,” “named patient sales” and “compassionate use sales” shall not be considered a First Commercial Sale in such country.
1.42“GCP” means the then-current standards, practices and procedures for good clinical practices promulgated or endorsed by any applicable Regulatory Authority, as may be updated from time to time, including applicable guidelines promulgated under the ICH guidelines.
1.43“Generic Product” means with respect to a Licensed Product in a country in the Territory, any pharmaceutical product that (a) contains the same active pharmaceutical ingredient at the same concentration as such Licensed Product, (b) is approved for use in such country pursuant to a Regulatory Approval process governing approval of a generic product of such Licensed Product based on the prior approval and data of the Licensed Product as determined by the applicable Regulatory Authority in such country; and (c) is sold in such country by a Third Party, which Third Party is (i) not authorized by Licensee, its Affiliates or Sublicensees to sell such product, and (ii) did not purchase such product in a chain of distribution that included any of Licensee, its Affiliates or Sublicensees.
1.44“Global Trial” means a multi-regional Clinical Trial that is designed to support Regulatory Approval for a Licensed Product in the Field in countries or regions both inside the Territory and outside the Territory through the conduct of Clinical Trials in multiple countries or
6
regions in the Territory and outside the Territory, in all circumstances conducted as part of one unified Clinical Trial.
1.45“Global Trial Development Costs” means any Development Cost incurred by or on behalf of a Party in the conduct of any Global Trial consistent with the Development Budget, including COGS, without mark-up, for the Manufacture of Licensed Products and placebo for any Global Trial.
1.46“GLP” means the then-current standards, practices and procedures for good laboratory practices promulgated or endorsed by any applicable Regulatory Authority, as may be updated from time to time, including applicable guidelines promulgated under the ICH guidelines.
1.47“GMP” means the then-current standards, practices and procedures for good manufacturing practices promulgated or endorsed by any applicable Regulatory Authority, as may be updated from time to time, including applicable guidelines promulgated under the ICH guidelines.
1.48“Governmental Authority” means any multi-national, national, federal, state, local, municipal, provincial or other governmental authority of any nature, including any governmental division, prefecture, subdivision, department, agency, bureau, branch, office, commission, council, court or other tribunal.
1.49“HSR Act” means the Hart-Scott-Rodino Antitrust Improvements Act of 1976, as amended.
1.50“HSR Clearance” means the following conditions, collectively: (a) the applicable waiting period required under the HSR Act (and any extension thereof) shall have expired or been earlier terminated, (b) no injunction (whether temporary, preliminary, or permanent) prohibiting consummation of this Agreement shall be in effect [*], and (c) no lawsuit or proceeding seeking to enjoin, restrain, make illegal or otherwise prohibit the consummation of the transactions contemplated by this Agreement shall be pending [*].
1.51“HSR Filing” means filings with the United States Federal Trade Commission and the Antitrust Division of the DOJ of a Notification and Report Form for Certain Mergers and Acquisitions (as that term is defined in the HSR Act) with respect to the subject matter of this Agreement, together with all required documentary attachments thereto.
1.52“ICH” means the International Council for Harmonization of Technical Requirements for Pharmaceuticals for Human Use.
1.53“IND” means an investigational new drug application, clinical trial application, clinical trial exemption or similar application or submission filed with or submitted to a Regulatory Authority in a country or jurisdiction that is necessary to commence Clinical Trials in such country or jurisdiction.
1.54“Indication” means, with respect to a Licensed Product, a separate disease, disorder or medical condition for which such Licensed Product can be used to treat or prevent. For clarity, (a) different histologic or genetic subtypes of the same disease or lines of therapy (e.g., a second line therapy versus a third line therapy) and (b) different patient subpopulations, in each case (a) and (b), shall not be deemed different Indications for purposes hereunder provided that overlapping but different diseases, including when a disease arises as a complication of a different disease shall be deemed different Indications for purposes hereunder. [*].
7
1.55“Initiation” means, with respect to a Licensed Product and a Clinical Trial, dosing of the first patient in such Clinical Trial for such Licensed Product.
1.56“Internal Costs” means costs incurred by a Party (or its Affiliate) for its employees in the conduct of any applicable activities under this Agreement and overhead costs of a Party reasonably allocable to such activities.
1.57“Invention” means any inventions or discoveries, including processes, manufacture, composition of matter, Know-How, methods, assays, designs, protocols, and formulas, and improvements or modifications thereof, patentable or otherwise, that are generated, developed, conceived or first reduced to practice (constructively or actually) by or on behalf of one Party or its Affiliates or their respective Sublicensees or the Parties jointly during the Term under this Agreement, including all rights, title and interests in and to the intellectual property rights therein and thereto.
1.58“JSC” means the Joint Steering Committee to be established by the Parties pursuant to Section 3.1 (Joint Steering Committee).
1.59“Know-How” means all proprietary or non-public technical, scientific, regulatory and other information, trade secrets, results, knowledge, techniques, materials and Data, in whatever form and whether or not confidential, patented or patentable, including invention disclosures, plans, processes, practices, methods, knowledge, know-how, skill, experience, ideas, concepts, test data (including pharmacological, toxicological and clinical test data), analytical and quality control data, formulae, specifications, marketing, pricing, distribution, cost, sales, and manufacturing data or descriptions. For clarity, Know-How does not include any Patents or the inventions claimed thereby.
1.60“Knowledge” means, with respect to Everest, the actual knowledge of the individuals set forth in Schedule 1.60 (Everest Knowledge Parties) (or any successor holding comparable authority to any of the foregoing as of the applicable date) and the knowledge any such individuals would have after a reasonable inquiry.
1.61“Licensed Compound” means (a) EVER001, (b) any compound disclosed or claimed in the Everest Patents set forth in Schedule 1.37 (List of Everest Patents) that disclose or claim the compound in clause (a) and (c) any metabolite, salt, ester, hydrate, solvate, isomer, enantiomer, free acid form, free base form, crystalline form, co-crystalline form, amorphous form, pro-drug (including ester prodrug) form, racemate, polymorph, chelate, stereoisomer, tautomer, or optically active form of the foregoing set forth in clause (a) or (b).
1.62“Licensed Product” means any pharmaceutical product in any presentation, dosage form, formulation, route of administration or package configuration containing the Licensed Compound as an active pharmaceutical ingredient, whether alone or in combination with Additional Active(s).
1.63“Licensee Know-How” means any and all Know-How that is (a) Controlled by Licensee or any of its Affiliates as of the Effective Date or during the Term, and (b) necessary or reasonably useful for the Exploitation of the Licensed Compound or Licensed Products, including any Know-How contained in Licensee Solely Owned Inventions or in Licensee’s interest in any Know-How contained in Jointly Owned Inventions. Notwithstanding the foregoing, Licensee Know-How shall not include (i) any Know-How Controlled by any Third Party that becomes an Affiliate of Licensee after the Effective Date as a result of a merger, acquisition or other similar transaction, unless such Know-How is used by Licensee in the Exploitation of the Licensed Compound or Licensed Products, and (ii) any Know-How that is related to any Additional Active or other proprietary compound or product Controlled by
8
Licensee or any of its Affiliates to the extent that it does not relate to Licensed Compounds or Licensed Products.
1.64“Licensee Patents” means any and all Patents that are (a) Controlled by Licensee or any of its Affiliates as of the Effective Date or during the Term, and (b) necessary or reasonably useful for the Exploitation of the Licensed Compound or Licensed Products, including Licensee Solely Owned Invention Patents and Licensee’s interest in the Jointly Owned Invention Patents. Notwithstanding the foregoing, Licensee Patents shall not include (i) any Patent Controlled by any Third Party that becomes an Affiliate of Licensee after the Effective Date as a result of a merger, acquisition or other similar transaction, and (ii) any Patents to the extent that claims in such Patents Cover any Additional Active or other proprietary compound or product Controlled by Licensee or any of its Affiliates and do not Cover the Licensed Compounds or Licensed Products.
1.65“Licensee Technology” means the Licensee Know-How and Licensee Patents.
1.66“MAA” means an application for the authorization for marketing of a product, including all amendments and supplements thereto, filed with the applicable Regulatory Authority to gain approval to market such product in the applicable country.
1.67“Mainland China” means the mainland of the People’s Republic of China.
1.68“Major European Country” means each of [*].
1.69“Major Market Country” means each of the United States [*].
1.70“Manufacture” means, with respect to a product, activities related to the manufacture of such product or any compound or component therein, including manufacturing supplies for Development or Commercialization, packaging, labeling, in-process and finished product testing, release of product or any component or ingredient thereof, quality assurance and quality control activities related to manufacturing and release of product, ongoing stability tests, storage, and shipment and regulatory activities directly related to any of the foregoing, but not including any Development or Commercialization activities.
1.71“MHRA” means the Medicines and Healthcare products Regulatory Agency of the United Kingdom or any successor agency thereto performing similar functions.
1.72“Net Sales” means, with respect to a Licensed Product, the gross amounts invoiced in arm’s-length transactions by or on behalf of Licensee or any of its Affiliates or Sublicensees (each, a “Selling Party”) from or on account of the sale of the Licensed Product by such Selling Party to Third Parties, less the sum of the following (to the extent such deduction is not already reflected in the amount invoiced and determined in accordance with the applicable Accounting Standards as consistently applied):
(a)credits, allowances and chargebacks, if any are actually granted, on account of recalls, damaged goods, and rejections or returns of items previously sold;
(b)import taxes, export taxes, excise taxes, sales taxes, value-added taxes, consumption taxes, duties or other taxes levied on, absorbed, determined or imposed with respect to such sales (excluding income or net profit taxes or franchise taxes of any kind), in each case, to the extent not reimbursed by any Third Party or refunded, recovered or credited back by the relevant tax authority;
9
(c)insurance, customs charges, freight, shipping and other transportation costs incurred in shipping of the Licensed Products to such Third Parties, in each case, to the extent not reimbursed by any Third Party;
(d)bad debt written off not exceeding [*] of gross invoiced amounts (provided that any bad debt subsequently collected shall be added to Net Sales); and
(e)discounts (including trade, quantity and cash discounts) actually granted and deducted solely on account of sales of the Licensed Product, and rebates actually paid to any Third Party on account of the purchase of the Licensed Product.
All of the foregoing deductions from the gross amounts invoiced for such sales of the Licensed Product shall be determined in accordance with the Accounting Standards. Net Sales shall not include Licensed Product provided for: (i) research and Clinical Trial purposes prior to and in support of Regulatory Approval; (ii) expanded access programs, compassionate use, named patient sales, patient assistance programs where the Licensed Products are provided at no cost or below cost; and (iii) test marketing programs or other similar programs or studies, or for sample or promotional purposes (provided that the Licensed Product is not otherwise generally available for purchase in the applicable country, and are provided at no cost or below cost). Subject to the foregoing, any sales income received by Licensee, its Affiliates or Sublicensees for Licensed Products prior to Regulatory Approval shall be Net Sales and subject to the royalties under Section 8.5 (Royalties).
Licensed Products shall be considered “sold” in accordance with the Accounting Standards. Net Sales shall be determined from the books and records of the Selling Party.
It is understood that any accruals for individual items reflected in Net Sales are periodically trued up and adjusted by each Selling Party consistent with its customary practices and in accordance with the Accounting Standards.
Sale or transfer of any Licensed Product between any of the Selling Parties shall not result in any Net Sales, and Net Sales shall be calculated based on any subsequent sales or dispositions to a Third Party in an arm’s length transaction. To the extent that any Selling Party receives consideration other than or in addition to cash in consideration for the sale of the Licensed Product to a Third Party, Net Sales shall include the fair market value of such non-cash consideration for such sale of Licensed Product. For clarity, sales by a Selling Party to a Third Party wholesaler or similar distributor, group purchasing organization, pharmacy benefit manager, or retail chain customer shall be considered sales to a Third Party and Net Sales hereunder.
Net Sales shall also include and be deemed to have been made with respect to any Licensed Products used by Licensee, or any of its Affiliates or Sublicensees, for its own commercial purposes, or transferred to any Third Party for less than the Selling Party is then charging in normal arm’s length sales transactions; and Net Sales in all such cases shall be deemed to have been made at the prices at which such Licensed Products are then being sold to the customers of such user or transferor (or of Licensee, if an Affiliate or Sublicensee is a user but not a seller) in arm’s length sales transactions.
10
For clarity, the foregoing paragraph shall not (a) apply to any Licensed Products provided for the purposes described in the exclusions from Net Sales set forth in the second paragraph of this Section 1.72 (Net Sales) (including research, Clinical Trial, expanded access, compassionate use, patient assistance, test marketing, sample or promotional purposes), (b) apply to any bona fide discounts, including volume-based discounts, formulary or market access pricing, or pricing required or mandated by a Governmental Authority, or (c) override or limit the deductions from Net Sales otherwise set forth in this Section 1.72 (Net Sales). For purposes of the foregoing paragraph, the deemed Net Sales price shall be calculated based on the weighted average net selling price charged by the applicable Selling Party to its customers in arm’s length sales transactions during the applicable Calendar Quarter.
To the extent that Licensee or any of its Affiliates, or Sublicensees, provides to the purchasing Third Party discounts or allowances that are applicable to purchases of the Licensed Product and one or more other products (such as in a “bundled sale” arrangement), such discounts and allowances shall be allocated between the Licensed Product (for purposes of the deductions used in calculating Net Sales as above) and such other products in an equitable and commercially reasonable manner that does not unfairly or inappropriately bias the level of discounting against the Licensed Product (as compared to the other products).
Net Sales of any Combination Product for the purpose of calculating royalties due under this Agreement shall be determined on a country-by-country basis for a given accounting period as follows: first, the Selling Party that owns or otherwise controls the Combination Product shall determine the actual Net Sales of such Combination Product (using the above provisions), and then: such Net Sales amount for the Combination Product shall be multiplied by the fraction A/(A+B), where “A” is the net selling price in such country of the Licensed Product, if sold separately for the same dosage as contained in the Combination Product, and “B” is the net selling price in such country of the Additional Active(s) in the combination if sold separately for the same dosage as contained in the Combination Product. All net selling prices of the components of such end-user product or service shall be calculated as the average net selling price of said components during the applicable accounting period for which the Net Sales are being calculated. In the event that, in any country, no separate sale of either the Licensed Product or Additional Active(s) included in such Combination Product are made during the accounting period in which the sale was made or if the net selling price for an active ingredient cannot be determined for an accounting period, Net Sales allocable to the Licensed Product in each such country shall be determined by mutual agreement reached in good faith by the Parties as soon as practicable based on an equitable method of determining the same that takes into account, on a country-by-country basis, all relevant factors (including variations in potency, the relative contribution and value to the end user of the Licensed Product and the Additional Active(s) in the combination).
1.73“Non-Renal Field” means any and all uses in the Field other than those within the Renal Field.
1.74“Ongoing Clinical Trial” means Everest’s ongoing Phase 1-b/Phase 2-a Clinical Trial of Licensed Product (also referred to as ES108001; NCT05800873).
11
1.75“Out-of-Pocket Costs” means costs and expenses paid to Third Parties (or payable to Third Parties and accrued in accordance with the Accounting Standards consistently applied) by a Party (or its Affiliate) directly incurred in the conduct of any applicable activities under this Agreement, including costs for independent contractors engaged as permitted under this Agreement.
1.76“Outside Disclosure Date” means (a) the [*] following the date Everest receives notice of HSR Clearance in accordance with Section 16.5(c) (HSR Filing) or (b) as the Parties otherwise agree in writing.
1.77“Party” means Licensee or Everest individually, and “Parties” means Licensee and Everest collectively.
1.78“Patents” means (a) all national, regional and international patent applications, including provisional and non-provisional applications, priority applications, and patent cooperation treaty (PCT) applications; (b) all patents, including supplemental protection certificates, that have issued or in the future issue from any of the foregoing, including utility models, design patents and certificates of invention; and (c) all divisionals, continuations, continuations-in-part, reissues, re-examination certificates, renewals, extensions or additions to any such patents and patent applications (as applicable).
1.79“Personal Data” means any data that constitutes “personal information,” “personal data” or similar term as defined and governed by applicable Data Protection Laws.
1.80“Phase II Clinical Trial” means, with respect to a Licensed Product, a Clinical Trial that is described as a phase II clinical trial in its protocol, or that otherwise would satisfy the requirements of 21 C.F.R. § 312.21(b) or corresponding counterparts in any applicable country in the Territory other than the United States. For clarity, the phase II part of a study designed as a continuous phase I/II study shall be considered a Phase II Clinical Trial for the purposes of this Agreement when the phase II part has been Initiated.
1.81“Phase III Clinical Trial” means, with respect to a Licensed Product, any (a) Clinical Trial that is described as a phase III clinical trial in its protocol, or that otherwise would satisfy the requirements of 21 C.F.R. §312.21(c) or corresponding counterparts in any applicable country in the Territory other than the United States or (b) other Clinical Trial, the results of which, together with prior data and information concerning such Licensed Product, are (at the time of Initiation) intended to be sufficient to support the submission of an MAA for and obtaining Regulatory Approval of such Licensed Product and to meet the evidentiary standard for demonstrating the safety, purity, efficacy, and potency of such product as required for such Regulatory Approval by the applicable Regulatory Authority. For clarity, (i) the phase III part of a study designed as a continuous phase II/III study shall be considered a Phase III Clinical Trial for the purposes of this Agreement when the phase III part has been Initiated and (ii) if a given Clinical Trial is not a Phase III Clinical Trial as of Initiation but is later converted to Phase III Clinical Trial, such Clinical Trial shall be deemed to have been Initiated as a Phase III Clinical Trial as of the date of the conversion.
1.82“Pivotal Clinical Trial” means a Clinical Trial of a Licensed Product in human subjects in any country, which trial is designed (a) to establish that the Licensed Product is safe and efficacious for its intended use; (b) to define warnings, precautions and adverse reactions that are associated with the Licensed Product in the dosage range to be prescribed; and (c) to be, either by itself or together with one or more other Clinical Trials having a comparable design and size, the final human Clinical Trial in support of Regulatory Approval of the Licensed Product. For the avoidance of doubt, it is the intent of the Parties that a Pivotal Clinical Trial shall be a Phase III Clinical Trial, provided, that any Phase II Clinical Trial that is subsequently used as a
12
registrational trial shall be deemed a Pivotal Clinical Trial only upon, and at such time as, an application for Regulatory Approval is accepted by the FDA, the National Medical Products Administration or a foreign equivalent regulatory authority in the other regions in the Territory using such Phase II Clinical Trial as the basis for such Regulatory Approval.
1.83“Planned Global Trials” means the Global Trials identified in Schedule 1.83 (Planned Global Trials).
1.84“PMDA” means the Pharmaceuticals and Medical Devices Agency of Japan or any successor agency thereto performing similar functions.
1.85“pMN” means primary membranous nephropathy.
1.86“Pricing and Reimbursement Approval” means, in any country where a Regulatory Authority authorizes reimbursement for, or approves or determines pricing or level of reimbursement for, pharmaceutical products, receipt (or, if required to make such authorization, approval or determination effective, publication) of such reimbursement authorization or pricing approval or determination.
1.87“Process” (and any inflection thereof) means any operation or set of operations performed on Data (including, without limitation, Personal Data), whether or not by automatic means, such as collection, recording, organization, storage, adaptation or alteration, retrieval, consultation, use, disclosure by transmission, dissemination or otherwise making available, alignment or combination, blocking, erasure or destruction.
1.88“Region” means each of the following groups of countries and jurisdictions: (a) the U.S. and Canada (“North America”); (b) the countries comprising the European Economic Area, and the United Kingdom (“Europe”); (c) Afghanistan, Australia, Bangladesh, Bhutan, India, Japan, Kazakhstan, Kyrgyzstan, Maldives, Mongolia, Nepal, New Zealand, North Korea, Pakistan, Sri Lanka, Tajikistan, Turkmenistan and Uzbekistan (collectively, “Asia”); and (d) all of the countries and jurisdictions in the Territory not covered by clauses (a), (b), or (c) (collectively, “ROW”).
1.89“Regulatory Approval” means the final or conditional approval of the applicable Regulatory Authority necessary to market and sell a pharmaceutical product in a country, region or other jurisdiction in the Territory, excluding separate Pricing and Reimbursement Approval, except to the extent necessary for such marketing or sale.
1.90“Regulatory Authority” means any supranational, country, federal, state or local regulatory agency, department, bureau or other governmental or regulatory authority having the administrative authority to regulate the development or marketing of pharmaceutical products in any country, region or jurisdiction.
1.91“Regulatory Exclusivity” means marketing or data exclusivity conferred by the applicable Regulatory Authority in a country, region or jurisdiction on the holder of a marketing approval for a pharmaceutical product in such country, region or jurisdiction, including, by way of example and not of limitation, regulatory data exclusivity, orphan drug exclusivity, new chemical entity exclusivity and pediatric exclusivity.
1.92“Regulatory Materials” means, with respect to a product, regulatory applications (including MAA), submissions, notifications, materials, communications, registrations, Regulatory Approvals or other filings made to, received from or otherwise conducted with a Regulatory Authority in order to Develop, Manufacture, market, sell or otherwise Commercialize such product in a particular country, region or jurisdiction.
13
1.93“Renal Field” means [*].
1.94“Retained Territory” means Mainland China, Hong Kong Special Administrative Region, Macau Special Administrative Region, Taiwan, Republic of Korea, Brunei Darussalam, the Kingdom of Cambodia, Indonesia, Laos, Malaysia, Myanmar, the Philippines, Singapore, Thailand, the Democratic Republic of Timor-Leste, and Vietnam.
1.95“Segregate” means [*].
1.96“Solely Owned Inventions” means Everest Solely Owned Invention and Licensee Solely Owned Invention, as applicable.
1.97“Sublicensee” means, (a) with respect to Licensee, a Third Party that has been granted a sublicense by Licensee under any of the rights or licenses granted to Licensee pursuant to this Agreement and (b) with respect to Everest, a Third Party that has been granted a license by Everest under the Everest Licensed IP to Exploit the Licensed Compound and Licensed Products in the Field in the Retained Territory. For clarity, a Third Party that was granted a further sublicense (including through multiple tiers) by a Sublicensee shall also be deemed a Sublicensee.
1.98“Territory” means worldwide except for the Retained Territory.
1.99“Third Party” means any entity other than the Parties and any of their Affiliates.
1.100“United States” or “U.S.” means the United States of America, including its territories and possessions as recognized by the United Nations from time to time, but in all cases including, for clarity, Puerto Rico.
1.101“U.S. Bulk Data Final Rule” means the Final Rule Implementing Executive Order 14117 of February 28, 2024, (Preventing Access to Americans’ Bulk Sensitive Personal Data and United States Government-Related Data by Countries of Concern), 90 Fed. Reg. 1636.
1.102“U.S. Dollars” or “$” means U.S. dollars, the lawful currency of the United States.
1.103“Upstream License Agreement” means [*].
1.104“Valid Claim” means a claim contained in (a) an issued and unexpired Patent, which claim has not been found to be unpatentable, invalid, revocable or unenforceable by a decision of a court or other authority of competent jurisdiction in the subject country or jurisdiction, which decision is unappealable or unappealed within the time allowed for appeal, and has not been admitted to be invalid or unenforceable through abandonment, reissue, disclaimer or otherwise, or (b) a Patent application that has not been irretrievably cancelled, withdrawn, abandoned or rejected; provided, however, that Valid Claim shall exclude any such claim in a Patent application that has not been granted within [*] from the earliest priority date for such application (except that any claim issued from such an excluded pending Patent application shall become a Valid Claim at and after its date of issuance).
1.105Additional Definitions. The following table identifies the location of definitions set forth in various Sections of the Agreement:
14
| Defined Terms | Section | ||||
| Additional Global Trial | 3.1(e) | ||||
| Alliance Manager | 3.10 | ||||
| Blocking IP | 8.7(b)(i) | ||||
| China Sufficient | 4.2 | ||||
| Combination Product Inventions | 12.1(d) | ||||
| Commercialization Plan | 7.2 | ||||
| Committee | 3.4 | ||||
| Data Breach | 11.4(a)(vii) | ||||
| Disclosing Party | 10.1 | ||||
| Dispute | 15.1 | ||||
| DOJ | 16.5(b) | ||||
| Effective Date | 13.1 | ||||
| Everest Additional Active | 12.1(d) | ||||
| Everest Additional Active Inventions | 12.1(d) | ||||
| Everest Indemnitees | 14.1 | ||||
| Everest Solely Owned Invention Patents | 12.1(b) | ||||
| Everest Solely Owned Inventions | 12.1(b) | ||||
| Everest Trademarks | 7.5(b) | ||||
| Execution Date | Introductory paragraph | ||||
| Exclusivity Period | 2.6 | ||||
| Executive Officers | 3.6(a) | ||||
| Force Majeure Event | 16.11 | ||||
| FTC | 16.5(b) | ||||
| Global Trademarks | 7.5(a) | ||||
| ICC | 15.1 | ||||
| Indemnitee | 14.3 | ||||
| Indemnitor | 14.3 | ||||
| Indirect Tax | 9.3(e) | ||||
| Infringement | 12.3(a) | ||||
| Initial Technology Transfer | 4.4 | ||||
| Joint Development Committee | 3.3 | ||||
| Jointly Owned Invention Patents | 12.1(c) | ||||
| Jointly Owned Inventions | 12.1(c) | ||||
| License Payments | 9.3(b) | ||||
| Licensee Additional Active | 12.1(d) | ||||
| Licensee Additional Active Inventions | 12.1(d) | ||||
| Licensee Indemnitees | 14.2 | ||||
| Licensee Solely Owned Invention Patents | 12.1(b) | ||||
| Licensee Solely Owned Inventions | 12.1(b) | ||||
| Losses | 14.1 | ||||
15
| Defined Terms | Section | ||||
| Manufacturing Technology Transfer Completion | 6.1 | ||||
| Ongoing Clinical Trial Development Costs | 4.3(a)(i) | ||||
| Other Trial Development Costs | 4.3(a)(ii) | ||||
| Permitted Development Overrun | 4.3(c) | ||||
| Pharmacovigilance Agreement | 5.8 | ||||
| Proposed Publication | 10.5(b) | ||||
| Receiving Party | 10.1 | ||||
| Remedial Action | 5.9 | ||||
| Review Period | 10.5(b) | ||||
| Royalty Floor | 8.7(d) | ||||
| Royalty Term | 8.6 | ||||
| SEC | 10.3(c) | ||||
| Subcommittee | 3.2 | ||||
| Term | 13.1 | ||||
| Territory Patents | 12.2(b) | ||||
| Third Party Claims | 14.1 | ||||
| Upfront Payment | 8.1 | ||||
| U.S. Bankruptcy Code | 13.5 | ||||
Article 2
LICENSE
LICENSE
2.1License Grant.
(a)Territory License Grant. Subject to the terms and conditions of this Agreement (including Everest’s retained rights in Section 2.3 (No Implied Licenses; Retained Rights)) and the Upstream License Agreement, Everest hereby grants to Licensee, during the Term, an exclusive (even as to Everest and its Affiliates), royalty-bearing license, with the right to sublicense through multiple tiers (in accordance with Section 2.2 (Sublicense Rights)), under Everest Licensed IP solely to Exploit the Licensed Compound and Licensed Products in the Field in the Territory.
(b)Retained Territory License Grant. Subject to the terms and conditions of this Agreement (including Everest’s retained rights in Section 2.3 (No Implied Licenses; Retained Rights)), Everest hereby grants to Licensee, during the Term, a non-exclusive license under the Everest Licensed IP to research, conduct non-clinical Development for, or Manufacture or have Manufactured the Licensed Compound and the Licensed Products in the Retained Territory solely for the purpose of Exploiting the Licensed Products in the Field in the Territory, provided, however, that Licensee shall not engage in such activities in the Retained Territory if such activities would be reasonably expected to materially adversely affect the Development or Commercialization of the Licensed Compound or Licensed Products in the Retained Territory.
2.2Sublicense Rights.
16
(a)Right to Sublicense. Subject to the terms and conditions of this Agreement and the Upstream License Agreement, Licensee shall have the right to grant sublicenses under Everest Licensed IP, to (i) an Affiliate, provided that such sublicense shall automatically terminate if such entity ceases to be an Affiliate of Licensee, or (ii) to a Third Party, in each case ((i) and (ii)), without Everest’s consent. Licensee will promptly provide written notice to Everest upon entering such sublicense.
(b)Sublicense Terms. If Licensee or any of its Affiliates grants a sublicense to any Third Party, then Licensee shall provide Everest with a copy of each such sublicense agreement (including any amendments thereof) within [*] after execution thereof (which copy may be redacted of any confidential or proprietary information that is not necessary for Everest to confirm or verify Licensee’s compliance with this Agreement or the Upstream License Agreement), which copy shall be deemed the Confidential Information of Licensee. For purposes of complying with the Upstream License Agreement, Everest has the right to notify the Upstream Licensors of the execution of a sublicense agreement by Licensee and provide a copy of such sublicense agreement to the Upstream Licensor subject to confidentiality and non-use obligations set forth in the Upstream License Agreement, and Everest shall be responsible for any breach of such confidentiality obligations by the Upstream Licensor. No such sublicense will diminish, reduce or eliminate any obligation of either Party under this Agreement or obligations to Upstream Licensors under the Upstream License Agreement. Any sublicense granted by Licensee under this Agreement shall be (i) in writing and (ii) subject and subordinate to, and consistent with, the terms and conditions of this Agreement and the Upstream License Agreement. It shall be a condition of any sublicense agreement that the Affiliate or Sublicensee, as applicable, agrees to be bound by the terms of this Agreement applicable to Affiliate or Sublicensee. Without limiting the foregoing, each sublicense agreement shall include the following additional terms and conditions: (A) the Sublicensee shall be bound by non-use and non-disclosure obligations consistent with those set forth in this Agreement; (B) the Sublicensee shall be obligated to maintain Development, Commercialization and financial records such that Licensee can comply with its obligations under Section 4.6 (Development Updates), Section 7.4 (Commercialization Reports) and Section 9.1 (Payment; Reports); and (C) any sublicense agreement shall contain appropriate intellectual property assignment and license provisions to give effect of the intellectual property ownership and license provisions in this Agreement.
(c)Right to Subcontract. Subject to the terms and conditions of this Agreement, Licensee may engage subcontractors for purposes of conducting Development, Manufacturing, Commercialization and other activities for Licensee under this Agreement, provided that any such subcontractor is bound by a written agreement that is consistent with the terms and conditions of this Agreement and the Upstream License Agreement. With respect to any activities of Licensee under this Agreement that are subcontracted to a subcontractor, Licensee will include in the applicable subcontract an assignment to Licensee of all rights in Patent and Know-How made by such subcontractor during the performance of the subcontracted activities, excluding any generally applicable modifications or improvements to such subcontractor’s background intellectual property rights, and in any event will include in the applicable subcontract a license to Licensee that is sublicensable to Everest and the Upstream Licensors, of any Patents and Know-How made by such subcontractor during the performance of the subcontracted activities. Notwithstanding Section 2.2(b) (Sublicense Terms), Licensee shall not be required to provide Everest with copies of agreements with subcontractors, provided that such subcontractor is not also a Sublicensee. For clarity, a subcontractor shall not be considered a Sublicensee under this Section 2.2(c) (Right to Subcontract) if Licensee or its Affiliate engages such subcontractor for conducting Development,
17
Manufacture, Commercialization and other activities under this Agreement on behalf of Licensee or its Affiliate solely on a fee-for-service basis and grants such subcontractor a limited right under the Everest Licensed IP solely as necessary to perform the subcontracted activities on behalf of Licensee or its Affiliate, and such right (i) is non-exclusive, non-transferable (except in connection with the permitted transfer of the applicable subcontract), and non-sublicensable (except to affiliates or further subcontractors of such subcontractor), (ii) is limited to the applicable subcontracted activities, (iii) does not permit the subcontractor to Exploit the Licensed Compounds or Licensed Products for its own account or for any other Third Party, and (iv) terminates upon completion or termination of the applicable subcontracted activities.
(d)Licensee’s Responsibility. Licensee shall be responsible for ensuring that the performance by any of its Affiliates, Sublicensees and subcontractors hereunder is in accordance with the applicable terms of this Agreement and the Upstream License Agreement. Licensee shall be responsible for any actions of its Affiliates, Sublicensees and subcontractors to the same extent as if such actions had been taken by Licensee itself, and Everest shall have the right to proceed directly against Licensee without any obligation to first proceed against such Affiliate, Sublicensee, or subcontractor. Licensee shall be fully responsible and liable for the failure of its Affiliates, Sublicensees or subcontractors to comply with the relevant obligations under this Agreement and shall, at [*] cost, be responsible for enforcing compliance by its Affiliates, Sublicensees, and subcontractors with the terms of the applicable sublicense agreement.
2.3No Implied Licenses; Retained Rights. No right or license under any Patents or Know-How of either Party is granted or shall be granted by implication. All such rights or licenses are or shall be granted only as expressly provided in the terms of this Agreement. Everest hereby expressly reserves all rights not expressly licensed to Licensee in Section 2.1 (License Grant), including (a) all rights under the Everest Licensed IP with respect to the Licensed Compound and Licensed Products in the Retained Territory, subject to Licensee’s non-exclusive license as set forth in Section 2.1(b) (Retained Territory License Grant), (b) all rights under the Everest Licensed IP to exercise its rights or perform its obligations under this Agreement, (c) all rights under the Everest Licensed IP to research, conduct non-clinical Development, or Manufacturing activities in the Territory solely for the purpose of Exploiting the Licensed Products in the Retained Territory, in each case whether directly or through its Affiliates, licensees or subcontractors.
2.4License to Everest. Licensee hereby grants to Everest a non-exclusive, fully paid, royalty-free license, with the right to sublicense through multiple tiers to any Affiliate of Everest or any other Third Party, (a) under the Licensee Technology, to fulfill, either itself, or through its Affiliates or subcontractors, its obligations under this Agreement, including its Manufacturing and supply obligations under Section 6.2 (Clinical Supply) and Section 6.3 (Commercial Supply) and (b) subject to Section 5.7(a), under the Licensee Solely Owned Inventions and Licensee Solely Owned Inventions Patents to Exploit the Licensed Compound or Licensed Products (excluding any Additional Active or other proprietary compound or product Controlled by Licensee or any of its Affiliates) solely in the Field in the Retained Territory.
2.5Upstream License Agreement. Licensee acknowledges and agrees that:
(a)(i) Everest obtained the rights to certain Everest Licensed IP from the Upstream Licensors under the Upstream License Agreement; (ii) the license grant to Licensee under Section 2.1 (License Grant) constitutes a sublicense under the Upstream License Agreement; and (iii) such sublicense is subject to the terms and conditions of the Upstream License Agreement;
18
(b)it has received from Everest a copy of the Upstream License Agreement;
(c)Licensee shall, and shall cause its Affiliates, and shall require its Sublicensees and subcontractors, to comply with the terms and conditions of the Upstream License Agreement; and
(d)without limiting the foregoing, Licensee shall maintain in confidence all Confidential Information (as such term is defined in the Upstream License Agreement) of the Upstream Licensors, and shall only use such Confidential Information as required or permitted by Article 8 (Confidentiality and Publicity) of the Upstream License Agreement.
2.6Non-Compete. From the Effective Date until the date that is ten (10) years thereafter (the “Exclusivity Period”), except pursuant to the terms and conditions of this Agreement with respect to any Licensed Compound or Licensed Products or with the advance, written agreement of the other Party, each Party shall not, and will cause its Affiliates and require its Sublicensees not to directly or indirectly, whether by itself or with or through any Third Party, Develop, Manufacture or Commercialize any Competing Product in the Field. Notwithstanding the foregoing, if a Party undergoes a Change of Control with a Third Party (such Third Party, together with its Affiliates existing prior to such Change of Control, the “Acquisition Party”), then the non-compete obligations set forth in this Section 2.6 (Non-Compete) shall not [*] if such Party and such Acquisition Party promptly Segregate such Competing Product [*]. If a Third Party becomes an Affiliate of a Party after the Effective Date through merger, acquisition, consolidation or other similar transaction and such transaction does not result in a Change of Control of such Party [*] then the non-compete obligations set forth in this Section 2.6 (Non-Compete) shall not [*] if such Party or its Affiliate, as applicable, Divests such Competing Product within [*] and at all times Segregates such Competing Product prior to such Divestiture. The exceptions set forth in this Section 2.6 (Non-Compete) shall apply to any Sublicensee, mutatis mutandis.
2.7[*].
Article 3
GOVERNANCE
GOVERNANCE
3.1Joint Steering Committee. Within [*] following the Effective Date, the Parties shall establish a JSC comprised of [*] representatives from each Party to provide oversight of the Parties’ activities with respect to the Development and Commercialization of the Licensed Products in the Territory and facilitate information exchange between the Parties under this Agreement. The JSC, as may be conducted through the applicable Subcommittee, shall in particular:
(a)review, discuss and coordinate the overall strategy for the Development of the Licensed Products in the Territory and outside the Territory;
(b)review, discuss and approve the initial Development Plan and any proposed amendments or revisions to the Development Plan(s), including with respect to any Global Trial;
(c)review, discuss and approve the initial Development Budget and any proposed amendments or revisions to the Development Budget;
(d)review, discuss and approve any study protocols relating to the Development Plan(s) (and any amendments thereto);
19
(e)review and discuss the feasibility of pursuing any Global Trial in the Field in addition to the Planned Global Trials (each, an “Additional Global Trial”);
(f)review, discuss and serve as a forum for the sharing of information between the Parties regarding the conduct of any Development activities by Licensee and Everest;
(g)oversee and coordinate the on-going disclosure, sharing or transfer of Everest Know-How pursuant to Section 4.4 (Technology Transfer) and of new Inventions generated in or related to the Development of the Licensed Products;
(h)review, discuss and serve as a forum to discuss the Commercialization of the Licensed Products in and branding strategies in the Field in the Territory and outside the Territory;
(i)review and discuss the Commercialization Plan for the Territory, in compliance with Applicable Laws (including antitrust and competition laws);
(j)coordinate supply of Licensed Products for the Territory in accordance with Article 6 (Manufacture & Supply);
(k)Establish other Subcommittees as necessary or advisable to further the purpose of this Agreement; and
(l)perform such other functions as appropriate to further the purposes of this Agreement, as expressly set forth in this Agreement or as determined by the Parties in writing.
3.2Subcommittees. From time to time during the Term, the JSC may establish and disband one or more subcommittee(s) (each, a “Subcommittee”) to oversee particular activities of the Parties, and the JSC may assign to such Subcommittee(s) duties or tasks independent of the duties of the JSC, or delegate part of such duties of the JSC to such Subcommittee(s) as it deems necessary and appropriate.
3.3Joint Development Committee. Without limiting the generality of Section 3.2 (Subcommittees), no later than [*] after the Effective Date, the JSC shall establish a joint development committee (the “Joint Development Committee” or “JDC”) comprised of an equal number of representatives from each Party. The JDC shall (i) oversee the Development of the Licensed Products and implementation and progress of the Development Plan in the Territory, including to discuss and propose to the JSC for approval the initial Development Plan and any amendments to the Development Plan(s), and (ii) share information and coordinate efforts on the Manufacturing of the Licensed Products for the Territory.
3.4Composition; Meetings. The JSC and each Subcommittee (each, a “Committee”) shall be composed of [*] representatives from each Party (or, for any Subcommittee, such other equal number of representatives of each of Licensee and Everest as the JSC may determine), each of whom shall be an officer or employee of the applicable Party with the requisite experience and seniority to make decisions on behalf of such Party, and each Party shall notify the other Party of its initial JSC representatives within [*] days after the Effective Date. Each Party shall designate a representative to be the co-chairperson of each Committee, who shall schedule meetings, prepare meeting agenda and meeting minutes and follow up on action items. These responsibilities shall alternate between the Parties or each co-chairperson, as applicable, with Everest’s co-chairperson taking the responsibility for the first meeting of each Committee. Each Party may change its representatives to any Committee from time to time in its
20
sole discretion, effective upon notice to the other Party of such change. Each Party’s representatives in each Committee shall possess appropriate experience with respect to the issues falling within the functions of such Committee and requisite seniority within such Party’s organization, and shall have the authority to make decisions on behalf of the Party such representative represents.
3.5Non-Member Attendance. Each Party may from time to time invite a reasonable number of participants, in addition to its representatives, to attend a meeting of any Committee (in a non-voting capacity) in the event that the planned agenda for such Committee meeting would require such participants’ expertise; provided that if either Party intends to have any Third Party (including any consultant or counsel) attend such a meeting, such Party shall provide prior written notice to the other Party, obtain approval from such other Party for such Third Party to attend, such approval not to be unreasonably withheld, conditioned or delayed, and ensure that such Third Party is bound by confidentiality and non-use obligations consistent with the terms of this Agreement.
3.6Decision-Making. The representatives of each Party shall have, collectively, [*] vote at each Committee on all matters brought before such Committee. The Committees may not take any action or decide any matter except at a meeting attended by at least [*] voting member representing each Party. Each Committee shall endeavor to make decisions by consensus.
(a)JSC Decisions. Except where consent or approval on any matter by a Party is expressly required herein, if after reasonable discussion and good faith consideration of each Party’s view on any matter within the decision-making authority of the JSC, the representatives of the Parties on the JSC cannot reach an agreement as to such matter within [*] after such matter was brought to the JSC for resolution or after such matter has been referred to the JSC, such disagreement shall be referred to the Chief Executive Officer of Everest and Chief Executive Officer of Licensee (collectively, the “Executive Officers”) for resolution. If the Executive Officers cannot resolve such matter within [*] after such matter has been referred to them, then:
(i)With respect to the initial Development Plan and any amendment to the Development Plan related to any Global Trial, Licensee shall be entitled to make final decisions, provided that (A) any Global Trial to be conducted pursuant to any Development Plan shall be designed to be China Sufficient in accordance with Section 4.2 (Development Plan), and (B) no Development Budget shall be binding on the Parties unless agreed to by the Parties (which, for clarity, shall not limit Licensee’s final decision-making authority with respect to any amendment to the Development Plan related to any Global Trial, provided that (x) Licensee pays for any increased costs associated with such amendment that are not included in a mutually-approved Development Budget and (y) such amendment does not increase Everest’s operational obligations with respect to such Global Trial);
(ii)With respect to any matter within the decision-making authority of the JSC other than the initial Development Plan and any amendment to the Development Plan related to any Global Trial:
(A)Licensee shall be entitled to make final decisions with respect to the Development or Commercialization of the Licensed Compound and Licensed Products in the Field in the Territory; provided, however, that Licensee shall not exercise such final decision-making authority if such decision would be reasonably expected to (x) materially adversely affect the Development or Commercialization of the Licensed
21
Products in the Retained Territory, or (y) increase Everest’s financial, operational or other obligations under the Development Plan; and
(B)Everest shall be entitled to make final decisions with respect to the Development or Commercialization of the Licensed Compound and Licensed Products in the Retained Territory, provided, however, that Everest shall not exercise such final decision making authority if such decisions would be reasonably expected to materially adversely affect the Development or Commercialization of Licensed Products in the Territory.
(iii)Limitations to Final Decision-Making Authority. Notwithstanding any provisions to the contrary in this Section 3.6 (Decision-Making), a Party’s exercise of its final decision-making authority under this Section 3.6 (Decisions-Making) must comply with the following:
(A)such Party’s decision must be consistent in all material respects with the terms and conditions of this Agreement;
(B)such Party may not impose any obligation on the other Party or waive any right or remedy of the other Party under this Agreement; and
(C)such Party may not require the other Party to violate any Applicable Law or ethical requirement.
(b)Subcommittee Decisions. Except where consent or approval on any matter by a Party is expressly required herein, if after reasonable discussion and good faith consideration of each Party’s view on any matter within the decision-making authority of the applicable Subcommittee, the representatives of the Parties on such Subcommittee cannot reach an agreement as to such matter within [*] after such matter was brought to such Subcommittee for resolution or after such matter has been referred to such Subcommittee, such disagreement shall be escalated to the JSC.
3.7Limitations on Authority. The JSC shall have only such powers as are expressly assigned to it in this Agreement, and such powers shall be subject to the terms and conditions of this Agreement. Without limiting the generality of the foregoing, the JSC shall not have the power to amend, modify, waive any provision of this Agreement, determine whether or not a Party has complied with any of its obligations under this Agreement, make any decision or approve any matter that is expressly stated to require the mutual agreement of the Parties or the written consent of one or both Parties and no decision of the JSC may be in contravention of any terms and conditions of this Agreement. For clarity, except as set forth in Section 3.1(b) (Joint Steering Committee), the JSC shall have no decision-making authority regarding any Development, Manufacturing, Commercialization or other activity in the Retained Territory.
3.8Meetings. Each Committee shall hold meetings at such times as it elects to do so, but will meet no less frequent than every [*]. Such meetings may be in person, via videoconference, or via teleconference. The location of in-person meetings shall be determined by the Parties. Each Party will bear its own expenses related to the attendance of the Committee meetings by its representatives. At least [*] prior to each Committee meeting, the co-chairpersons of the applicable Committee or their designees shall prepare and circulate agenda items for discussion at such meeting, together with appropriate information related thereto. Reasonably detailed written minutes shall be kept for all Committee meetings. Meeting minutes shall be prepared on an alternating basis by the co-chairpersons of the applicable Committee or
22
their designees and sent to each member of the applicable Committee for review and approval within [*] after the meeting. Minutes shall be deemed approved by a member of the applicable Committee unless such member objects to the accuracy of such minutes within [*] of receipt.
3.9Discontinuation of Committee. A Committee shall continue to exist until the Parties mutually agree to disband such Committee. Once the JSC is disbanded, all other Committees shall disband, the JSC shall have no further obligations under this Agreement and, thereafter, the Alliance Managers shall be the points of contact for the exchange of information under this Agreement, and the Parties shall reach decision directly on matters that are subject to the decision of the JSC as set forth in Section 3.1 (Joint Steering Committee) subject to the same respective decision-making rights and limitations set forth in Section 3.6 (Decision-Making) and the other terms and conditions of this Agreement.
3.10Alliance Managers. Each Party shall appoint an individual, who is an employee of such Party, to act as its alliance manager under this Agreement within [*] after the Effective Date (the “Alliance Manager”). The Alliance Managers shall: (a) serve as the primary points of contact between the Parties for the purpose of providing the other Party with information on the progress of a Party’s activities under this Agreement; (b) be responsible for facilitating the flow of information and otherwise promoting communication, coordination and collaboration between the Parties; (c) facilitate the prompt resolution of any disputes; and (d) attend JSC and Subcommittee meetings (in each case, as a non-voting participant); provided that the Alliance Managers shall not count toward the number of representatives that each Party may have on each such Committee. An Alliance Manager may also bring any matter to the attention of the JSC, if such Alliance Manager reasonably believes that such matter warrants such attention. Each Party may replace its Alliance Manager at any time upon written notice to the other Party.
Article 4
DEVELOPMENT
DEVELOPMENT
4.1Overview; Diligence. As between the Parties, Licensee shall be responsible for the Development of Licensed Products in the Field in the Territory. Subject to this Article 4 (Development) and Section 3.6 (Decision-Making), Licensee (itself or through its Affiliates and Sublicensees) shall use Commercially Reasonable Efforts to (i) Develop and, seek and obtain Regulatory Approval for at least one (1) Licensed Product in the Field in each of the Major Market Countries in accordance with this Agreement and the Development Plan. Each Party shall (a) perform the Development activities assigned to it under the Development Plan and in compliance with all Applicable Laws (including GCP) and (b) use Commercially Reasonable Efforts to perform the activities assigned to it under the Development Plan pursuant to the timeline set forth in the Development Plan, provided that subclause (b) shall not apply to any activities to be conducted by Everest solely for the benefit of the Retained Territory. Everest shall (i) perform and Complete the Ongoing Clinical Trial, unless suspension or termination of the Ongoing Clinical Trial is required to comply with the order of any Regulatory Authority, at [*] cost and expense, and (ii) perform and complete any CMC development and manufacturing process activities allocated to Everest within the then-current Development Plan that are necessary to conduct the first Phase III Clinical Trial for a Licensed Product; provided that for any such activities in clause (ii), Licensee shall reimburse [*] of Everest’s costs for such activities up to [*] and Everest shall bear all other costs in connection with such activities.
4.2Development Plan. The JSC shall review, discuss, and approve an initial Development Plan (including an initial Development Budget) promptly following the Effective Date (within [*] of the Effective Date), for the Development activities, including with respect to each Planned Global Trial, to be conducted by or on behalf of Licensee and Everest as part of a global Development program in order to obtain Regulatory Approvals for the Licensed Products in the Territory and outside the Territory. Once approved by the JSC, the Development Plan
23
shall be signed by both Parties and incorporated under this Agreement by reference. The Development Plan shall include, at a minimum, (a) all Indications then being pursued by the Parties; (b) a description of the Development and regulatory activities to be conducted by each Party, and the relevant deliverables; (c) a description of any Clinical Trial of any Licensed Product to be conducted by or on behalf Everest to obtain and maintain Regulatory Approval for such Licensed Product in the Field outside of the Territory, including the relevant Indication, dose and dosing schedule for Licensed Product and inclusion/exclusion criteria, in each case, for such Clinical Trial; (d) a Development Budget for the Development Costs subject to Development Cost Sharing and Reimbursement, including Other Trial Development Costs and Development Costs to be incurred by Everest for Development activities to be conducted by Everest thereunder; and (e) an estimated timeline for the performance of the Development activities to be conducted thereunder. From time to time during the Term, either Party may propose written amendments and updates to the then-current Development Plan and Development Budget, including with respect to any global Development activities to be conducted by or on behalf of the Parties for any Additional Global Trials in the Field in the Territory and outside the Territory, and shall submit such amendments and updates to the JSC for review and approval. The Parties agree that any Planned Global Trial and any Additional Global Trial to be conducted pursuant to the Development Plan shall be designed to support the Regulatory Approvals of the Licensed Products both in the U.S. and Mainland China (for example, with respect to Mainland China, the Development Plan shall require that a set percentage of total patients enrolled in such Global Trial shall be contributed from Mainland China consistent with the regulatory requirements in Mainland China for the results of such Global Trial to be sufficient to support Regulatory Approval in Mainland China without the need for the conduct of any additional Clinical Trial) (“China Sufficient”). The Parties further agree that, Everest may, at its sole discretion, decline to participate in any Additional Global Trial proposed by Licensee, and, to the extent Everest has declined to participate in any proposed Phase II Clinical Trial for an Indication in the Field, such that such Phase II Clinical Trial will not be deemed a Global Trial under this Agreement, Everest may elect, at its discretion, to participate in any subsequent Phase III Clinical Trial or Pivotal Clinical Trial for the same Indication in the Field under the Development Plan and such subsequent Phase III Clinical Trial or Pivotal Clinical Trial will be deemed a Global Trial under this Agreement, provided upon such election, Everest is obligated to comply with the payment obligation set forth in Section 5.7(a) in relation to such Phase II Clinical Trial. Each amendment or update to the Development Plan will become effective and will supersede such previous Development Plan upon approval by the JSC. Each Party shall be responsible for conducting any Development activities assigned to it under such Development Plan in compliance with Applicable Law. For the avoidance of doubt, neither Everest nor Licensee shall conduct any material Development activities for Licensed Products that are not set forth in the then-current Development Plan.
4.3Development Costs.
(a)Development Cost Sharing.
(i)Ongoing Clinical Trial Development Costs. Everest will be solely responsible for any Development Costs incurred in connection with the Ongoing Clinical Trial (“Ongoing Clinical Trial Development Costs”).
(ii)Other Trial Development Costs. With respect to any Global Trial Development Costs and Basket Renal Trial Development Costs, including any related Permitted Development Overrun, (collectively, the “Other Trial Development Costs”) incurred by or on behalf of either Party in connection with the conduct of any Global Trial or Basket Renal Trial consistent with the Development Budget, the Parties shall share such Other Trial Development Costs such that Everest shall be responsible for [*] of total Other Trial Development
24
Costs, and Licensee shall be responsible for [*] of total Other Trial Development Costs. For the avoidance of doubt, the Other Trial Development Cost sharing structure as set forth in this Section 4.3(a)(ii) (Other Trial Development Costs) shall (A) apply to any Global Trial for any Indication to be pursued by the Parties under this Agreement and (B) include the COGS (without mark-up) for Manufacturing Licensed Products for any Global Trial or the Basket Renal Trial.
(iii)Other Development Costs. Except as set forth in Section 4.1 (Overview; Diligence); Section 4.3(a)(i) (Ongoing Clinical Trial Development Costs), Section 4.3(a)(ii) (Other Trial Development Costs) and Section 4.3(c) (Permitted Development Overrun), Licensee shall be solely responsible for any Development Costs incurred by or on behalf of either Party for the conduct of Development activities pursuant to the Development Plan for any Indication. To the extent that Everest incurs any Development Cost, other than costs as set forth in Section 4.1 (Overview; Diligence), Ongoing Clinical Trial Development Costs or Other Trial Development Costs, including any Permitted Development Overrun, consistent with the Development Budget, Licensee shall reimburse Everest for such Development Costs pursuant to Section 4.3(d) (Cost Reconciliation) excluding any overspend that is not a Permitted Development Overrun.
(b)Development Budget. The JSC shall review, discuss, and approve an initial Development Budget for the conduct of Development activities (including for each Planned Global Trial) pursuant to the initial Development Plan that are expected to incur Development Costs, which are subject to the Development Cost Sharing and Reimbursement. Each Party may, from time to time during the Term, provide the JSC (for review, discussion and approval) an amendment to the Development Budget in connection with any amendment to the Development Plan, reflecting any additional proposed Development activities that were not previously included as Development activities in the then-current Development Plan and Development Budget for review and approval. For clarity, only Development Costs that are incurred pursuant to the Development Budget, including any Permitted Development Overrun, shall be subject to Development Cost Sharing and Reimbursement.
(c)Permitted Development Overrun. Each Party shall use Commercially Reasonable Efforts to ensure that the actual Development Costs that are subject to Development Cost Sharing and Reimbursement incurred during the performance of Development activities under the Development Plan for a given Calendar Year do not exceed [*] of the budgeted Development Costs allocated to such Party for such Calendar Year as set forth in the applicable Development Budget. If, in any Calendar Year, a Party anticipates exceeding the budgeted Development Costs set forth in the Development Budget for the Development activities allocated to it thereunder for such Calendar Year, then such Party shall provide to the JSC a full explanation for anticipating exceeding the amount set forth in such Development Budget for such Development activities for such Calendar Year and propose an amendment to the applicable Development Budget to include such anticipated costs and expenses. If and to the extent that any such overspend (i) (A) does not exceed [*] of the budgeted Development Costs set forth in the applicable Development Budget as first approved by the JSC for such Calendar Year; and (B) does not result from the gross negligence or willful misconduct of, or breach of this Agreement by, such Party or any of its Affiliates or Sublicensees or (ii) is otherwise approved by the Parties (each, a “Permitted Development Overrun”), then such Permitted Development Overrun shall be included in the total Development Costs in the Development Budget and shared by the Parties as set forth in Section 4.3(a)(ii) (Other Trial Development Costs) or reimbursed by Licensee to Everest pursuant to Section 4.3(a)(iii) (Other Development
25
Costs), as applicable. To the extent that any overspend is not a Permitted Development Overrun, such excess costs and expenses shall be solely borne by the Party responsible for performing such Development activities.
(d)Development Cost Reconciliation. The Parties shall negotiate in good faith a reasonable procedure pursuant to which, on a [*] basis, the Parties shall reconcile any payment for Development Costs that are subject to Development Cost Sharing and Reimbursement, such that Everest and Licensee shall share such Development Costs as set forth in Section 4.3(a)(ii) (Other Trial Development Costs) and Section 4.3(a)(iii) (Other Development Costs). The Parties agree to, in connection with such Development Cost reconciliation, provide reasonable documentation to the other Party supporting the Development Costs actually incurred by such Party. In addition, each Party will provide to the other Party, within [*] after the end of each [*], a good faith estimate of the Development Costs incurred by such Party during the preceding [*] that are subject to Development Cost Sharing and Reimbursement.
4.4Technology Transfer. Within [*] after Everest’s receipt of the Upfront Payment, Everest shall, at its cost, provide to Licensee one (1) electronic copy of the documents embodying Everest Know-How Controlled by and in the possession of Everest or any of its Affiliates as of the Effective Date and set forth in Schedule 4.4 (List of Initial Technology Transfer) in its original language (“Initial Technology Transfer”) except any Everest Know-How solely related to Manufacturing, which shall be transferred pursuant to Section 6.1 (Manufacturing Technology Transfer). A description such documents, and any English translations thereof to be provided by Everest, for the Initial Technology Transfer is attached as Schedule 4.4 (List of Initial Technology Transfer) hereto. On a [*] basis, Everest shall keep Licensee reasonably informed of the existence of any additional Everest Know-How that becomes Controlled by Everest after the Effective Date and during the Term, to the extent that such Everest Know-How has not previously been provided or made available to Licensee. The Parties shall discuss the means of the transfer of such additional Everest Know-How and Everest shall provide electronic copies of such additional Everest Know-How to Licensee in a manner in line with that for the Initial Technology Transfer and at no additional cost to Licensee. In addition, as reasonably requested by Licensee, Everest shall provide reasonable assistance to Licensee so as to assist Licensee to practice the Everest Licensed IP in connection with the Exploitation of the Licensed Compound and Licensed Products in the Field in the Territory.
4.5Development Records. Each Party shall maintain or cause to be maintained complete, current and accurate records of all activities conducted by or on behalf of it, its Affiliates or Sublicensees pursuant to this Agreement, and all Know-How and other information resulting from such activities. Such records shall fully and properly reflect all work done and results achieved in the performance of the Development activities in good scientific manner appropriate for regulatory and patent purposes. Each Party shall, and shall ensure that its Affiliates and Sublicensees shall, document all nonclinical studies and Clinical Trials in formal written study records in accordance with all Applicable Laws. Subject to Section 5.7 (Data Access; Right of Reference) and to the extent permitted by the Applicable Laws, each Party shall have the right to review and copy such records of the other Party at reasonable times and to obtain access to review the original to the extent necessary or useful for regulatory, patent or other reasonable purposes under this Agreement upon reasonable notice to such other Party and at a time and location mutually acceptable to the Parties.
4.6Development Updates. Each Party shall keep the other Party reasonably and timely informed of the progress and results of its and its Affiliates’ and Sublicensees’ Development activities under this Agreement (including prompt reporting of available non-clinical and clinical data). Without limiting the generality of the foregoing, at each JSC meeting, each Party will provide to the JSC an update on the status, progress and results of the
26
Development activities conducted by or on behalf of such Party and its Affiliates and (sub)licensees pursuant to this Agreement. Such updates shall be provided at a level of detail reasonably sufficient to enable each Party to determine the other Party’s compliance with its diligence obligations under Section 4.1 (Overview; Diligence). Meeting minutes prepared pursuant to Section 3.8 (Meetings) shall memorialize such update with respect to such Development activities. Each Party shall promptly respond to the other Party’s reasonable questions or requests for additional information relating to such Development activities.
4.7Clinical Trial Audits. Licensee or its representatives may [*] itself or through a Third Party expert, selected by Licensee and reasonably acceptable to Everest, conduct an audit of any Clinical Trial site in the Territory engaged by Everest or its Affiliates or its or their subcontractors to perform Everest’s obligations with respect to any Global Trial to ensure that the applicable Global Trials are conducted in compliance with the Development Plan and Applicable Laws. Licensee may conduct such audit no more than once per [*] (unless an additional audit is warranted for cause) upon [*] prior written notice to Everest. No later than [*] after the completion of such audit, Licensee or such Third Party expert, as applicable, shall provide Everest with a written summary of its findings of any deficiencies or other areas of remediation that it identifies during any such audit. Everest shall use Commercially Reasonable Efforts to respond or remediate any such deficiencies following its receipt of such report. In the event that Licensee or any of its Affiliates or Sublicensees conducts an audit of any Clinical Trial site in the Territory, to the extent permitted under Applicable Laws and subject to any confidentiality obligations of Licensee, Licensee shall promptly share the results and findings of such audit with Everest following its completion.
Article 5
REGULATORY
REGULATORY
5.1Overview of Conduct of Regulatory Activities. Licensee (itself and through its Affiliates and Sublicensees, as applicable) shall be responsible, at its expense, for all regulatory activities with respect to the Licensed Products in the Field in the Territory. Subject to Section 5.2 (Ownership of Regulatory Approvals and Regulatory Materials), Licensee shall be the holder of all Regulatory Approvals and Regulatory Materials for the Licensed Products in the Territory. Upon a Party’s reasonable request, the other Party shall reasonably cooperate with the requesting Party to prepare Regulatory Materials and obtain Regulatory Approvals for the Licensed Products in the requesting Party’s territory.
5.2Ownership of Regulatory Approvals and Regulatory Materials. As between the Parties, Licensee shall be the holder of all Regulatory Approvals and Regulatory Materials for the Licensed Compound and Licensed Products in the Field in the Territory. To the extent permissible under the Applicable Laws, Everest shall transfer and assign, or shall cause the transfer or assignment, to Licensee or its designees or Affiliates, the entire right, title, and interest in and to all Regulatory Approvals, if any, and transfer any Regulatory Materials with respect to the Licensed Compound and Licensed Products in the Territory that are owned and assignable by Everest or any of its Affiliates, unless the Parties agree otherwise.
5.3Review of Regulatory Materials. Everest shall provide to Licensee (a) for review English language drafts of all material Regulatory Materials, including drafts of MAAs, for the Licensed Compound or Licensed Products outside of the Territory no later than [*] prior to the planned submission, provided, however, that as part of such drafts, Everest will not be obligated to provide in English any portion of such Regulatory Materials that is prepared based on Regulatory Materials previously provided by Licensee to Everest and (b) any material Regulatory Materials, including MAAs, for the Licensed Compound or Licensed Products outside of the Territory submitted to or received from any Regulatory Authority outside of the Territory within [*] after submission or receipt thereof.
27
5.4Notice of Meetings. Everest shall provide Licensee with notice of any meeting or discussion with any Regulatory Authority outside of the Territory related to the Licensed Compound or a Licensed Product no later than [*] after receiving notice thereof. If any such meeting or discussion would reasonably be expected to materially affect the Exploitation or regulatory status of Licensed Compounds or Licensed Products in the Territory, Everest may, in its discretion, not to be unreasonably withheld, conditioned or delayed, invite Licensee or its designee to attend and participate in such meeting or discussion on an as-needed basis, to the extent permitted by Applicable Laws or Regulatory Authority. If Licensee does not attend such meeting or discussion, then Everest shall provide to Licensee a written summary thereof in English promptly following the issuance or approval of the corresponding official minutes by the applicable Regulatory Authority.
5.5Regulatory or Third Party Action or Inspection. Everest shall promptly notify Licensee in writing if Everest becomes aware of any Regulatory Authority inspections relating to any Licensed Compound or Licensed Product outside of the Territory. If any such inspection would reasonably be expected to materially affect the Exploitation or regulatory status of Licensed Compounds or Licensed Products in the Territory, to the extent permitted by Applicable Laws or Regulatory Authority, Everest may, in its discretion, not to be unreasonably withheld, conditioned or delayed, invite Licensee to be present at such inspections and to review and comment on any responses that may be required in connection with such inspections. If Everest does not receive prior notice of any such inspection, Everest shall notify Licensee in writing as soon as practicable after such inspection and shall provide Licensee with copies of all materials, correspondence, statements, forms and records received or generated pursuant to any such inspection. In addition to such obligations with respect to Regulatory Authority inspections, Everest shall promptly notify Licensee in writing of any information it receives regarding any threatened or pending action or communication by or from any Third Party, including a Regulatory Authority outside of the Territory, that would reasonably be expected to materially affect the Exploitation or regulatory status of Licensed Compounds or Licensed Products in the Territory.
5.6Interactions with Regulatory Authorities. Except as set forth herein or as otherwise agreed by the Parties, (a) neither Party shall communicate with any Regulatory Authority having jurisdiction outside of its respective territory with respect to the Licensed Compound or Licensed Products, unless required by such Regulatory Authority, in which case such Party shall notify the other Party of such requirement within [*] of such communication, and (b) neither Party shall submit any Regulatory Materials nor seek Regulatory Approval for the Licensed Products in the other Party’s respective territory.
5.7Data Access; Right of Reference; Access to Regulatory Materials.
(a)Each Party shall own any and all Data generated in all Clinical Trials by such Party in its respective territory under the Development Plan. Except to the extent prohibited by Applicable Law, Everest shall and does hereby grant to Licensee and its Affiliates a right to access and utilize the Data Controlled by Everest, its Affiliates or Sublicensees for the purpose of Exploiting the Licensed Compound or Licensed Products in the Field in the Territory [*]. Except to the extent prohibited by Applicable Law, Licensee shall and does hereby grant to Everest and its Affiliates and other licensees a right to access and utilize the Data Controlled by Licensee or its Affiliates or Sublicensees, solely for the purpose of Exploiting the Licensed Compound or Licensed Products in the Field in the Retained Territory [*], provided that, with respect to any Data Controlled by Licensee or its Affiliates or Sublicensees and generated from any Global Trial proposed by Licensee and in which Everest declined to participate, Licensee shall not, and does not, grant to Everest and its Affiliates and other licensees the right to access and utilize such Data for the purpose set forth in this Section 5.7(a), unless and until
28
Everest makes a payment to Licensee for an amount equal to [*] of the Other Trial Development Costs incurred in the activities that generated such Data for which Everest would have been responsible pursuant to Development Cost Sharing and Reimbursement set forth in this Agreement, had Everest agreed to participate in such Global Trial, provided that, in any case, Licensee shall share with Everest and shall, and does hereby, grant to Everest a right of reference to, any safety data generated from any Clinical Trials [*].
(b)Each Party shall, and does hereby, grant to the other Party a right of reference to all Regulatory Materials related to the Licensed Compound and Licensed Products in the Field Controlled by such Party or its Affiliates or Sublicensees, including all applications for Regulatory Approval, in such first Party’s territory, for use in regulatory filings related to the Licensed Compound and Licensed Products, including applications for Regulatory Approval, in such other Party’s territory; provided that, with respect to any such Regulatory Materials for or based on any Global Trial proposed by Licensee and for which Everest declined to participate, except for safety data as set forth in Section 5.7(a), Licensee shall not, and does not, grant to Everest and its Affiliates the right of reference set forth in this Section 5.7(b) unless and until Everest makes a payment to Licensee for an amount equal to [*] of the Other Trial Development Costs incurred in the activities that generated such Data for which Everest would have been responsible pursuant to Development Cost Sharing and Reimbursement set forth in this Agreement, had Everest agreed to participate in such Global Trial. If requested by the other Party, such first Party shall, and shall cause its Affiliates or Sublicensees to, provide a signed statement to this effect in accordance with Applicable Laws for submission to the applicable Governmental Authorities.
(c)Notwithstanding any other term or condition of this Agreement, neither Licensee, nor any of its Affiliates nor Sublicensees, shall have any obligation under this Agreement to transfer or provide “access” (as defined at 28 C.F.R. § 202.201) to Everest, Everest’s Affiliates or Everest’s Sublicensees any Personal Data or other data or materials if such transfer or provision of access is (i) a “covered data transaction” as defined under the U.S. Bulk Data Final Rule and not exempt under 28 C.F.R. § 202.510, § 202.511, or another exemption under the U.S. Bulk Data Final Rule, or (ii) if with a “foreign person” (as defined at 28 C.F.R. § 202.221) that is a non-covered person, not compliant with the onward transfer requirements under 28 C.F.R. § 202.302. If neither clause (i) nor clause (ii) apply to the transfer or provision of access of Personal Data or other data or materials, then Licensee shall implement such transfer or provision of access to Everest, Everest’s Affiliates or Everest’s Sublicensees in accordance with this Agreement, provided that Licensee shall have the right to take all reasonable measures to ensure its compliance with the U.S. Bulk Data Final Rule. To the extent that Licensee determines not to transfer Personal Data or other data or materials pursuant to this Section 5.7(c), it shall promptly notify Everest [*]. Everest shall provide all reasonably necessary assistance in complying with Licensee’s obligations under the U.S. Bulk Data Final Rule to the extent applicable to the Parties’ obligations and rights under this Agreement.
5.8Pharmacovigilance. Prior to [*], the Parties shall enter into a pharmacovigilance agreement regarding the Licensed Compound and Licensed Products (the “Pharmacovigilance Agreement”), which shall set forth standard operating procedures governing the collection, investigation, adverse events reporting, and exchange of safety information sufficient to permit each Party to comply with its regulatory and other legal obligations and business needs within the applicable timeframes. In the event of any inconsistency between the terms of this Agreement and the Pharmacovigilance Agreement, the terms of this Agreement shall prevail and govern, except to the extent such conflicting terms relate directly to the pharmacovigilance
29
responsibilities of the Parties (including the exchange of safety data), in which case the terms of the Pharmacovigilance Agreement shall prevail and govern. Prior to the Parties entering into the Pharmacovigilance Agreement, Everest is and shall continue to be the owner of the global safety database for the Licensed Products. After the Parties enter into the Pharmacovigilance Agreement, Everest shall promptly transfer the global safety database for the Licensed Products to Licensee, and Licensee shall be the owner of and responsible for maintaining the global safety database and shall enter into such database all pharmacovigilance and other drug safety data for the Licensed Products used in the Territory as required by Applicable Laws (including any such data collected by its Sublicensees, as applicable), and Everest will notify Licensee of all pharmacovigilance and other drug safety data for the Licensed Products used in the Retained Territory (including any such data collected by its Sublicensees, as applicable) as set forth in the Pharmacovigilance Agreement. Each Party shall have the right to assess all drug safety data from the global safety database necessary for such Party to comply with all Applicable Laws in such Party’s territory.
5.9Remedial Actions. Each Party shall notify the other Party [*], and [*] confirm such notice in writing, if it obtains information indicating that any Licensed Product may be subject to any recall, corrective action or other regulatory action by any Governmental Authority or Regulatory Authority (a “Remedial Action”). The Parties shall assist each other in gathering and evaluating such information as is necessary to determine the necessity of conducting a Remedial Action. The Parties shall collaborate in good faith and endeavor to reach an agreement on Remedial Action related decisions, provided that each Party shall have the right to determine to conduct a Remedial Action in its respective territory.
Article 6
MANUFACTURE & SUPPLY
MANUFACTURE & SUPPLY
6.1Manufacturing Technology Transfer. Following the Effective Date, upon Licensee’s reasonable advance request, Everest shall transfer the Everest Know-How that are necessary or reasonably useful for the Manufacture of the Licensed Compound to Licensee in accordance with a written manufacturing technology transfer plan to be reasonably agreed by the Parties and provide reasonable technical assistance to Licensee in a timely manner to enable Licensee to Manufacture the Licensed Product by itself or through a CMO (the date on which Licensee is able to Manufacture the Licensed Products by itself or through a CMO, the “Manufacturing Technology Transfer Completion”). The first [*] of such technical assistance shall be provided at [*] cost and any additional technical assistance shall be provided at [*] cost. For the avoidance of doubt, the foregoing shall only apply to any Internal Costs incurred by Everest in connection with such technical assistance, and any Out-of-Pocket Costs, including any travel and lodging expenses, incurred by Everest in connection with such technical assistance shall be borne by [*]; provided that Everest shall obtain Licensee’s prior written approval for any such Out-of-Pocket Costs.
6.2Clinical Supply. Within [*] after the Effective Date, the Parties shall negotiate in good faith and enter into a written agreement, pursuant to which, Everest shall, by itself or through one or more CMO(s), provide clinical supply of the Licensed Products to Licensee for any clinical Development at a price equal to Everest’s COGS plus a markup of [*] prior to the Manufacturing Technology Transfer Completion; provided that the price of clinical supply of the Licensed Products for any Global Trial or the Basket Renal Trial shall be subject to the Development Cost Sharing and Reimbursement pursuant to Section 4.3(a)(ii) (Other Trial Development Costs).
6.3Commercial Supply. To the extent reasonably requested by a Party, the Parties shall negotiate in good faith and may, if the Parties agree on mutually acceptable terms, enter into a written agreement, pursuant to which, either Party shall, by itself or through one or more
30
CMO(s), provide commercial supply of the Licensed Products to other Party at a price equal to the supplying Party’s COGS plus a markup of [*].
Article 7
COMMERCIALIZATION MATTERS
COMMERCIALIZATION MATTERS
7.1Overview; Diligence. Subject to the terms and conditions of this Agreement (including the diligence obligations set forth below), Licensee (itself and through its Affiliates and Sublicensees, as applicable) shall be solely responsible for the Commercialization of the Licensed Products in the Field in the Territory, including: (a) developing and executing a commercial launch and pre-launch plan, (b) negotiating with applicable Governmental Authorities regarding the price and reimbursement statuses of the Licensed Products; (c) marketing, advertising and promotion; (d) booking sales and distribution and performance of related services; (e) handling all aspects of order processing, invoicing and collection, inventory and receivables; (f) providing customer support, and performing other related functions; and (g) conforming its practices and procedures to Applicable Laws relating to the marketing, detailing and promotion of the Licensed Products in the Field in the Territory. Licensee shall bear all of the costs and expenses incurred in connection with such Commercialization activities. Licensee shall use Commercially Reasonable Efforts to Commercialize the Licensed Products in each Major Market Country.
7.2Commercialization Plan. No later than [*] before the anticipated date of the submission of the first MAA for a Licensed Product in the Territory, Licensee shall submit to the JSC for review and discussion a written strategic plan that sets forth a summary of all major Commercialization activities planned for such Licensed Product in the Territory (the “Commercialization Plan”). Thereafter, from time to time, but at least [*], Licensee shall prepare updates or amendments to the Commercialization Plan to reflect changes in such plan, including those in response to changes in the marketplace, relative success of the Licensed Product, and other relevant factors influencing such plan and activities, and submit such updated or amended plan to the JSC for review and discussion.
7.3Coordination of Commercialization Activities. The Parties recognize that they may benefit from the coordination of certain activities in support of the Commercialization of a Licensed Product across their territories. As such, the Parties may coordinate such activities where appropriate, including scientific and medical communication and Licensed Product positioning. For clarity, Licensee shall not conduct any Commercialization of any Licensed Product outside the Territory without Everest’s express prior written consent.
7.4Commercialization Reports. Licensee shall update the JSC at each regularly scheduled JSC meeting regarding its material Commercialization activities with respect to Licensed Products in the Field in the Territory by providing a written report (by means of a slide presentation or otherwise) summarizing the material Commercialization activities undertaken by or on behalf of Licensee for the Licensed Products in the Field in the Territory sufficient to enable Everest to determine Licensee’s compliance with this diligence obligations pursuant to Section 7.1 (Overview; Diligence) and the future material Commercialization activities it expects to initiate for the Licensed Products during the following [*] period.
7.5Trademarks.
(a)The Parties shall discuss in good faith the use of any global trademarks to be used in connection with the Commercialization of the Licensed Products in the Field throughout the world (the “Global Trademarks”). Licensee shall own all rights in the Global Trademarks and shall be responsible for the registration, filing, maintenance and enforcement of the Global Trademarks in any country in the world at [*] cost and
31
expense. Everest has the right, but not the obligation, to market and sell the Licensed Products outside the Territory under such Global Trademarks. If Everest decides, in its sole discretion, to use the Global Trademarks, the Parties shall negotiate in good faith and [*] enter into a license agreement, pursuant to which Licensee shall grant to Everest a [*] non-exclusive, sub-licensable license to use the Global Trademarks solely in connection with the Commercialization of the Licensed Products in the Field outside the Territory during the Term of this Agreement.
(b)In addition to (or in lieu of) the Global Trademarks, Everest shall have the right to brand the Licensed Products in the Field outside the Territory with trademarks selected by Everest, including those associated with Everest’s name or corporate identity (the “Everest Trademarks”); provided that Everest shall not, and shall cause its Affiliates and their respective (sub)licensees to not, select or make any use of trademarks that are confusingly similar to any trademarks or housemarks of Licensee or its Affiliates (including the corporate name of Licensee or any of its Affiliates). Everest shall own all rights in the Everest Trademarks, and have the sole right (but not the obligation) to register, maintain and enforce the Everest Trademarks outside the Territory as it determines appropriate, at [*] cost and expense.
7.6No Diversion. Each Party hereby covenants and agrees that during the Term, and except as expressly permitted by this Agreement, it shall not (and shall cause its Affiliates and Sublicensees and subcontractors not to), either itself or through a Third Party, develop, use, market, promote, import, export, sell or actively offer for sale the Licensed Products in the Field in the other Party’s territory. Without limiting the generality of the foregoing, except as mutually agreed by the Parties, each Party shall not (a) engage in any advertising activities relating to the Licensed Products in the Field directed primarily to customers in the other Party’s territory, or (b) actively or intentionally solicit orders from any prospective purchaser it knows is purchasing Licensed Products in the Field for the other Party’s territory. To the extent permitted by Applicable Laws, including applicable antitrust laws, if a Party receives any order for Licensed Products in the Field from a prospective purchaser located in a country or jurisdiction in the other Party’s territory, such Party shall promptly refer that order to the other Party and shall not accept any such order or deliver or tender (or cause to be delivered or tendered) the Licensed Products under such order. If a Party is aware that a customer or distributor is actively engaged itself or through a Third Party in the sale or distribution of the Licensed Products in the other Party’s territory, then such Party shall (i) promptly notify the other Party regarding such activities and (ii) cease sales to such customer or distributor, unless otherwise agreed in writing by the Parties.
Article 8
FINANCIAL TERMS
FINANCIAL TERMS
8.1Upfront Payment. In partial consideration of Everest’s granting of the licenses and rights to Licensee hereunder, Licensee shall make a one-time, non-refundable, non-creditable payment to Everest of one hundred twelve million, five hundred thousand U.S. Dollars ($112,500,000) (the “Upfront Payment”) within [*] after the Effective Date.
8.2Development and Regulatory Milestone Payments. With respect to the milestone events set forth in the table below, promptly following the first achievement of the corresponding milestone event by the first Licensed Product, Licensee shall notify Everest within [*] of such achievement, and Licensee shall pay to Everest the corresponding non-refundable, non-creditable milestone payment within [*] after the achievement of the applicable milestone event: [*]
32
8.3Pricing Approval Milestone Payment. Licensee shall pay to Everest the additional one-time, non-refundable, non-creditable pricing approval milestone payment of [*] upon the receipt of any Pricing and Reimbursement Approval for [*]. For clarity, no milestone payments under this Section 8.3 (Pricing Approval Milestone Payment) shall be payable in connection with any Indication other than [*] or any [*] Licensed Product for [*].
8.4Commercial Milestone Payments. Licensee shall pay to Everest the additional one-time, non-refundable, non-creditable commercial milestone payments set forth in the table below after the first achievement of each commercial milestone event described below: [*]
Within [*] after the end of the [*] (but within [*] after the end of the [*]) in which any commercial milestone event set forth above in this Section 8.4 (Commercial Milestone Payments) for which a commercial milestone payment payable pursuant to this Section 8.4 (Commercial Milestone Payments) is achieved, Licensee shall deliver a written notice to Everest of such achievement, and Licensee shall pay to Everest the corresponding commercial milestone payment within [*] after the end of such [*] (but within [*] after the end of the [*]). For clarity, each of the commercial milestone payments set forth above in this Section 8.4 (Commercial Milestone Payments) shall be (a) payable only once and (b) additive such that if multiple commercial milestone events specified above are achieved in the same [*], then the commercial milestone payments for all such commercial milestone events shall be payable by Licensee.
8.5Royalties. Licensee shall pay tiered royalties to Everest, on a Licensed Product-by-Licensed Product and country-by-country basis, on Annual Net Sales of each Licensed Product in the Territory as set forth below, calculated by multiplying the applicable royalty rate by the corresponding portion of Annual Net Sales of such Licensed Product in the Territory: [*]
8.6Royalty Term. Royalties under Section 8.5 (Royalties) shall be payable, on a country-by-country and Licensed Product-by-Licensed Product basis, during the period beginning on the date of the First Commercial Sale of such Licensed Product in such country in the Territory and continuing until the latest of: (a) the expiration of the last-to-expire Valid Claim of the Everest Patents Covering the composition of matter, formulations, or all approved methods of use set forth in the Regulatory Approval of such Licensed Compound or Licensed Product in such country; (b) the expiration of the Regulatory Exclusivity of the applicable Licensed Product in such country; and (c) ten (10) years following the First Commercial Sale of such Licensed Product in such country (the “Royalty Term”), provided that, subject to the exclusion to Net Sales under Section 1.72 (“Net Sales”), any sales income received by Licensee, its Affiliates or Sublicensees for the Licensed Products accrued prior to First Commercial Sales will count toward the calculation of royalties even though such sales occur prior to the First Commercial Sale, but such royalties will not be payable until after First Commercial Sale and such sales will not be considered as a First Commercial Sale.
8.7Royalty Payment Reduction.
(a)Generic Product. On a country-by-country and Licensed Product-by-Licensed Product basis in the Territory, if a Generic Product with respect to such Licensed Product is being sold in such country, then the Net Sales of such Licensed Product in such country shall be reduced as follows for the purposes of determining royalties due under Section 8.5 (Royalties): [*]
Market share data shall be based on data reported by [*].
33
(b)Third Party Licenses.
(i)If either Party identifies any Third Party Patent (or Patent and related Know-How) that, absent a license or agreement with such Third Party, is necessary for the Exploitation of a Licensed Product (and such Third Party Patent (or Patent and related Know-How) does not solely Cover (or solely relate to) any Additional Active incorporated in a Combination Product ) in the Field, whether in the Territory or outside of the Territory (“Blocking IP”), it shall notify the other Party in writing. Licensee, its Affiliates or any of their Sublicensees shall have the first right, but not the obligation, to enter into an agreement with such Third Party to acquire or obtain a license, covenant not to sue or other similar right to any Blocking IP. Any royalties payable by Licensee to Everest with respect to such Licensed Product shall be reduced by an amount equal to [*] of the amounts actually paid to such Third Party by Licensee, its Affiliates or Sublicensees on account of the Exploitation of such Licensed Product in the Territory. Upon Everest’s written request, and subject to the terms and conditions of Licensee’s agreement with the relevant Third Party, Licensee will grant a sublicense to Everest under the Blocking IP to Exploit Licensed Products in the Field outside of the Territory; provided that Everest shall be responsible for all amounts payable to such Third Party in connection with such sublicense granted to Everest. At least [*] prior to entering into such agreement, Licensee will provide Everest with the opportunity to review such Blocking IP owned or Controlled by such Third Party; and (ii) in the event Everest (or an Upstream Licensor pursuant to the Upstream License Agreement) reasonably disputes whether such Third Party license, covenant not to sue or other similar right is necessary during such [*] period, (A) the matter will be referred to the chief patent counsels of or patent attorneys engaged by Everest (or Upstream Licensor if the Upstream Licensor disputes) and Licensee, (B) the [*] shall meet promptly to discuss and resolve the matter within [*] of such matter being referred to them for resolution, and (C) if the [*] cannot agree on a resolution to the matter within such [*] period, then the Parties shall refer such matter for resolution to [*] mutually agreed upon by the Parties who has at least [*] for resolution within [*] of such matter first being referred to such [*], and such [*] decision on the matter shall be binding upon the Parties (and, for clarity, such matter shall not be subject to the dispute resolution procedures set forth in Article 15 (Dispute Resolution)). If such Third Party license is found unnecessary for Licensee to Exploit the Licensed Product in the Territory, such finding shall not prevent Licensee from entering into such Third Party license; provided that the royalty reduction mechanism described in this Section 8.7(b)(i) will not apply with respect to such Third Party license. Within [*] following the execution of any such Third Party agreement, Licensee shall provide Everest with a true and complete copy of such Third Party agreement (which copy may be redacted of any confidential or proprietary information that is not necessary for Everest to confirm or verify Licensee’s compliance with this Agreement).
(ii)If Licensee, its Affiliates and their Sublicensees elect not to enter into an agreement with respect to any Blocking IP pursuant to this Section 8.7(b) (Third Party Licenses), Everest may elect to enter into an agreement with respect to such Blocking IP; provided that Licensee, its Affiliates and their Sublicensees shall remain free to independently defend against any infringement claim brought against any of them by such Third Party, negotiate and enter into an agreement to obtain a license or other rights to such Blocking IP in connection with such claim; provided that Everest will use reasonable efforts to negotiate the right for Everest to grant Licensee a sublicense to such Blocking IP hereunder. If Everest obtains a
34
license to any Blocking IP and grants a sublicense to Licensee under such Blocking IP, Licensee shall be responsible for the amounts payable to such Third Party in connection with such sublicense granted to Licensee; provided that any royalties payable by Licensee to Everest shall be reduced by an amount equal to [*] of the amounts actually paid to such Third Party by Licensee with respect to the applicable Licensed Product.
(c)Lack of Patent Protection. On a country-by-country and Licensed Product-by-Licensed Product basis in the Territory, if at any time during the Royalty Term with respect to a Licensed Product, there are no Valid Claims under the Everest Patents Covering the relevant Licensed Compound or such Licensed Product in such country in the Territory, then the Net Sales of such Licensed Product in such country shall be reduced by [*] for the purposes of determining the royalties due for such Licensed Product in such country under Section 8.5 (Royalties).
(d)Royalty Floor. Notwithstanding the foregoing, for any Calendar Quarter during the Royalty Term for a Licensed Product in a particular country in the Territory, the operation of all deductions set forth in Section 8.7(a) (Generic Product), Section 8.7(b) (Third Party License Payments) and Section 8.7(c) (Lack of Patent Protection), individually or in combination, shall not reduce the final royalty payment to less than [*] of the royalties otherwise payable to Everest for such Licensed Product pursuant to Section 8.5 (Royalties) prior to the application of any reductions pursuant to Section 8.7(a) (Generic Product), Section 8.7(b) (Third Party License Payments) or Section 8.7(c) (Lack of Patent Protection), in each case, in such country during such Calendar Quarter (“Royalty Floor”).
Article 9
PAYMENT; RECORDS; AUDITS
PAYMENT; RECORDS; AUDITS
9.1Payment; Reports. Royalties shall be calculated and reported for each Calendar Quarter and shall be paid within [*] after the end of each Calendar Quarter; provided that (i) Everest shall use commercially reasonable efforts to amend the Upstream License Agreement to provide for Everest to submit royalty reports for Net Sales under this Agreement to the Upstream Licensors at least [*] after the end of a Calendar Quarter (and to submit royalty reports at least [*] after the end of the last Calendar Quarter in a Calendar Year) and, upon execution of such amendment, Licensee shall calculate, report and pay to Everest the royalties for each Calendar Quarter within [*] after the end of each Calendar Quarter (and within [*] after the end of the last Calendar Quarter in a Calendar Year) and (ii) if (a) Everest fails to execute the amendment to the Upstream License Agreement with the Upstream Licensors as contemplated under subclause (i), (b) royalties are calculated based on [*] for a given Calendar Quarter in accordance with Licensee’s standard accounting practices, and (c) a recalculation of royalties for such Calendar Quarter by Licensee based on its [*] in accordance with Licensee’s standard accounting timelines indicates a different royalty amount due for such Calendar Quarter, then (A) Licensee shall notify Everest of such discrepancy promptly after such recalculation (in any event within [*] after such recalculation), (B) Licensee shall be entitled to reduce royalty payment(s) for any future Calendar Quarter(s) by the amount of any overpayment and (C) Licensee shall pay Everest the amount of any underpayment for such Calendar Quarter and any interest payable for such amount under the Upstream License Agreement within [*] after such recalculation, and such payment shall not be subject to interest as set forth in Section 9.5 (Late Payments). Each payment shall be accompanied by a report of Net Sales of the Licensed Products by Licensee, its Affiliates and Sublicensees in sufficient detail to permit confirmation of the accuracy of the
35
payment made, including (1) the amount of gross sales and Net Sales on a Licensed Product-by-Licensed Product and country- by-country basis during such Calendar Quarter (including such amounts expressed in local currency and as converted to U.S. Dollars and the exchange rate used), (2) the type and amount of permitted deductions from gross sales to determine Net Sales and the total amount of such deductions, (3) the applicable royalty rates for each Licensed Product in each country or country in the Territory after applying any permitted deductions set forth above, and (4) a calculation of the royalties due to Everest for such Calendar Quarter. Except as may otherwise be expressly provided herein, Licensee shall not have the right to set off, withhold or make any deduction from any payment of royalties or other payments due to Everest hereunder for any reason whatsoever.
9.2Exchange Rate; Manner and Place of Payment. All payments hereunder shall be payable in U.S. Dollars. When conversion of payments from any foreign currency is required, such conversion shall be at an exchange rate equal to the weighted average of the rates of exchange for the currency of the country from which such payments are payable as published by The Wall Street Journal, Western U.S. Edition during the Calendar Quarter for which a payment is due. All payments owed under this Agreement shall be made by wire transfer in immediately available funds to a bank and account designated in writing by Everest, unless otherwise specified in writing by Everest.
9.3Taxes.
(a)Taxes on Income. Each Party shall be solely responsible for the payment of all taxes imposed on, or measured by, net income derived by such Party in the Territory in connection with the payments and activities contemplated hereunder.
(b)Tax Withholdings. [*].
(c)Tax Indemnification. Everest agrees that in the case that no withholding is made by Licensee in respect of the Upfront Payment under the Agreement and a tax authority makes a written claim or otherwise asserts in writing that a withholding tax should have been imposed and paid over to such tax authority with respect to the Upfront Payment (a “Claim”), Everest will bear and indemnify Licensee in respect of any withholding tax that is due in connection with such Claim (a “Tax Loss”), unless such Tax Losses resulted from the gross negligence or knowing willful misconduct of Licensee. Licensee will provide prompt notice of the receipt of such Claim and will keep Everest informed of correspondence with the tax authority. Licensee and Everest shall cooperate in good faith to contest, settle or pay over such amounts due to the tax authority as the Parties mutually agree; provided that (i) no settlement or payment shall be made to such tax authority without the consent of Everest, which consent shall not be unreasonably withheld, conditioned or delayed and (ii) Everest shall reimburse promptly Licensee for Licensee’s costs and expenses incurred in connection with the contest of any such Claim.
(d)Tax Cooperation. Each Party agrees to use commercially reasonable efforts to cooperate with the other Party in claiming refunds, reductions, or exemptions from tax payments, deductions or withholdings under any relevant agreement or treaty that is in effect. Everest shall provide Licensee with any tax forms or other documentation reasonably necessary in order for Licensee not to withhold or to withhold tax at a reduced rate under an applicable bilateral income tax treaty. If the taxes originally paid or otherwise borne by a Party are in whole or in part subsequently determined not to have been chargeable, each Party will take commercially reasonable steps to obtain a refund of these undue taxes from the applicable Governmental Authority or other tax authority and any amount of undue taxes repaid by such authority to the
36
receiving Party, as well as any tax forms or official documents issued by applicable Governmental Authority or other tax authority shall be transferred to the paying Party within [*] of receipt. In addition, prior to making any deduction or withholding in relation to a withholding tax, if a paying Party makes a determination on whether a certain type of payment to be made by such Party will or will not require withholding, such paying Party will promptly upon such determination provide notice to the other Party of such determination. If the other Party provides notice to the paying Party within [*] of receipt of such notice in the immediately preceding sentence that it disagrees with such determination, then the Parties will promptly meet and discuss in good faith each Party’s basis for making their determination.
(e)Indirect Tax. All transfer, documentary, sales, use, stamp, registration, value-added, consumption, goods and services tax or other similar tax in respect of any License Payments (each an “Indirect Tax”) shall be borne by Licensee to the extent that a valid Indirect Tax invoice or other applicable documentation has been provided by Everest to Licensee in accordance with Applicable Law; provided that, any Indirect Tax recoverable by Everest shall not be borne by Licensee. In the case and to the extent that any Indirect Tax in connection with the transactions contemplated hereby is recoverable by Everest, Everest shall reimburse such recovered Indirect Tax at the applicable Indirect Tax rate to Licensee within a reasonable time frame.
9.4Records; Audits. Each Party shall keep, and require its Affiliates and Sublicensees to keep, complete, fair and true books of accounts and records for the purpose of determining the amounts payable to the other Party pursuant to this Agreement. Such books and records shall be kept for at least [*] following the end of the Calendar Year to which they pertain. Each Party shall have the right to cause an independent, certified public accountant reasonably acceptable to the other Party to audit such records to confirm Net Sales, royalties and other payments for a period covering not more than the preceding [*]; provided that (a) such audit shall not be more frequent than once in any [*] period, and (b) once such accountant has conducted a review and audit of any records pursuant to this Section 9.4 (Records; Audits) in respect of any given period, it may not subsequently re-inspect such records with respect to such period, unless, in each case of (a) and (b), for cause. Such audits may be exercised during normal business hours upon reasonable prior written notice to the audited Party. Prompt adjustments shall be made by the Parties to reflect the results of such audit. The auditing Party shall bear the full cost of such audit unless such audit discloses an underpayment by or an overpayment to the other Party of more than [*] of the amount of royalties or other payments due under this Agreement for any applicable Calendar Quarter, in which case, the audited Party shall bear the cost of such audit and shall promptly remit to the auditing Party the amount of any underpayment or overpayment. At Licensee’s discretion, any overpayment by Licensee revealed by an audit shall be fully-creditable against future payment owed by Licensee to Everest (and if no further payments are due, shall be refunded by Everest at the request of Licensee).
9.5Late Payments. In the event that Licensee fails to make any payment due under this Agreement, simple interest shall thereafter accrue on the sum due from the due date until the date of payment at [*]; provided, however, that in no event shall such rate exceed the maximum legal annual interest rate. The payment of such interest shall not limit Everest from exercising any other rights it may have as a consequence of such late payment.
Article 10
CONFIDENTIALITY
CONFIDENTIALITY
10.1Confidential Information. Except to the extent expressly authorized by this Agreement or otherwise agreed in writing by the Parties, each Party (in such capacity, the “Receiving Party”) agrees that, during the Term and for [*] thereafter, it shall keep confidential
37
and shall not publish or otherwise disclose to any Third Party, and shall not use for any purpose other than as expressly provided for in this Agreement or any other written agreement between the Parties, any Confidential Information, including any Know-How, furnished or made available to it by or on behalf of the other Party (in such capacity, the “Disclosing Party”). The Receiving Party shall use at least the same standard of care as it uses to protect proprietary or confidential information of its own (but in no event less than reasonable care) to ensure that its, and its Affiliates’, employees, agents, contractors, consultants and other representatives do not disclose or make any unauthorized use of the Confidential Information. The Receiving Party shall promptly notify the Disclosing Party upon discovery of any unauthorized use or disclosure of the Disclosing Party’s Confidential Information. Everest Solely Owned Inventions and Everest Additional Active Inventions are the Confidential Information of Everest. Licensee Solely Owned Inventions and Licensee Additional Active Inventions are the Confidential Information of Licensee. The terms and conditions of this Agreement and Jointly Owned Inventions (including Combination Product Inventions) are the Confidential Information of both Parties.
10.2Exceptions. Confidential Information shall not include any information which the Receiving Party can prove by competent evidence: (a) is at the time of disclosure, or thereafter becomes, through no act or failure to act on the part of the Receiving Party, generally known or available to the public or part of the public domain; (b) is known by the Receiving Party or any of its Affiliates at the time of receiving such information, as evidenced by its written records; (c) is rightfully hereafter furnished to the Receiving Party or any of its Affiliates by a Third Party, as a matter of right and without restriction on disclosure; or (d) is independently discovered or developed by the Receiving Party or any of its Affiliates, without the use of or reference to Confidential Information of the Disclosing Party, as evidenced by its written records. Notwithstanding the foregoing, (i) no element within the Confidential Information shall be deemed to fall within the foregoing exceptions merely because it is embraced by more general information or data that is part of the public domain or in the possession of Receiving Party prior to disclosure thereof by Disclosing Party; (ii) no combination of elements within the Confidential Information shall be deemed to fall within the foregoing exceptions merely because the individual elements of such combination fall within an exception, unless the entire combination itself, or the entire principle of use or operation of such combination (if any), is part of the public domain or in the possession of Receiving Party prior to disclosure thereof by Disclosing Party; and neither the act of using Confidential Information in a Clinical Trial nor the filing of Confidential Information with a Governmental Authority shall, for purposes of this Agreement, be deemed to place such Confidential Information in the public domain.
10.3Authorized Disclosure. Notwithstanding the provisions of Section 10.1 (Confidential Information), the Receiving Party may disclose Confidential Information of the Disclosing Party as expressly permitted by this Agreement, or if and to the extent such disclosure is reasonably necessary in the following instances:
(a)filing or prosecuting or defending Patents as permitted by this Agreement;
(b)disclosure in connection with litigation directly related to the Licensed Compound or Licensed Products;
(c)disclosure required in connection with any judicial or administrative process relating to or arising from this Agreement (including any enforcement hereof) or to comply with applicable court orders or Applicable Law, including without limitation applicable securities laws (including disclosure requirements of the U.S. Securities and Exchange Commission (“SEC”) or a similar regulatory agency in a country other than the United States, or any applicable stock exchange (such as Nasdaq or The Stock Exchange of Hong Kong Limited) on which securities issued by a Party or its Affiliates are traded);
38
(d)disclosure to Affiliates, Sublicensees, employees, contractors, consultants or agents of the Receiving Party who have a need to know such information in order for the Receiving Party to exercise its rights or fulfill its obligations under this Agreement; provided, in each case, that any such Affiliate, Sublicensee, employee, contractor, consultant or agent is bound by terms of confidentiality and non-use as restrictive as those set forth in this Article 10 (Confidentiality);
(e)disclosure by either Party or their Affiliates or their Sublicensees (or licensees or sublicensees with respect to Everest) as reasonably necessary or useful to obtain or maintain approval to conduct Clinical Trials for Licensed Products in the Field in the Territory (with respect to Licensee) and in the Field outside the Territory (with respect to Everest), or to obtain and maintain Regulatory Approval; provided that, Everest and its Affiliates and its and their licensees or sublicensees shall not disclose any Data (other than safety data) generated from any Global Trial proposed by Licensee in which Everest declined to participate unless Everest makes a payment to Licensee as set forth in Section 5.7(a);
(f)subject to confidentiality obligations that are consistent with this Agreement, disclosure by either Party or its Affiliates or its or their licensees or sublicensees, as applicable, to exercise such Party’s rights under this Agreement;
(g)such disclosure is made to such Party’s attorneys, independent accountants or financial advisors for the sole purpose of enabling such attorneys, independent accountants or financial advisors to provide advice to the Receiving Party, on the condition that such attorneys, independent accountants and financial advisors are bound by confidentiality and non-use obligations consistent with the confidentiality and non-use provisions of this Agreement as they apply to the Receiving Party (provided, however, that in the case of financial advisers, including investment bankers, the term of confidentiality may be shortened to [*] from the date of disclosure and in the case of attorneys, no written agreement shall be required);
(h)disclosure to existing investors, acquirors or collaborators or potential bona fide investors, acquirors, licensees, Sublicensees or collaborators in connection with due diligence or similar investigations by such Third Parties; provided, in each case, that any such existing or potential investor, acquiror, licensee, Sublicensee, collaborator or financing partner (including any royalty financing partner) agrees to be bound by confidentiality and non-use obligations consistent with those contained in this Agreement as they apply to the Receiving Party (but of duration customary in confidentiality agreements entered into for similar purpose);
(i)in the case of Everest, disclosure to the Upstream Licensors solely to the extent necessary to perform its obligations or confirm its compliance with the Upstream License Agreement, subject to confidentiality and non-use obligations set forth in the Upstream License Agreement; and
(j)to the extent mutually agreed to by the Parties in writing.
Notwithstanding the foregoing, in the event a Party is required to make a disclosure of the other Party’s Confidential Information pursuant to Section 10.3(b) or Section 10.3(c), it shall, except where impracticable or prohibited by Applicable Laws, give reasonable advance notice to the other Party of such disclosure and cooperate with the other Party to seek and obtain confidential treatment of such information to the extent legally permissible and shall only disclose that portion of the other Party’s Confidential Information that is required to be disclosed.
39
10.4Public Announcements.
(a)Press Releases and Publicity.
(i)The Parties have agreed upon the content of one or more press releases that shall be issued substantially in the form attached hereto as Schedule 10.4 (Initial Press Release), the release of which the Parties shall coordinate in order to accomplish such release promptly after the Execution Date. Thereafter, except as required by applicable securities laws (including disclosure requirements of the SEC or any stock exchange on which securities issued by a Party or its Affiliates are traded), neither Party shall make any other public announcement or statement, whether oral or written, concerning this Agreement or the subject matter hereof without the prior written consent of the other Party, which shall not be unreasonably withheld or delayed; provided that each Party may make any public statement in response to questions by the press, analysts, investors or those attending industry conferences or financial analyst calls, or issue press releases, so long as any such public statement or press release is not inconsistent with prior public disclosures or public statements approved by the other Party pursuant to this Section 10.4 (Press Release and Publicity), and the relevant information remains accurate as of such time, and which do not reveal non-public information about the other Party. In the event of a required public announcement, to the extent practicable under the circumstances, the Party making such announcement shall provide the other Party with a copy of the proposed text of such announcement sufficiently in advance of, and in any event at least [*] prior to, the scheduled release to afford such other Party a reasonable opportunity to review and comment upon the proposed text. Neither Party shall be required to seek the permission of the other Party to disclose any information regarding the terms of this Agreement that has already been publicly disclosed by either Party in accordance with this Section 10.4(a)(i); provided that such information remains accurate as of the time of such disclosure. After a publication has been made available to the public, each Party may post such publication or a link to it on its corporate web site without the prior written consent of the other Party.
(ii)Notwithstanding anything to the contrary in this Section 10.4 (Press Release and Publicity), Everest has the right to publicly disclose the achievement of any milestones under this Agreement.
(b)Filing of this Agreement. The Parties shall coordinate in advance with each other in connection with the filing of this Agreement (including redaction of certain provisions of this Agreement) with the SEC or any stock exchange or governmental agency on which securities issued by a Party or its Affiliate are traded, and each Party shall use reasonable efforts to seek confidential treatment for the terms proposed to be redacted; provided that each Party shall ultimately retain control over what information to disclose to the SEC or any stock exchange or other governmental agency, as the case may be, and provided further that the Parties shall use their reasonable efforts to file redacted versions with any governing bodies which are consistent with redacted versions previously filed with any other governing bodies. Except as set forth in this Section 10.4(b) (Filing of this Agreement), neither Party (nor its Affiliates) shall be obligated to consult with or obtain approval from the other Party with respect to any filings to the SEC or any stock exchange or other governmental agency.
10.5Publication.
40
(a)The Parties shall collaborate in good faith to jointly publish or publicly disclose any Data, results of or other information regarding any global Development activities (including Global Trials) performed under this Agreement by or on behalf of either Party with respect to the Licensed Compound or Licensed Products; provided that (i) if the Parties cannot agree on a joint publication plan for such global Development activities, then Licensee shall have the first right to publish any Data, results of or other information regarding such global Development activities and (ii) if a Party declines to be a co-author of such publication, the other Party is permitted to publish or publicly disclose any data, results of or other information generated within such other Party’s territory, in each case, (i) and (ii), subject to the review by the non-publishing Party pursuant to Section 10.5(b). For clarity, as between the Parties, Licensee shall have the first right to publish or otherwise disclose the results of any Clinical Trial conducted under this Agreement that is not a Global Trial.
(b)If either Party shall independently publish or publicly present any Clinical Trial data, nonclinical data or any associated results or conclusions generated by such Party in connection with the Development of the Licensed Compound or Licensed Products in its territory under this Agreement pursuant to Section 10.5(a) (each such proposed presentation or publication, a “Proposed Publication”), it shall submit such Proposed Publication to the non-publishing Party for review and comment in accordance with this Section 10.5(b). The publishing Party shall provide the non-publishing Party with a copy of such Proposed Publication at least [*] prior to the earlier of its presentation or intended submission for publication (such applicable period, the “Review Period”). The publishing Party agrees that it shall not submit or present any Proposed Publication (i) until the non-publishing Party has provided written comments during such Review Period on the material in such Proposed Publication or (ii) until the applicable Review Period has elapsed without written comments from the non-publishing Party, in which case the publishing Party may proceed, and the Proposed Publication shall be considered approved in its entirety. If the publishing Party receives written comments from the non-publishing Party during the applicable Review Period, it shall consider the comments of the non-publishing Party in good faith, but shall retain the sole authority to submit the manuscript for such Proposed Publication; provided that the publishing Party shall (A) delete any Confidential Information of the non-publishing Party that the non-publishing Party identifies for deletion in the non-publishing Party’s written comments, and (B) delay such Proposed Publication for a period of up to an additional [*] after the end of the applicable Review Period to enable the non-publishing Party to draft and file Patent applications with respect to any subject matter to be made public in such Proposed Publication and to which the non-publishing Party has the applicable intellectual property rights to file such Patent application. The publishing Party shall provide the non-publishing Party a copy of the Proposed Publication at the time of the submission or presentation. The publishing Party shall require its Affiliates or Sublicensees to comply with the obligations of this Section 10.5(b) as if they were the publishing Party, and shall be liable for their non-compliance. Licensee agrees to acknowledge the contribution of Everest and Upstream Licensors, and its and their employees in all publications as scientifically appropriate, consistent with customary scientific publication standards and based on the actual contributions of Everest and Upstream Licensors, and its and their employees with respect to such publications. For clarity, such acknowledgement shall not provide Everest with any review or approval of such publications except as set forth in this Section 10.5(b) or provide Everest, the Upstream Licensors or its or their employees with any authorship rights in such publications.
10.6Publication and Listing of Clinical Trials. With respect to the listing of Clinical Trials or the publication of Clinical Trial results for the Licensed Products and to the extent applicable to a Party’s activities conducted under this Agreement, each Party will comply
41
with (a) the Pharmaceutical Research and Manufacturers of America (PhRMA) Guidelines on the listing of Clinical Trials and the publication of Clinical Trial results, and (b) any Applicable Laws or applicable court order, stipulations, consent agreements, and settlements entered into by such Party. The Parties agree that any such listings or publications made pursuant to this Section 10.6 (Publication and Listing of Clinical Trials) shall be considered a publication hereunder and shall be subject to Section 10.5 (Publication).
10.7Prior Non-Disclosure Agreement. As of the Execution Date, the terms of this Article 10 (Confidentiality) shall supersede the Confidentiality Agreement. Any information disclosed pursuant to the Confidentiality Agreement shall be deemed Confidential Information for purposes of this Agreement.
10.8Equitable Relief. Given the nature of the Confidential Information and the competitive damage that would result to a Party upon unauthorized disclosure, use or transfer of its Confidential Information to any Third Party, the Parties agree that monetary damages would not be a sufficient remedy for any breach of this Article 10 (Confidentiality). In addition to all other remedies, a Party shall be entitled to seek specific performance and injunctive and other equitable relief as a remedy for any breach or threatened breach of this Article 10 (Confidentiality).
Article 11
REPRESENTATIONS AND WARRANTIES; LIMITATION OF LIABILITY
REPRESENTATIONS AND WARRANTIES; LIMITATION OF LIABILITY
11.1Mutual Representations and Warranties. Each Party represents and warrants to the other that, as of the Execution Date and the Effective Date:
(a)it is duly organized and validly existing under the laws of its jurisdiction of incorporation or formation, and has full corporate or other power and authority to enter into this Agreement and to carry out the provisions hereof;
(b)it is duly authorized to execute and deliver this Agreement and to perform its obligations hereunder, and the person or persons executing this Agreement on its behalf has been duly authorized to do so by all requisite corporate or partnership action; and
(c)this Agreement is legally binding upon it, enforceable in accordance with its terms, and does not (i) conflict with any agreement, instrument or understanding, oral or written, to which it is a party or by which it may be bound, (ii) conflict with or result in a breach of any provision of its organizational documents, or (iii) violate any Applicable Laws.
11.2Additional Everest Representations and Warranties. Everest represents and warrants to Licensee, as of the Execution Date and, subject to any exceptions that Everest notifies Licensee of pursuant to Section 11.8 (Bring Down of Additional Everest Representations and Warranties), the Effective Date, as follows:
(a)Everest (i) has sufficient legal or beneficial title or ownership or license, free and clear from any mortgages, pledges, liens, security interests, encumbrances, charges or claim of any kind, of the Everest Licensed IP to grant the license that it purports to grant in Section 2.1 (License Grant); and (ii) has not granted any right to any Third Party with respect to the Everest Licensed IP that would conflict with the license or rights granted to Licensee hereunder;
42
(b)Schedule 1.37 (List of Everest Patents) sets forth a complete and accurate list of all Everest Patents as of the Execution Date or the Effective Date, as applicable;
(c)Everest and, to Everest’s Knowledge, the Upstream Licensors have not received any written notice that any Third Party has taken any action before any applicable patent office in the Territory, claiming ownership of any Everest Licensed IP;
(d)Everest and, to Everest’s Knowledge, the Upstream Licensors have not received any written notice from any Third Party asserting that the issued Patents within the Everest Patents are invalid or unenforceable;
(e)Everest has, under all Applicable Laws, provided all notices to and obtained all consents necessary to transfer, disclose or otherwise make available to Licensee all Everest Know-How (including Personal Data) Controlled by Everest or any of its Affiliates as of the Execution Date or Effective Date, as applicable, as required under this Agreement without Licensee’s further action;
(f)no litigation, discovery process, interference, reissue, reexamination, interference, invalidity, opposition, post-grant review, or inter partes review is pending or threatened in writing with respect to any Everest Patent; and
(g)to Everest’s Knowledge, the Everest Licensed IP includes all Know-How and Patents owned, in-licensed by, held for use or otherwise Controlled by Everest or its Affiliates that are necessary or reasonably useful to Develop, Manufacture or Commercialize the Licensed Compound or Licensed Products in the Field in the Territory as such Development, Manufacture or Commercialization is currently being conducted or contemplated to be conducted under this Agreement;
(h)each of the Everest Patents is subsisting and, to Everest’s Knowledge, is valid and enforceable;
(i)to Everest’s Knowledge, the conduct of any Development, Manufacture or Commercialization with respect to the Licensed Compound or Licensed Products by or on behalf of Everest or any of its Affiliates, as conducted on or prior to the Execution Date or the Effective Date, as applicable, or as contemplated (as of the Execution Date or the Effective Date, as applicable) to be conducted during the Term in the Territory, in each case, does not infringe any valid and enforceable claim of any issued Patent of any Third Party in the Territory or misappropriate any other intellectual property rights of any Third Party in the Territory;
(j)to Everest’s Knowledge, no Third Party has infringed, misappropriated or otherwise violated or is infringing, misappropriating or otherwise violating any Everest Licensed IP in the Field in the Territory;
(k)no Governmental Authority provided funding for the development of any of the Everest Licensed IP or conduct of any Clinical Trial gives such Governmental Authority (i) any ownership interest in or right to prosecute or enforce any Everest Licensed IP or (ii) any rights to exercise any Everest Licensed IP to Exploit the Licensed Compound or Licensed Products in a manner that conflicts with, or limits the scope of, the rights and licenses granted to Licensee herein;
(l)to Everest’s Knowledge (i) there have been no acts or omissions of Everest that would constitute inequitable conduct, fraud, or misrepresentation to the applicable patent office with respect to any Everest Patents and (ii) the inventorship of
43
the Everest Patents is properly identified on each patent or patent application within the Everest Patents;
(m)Everest and its Affiliates have obtained from their employees and individual consultants who are inventors of the Everest Licensed IP, assignments of all rights in and to the Everest Licensed IP;
(n)Everest and its Affiliates have complied in all material respects with all Applicable Laws in the conduct of its and their Development and Manufacture of the Licensed Compound and Licensed Products in the Field, including in the conduct of the Ongoing Clinical Trial;
(o)Everest has provided to Licensee all material documentation, Data, and information under its Control relating to the Licensed Compound or any Licensed Product and the use thereof in the Field;
(p)all Regulatory Materials submitted to any Regulatory Authority by Everest or its Affiliates with respect to the Licensed Compound or any Licensed Product were, at the time of filing, true, complete, and accurate in all material respects, and Everest has disclosed all material facts required to be disclosed with respect to any Licensed Product to each applicable Regulatory Authority;
(q)Everest has provided to Licensee all material adverse event information with respect to the Licensed Compound or any Licensed Product known to Everest or its Affiliates; and
(r)Everest and its Affiliates are not, and have not been, debarred or disqualified by any Regulatory Authority and, to Everest’s Knowledge, (A) none of Everest’s or its Affiliates’ employees or contractors who have been or will be involved in the Exploitation of the Licensed Compound or Licensed Products are, or have been, debarred or disqualified by any Regulatory Authority; and (B) Everest and its Affiliates have not engaged in any conduct or activity that would be likely to lead to any debarment action by the FDA or other Regulatory Authority;
11.3Additional Licensee Representations and Warranties. Licensee represents and warrants to Everest that, as of the Effective Date, Licensee has the right to grant the license that it purports to grant in Section 2.4 (License to Everest).
11.4Mutual Covenants. In addition to any covenants made by the Parties elsewhere in this Agreement, each Party hereby covenants to the other Party that:
(a)with respect to Data Processed in each Party’s respective Control as part of such Party’s performance of this Agreement, each Party shall:
(i)comply with its respective obligations under all Applicable Laws (including arising out of or relating to the transfer of Data in connection with the granting of access of Data that such Party is required to provide to the other Party pursuant to Section 5.7 (Data Access; Right of Reference; Access to Regulatory Materials));
(ii)at all times during the Term, provide all notices and obtain all authorizations and consents required under all Applicable Laws (in connection with any applicable Clinical Trial) designed to permit the granting of access of Data that such Party is required to provide to the other Party pursuant to Section
44
5.7 (Data Access; Right of Reference; Access to Regulatory Materials) and to enable the other Party to Process such Data as required in order to (A) comply with its obligations under this Agreement and Applicable Laws; and (B) obtain the benefit of its rights under this Agreement;
(iii)not act in a manner such that it is subject to any prohibition or restriction which shall (A) prevent or restrict it from disclosing, transferring or otherwise making available its Data to the other Party pursuant to Section 5.7 (Data Access; Right of Reference; Access to Regulatory Materials); or (B) prevent or restrict it from Processing such Data as envisaged under this Agreement;
(iv)each Party shall obtain all the necessary authorizations, consents and approvals in order for such Party to grant access to, transfer and otherwise make available its Data to the other Party in accordance with this Agreement, including obtaining any necessary Clinical Trial participant authorizations and consents, and obtaining the necessary approvals from and completing all necessary filing procedures with, as applicable, the Governmental Authorities in the Territory;
(v)implement and maintain commercially reasonable administrative, technical, and physical safeguards designed to (A) maintain the security, integrity, availability and privacy of the Data; (B) protect against Data Breaches;
(vi)reasonably cooperate with the other Party to enable the other Party to fulfill its obligations, as applicable, under Applicable Laws in relation to Personal Data (including, without limitation, negotiating in good faith to enter into such additional agreements as necessary to comply with such Applicable Laws [*] and obtaining approvals from relevant Governmental Authorities to transfer Personal Data across national borders); and
(vii)in the event of an actual accidental, unauthorized or unlawful destruction, loss, alteration, disclosure, acquisition of or access to Data (any, a “Data Breach”) in a Party’s Control, promptly notify the other Party and reasonably cooperate with the other Party in responding to such Data Breach.
(b)it shall not knowingly, during the Term, employ or use the services of any person who is debarred or disqualified in connection with activities relating to the Licensed Compound or Licensed Products; and in the event that it becomes aware of the debarment or disqualification or threatened debarment or disqualification of any person providing services to it with respect to any activities relating to the Licensed Compound or Licensed Products, it shall immediately notify the other Party in writing and it shall cease employing, contracting with, or retaining any such person to perform any services relating to the Licensed Compound or Licensed Products.
11.5Licensee Covenants. In addition to any covenants made by Licensee elsewhere in this Agreement, Licensee hereby covenants to Everest as follows:
(a)it shall conduct its obligations with respect to the Development Plan in accordance with the study design set forth therein, each as may be amended from time to time;
(b)it shall, and shall cause its Affiliates and require its Sublicensees and subcontractors to, comply with all applicable provisions of the Upstream License
45
Agreement, and it shall not materially breach (or cause a material breach by Everest) of the Upstream License Agreement or take any other action in a manner that would give rise to the right of any Upstream Licensor to terminate the Upstream License Agreement; and
(c)it shall conduct, and shall cause its Affiliates and Sublicensees to conduct all activities under this Agreement (including as set forth in the Development Plan and Commercialization Plan) in compliance with all Applicable Law and other good scientific, laboratory, manufacturing and clinical practices under the Applicable Law of the region in which such activities are conducted, and any requirements of any Regulatory Authority and Governmental Authority health care programs having jurisdiction, each as may be amended from time to time.
11.6Everest Covenants. In addition to any covenants made by Everest elsewhere in this Agreement, Everest hereby covenants to Licensee as follows:
(a)it shall conduct its obligations with respect to the Development Plan in accordance with the study design set forth therein, as may be amended from time to time;
(b)it shall, and shall cause its Affiliates to, (i) maintain the Upstream License Agreement in full force and effect, (ii) materially comply with all provisions of the Upstream License Agreement, (iii) not materially breach or take any other action in a manner that would give rise to the right of any Upstream Licensor to terminate the Upstream License Agreement, (iv) not amend, modify, terminate, assign or transfer the Upstream License Agreement in a manner that could materially limit Licensee’s rights under this Agreement or increase Licensee’s obligations under this Agreement and (v) provide Licensee with copies of all amendments and waivers to the Upstream License Agreement promptly after execution;
(c)it shall provide all notices and perform all actions necessary to obtain the consents required under all Applicable Laws to transfer, disclose or otherwise make available to Licensee all Everest Know-How, as required under this Agreement; and
(d)it shall conduct, and shall cause its Affiliates and Sublicensees to conduct all activities under this Agreement (including as set forth in the Development Plan) in compliance with all Applicable Law and other good scientific, laboratory, manufacturing and clinical practices under the Applicable Law of the region in which such activities are conducted, and any requirements of any Regulatory Authority and Governmental Authority health care programs having jurisdiction, each as may be amended from time to time.
11.7Interim Covenants. Without limitation to any other provision of this Agreement:
(a)from the Execution Date until the Effective Date, Everest shall, and shall cause each of its Affiliates to (i) conduct its business with respect to the Licensed Compounds, the Licensed Products and the intellectual property rights to be licensed hereunder in the ordinary course consistent with past practice and in accordance with all Applicable Laws, and (ii) refrain from taking action or omitting to take any action that is reasonably expected to materially restrict or impair the rights to be granted to Licensee hereunder or prevent either Party’s ability to perform its obligations under this Agreement, including (A) licensing, transferring or otherwise disposing of any Everest Licensed IP, (B) entering into, modifying or amending any material contract (including the Upstream License Agreement) related to the Licensed Compounds, Licensed Products or the intellectual property rights granted hereunder in a manner that would materially
46
limit or impair Licensee’s rights or increase Licensee’s obligations under this Agreement or (C) creating or assuming any encumbrance, lien or claim of ownership by any Third Party with respect to any Everest Licensed IP;
(b)without limiting the generality of the foregoing, from the Execution Date until the Effective Date, except (i) to the extent prohibited by Applicable Laws or (ii) with the prior written approval of Licensee, Everest shall, and shall cause its Affiliates to: (A) ensure that all Manufacturing and Development (including the Ongoing Clinical Trial) of the Licensed Compounds and Licensed Products is conducted in material compliance with all Applicable Laws and (B) not employ or otherwise use in any capacity in the Manufacturing or Development (including the Ongoing Clinical Trial) of the Licensed Compounds or Licensed Products in the Field in the Territory, any person that is debarred under Section 21 USC 335a or any foreign equivalent thereof;
(c)from the Execution Date until the Effective Date, Everest shall (i) maintain the Upstream License Agreement in full force and effect, (ii) materially comply with all provisions of the Upstream License Agreement and (iii) not materially breach or take any other action in a manner that would give rise to the right of any Upstream Licensor to terminate the Upstream License Agreement; and
(d)Everest shall notify Licensee promptly after Everest or any of its Affiliates (i) Initiates any Clinical Trials or conducts any material Development of Licensed Compounds or Licensed Products in the Field in the Territory, (ii) discontinues, terminates or suspends the Ongoing Clinical Trial, or otherwise makes any material change to the Ongoing Clinical Trial, or (iii) becomes aware of any act, omission or event that occurred on or after the Execution Date that would cause the material breach of any covenant set forth in this Section 11.7 (Interim Covenants).
11.8Bring Down of Additional Everest Representations and Warranties. Following Everest’s receipt of notice of HSR Clearance in accordance with Section 16.5(c) (HSR Filing) and not later than the Outside Disclosure Date, Everest shall provide Licensee with a certificate (a) bringing-down the representation and warranties in Section 11.2 (Additional Everest Representations and Warranties) as of the Effective Date, and (b) certifying that none of the covenants set forth in Section 11.7 (Interim Covenants) have been breached, which certificate shall, if applicable, include any updates to the Schedules and identify any exceptions to the representations and warranties as of the Effective Date. Everest acknowledges and agrees that any disclosure made by Everest pursuant to Section 11.7 (Interim Covenants) and/or this Section 11.8 (Bring Down of Additional Everest Representations and Warranties) shall not cure (x) any breach of the representations and warranties in Section 11.2 (Additional Everest Representations and Warranties) as of the Execution Date or the Effective Date or (y) a breach of any covenant of Everest set forth in Section 11.7 (Interim Covenants).
11.9Performance by Affiliates, Sublicensees and Subcontractors. Licensee, may perform some or all of its obligations under this Agreement through one or more Affiliates, subcontractors or Sublicensees; provided, in each case, that (a) none of Everest’s rights hereunder are diminished or otherwise adversely affected as a result of such delegation or subcontracting, and (b) each such Affiliate, subcontractor or Sublicensee undertakes in writing to comply with the obligations of confidentiality and non-use regarding Confidential Information and obligations regarding ownership of Inventions which are consistent with those set forth in Article 10 (Confidentiality) and Section 12.1 (Ownership); and provided, further, that Licensee shall at all times be fully responsible and liable for the performance and payment of such Affiliate, subcontractor or Sublicensee.
47
11.10Disclaimer. EXCEPT AS EXPRESSLY SET FORTH IN THIS AGREEMENT, THE TECHNOLOGY AND INTELLECTUAL PROPERTY RIGHTS PROVIDED BY EACH PARTY TO THE OTHER PARTY HEREUNDER ARE PROVIDED “AS IS,” AND EACH PARTY EXPRESSLY DISCLAIMS ANY AND ALL WARRANTIES OF ANY KIND, EXPRESS OR IMPLIED, INCLUDING THE WARRANTIES OF DESIGN, MERCHANTABILITY, FITNESS FOR A PARTICULAR PURPOSE, WITH RESPECT TO OBTAINING SUCCESSFUL RESULTS, NON-INFRINGEMENT OF THE INTELLECTUAL PROPERTY RIGHTS OF THIRD PARTIES, OR WARRANTIES ARISING FROM A COURSE OF DEALING, USAGE OR TRADE PRACTICES.
Article 12
INTELLECTUAL PROPERTY
INTELLECTUAL PROPERTY
12.1Ownership.
(a)Background IP. As between the Parties, (i) Everest shall retain all right, title and interest in and to any Patents, Know-How, and other intellectual property rights Controlled by Everest or any of its Affiliates on or prior to the Effective Date or during the Term independent of the activities hereunder, and (ii) Licensee shall retain all right, title and interest in and to any Patents, Know-How, and other intellectual property rights Controlled (other than pursuant to this Agreement) by Licensee or any of its Affiliates on or prior to the Effective Date or during the Term independent of the activities hereunder.
(b)Solely Owned Inventions and Patents. Except as set forth in Section 12.1(d) (Combination Products), as between the Parties, any and all Inventions, Data, and other Know-How generated, developed, conceived or first reduced to practice (constructively or actually) solely by Everest and its Affiliates during the course of the performance of activities under this Agreement (“Everest Solely Owned Inventions”) and any Patents filed claiming or disclosing such Everest Solely Owned Inventions (“Everest Solely Owned Invention Patents”) shall be solely owned by Everest. Any and all Inventions, Data, and other Know-How generated, developed, conceived or first reduced to practice (constructively or actually) solely by Licensee and its Affiliates during the course of the performance of activities under this Agreement (“Licensee Solely Owned Inventions”) and any Patents filed claiming or disclosing such Licensee Solely Owned Inventions (“Licensee Solely Owned Invention Patents”) shall be solely owned by Licensee. Everest’s Solely Owned Inventions shall be included in the Everest Licensed IP (if within the scope of such definition) and included in the licenses and rights granted to Licensee by Everest hereunder; Licensee’s Solely Owned Inventions shall be included in the Licensee Technology (if within the scope of such definition) and included in the licenses and rights granted to Everest by Licensee hereunder.
(c)Joint Inventions and Patents. Except as set forth in Section 12.1(d) (Combination Products), as between the Parties, any and all Inventions, Data and other Know-How generated, developed, conceived or first reduced to practice by Everest and its Affiliates on the one hand, and Licensee and its Affiliates on the other hand, during the course of the performance of activities under this Agreement (“Jointly Owned Invention”), and any Patents filed claiming or disclosing such Jointly Owned Inventions (“Jointly Owned Invention Patents”) shall be jointly owned by the Parties. Each Party shall have the right to practice or transfer the Jointly Owned Invention (including licensing or sublicensing thereunder) without any duty of seeking or need to seek consent by or accounting to the other Party, subject to the terms of this Agreement.
(d)Combination Products. Notwithstanding Sections 12.1(b) (Solely Owned Inventions and Patents) and 12.1(c) (Joint Inventions and Patents), if the Parties
48
collaborate under this Agreement on Development activities or a Global Trial for a Combination Product comprising an Additional Active that is owned or in-licensed by Licensee or its Affiliates (“Licensee Additional Active”) or Combination Product comprising an Additional Active that is owned or in-licensed by Everest or its Affiliates (“Everest Additional Active”), then any and all Inventions, Data and other Know-How generated, developed, conceived or first reduced to practice under this Agreement that (i) specifically relate to such Licensee Additional Active shall be solely owned by Licensee (“Licensee Additional Active Inventions”), (ii) specifically relate to such Everest Additional Active shall be solely owned by Everest (“Everest Additional Active Inventions”), or (iii) specifically relate to such Combination Product comprising the Licensee Additional Active or the Everest Additional Active, as applicable, shall be jointly owned by the Parties (“Combination Product Inventions”). Everest hereby assigns, and agrees to assign, to Licensee all of its right, title and interest in and to Licensee Additional Active Inventions. Licensee hereby assigns, and agrees to assign, to Everest all of its right, title and interest in and to Everest Additional Active Inventions. Each Party hereby assigns, and agrees to assign, to the other Party an undivided one-half interest in and to Combination Product Inventions. Combination Product Inventions shall be Jointly Owned Inventions, and any Patents filed claiming or disclosing such Combination Product Inventions shall be Jointly Owned Invention Patents.
(e)Disclosures; Cooperation. Each Party shall ensure that each of its Affiliates, Sublicensees and subcontractors under this Agreement has a contractual obligation to disclose to such Party all Inventions, Data and other Know-How generated, invented, discovered, developed, made or otherwise created by them or their employees, agents or independent contractors, and to grant sufficient rights with respect thereto, so that such Party can comply with its obligations under this Section 12.1 (Ownership).
(f)Inventorship Determination. The Parties agree that determination of whether any Inventions, Data, and other Know-How are conceived, discovered, developed or otherwise made by a Party or its Affiliates for the purpose of allocating proprietary rights (including Patent, copyright, or other intellectual property rights) therein, shall, for purposes of this Agreement, be made in accordance with Patent law of the United States (without regard to conflict of laws) as such law exists as of the date of such conception, discovery, development, or making irrespective of where such conception, discovery, development, or making occurs. If United States Patent law otherwise would not apply to the discovery, generation, creation or conception of any Inventions, Data, and other Know-How hereunder, each Party shall, and does hereby, assign, and shall cause its Affiliates, and Sublicensees and its and their employees and contractors to so assign, to the other Party, without additional compensation from the other Party, such right, title and interest in and to any Inventions, Data, and other Know-How as well as any intellectual property rights with respect thereto, as is necessary to fully effect, as applicable, (i) the sole ownership provided for in Section 12.1(b) (Solely Owned Inventions and Patents) and (ii) the joint ownership provided for in Section 12.1(c) (Joint Inventions and Patents).
12.2Patent Prosecution and Maintenance.
(a)Definition. For purposes of this Section 12.2 (Patent Prosecution and Maintenance), the terms “prosecute,” “prosecuting” and “prosecution,” when used in reference to any Patent, shall be deemed to include, without limitation, control of any interferences, reissue proceedings, post-grant proceedings, oppositions and reexaminations with respect to such Patent and the right to seek patent term extensions and supplementary protection certificates with respect to such Patent.
49
(b)Licensee’s First Right. Licensee shall have the first right (which right, for clarity, may be allocated to and exercised through its Affiliates and Sublicensees), but not the obligation, to control the prosecution and maintenance of the Everest Patents and Jointly Owned Invention Patents, in each case, in the Territory (together, the “Territory Patents”), at [*] cost. Licensee shall keep Everest reasonably informed of the status of the prosecution and maintenance of any such Territory Patents, including that Licensee shall provide Everest with (a) copies of all correspondence and material communication it sends to or receives from any patent authority in connection with the prosecution and maintenance of such Territory Patents, (b) drafts of any material applications, filings or responses to be made or provided to any patent authorities in connection with such Territory Patents sufficiently in advance of submitting such filings or responses so as to allow for a reasonable opportunity for Everest to review and comment thereon, and Licensee shall consider in good faith all such reasonable comments of Everest and (c) a copy of applications as filed, together with notice of the filing date and serial number of each application. If Licensee determines to abandon or otherwise not prosecute and maintain any such pending or issued Territory Patents, then Licensee shall inform Everest of such decision promptly (but in no event later than [*] prior to allowing the applicable subject matter or such Territory Patents to lapse or otherwise become unpatentable, abandoned, unenforceable or dedicated to the public), and Everest shall thereafter have the right (which right, for clarity, may be allocated to and exercised through its Affiliates or Upstream Licensor), but not the obligation, to continue the prosecution and maintenance of such Territory Patents, at [*] cost and expense. Upon transfer of Licensee’s responsibility for prosecuting and maintaining any of the Territory Patents to Everest under this Section 12.2(b) (Licensee’s First Right), Licensee will promptly deliver to Everest copies of all necessary files related to such Patents with respect to which responsibility has been transferred and, at Everest’s written request, will take all actions and execute all documents reasonably necessary for Everest (or Upstream Licensors as applicable) to assume such prosecution, maintenance and defense.
(c)Everest’s Right. Everest shall have (i) the sole right, but not the obligation, to control the prosecution and maintenance of (A) the Everest Patents outside of the Territory, (B) any Everest Solely Owned Invention Patents that are not Everest Patents, and (C) any Patents filed claiming or disclosing Everest Additional Active Inventions and (ii) the first right, but not the obligation, to control the prosecution and maintenance of the Jointly Owned Invention Patents outside of the Territory, in each case (i) and (ii), at [*] cost. Everest shall keep Licensee reasonably informed of the status of the prosecution and maintenance of any such Jointly Owned Invention Patents outside of the Territory, including that Everest shall provide Licensee with (a) copies of all material correspondence received from any patent authority in connection with the prosecution and maintenance of such Jointly Owned Invention Patents and (b) drafts of any material filings or responses to be made to any patent authorities in connection with such Jointly Owned Invention Patents in advance of submitting such filings or responses so as to allow for a reasonable opportunity for Licensee to review and comment thereon, and Everest shall consider in good faith all such reasonable comments of Licensee. If Everest determines to abandon or otherwise not prosecute and maintain any such pending or issued Jointly Owned Invention Patents outside of the Territory, then Everest shall inform Licensee of such decision promptly (but in no event later than [*] prior to allowing the applicable subject matter or such Jointly Owned Invention Patents to lapse or otherwise become unpatentable, abandoned, unenforceable or dedicated to the public), and Licensee shall thereafter have the right, but not the obligation, to continue the prosecution and maintenance of such Jointly Owned Invention Patents outside of the Territory, at [*] cost and expense.
50
(d)Licensee’s Sole Right. Licensee shall have the sole right, at [*] costs, to control the prosecution and maintenance of any Licensee Solely Owned Invention Patents and any Patents filed claiming or disclosing Licensee Additional Active Inventions.
(e)Cooperation of the Parties. Prior to the filing of any Jointly Owned Invention Patent, the prosecuting Party shall consult with the other Party and give the other Party reasonable opportunity to review and comment on any draft Patent application in advance of its filing or submission and consider the other Party’s comments and suggestions in good faith.
(f)Patent Term Extension. Licensee shall have the sole right (which right, for clarity, may be allocated to and exercised through its Affiliates and Sublicensees) to obtain, and to determine whether to obtain, patent term extensions, supplementary protection certificates, and equivalents thereof with respect to the Territory Patents to the extent applicable to any Licensed Product in the Territory, and Everest shall reasonably cooperate with Licensee in connection therewith, including providing reasonable assistance to Licensee (including executing any documents as may reasonably be required). If elections with respect to obtaining such patent term extensions, supplementary protection certificates, and equivalents thereof are to be made in connection therewith, Licensee shall have the sole right to make the election, and Everest agrees to abide by such election.
12.3Infringement by Third Parties.
(a)Notice. In the event that either Everest or Licensee becomes aware of (i) any infringement or threatened infringement by a Third Party of any Territory Patent, or any related declaratory judgment or equivalent action alleging the invalidity, unenforceability or non-infringement of any Territory Patent (the “Infringement”) or (ii) any infringement or threatened infringement by a Third Party of any Everest Patent or Jointly Owned Invention Patent that Covers the Licensed Compound or Licensed Products in the Retained Territory, or any related declaratory judgment or equivalent action alleging the invalidity, unenforceability or non-infringement of any such Patent, it shall notify the other Party in writing to that effect.
(b)Territory Patents. With respect to any Territory Patent, as between the Parties, Licensee shall have the first right, but not the obligation, to bring and control any action or proceeding with respect to the Infringement of any such Patent at [*] expense and by counsel of its own choice, and Everest shall have the right to join or to be represented in any such action by counsel of its own choice. Licensee will notify and keep Everest apprised in writing of any such action or proceeding and will consider Everest’s reasonable interests and requests regarding such action or proceeding. Licensee shall not enter into any settlement that would reasonably be expected to have a material and adverse effect on the scope, validity or enforceability of any Territory Patent in any enforcement action hereunder without the Everest’s prior written consent, such consent not to be unreasonably withheld, conditioned or delayed. If Licensee (i) does not intend to prosecute such Infringement, or ceases to diligently pursue an action or proceeding against such Infringement, or (ii) fails to bring any such action or proceeding with respect to such Infringement in the Territory within [*] upon becoming aware of the alleged Infringement, then Licensee will notify Everest and Everest shall have the right (which right, for clarity, may be allocated to and exercised through its Affiliates or Upstream Licensors), but not the obligation, to bring and control any such action in the Territory at [*] expense and by counsel of its own choice. Everest shall not enter into any settlement admitting the invalidity of, or otherwise having a material and adverse effect on the scope, validity or enforceability of any Territory Patent in any enforcement action
51
hereunder without Licensee’s prior written consent, such consent not to be unreasonably withheld, conditioned or delayed.
(c)Everest Patents, Solely Owned and Additional Active Inventions Patents. Everest shall have the sole right, at [*] costs, to enforce (i) Everest Patents and Jointly Owned Invention Patents outside of the Territory, (ii) Everest Solely Owned Invention Patents that are not Everest Patents, and (iii) any Patents filed claiming or disclosing Everest Additional Active Inventions, in each case ((i)-(iii)), against any infringement.
(d)Licensee Solely Owned and Additional Active Invention Patents. Licensee shall have the sole right, at [*] costs, to enforce any Licensee Solely Owned Invention Patents and any Patents filed claiming or disclosing Licensee Additional Active Inventions against any infringement.
(e)Cooperation; Award. In the event a Party brings an infringement action in accordance with this Section 12.3 (Infringement by Third Parties), the other Party shall cooperate fully, including, if required to bring such action, the furnishing of a power of attorney or being named as a party. Except as provided for explicitly elsewhere in this Section 12.3 (Infringement by Third Parties), neither Party shall enter into any settlement or compromise of any action under this Section 12.3 (Infringement by Third Parties) which would in any manner alter, diminish, or be in derogation of the other Party’s rights under this Agreement without the prior written consent of such other Party, which shall not be unreasonably withheld. Except as otherwise agreed by the Parties in connection with a cost-sharing arrangement, any recovery realized by a Party as a result of any action or proceeding pursuant to this Section 12.3 (Infringement by Third Parties), whether by way of settlement or otherwise, shall be applied [*].
12.4Infringement of Third Party Rights. In the event that a claim is brought against either Party alleging the infringement, violation or misappropriation of any Third Party intellectual property right based on the Exploitation of the Licensed Products in the Territory, each Party shall promptly notify the other Party in writing of such claim, and the Parties shall promptly meet to discuss the defense of such claim, and the Parties shall, as appropriate, enter into a joint defense agreement with respect to the common interest privilege protecting communications regarding such claim in a form reasonably acceptable to the Parties. Unless otherwise agreed in the joint defense agreement, each Party shall have the right to defend any such claims against it in the Territory at [*] expense as it reasonably determines appropriate. Upon the request of the Party controlling the response to the infringement claim under this Section 12.4 (Infringement of Third Party Rights), the other Party will reasonably cooperate with the controlling Party in the reasonable defense of such infringement claim. The other Party will have the right to consult with the controlling Party concerning such infringement claim and to participate in and be represented by independent counsel in any associated litigation. The Party controlling the defense against an infringement claim under this Section 12.4 (Infringement of Third Party Rights) agrees not to settle such infringement action without the prior written consent of the other Party, which will not be unreasonably withheld or delayed; provided that, the Party controlling the defense may settle such infringement claim without the prior written consent of the other Party, if such settlement will only involve the payment of monetary damages by the other Party, in which case the Party controlling the defense will be liable for the payment of any monetary damage under such settlement. The terms in Section 12.3(e) (Cooperation; Award) shall apply, mutatis mutandis, with respect to the sharing and allocation of expenses and recoveries between the Parties for any action described in this Section 12.4 (Infringement of Third Party Rights).
52
12.5Marking. To the extent required by Applicable Law, Licensee shall, and shall cause its Affiliates and Sublicensees to, mark the Licensed Products sold under this Agreement with the number of each issued Everest Patent or Jointly Owned Invention Patent that applies to the Licensed Products.
12.6Joint Research Agreement. The Parties acknowledge and agree that this Agreement constitutes a “joint research agreement” within the meaning of 35 U.S.C. § 100(h). Either Party shall be entitled to invoke the prior art disqualification provisions of 35 U.S.C. § 102(b)(2)(C) by naming the Parties to this Agreement in any application for patent, either at the time of filing or by way of amendment, as contemplated by 35 U.S.C. § 102(c)(3).
Article 13
TERM; TERMINATION
TERM; TERMINATION
13.1Term. Notwithstanding anything to the contrary herein, this Agreement (other than Article 10 (Confidentiality), Section 11.1 (Mutual Representations and Warranties), Section 11.2 (Additional Everest Representations and Warranties), Section 11.7 (Interim Covenants), Section 11.8 (Bring Down of Additional Everest Representations and Warranties), this Section 13.1 (Term), Section 13.2(e) (Termination for Failure or Delay to Obtain HSR Clearance or Change in Applicable Laws), Section 13.2(f) (Termination for Material Change in Representations and Warranties or Breach of Covenant), Section 14.6 (Limitation of Liability), Article 15 (Dispute Resolution) and Article 16 (Miscellaneous), which are binding and effective as of the Execution Date) shall become effective subject to and conditional upon performance by the Parties of the following actions and occurrence of the following circumstances: (a) HSR Clearance; (b) Everest’s delivery of a certificate in accordance with Section 11.8 (Bring Down of Additional Everest Representations and Warranties), (c) no injunction (whether temporary, preliminary, or permanent) prohibiting consummation of this Agreement shall be in effect, (d) after the Execution Date and on or before the later of (x) the date of HSR Clearance and (y) the date of Everest’s delivery of the bring-down certificate pursuant to Section 11.8 (Bring Down of Additional Everest Representations and Warranties), or, if Everest made any additional disclosures in such certificate, the date of the expiration of the [*] period following Licensee’s receipt of such certificate as set forth in Section 13.2(f) (Termination for Material Change in Representations and Warranties or Breach of Covenant), there has not been a change in Applicable Laws that prohibits Licensee from making the Upfront Payment in accordance with Section 8.1 (Upfront Payment), prohibits the grant of any rights or licenses under Section 2.1 (License Grant) or Section 2.4 (License to Everest), prohibits granting of the rights of reference under Section 5.7(b) or prohibits the technology transfer under Section 4.4 (Technology Transfer) or Section 6.1 (Manufacturing Technology Transfer), and (e) either (i) if Everest made any additional disclosures to Licensee pursuant to Section 11.8 (Bring Down of Additional Everest Representations and Warranties), Licensee having not exercised its termination right under Section 13.2(f) (Termination for Material Change in Representations and Warranties or Breach of Covenant), upon the [*] after expiration of the period set forth in Section 13.2(f) (Termination for Material Change in Representations and Warranties or Breach of Covenant) or (ii) if Everest did not make any disclosures to Licensee in the certificate provided pursuant Section 11.8 (Bring Down of Additional Everest Representations and Warranties), upon the [*] after Everest providing such certificate (such date, in either (e)(i) or (e)(ii), the “Effective Date”), and, in each case (e)(i) and (e)(ii) whereupon the full Agreement and all its terms and provisions shall be automatically effective and binding on both Parties. The term of this Agreement (the “Term”) shall commence on the Effective Date, and unless terminated earlier as provided in this Article 13 (Term; Termination), shall expire, on a Licensed Product-by-Licensed Product and country-by-country basis, upon the expiration of the Royalty Term with respect to such Licensed Product in such country.
13.2Termination.
53
(a)Termination by Licensee for Convenience. Licensee shall have the right to terminate this Agreement in its entirety or on a Region-by-Region basis, at any time, for any or no reason, upon [*] written notice to Everest.
(b)Material Breach. A Party shall have the right to terminate this Agreement upon written notice to the other Party if such other Party is in material breach of this Agreement (including with respect to Licensee, Licensee’s failure to make the Upfront Payment pursuant to Section 8.1 (Upfront Payment)) and has not cured such breach within [*] (or [*] with respect to any breach of undisputed payment obligations hereunder) after written notice from the terminating Party requesting cure of such breach. Any such termination shall become effective at the end of the applicable curing period unless the breaching Party has cured such breach prior to the end of such curing period. If the alleged breaching Party disputes in good faith the existence or materiality of a breach specified in a notice provided by the other Party, and such alleged breaching Party provides the other Party notice of such dispute within [*], then the cure period set forth above shall be tolled, and the other Party shall not have the right to terminate this Agreement under this Section 13.2(b) (Material Breach) unless and until it is determined pursuant to Article 15 (Dispute Resolution) that the alleged breaching Party has materially breached this Agreement and fails to cure such breach within the remainder of the applicable cure period set forth above following such decision.
(c)Patent Challenge. Everest shall have the right to terminate this Agreement in its entirety with [*] upon written notice to Licensee if Licensee or any of its Affiliates or Sublicensees, directly or indirectly through any Third Party, commences, voluntarily participates in, or voluntarily provides support to, any action or proceeding in any forum of competent jurisdiction in the Territory (whether a court, a patent office, or an arbitral tribunal) to challenge the patentability, validity, enforceability, or scope of any Everest Patent, or Everest’s or any Upstream Licensor’s ownership of any Everest Licensed IP. Notwithstanding the foregoing, Everest shall not have the right to terminate the Agreement under this Section 13.2(c) (Patent Challenge) if (i) such action or proceeding (A) is initiated as a defense against a claim, action or proceeding asserted by or on behalf of Everest or of its Affiliates or its or their licensees or Sublicensees against Licensee or any of its Affiliates or its or their Sublicensees, or otherwise in connection with an assertion of a cross-claim or a counterclaim, provided such claim, action, proceeding or assertion does not arise from Licensee’s Exploitation of Everest Licensed IP, Licensed Compound or Licensed Product outside of the scope of license granted hereunder, (B) is brought by a Sublicensee and Licensee has terminated the relevant agreement with such Sublicensee within such [*] period, or (C) is dismissed within such [*] period and not thereafter continued. For clarity, Everest shall not have the right to terminate the Agreement under this Section 13.2(c) (Patent Challenge) if Licensee or its Affiliate or Sublicensee is merely making arguments that distinguish the inventions claimed in any Patents owned or controlled by Licensee or its applicable Affiliate or Sublicensee from those claimed in any Everest Patent in the ordinary course of ex parte prosecution of such Patent (or similar process or procedure).
(d)Cessation. If none of Licensee or its Affiliates or Sublicensees have conducted any material Development, Manufacture or Commercialization activities with respect to any Licensed Product in the Territory for a continuous period of twelve (12) months or longer (the “Cessation Period”), then Everest shall have the right to terminate this Agreement with [*] prior written notice to Licensee. Such Cessation Period shall be tolled to the extent that such cessation is caused by (i) a Force Majeure Event, (ii) [*], (iii) written guidance from, or action by, a Regulatory Authority, including with respect to any clinical hold or a recall or withdrawal, with respect to any Licensed Compound or Licensed Product, (iv) any injunction or other operation of Applicable Law, or (v) any
54
actual claim that the Development, Manufacture, Commercialization or other Exploitation of any Licensed Compound or Licensed Product under this Agreement infringes or misappropriates any Patent or Know-How of any Third Party.
(e)Termination for Failure or Delay to Obtain HSR Clearance or Change in Applicable Laws. Either Party may terminate this Agreement upon written notice to the other Party in the event that (i) either Party receives a request for additional information or documentary material, as described in 16 C.F.R. § 803.20, (ii) HSR Clearance has not occurred within [*] following the submission of all required HSR Filings in accordance with Section 16.5 (HSR Filing) or (iii) after the Execution Date and on or before the later of (x) the date of HSR Clearance and (y) the date of Everest’s delivery of the bring-down certificate pursuant to Section 11.8 (Bring Down of Additional Everest Representations and Warranties), or, if Everest made any additional disclosures in such certificate, the date of the expiration of the [*] period following Licensee’s receipt of such certificate as set forth in Section 13.2(f) (Termination for Material Change in Representations and Warranties or Breach of Covenant), there has been a change in Applicable Laws that prohibits Licensee from making the Upfront Payment in accordance with Section 8.1 (Upfront Payment), prohibits the grant of any rights or licenses under Section 2.1 (License Grant) or Section 2.4 (License to Everest), prohibits granting of the rights of reference under Section 5.7(b) or prohibits the technology transfer under Section 4.4 (Technology Transfer) or Section 6.1 (Manufacturing Technology Transfer), provided that in the event this Agreement is terminated in accordance with this Section 13.2(e) (Termination for Failure or Delay to Obtain HSR Clearance or Change in Applicable Laws), each Party shall be permitted to issue a press release or make other public announcement stating that this Agreement was terminated, as applicable, due to a delay in obtaining HSR Clearance or such change in Applicable Laws; provided that the Party making such announcement shall provide the other Party with a copy of the proposed text of such announcement sufficiently in advance of, and in any event at least [*] prior to, the scheduled release to afford such other Party a reasonable opportunity to review and comment upon the proposed text.
(f)Termination for Material Change in Representations and Warranties or Breach of Covenant. Without limiting Licensee’s rights under Section 13.2(b) (Material Breach), if (i) any actions or omissions by Everest or its Affiliates on or after the Execution Date cause any of the representations or warranties by Everest in Section 11.1 (Mutual Representations and Warranties) or Section 11.2 (Additional Everest Representations and Warranties) to become untrue or misleading as of the Effective Date (or as otherwise disclosed in the certificate provided pursuant to Section 11.8 (Bring Down of Additional Everest Representations and Warranties)), or (ii) Everest or any of its Affiliates materially breaches any covenant set forth in Section 11.7 (Interim Covenants) (other than Section 11.7(d)), and, in each case ((i) or (ii)), such occurrence [*], then Licensee may terminate this Agreement in its entirety upon written notice to Everest not later than [*] after its receipt of the bring-down certificate provided or to be provided pursuant to Section 11.8 (Bring Down of Additional Everest Representations and Warranties) [*]. Notwithstanding anything to the contrary herein, if Licensee terminates this Agreement in its entirety pursuant to this Section 13.2(f) (Termination for Material Change in Representations and Warranties or Breach of Covenant), then Licensee shall have no obligation to make the Upfront Payment or any other payment to Everest under this Agreement. [*]
(g)Bankruptcy. A Party shall have the right to terminate this Agreement upon written notice to the other Party upon the filing or institution of bankruptcy, reorganization for the benefit of creditors, dissolution, liquidation or winding up of such other Party, or the making or seeking to make or arrange an assignment of a substantial
55
portion of such other Party’s assets for the benefit of creditors of such other Party, or the initiation of proceedings in voluntary or involuntary bankruptcy against such other Party, or being adjudged bankrupt, or the appointment of a receiver or trustee of such other Party’s property, in each case that is not discharged within [*].
(h)Termination by Licensee for Failure to Complete Initial Technology Transfer. Licensee shall have the right to terminate this Agreement in its entirety upon [*] written notice to Everest if Everest fails to complete the Initial Technology Transfer within [*] after Everest’s receipt of the Upfront Payment, and Everest shall refund the Upfront Payment to Travere within [*] after the effective date of such termination.
13.3Effects of Expiration or Termination.
(a)Effect of Expiration. Upon expiration (but not earlier termination) of the Royalty Term for a given Licensed Product in a given country, and provided that Licensee has paid all payments payable with respect to such Licensed Product in such country, the license grant with respect to such Licensed Product in such country shall survive on an exclusive, fully paid, irrevocable, perpetual basis, and all other rights and obligations of the Parties with respect to such Licensed Product in such country under this Agreement shall terminate, except as provided otherwise in this Section 13.3 (Effects of Expiration or Termination) or in Section 13.4 (Accrued Obligations; Survival).
(b)Effect of Termination. Upon any termination of this Agreement, the following provisions shall apply:
(i)the license grant to Licensee under Section 2.1 (License Grant) shall automatically terminate and revert to Everest, and all other rights and obligations of the Parties under this Agreement shall terminate, except as provided otherwise in this Section 13.3 (Effects of Expiration or Termination) or in Section 13.4 (Accrued Obligations; Survival). For clarity, if any milestone event is achieved during the termination notice period, then the corresponding milestone payment is accrued, and Licensee shall remain responsible for the payment of such milestone payment even if the due date of such milestone payment may occur after the effective date of termination.
(ii)Any sublicense granted by Licensee or its Affiliates shall automatically survive as a direct license from Everest to the applicable Sublicensee, to the extent applicable to the scope and territory of such sublicense; provided that (A) such Sublicensee is not then in breach of its sublicense agreement (and, in the case of termination by Everest for breach by Licensee, that such Sublicensee and any further sublicensees did not cause the breach that gave rise to the termination by Everest), (B) such Sublicensee will be required to perform the same obligations contained in this Agreement as applicable to such Sublicensee, including to pay to Everest the same amounts in consideration for such direct grant as Everest would have otherwise received from Licensee pursuant to this Agreement on account of such Sublicensee’s Exploitation of the relevant Licensed Products had this Agreement not been terminated, and (C) Everest will not be bound by any grant of rights broader than, and will not be required to perform any obligation other than those obligations contained in, this Agreement as applicable to such Sublicensee.
(iii)Each Party shall provide information and execute documents as may be reasonably requested by the other Party to the extent reasonably necessary in order to give effect to this Section 13.3 (Effects of Expiration or Termination).
56
(c)Additional Effects of Termination.
(i)Subject to Section 13.3(c)(ii), upon termination of this Agreement other than pursuant to Section 13.2(e) (Termination for Failure or Delay to Obtain HSR Clearance or Change in Applicable Laws) or Section 13.2(f) (Termination for Material Change in Representations and Warranties or Breach of Covenant), the following provisions shall apply:
(A)As promptly as practicable (and in any event within [*]) after such termination, Licensee shall: (I) to the extent not previously provided to Everest, deliver to Everest true, correct and complete copies of all regulatory filings and registrations (including Regulatory Approvals) for the Licensed Products in the Territory held in the name of Licensee or its Affiliate, and disclose to Everest any Licensee Solely Owned Invention and Jointly Owned Invention (including all preclinical, clinical data, efficacy data and safety data) not previously disclosed to Everest and (II) transfer or assign, or cause to be transferred or assigned, to Everest or its designee (or to the extent not so assignable, take all reasonable actions to make available to Everest or its designee the benefits of) all regulatory filings and registrations (including Regulatory Approvals) for the Licensed Products in the Territory, held in the name of Licensee or its Affiliate, or as directed by Everest in its sole discretion, provide Everest with a right of reference with respect to any such regulatory filings and registrations.
(B)Licensee shall wind-down any ongoing Clinical Trials of any Licensed Product in the Field in the Territory in compliance with all Applicable Law and ethical standards, or, at Everest’s election and request, promptly transfer the conduct of such Clinical Trials to Everest or its designees. For any Clinical Trials to be transferred to Everest or its designees, Licensee shall continue to conduct such Clinical Trials after the effective date of termination until such transfer is completed, so that the termination of this Agreement will not cause any unreasonable interruption by Licensee of any such Clinical Trials; provided that (A) the Parties’ rights and obligations under Article 14 (Indemnification) will apply to such transferred Clinical Trials (which from the effective date of termination will be deemed to be conducted on behalf of Everest and not by Licensee, its Affiliates or Sublicensees).
(C)At Everest’s option and request, for a period of [*] after the effective date of termination, Licensee shall reasonably cooperate to introduce Everest to any Third Party contractor that provides services to Licensee or its Affiliate related to the Development, Manufacture, or Commercialization of the Licensed Compound or Licensed Products.
(D)At Everest’s election and request to be provided within [*] after the effective date of termination, Licensee shall transfer to Everest or its designee some or all inventory of the Licensed Products (including all final product, bulk drug substance, intermediates, works-in-process, formulation materials, reference standards, drug product clinical reserve samples, packaged retention samples, and the like) then in the possession or control of Licensee, its Affiliates or Sublicensees; provided that, Everest shall pay Licensee a price equal to Licensee’s purchase price from Everest or Licensee’s COGS of such transferred Licensed Products, as
57
applicable. The definition of COGS shall mutatis mutandis apply to the calculation and determination of such Licensee’s COGS.
(E)Licensee shall, and it hereby does, effective as of the date set forth in Section 13.3(c)(ii), grant to Everest an exclusive license, with the right to sublicense through multiple tiers of sublicense, under the Licensee Solely Owned Inventions, Licensee Solely Owned Invention Patents, and Licensee’s interest in any Jointly Owned Inventions and Jointly Owned Invention Patents, in each case, to Develop, Manufacture and Commercialize the Licensed Compounds and Licensed Products (excluding any Additional Active or other proprietary compound or product Controlled by Licensee or any of its Affiliates) in the Field worldwide.
(F)Licensee shall transfer and assign to Everest all Global Trademarks owned by Licensee or its Affiliates that solely relate to any Licensed Product and any applications therefor, and in-process applications for generic names for any Licensed Product owned by Licensee or its Affiliates.
(ii)Upon termination of this Agreement by Licensee pursuant to Section 13.2(b) (Material Breach) or Section 13.2(g) (Bankruptcy), the Parties will engage in good faith negotiations for a period of [*] and agree upon commercially reasonable financial terms to compensate Licensee for the grant of the license set forth in Section 13.3(c)(i)(E) and its conduct of the actions set forth in Section 13.3(c)(i) above. Upon termination of this Agreement by Everest pursuant to Section 13.2(b) (Material Breach), Section 13.2(c) (Patent Challenge), Section 13.2(d) (Cessation) or Section 13.2(g) (Bankruptcy) or by Licensee pursuant to Section 13.2(a) (Termination for Convenience), Licensee shall carry out the activities set forth in Section 13.3(c)(i) at [*] to Everest, except as set forth in Section 13.3(c)(i)(D), and the license set forth in Section 13.3(c)(i)(E) shall be effective as of the date of termination and [*] irrevocable and perpetual.
(d)Confidential Information. Upon expiration or termination of this Agreement in its entirety, except to the extent that a Party retains a license from the other Party as provided in this Article 13 (Term; Termination), each Party shall promptly return to the other Party, or delete or destroy, all relevant records and materials in such Party’s possession or control containing Confidential Information of the other Party; provided that such Party may keep one copy of such materials for archival purposes only subject to continuing confidentiality obligations under Article 10 (Confidentiality).
(e)Partial Termination. For the avoidance of doubt, if this Agreement is terminated with respect to only one or more countries, then this Section 13.3 (Effects of Expiration or Termination) shall apply only to such terminated Region(s), and such Region(s) shall be removed from the Territory for all purposes of this Agreement.
13.4Accrued Obligations; Survival. Neither expiration nor any termination of this Agreement shall relieve either Party of any obligation or liability accruing prior to such expiration or termination, nor shall expiration or any termination of this Agreement preclude either Party from pursuing all rights and remedies it may have under this Agreement, at law or in equity, with respect to breach of this Agreement. In addition, the Parties’ rights and obligations under Sections 2.3 (No Implied Licenses; Retained Rights), 9.3(c) (Tax Indemnification), 9.4 (Records; Audits), 11.10 (Disclaimer), 12.1 (Ownership), 13.2(h) (Termination by Licensee for Failure to Complete Initial Technology Transfer) (following termination by Licensee pursuant to
58
Section 13.2(h)), 13.3 (Effects of Expiration or Termination), 13.4 (Accrued Obligations; Survival) and 16.15 (Parent Guarantee) (solely with respect to (i) any of Everest’s obligations, representations, warranties and covenants accruing prior to such expiration or termination and unfulfilled as of such expiration or termination, and (ii) all other obligations of Everest under provisions of this Agreement that survive expiration or termination pursuant to this Section 13.4 (Accrued Obligations; Survival)); with respect to any Development Costs that have accrued prior to the effective date of termination, but not reimbursed or reconciled as of the effective date of termination, 4.1 (Overview; Diligence) and 4.3 (Development Costs); with respect to any amounts payable to Everest as of, but not paid by the effective date of expiration or termination, Sections 9.1 (Payment; Reports), 9.2 (Exchange Rate; Manner and Place of Payment), 9.3(a), (b), (d) and (e) (Taxes) and 9.5 (Late Payments); and Article 1 (Definitions), Article 8 (Financial Terms) (but solely with respect to any amounts payable to Everest as of, but not paid by the effective date of expiration or termination), 10 (Confidentiality), 14 (Indemnification) (excluding Section 14.5 (Insurance)), 15 (Dispute Resolution) (excluding Section 15.5 (Continued Performance)) and 16 (Miscellaneous) (excluding Section 16.15 (Parent Guarantee) except as set forth in this Section 13.4 (Accrued Obligations; Survival)) of this Agreement shall survive expiration or any termination of this Agreement.
13.5Rights Upon Bankruptcy. All rights and licenses granted under or pursuant to this Agreement by one Party to the other Party are, and otherwise shall be deemed to be, for purposes of Section 365(n) of Title 11 of the United States Code (“U.S. Bankruptcy Code”) or comparable provision of applicable bankruptcy or insolvency laws, licenses of right to “intellectual property” as defined under Section 101 of the U.S. Bankruptcy Code or comparable provision of applicable bankruptcy or insolvency laws. In the event that a case under the U.S. Bankruptcy Code or comparable provision of applicable bankruptcy or insolvency laws is commenced by or against a Party, the other Party shall have all of the rights and elections set forth in Section 365(n) or comparable provision of applicable bankruptcy or insolvency laws to the maximum extent permitted thereby. The Parties further agree that, in the event of the commencement of a bankruptcy proceeding by or against a Party under U.S. Bankruptcy Code or any comparable provision of applicable bankruptcy or insolvency laws, the other Party shall be entitled to a complete duplicate of (or complete access to, as appropriate) any such intellectual property and all embodiments of such intellectual property, which, if not already in such other Party’s possession, shall be promptly delivered to such other Party (i) upon any such commencement of a bankruptcy proceeding upon such other Party’s written request therefor, unless such Party elects to continue to perform all of its obligations under this Agreement, or (ii) if not delivered under clause (i), following the rejection of this Agreement by such Party upon written request therefor by such other Party.
Article 14
INDEMNIFICATION
INDEMNIFICATION
14.1Indemnification of Everest. Licensee shall indemnify and hold harmless each of Everest and its Affiliates and its and their respective directors, officers, employees, consultants, agents and successors and assigns of any of the foregoing (the “Everest Indemnitees”) from and against any and all losses, damages, liabilities, expenses and costs, including reasonable legal expense and attorneys’ fees (“Losses”), incurred by any Everest Indemnitee as a result of any claims, demands, actions, suits or proceedings brought by a Third Party (“Third Party Claims”) to the extent arising directly or indirectly out of: (a) the practice by Licensee or its Affiliates or Sublicensees of the license grant to Licensee under Section 2.1 (License Grant); (b) the research, Development, Manufacture, use, handling, storage, Commercialization or other Exploitation of the Licensed Compound or the Licensed Products by Licensee or its Affiliates or Sublicensees; (c) the performance by Licensee or its Affiliates or Sublicensees of the activities as the legal and beneficial owner of the Regulatory Approvals and Regulatory Materials for the Licensed Products in the Territory; (d) the negligence or willful misconduct of any Licensee Indemnitee;
59
or (e) any breach of any representations, warranties or covenants by Licensee under this Agreement; except, in each case, to the extent such Third Party Claims fall within the scope of the indemnification obligations of Everest set forth in Section 14.2 (Indemnification of Licensee).
14.2Indemnification of Licensee. Everest shall indemnify and hold harmless each of Licensee and its Affiliates and their respective directors, officers, employees, consultants, agents and successors and assigns of any of the foregoing (the “Licensee Indemnitees”), from and against any and all Losses incurred by any Licensee Indemnitee as a result of any Third Party Claims to the extent arising directly or indirectly out of: (a) the practice by Everest or its Affiliates of the license granted to Everest under Section 2.4 (License to Everest); (b) the research, Development, Manufacture, use, handling, storage, Commercialization or other Exploitation of the Licensed Compound or Licensed Products by Everest or its Affiliates or (sub)licensees (other than Licensee or its Affiliates or Sublicensees); (c) performance of the activities as the legal and beneficial owner of the Regulatory Approvals and Regulatory Materials for the Licensed Products (i) outside of the Territory or (ii) in the Territory prior to transfer to Licensee or its designee pursuant to Section 5.2 (Ownership of Regulatory Approvals and Regulatory Materials); (d) the negligence or willful misconduct of any Everest Indemnitee; or (e) any breach of any representations, warranties or covenants by Everest under this Agreement; except, in each case, to the extent such Third Party Claims fall within the scope of the indemnification obligations of Licensee set forth in Section 14.1 (Indemnification of Everest).
14.3Procedure. If any Everest Indemnitee or Licensee Indemnitee intends to claim indemnification under this Article 14 (Indemnification) (the “Indemnitee”), Everest or Licensee, as the case may be, shall promptly notify the indemnifying Party (the “Indemnitor”) in writing of any Third Party Claim, in respect of which the Indemnitee intends to claim such indemnification, and the Indemnitor shall have sole control of the defense or settlement thereof. The Indemnitee may participate at its expense in the Indemnitor’s defense of and settlement negotiations for any Third Party Claim with counsel of the Indemnitee’s own selection. The indemnity obligation as set forth in this Article 14 (Indemnification) shall not apply to amounts paid in settlement of any action with respect to a Third Party Claim, if such settlement is effected without the consent of the Indemnitor, which consent shall not be withheld or delayed unreasonably. The failure to deliver written notice to the Indemnitor within a reasonable time after the commencement of any action with respect to a Third Party Claim shall only relieve the Indemnitor of its indemnification obligations under this Article 14 (Indemnification) if and to the extent the Indemnitor is actually prejudiced thereby. Everest or Licensee, as the case may be, and other Indemnitees shall cooperate fully with the Indemnitor and its legal representatives in the investigation of any action with respect to a Third Party Claim covered by these indemnification provisions, at the Indemnitor’s expense. The Indemnitor shall not settle any Third Party Claim without the prior written consent of the Indemnitee if the settlement is reasonably expected to: (a) result in or impose any obligation (including any payment obligation) on the Indemnitee, or (b) result in any admission of wrong-doing or fault by the Indemnitee.
14.4Mitigation of Loss. Each of Everest or Licensee, as applicable, will take and will procure that its Affiliates and Sublicensees take all such reasonable steps and actions as are necessary or as the Indemnitor may reasonably require in order to mitigate any Third Party Claims (or potential losses or damages) under this Article 14 (Indemnification). Nothing in this Agreement will or will be deemed to relieve any Party of any common law or other duty to mitigate any losses incurred by such Party.
14.5Insurance. Each Party, at its own expense, shall maintain product liability and other appropriate insurance in an amount consistent with sound business practice and reasonable in light of its obligations under this Agreement during the Term. Each Party shall provide a
60
certificate of insurance evidencing such coverage to the other Party upon request. If a Party wishes to fulfill its obligations under this Section 14.5 (Insurance) through self-insurance, it shall obtain the prior written consent of the other Party, such consent not to be unreasonably withheld, conditioned or delayed.
14.6Limitation of Liability. NEITHER PARTY SHALL BE LIABLE TO THE OTHER FOR ANY SPECIAL, CONSEQUENTIAL, INCIDENTAL, PUNITIVE, OR INDIRECT DAMAGES ARISING FROM OR RELATING TO ANY BREACH OF THIS AGREEMENT, REGARDLESS OF ANY NOTICE OF THE POSSIBILITY OF SUCH DAMAGES. NOTWITHSTANDING THE FOREGOING, NOTHING IN THIS SECTION 14.6 (LIMITATION OF LIABILITY) IS INTENDED TO OR SHALL LIMIT OR RESTRICT THE INDEMNIFICATION RIGHTS OR OBLIGATIONS OF ANY PARTY UNDER SECTION 14.1 (INDEMNIFICATION OF EVEREST) OR SECTION 14.2 (INDEMNIFICATION OF LICENSEE), OR DAMAGES (A) AVAILABLE FOR (I) A PARTY’S BREACH OF ITS CONFIDENTIALITY OBLIGATIONS IN ARTICLE 10 (CONFIDENTIALITY) OR (II) A PARTY’S BREACH OF ITS OBLIGATIONS UNDER SECTION 2.6 (NON-COMPETE) OR (B) RESULTING FROM GROSS NEGLIGENCE, WILLFUL MISCONDUCT, FRAUD OR FRAUDULENT MISREPRESENTATION BY EITHER PARTY.
Article 15
DISPUTE RESOLUTION
DISPUTE RESOLUTION
15.1Disputes. Subject to Section 15.3 (Excluded Claims) and except as provided in Section 3.6 (Decision-Making), upon the written request of either Party to the other Party, any claim, dispute, or controversy arising out of, relating to or in connection with this Agreement as to the breach, enforcement, interpretation or validity thereof (a “Dispute”) shall be referred to the Executive Officers for resolution. In the event such executives are unable to resolve such Dispute within [*] after the initial written request, then, upon the written demand of either Party and subject to Section 15.2 (Arbitration) below, the Dispute shall be finally resolved by binding arbitration under the Rules of Arbitration of the International Chamber of Commerce (“ICC”) in effect at the time such Dispute is submitted to arbitration, except as may be modified herein, applying the substantive law specified in Section 16.1 (Governing Law).
15.2Arbitration.
(a)Procedure. The arbitration shall be conducted by a panel of [*] arbitrators experienced in the business of pharmaceuticals. If the issues in dispute involve scientific, technical or commercial matters, the arbitrators chosen hereunder may, at their option, engage experts having educational training or industry experience sufficient to demonstrate a reasonable level of relevant scientific, medical and industry knowledge, as necessary to advise them with respect to the Dispute. If any expert is appointed, the Parties shall have the right review such expert(s) reports to the arbitrators and to examine such expert at an oral hearing. Within [*] after initiation of arbitration, each of the Parties shall nominate [*] after the [*] arbitrator’s appointment. If a Party fails to select an arbitrator or if the [*] arbitrators nominated by the Parties are unable or fail to agree upon the [*] arbitrator within the time period specified above, any unselected arbitrator or [*] arbitrator, as the case may be, shall be appointed by the ICC. The seat of arbitration will be New York, New York. The language of the arbitration will be English. Except to the extent necessary to confirm or challenge an award or as may be required by law, or to protect or pursue a legal right, neither a Party nor the arbitrators may disclose the existence, content, or results of arbitration without the prior written consent of both Parties. Notwithstanding Section 16.1 (Governing Law) with respect to the applicable substantive law, any arbitration and this agreement to arbitrate shall be governed by the
61
Federal Arbitration Act, 9 U.S.C. § 1 et. seq. Any dispute concerning the propriety of commencing arbitration, or the scope or applicability of this agreement to arbitrate shall be determined by the arbitrators.
(b)Arbitrators’ Award. Prior to the arbitrators being appointed, either Party, without waiving any remedy under this Agreement, may seek from any court having jurisdiction any temporary injunctive or provisional relief necessary to protect the rights or property of such Party until final resolution of the issue by the arbitrators or other resolution of the controversy between the Parties. Once the arbitrators have been appointed, either Party may apply to the arbitrators for interim injunctive relief until the arbitration award is rendered or the controversy is otherwise resolved, and either Party may apply to a court of competent jurisdiction to enforce interim injunctive relief granted by the arbitrators. The arbitrators shall, within [*] after the later of the last arbitration hearing date or the final post-hearing submission, issue a written award, which shall include a statement of decision describing the essential findings and conclusions on which the award is based, including the calculation of any damages awarded. No award issued by the arbitrators shall be rendered invalid or improper by reasons of it being delivered after the expiry of this time limit. The award rendered by the arbitrators shall be final and non-appealable, and judgment may be entered upon it in any court of competent jurisdiction.
(c)Costs. Each Party shall bear its own attorney’s fees, costs, and disbursements arising out of the arbitration, and shall pay an equal share of the fees and costs of the arbitrators [*].
15.3Excluded Claims. Nothing contained in this Agreement shall deny either Party the right to bring an action in any court of competent jurisdiction to resolve any dispute, controversy or claim that concerns (a) the validity, enforceability or infringement of any Patents or other intellectual property rights, or (b) any antitrust, anti-monopoly or competition law or regulation, whether or not statutory, and no such claim shall be subject to arbitration as a Dispute pursuant to Section 15.2 (Arbitration).
15.4Interim Relief. Each Party shall be entitled to take action in any court with competent jurisdiction to apply for and be granted emergency or other interim relief and otherwise enforce by injunction or other provisional equitable relief, without prejudice to any other rights and remedies that it may have under this Agreement.
15.5Continued Performance. Provided that this Agreement has not been terminated in its entirety, the Parties agree to continue performing under this Agreement in accordance with its provisions, pending the final resolution of any Dispute.
Article 16
MISCELLANEOUS
MISCELLANEOUS
16.1Governing Law. This Agreement and any disputes, claims, or actions related thereto shall be governed by and construed in accordance with the laws of the State of New York, without regard to the conflicts of law provisions thereof. The application of the U.N. Convention on Contracts for the International Sale of Goods (1980) is excluded.
16.2Entire Agreement; Amendment. This Agreement, including the Schedules hereto, sets forth all of the agreements and understandings between the Parties with respect to the subject matter hereof and thereof, and supersedes and terminates all prior agreements and understandings between the Parties with respect to the subject matter hereof and thereof, including the Confidentiality Agreement. There are no other agreements or understandings with
62
respect to the subject matter hereof, either oral or written, between the Parties. Except as expressly set forth in this Agreement, no subsequent amendment, modification or addition to this Agreement shall be binding upon the Parties unless reduced to writing and signed by the respective authorized officers of the Parties.
16.3Further Assurances. Each Party shall duly execute and deliver, or cause to be duly executed and delivered, such further instruments and do and cause to be done such further acts and things, including the filing of such assignments, agreements, documents, and instruments, as may be necessary or as any other Party may reasonably request in connection with this Agreement or to carry out more effectively the provisions and purposes hereof, or to better assure and confirm unto such other Party its rights and remedies under this Agreement.
16.4Relationship Between the Parties. The Parties’ relationship, as established by this Agreement, is solely that of independent contractors. This Agreement does not create any partnership, joint venture or similar business relationship or legal entity of any type between the Parties. Neither Party is a legal representative of the other Party, and neither Party can assume or create any obligation, representation, warranty or guarantee, express or implied, on behalf of the other Party for any purpose whatsoever. Neither Party shall treat or report the relationship arising under this Agreement as a partnership for United States tax purposes unless otherwise required pursuant to a determination within the meaning of Section 1313 of the Internal Revenue Code of 1986, as amended.
16.5HSR Filing.
(a)Prior to consummation of this Agreement, each Party shall make any required HSR Filing in good faith and shall use commercially reasonable efforts to obtain the necessary approvals or clearances, or the expiration or termination of the applicable waiting period (including any extension thereof, including under any agreement between a Party and the DOJ, FTC, or other Governmental Authority agreeing not to consummate the Agreement prior to a certain date); provided, however, that each Party shall cooperate as may be reasonably requested to ensure any such required approval, clearance, or expiration or termination of applicable waiting periods are obtained promptly.
(b)Subject to this Section 16.5 (HSR Filing), each Party shall, as promptly as practicable (but no later than [*] after the Execution Date (unless a later date is mutually agreed between outside antitrust counsel on behalf of the Parties, which agreement shall not be unreasonably withheld)), prepare and file the HSR Filings. The Parties shall cooperate in the process to obtain HSR Clearance, including by furnishing to each other’s counsel such necessary information and reasonable assistance as the other may reasonably request in connection with its preparation of any filing or submission that is necessary under the HSR Act and to respond as promptly as reasonably practicable and advisable to requests from the United States Federal Trade Commission (“FTC”), the Antitrust Division of the United States Department of Justice (“DOJ”), or other Governmental Authority for information reasonably requested by them in connection with such filings. Each Party shall be responsible for its own fees, costs, and expenses associated with the HSR Filing, except that Licensee shall pay the filing fee required under the HSR Act. Notwithstanding anything to the contrary in this Agreement, nothing in this Section 16.5 (HSR Filing) will require either Party (or any of their respective Affiliates) to propose, negotiate, commit to or effect, by consent decree, hold separate order, or otherwise (i) the sale, divestiture, disposition, licensing or sublicensing of any of a Party’s or its Affiliates’ assets, properties, or business; or (ii) behavioral limitations, conduct restrictions, or commitments with respect to such assets, properties, or business, or of any of the rights or obligations of a Party under this Agreement; and each Party and its Affiliates shall have no obligation to contest, administratively or in court, any ruling,
63
order, or other action of the DOJ, FTC, any other Governmental Authority or any Third Party seeking to prohibit or otherwise challenging the transactions contemplated by this Agreement, or defend through litigation any claim asserted in court by any Third Party respecting the transactions contemplated by this Agreement.
(c)Subject to this Section 16.5 (HSR Filing), the Parties shall instruct their respective counsel to cooperate with each other and use commercially reasonable efforts to facilitate and expedite the identification and resolution of any issues arising under the HSR Act and to cause the Effective Date of this Agreement to occur as soon as practical. Such commercially reasonable efforts and cooperation include counsel’s undertaking to (i) keep each other informed of communications, inquiries, and requests from and to personnel of the reviewing Governmental Authority, including the DOJ or FTC, including by providing copies thereof to the other Party (subject to reasonable redactions for privilege or confidentiality concerns), and (ii) confer with each other regarding appropriate contacts with and response to personnel of the DOJ, FTC, or other Governmental Authority, and the content of any such contacts or presentations. Each Party shall consult with the other Party, to the extent practicable in advance of participating in any substantive meeting or discussion with the relevant Governmental Authority, including the DOJ or FTC, with respect to any such filings, applications, investigation, or other inquiry and, to the extent permitted by such Governmental Authority, give the other Party the opportunity to attend and participate in such meeting or discussion. Each Party shall give the other Party the opportunity to review in advance, and shall consider in good faith the other Party’s reasonable comments in connection with, the content of any presentations, white papers, or other written materials (excluding, for clarity, any HSR Filing) to be submitted to the DOJ, FTC, or other Governmental Authority with respect to this Agreement and the transactions contemplated thereby (subject to redaction of commercially sensitive information) and upon request shall provide the other Party a copy of any such materials (excluding, for clarity, any HSR Filing and documentary attachments thereto) as submitted to the DOJ, FTC, or other Governmental Authority. Any materials required to be provided pursuant to this Section 16.5(c) may be restricted to each Party’s outside antitrust counsel and redacted to preserve privilege. Within [*] of HSR Clearance, Licensee shall provide Everest with notice of the occurrence of HSR Clearance. Notwithstanding anything contained in this Section 16.5 (HSR Filing), Licensee shall have the final-decision making authority with respect to the strategy for obtaining HSR Clearance.
16.6Non-Waiver. The failure of a Party to insist upon strict performance of any provision of this Agreement or to exercise any right arising out of this Agreement shall neither impair that provision or right nor constitute a waiver of that provision or right, in whole or in part, in that instance or in any other instance. Any waiver by a Party of a particular provision or right shall be in writing, shall be as to a particular matter and, if applicable, for a particular period of time and of a particular scope, and shall be signed by such Party.
16.7Assignment. Except as expressly provided hereunder, neither this Agreement nor any rights or obligations hereunder may be assigned by a Party without the prior written consent of the other Party, except to (a) an Affiliate; provided that this Agreement shall be assigned in whole, and the assigning Party shall remain liable and responsible to the non-assigning Party hereto for the performance and observance of all such duties and obligations by such Affiliate; (b) a Third Party in connection with the sale of all or substantially all of its assets to which this Agreement relates, whether in a sale of stock, sale of assets, business combination, reorganization, or other transaction or series of related transactions. Everest may assign without the prior consent of Licensee its right to receive payments under this Agreement or grant any security interest in its right, title and interest in this Agreement, in whole or in part and in their entirety or in portions, to an investor in connection with a financing transaction, including a
64
royalty financing; provided that Everest has given Licensee [*] prior written notice before entering into any agreement providing for such assignment. The rights and obligations of the Parties under this Agreement shall be binding upon and inure to the benefit of the permitted assigns of the Parties. Any assignment not in accordance with this Agreement shall be void.
16.8Third Party Beneficiaries. This Agreement is not intended to and shall not be construed to give any Third Party any interest or rights (including any third party beneficiary rights) with respect to or in connection with any agreement or provision contained herein or contemplated hereby.
16.9Severability. If, for any reason, any part of this Agreement is adjudicated invalid, unenforceable or illegal by a court of competent jurisdiction, such adjudication shall not affect or impair, in whole or in part, the validity, enforceability or legality of any remaining portions of this Agreement. All remaining portions shall remain in full force and effect as if the original Agreement had been executed without the invalidated, unenforceable or illegal part. The Parties shall use their commercially reasonable efforts to replace the invalid, illegal or unenforceable provision(s) with valid, legal and enforceable provision(s) in a way that, to the extent practicable and legally permissible, implements the original intent of the Parties.
16.10Notices. Any notice to be given under this Agreement must be in writing and delivered either in person, by any method of mail (postage prepaid) requiring return receipt, or by overnight courier or facsimile confirmed thereafter by any of the foregoing, to the Party to be notified at its address(es) given below, or at any address such Party has previously designated by prior written notice to the other. Notice shall be deemed sufficiently given for all purposes upon the earliest of: (a) the date of actual receipt; (b) if delivered by overnight courier, [*] after delivery; or (c) if sent by email, upon electronic confirmation of receipt.
65
| if to Licensee: | Travere Therapeutics, Inc. | ||||
3611 Valley Centre Drive, Suite 300 San Diego, CA 92130 United States | |||||
| Attention: General Counsel | |||||
| Email: [*] | |||||
| with a copy to (which shall not constitute notice): | Cooley LLP | ||||
10265 Science Center Drive San Diego, CA 92121 United States | |||||
| Attention: Jason Kent; Charity R. Williams | |||||
| Email: [*] | |||||
| if to Everest: | Everest Medicines (Singapore) Pte. Ltd. 36 Robinson Road, #20-01 City House, Singapore 068877 Attention: Ian Woo and Jason Brown Email: [*] | ||||
| with a copy to (which shall not constitute notice): | Everest Medicines (Singapore) Pte. Ltd. 17F, AIA Financial Center, 866 Dongchangzhi Road, Hongkou District, Shanghai 20083, P.R. China Attention: Legal Department Email: [*] and Ropes & Gray LLP | ||||
One Maritime Plaza, Suite 1800 300 Clay Street | |||||
| San Francisco, CA 94111 USA | |||||
| Attention: Geoffrey Lin | |||||
Email:.[*] | |||||
16.11Force Majeure. Each Party shall be excused from liability for the failure or delay in performance of any obligation under this Agreement by reason of any event beyond
66
such Party’s reasonable control including but not limited to acts of God, fire, flood, explosion, earthquake, or other natural forces, regional or worldwide epidemic, pandemic, war, civil unrest, acts of terrorism, accident, destruction or other casualty (a “Force Majeure Event”). Such excuse from liability shall be effective only to the extent and duration of the event(s) causing the failure or delay in performance and provided that the Party has not caused such event(s) to occur. Notice of a Party’s failure or delay in performance due to a Force Majeure Event must be given to the other Party as soon as reasonably practicable after its occurrence. The Party affected by a Force Majeure Event shall use reasonable efforts to resume performance of its obligations and shall keep the other Party informed of actions related thereto.
16.12Cumulative Remedies. No remedy referred to in this Agreement is intended to be exclusive unless explicitly stated to be so, and each shall be cumulative and in addition to any other remedy referred to in this Agreement or otherwise available under law.
16.13Interpretation. The headings of clauses contained in this Agreement preceding the text of the sections, subsections and paragraphs hereof are inserted solely for convenience and ease of reference only and shall not constitute any part of this Agreement, or have any effect on its interpretation or construction. All references in this Agreement to the singular shall include the plural where applicable. Unless otherwise specified, references in this Agreement to any Article shall include all Sections, subsections and paragraphs in such Article, references to any Section shall include all subsections and paragraphs in such Section, and references in this Agreement to any subsection shall include all paragraphs in such subsection. The word “including” and similar words means “including without limitation,” whether or not specifically stated. The word “or” means “and/or” unless the context dictates otherwise because the subjects of the conjunction are mutually exclusive. The words “herein,” “hereof” and “hereunder” and other words of similar import refer to this Agreement as a whole and not to any particular Section or other subdivision. The words “pharmaceuticals” or “drugs” include biologics unless expressly indicated otherwise. All references to days in this Agreement means calendar days, unless otherwise specified. All references to any Applicable Law in this Agreement mean such Applicable Law as amended, restated, supplanted or otherwise modified from time to time. Ambiguities and uncertainties in this Agreement, if any, shall not be interpreted against either Party, irrespective of which Party may be deemed to have caused the ambiguity or uncertainty to exist. This Agreement has been prepared in the English language, and the English language shall control its interpretation. In addition, all notices required or permitted to be given hereunder and all written, electronic, oral or other communications between the Parties regarding this Agreement, shall be in the English language.
16.14Construction. The Parties hereto acknowledge and agree that: (a) each Party and its counsel reviewed and negotiated the terms and provisions of this Agreement and have contributed to its revision; (b) the rule of construction to the effect that any ambiguities are resolved against the drafting Party shall not be employed in the interpretation of this Agreement; and (c) the terms and provisions of this Agreement shall be construed fairly as to all Parties hereto and not in favor of or against any Party, regardless of which Party was generally responsible for the preparation of this Agreement.
16.15Parent Guarantee. Parent hereby unconditionally guarantees all of the obligations, representations, warranties and covenants of Everest under this Agreement. In the event of any breach of this Agreement by Everest, Parent hereby assumes full responsibility for the obligations, representations, warranties and covenants of Everest hereunder and will take all necessary actions to remedy any breach. Parent hereby agrees to pay and perform all obligations of Everest under this Agreement, including but not limited to financial obligations, indemnification and performance of any actions required under this Agreement, which responsibility shall be enforceable by Licensee against Parent without any requirement that Licensee first seek payment or performance from Everest. The Parent will promptly provide
67
written notice to Everest in the event that it ceases to control Everest. This clause shall be binding upon and inure to the benefit of the Parties and Parent and their respective successors and assigns.
16.16Counterparts; Electronic Signatures. This Agreement may be executed in two (2) or more counterparts, including by transmission of facsimile or PDF copies of signature pages to the Parties or their representative legal counsel, each of which shall be deemed an original document, and all of which, together with this writing, shall be deemed one and the same instrument. Electronic, facsimile or PDF image signatures shall be treated as original signatures, with the understanding that each Party expressly agrees that such Party shall be bound by its own electronically transmitted signature and shall accept the electronically transmitted signature of the other Party (including through the use of eSignature platforms such as DocuSign®). No Party shall raise the use of electronic delivery to transmit a signature or the fact that any signature or agreement or instrument was transmitted or communicated through the use of electronic delivery as a defense to the formation of a contract.
[REMAINDER OF THIS PAGE INTENTIONALLY LEFT BLANK]
68
In Witness Whereof, the Parties hereto have duly executed this Exclusive License and Collaboration Agreement as of the Execution Date.
| Travere Therapeutics, Inc. | Everest Medicines (Singapore) Pte. Ltd. | ||||
By: /s/ Eric Dube | By: /s/ Ian Woo | ||||
Name: Eric Dube | Name: Ian Woo | ||||
Title: CEO | Title: Director | ||||
In Witness Whereof, Parent, solely for purpose of Section 16.15 (Parent Guarantee), has duly executed this Exclusive License and Collaboration Agreement as of the Execution Date.
| Everest Medicines Limited | |||||
By: /s/ Ian Woo | |||||
Name: Ian Woo | |||||
Title: President and Chief Financial Officer | |||||
Schedule 1.34
EVER001
[*]
Schedule 1.37
List of Everest Patents
[*]
Schedule 1.60
Everest Knowledge Parties
[*]
Schedule 1.83
Planned Global Trials
[*]
Schedule 4.4
List of Initial Technology Transfer
[*]
Schedule 10.4
Initial Press Release
Travere Therapeutics Enters Into Exclusive Licensing Agreement with Everest Medicines for Civorebrutinib a Potential Best-in-Class BTK Inhibitor for Rare Kidney Diseases
Civorebrutinib is an investigational oral, covalent reversible BTK inhibitor designed to provide differentiated efficacy, safety and convenience in immune-mediated kidney diseases
Civorebrutinib adds pipeline-in-a-product potential across multiple rare kidney diseases, expanding and diversifying Travere’s pipeline
Company to host conference call June 2, 2026 at 8:30 a.m. ET
SAN DIEGO, June 2, 2026 – Travere Therapeutics, Inc., (Nasdaq: TVTX) today announced that it has entered into an exclusive licensing and collaboration agreement with Everest Medicines for the development and commercialization of civorebrutinib (also known as EVER001), a potential best-in-class oral, covalent reversible Bruton’s tyrosine kinase (BTK) inhibitor in all markets outside China and certain countries in East and Southeast Asia.
“Civorebrutinib represents a strategic and complementary addition to our rare kidney disease portfolio, with the potential to become a best-in-class therapy across multiple immune-mediated rare kidney diseases,” said Eric Dube, Ph.D., president and chief executive officer of Travere Therapeutics. “Patients living with rare kidney diseases still face significant unmet need, and we believe the progress made to date in IgAN and FSGS is only the beginning of what is possible for these communities. Travere has helped to deliver important firsts in these diseases, and we believe our expertise, infrastructure and deep commitment to the rare kidney community position us well to continue advancing innovation for patients. With proof-of-concept data in primary membranous nephropathy, a differentiated profile as an oral, reversible BTK inhibitor, and expected broad mechanistic applicability across diseases such as immune-mediated FSGS, minimal change disease and beyond, we believe civorebrutinib has the potential to meaningfully advance the treatment paradigm for rare kidney disease patients.”
“This collaboration with Travere brings together deep expertise in kidney disease development and commercialization and we look forward to advancing civorebrutinib in primary membranous nephropathy, immune-mediated FSGS, and minimal change disease, delivering transformative therapies for patients with serious kidney diseases worldwide,” said Mr. Yifang Wu, Chairman of the Board of Everest Medicines. “As a differentiated, potential best-in-class therapy, civorebrutinib has demonstrated encouraging efficacy in primary membranous nephropathy. With its highly selective and reversible covalent mechanism of action, it is well positioned to advance in development across multiple immune-mediated kidney indications. Everest remains committed to our dual-engine strategy of business development partnerships and in-house R&D. This collaboration will accelerate the global development and potential commercialization of civorebrutinib, expanding its clinical and future commercial value in autoimmune kidney diseases and the ability to deliver more innovative treatment options to patients.”
Civorebrutinib is an investigational oral, covalent reversible BTK inhibitor designed to provide differentiated efficacy, safety and convenience for patients with rare, immune-mediated kidney diseases, including primary membranous nephropathy (PMN), with planned evaluation in focal segmental glomerulosclerosis (FSGS), minimal change disease (MCD) and potentially additional indications. BTK is a key mediator of B-cell receptor signaling and plays an important role in B-cell activation, maturation, proliferation, and differentiation into antibody-producing cells.
In immune-mediated kidney diseases, B-cell activation and autoantibody production are believed to contribute directly to kidney injury. Civorebrutinib has demonstrated proof of concept in a Phase 1/2 clinical trial of patients with PMN. The previously reported Phase 1/2 data demonstrated rapid and sustained reductions in anti-PLA2R autoantibodies and proteinuria, with high rates of immunologic and clinical remission and stable kidney function through 52 weeks of follow-up. Civorebrutinib has been generally well tolerated throughout the development program to date.
As innovation in rare kidney diseases continues to accelerate, patients still face significant unmet need and limited treatment options across many serious conditions. Civorebrutinib has the potential to serve as a pipeline-in-a-product across multiple immune-mediated kidney diseases. Travere plans to investigate civorebrutinib in PMN, immune-mediated FSGS and MCD, with the potential for additional indications. These diseases share immune-mediated mechanisms that can lead to glomerular damage, resulting in proteinuria and impaired kidney function that may ultimately require dialysis or transplant. Civorebrutinib may also broaden future treatment approaches in FSGS, where both nephroprotective and targeted immune control approaches may play important roles.
Under the terms of the agreement, Everest will receive an upfront payment of $112.5 million in exchange for granting Travere exclusive development and commercialization rights for civorebrutinib in all markets outside of China and certain countries in East and Southeast Asia. Everest is also eligible to receive up to approximately $1.03 billion in additional cash payments tied to specified clinical development, regulatory and commercial milestones across up to five indications. Travere will also pay tiered royalties on future sales it its licensed territories, ranging from high single-digit to double-digit percentages based on annual net sales thresholds. The license agreement will become effective upon satisfaction of customary conditions, including expiration or termination of the applicable waiting period under the Hart-Scott-Rodino Antitrust Improvements Act of 1976, as amended.
Conference Call Information
Travere Therapeutics will host a conference call and webcast today, Tuesday, June 2, 2026, at 8:30 a.m. ET. To participate in the conference call, dial +1 (833) 461-5787 (U.S.) or +1 (585) 542-9983 (International), conference ID 574 733 925 shortly before 8:30 a.m. ET. The webcast can be accessed on the Investor page of Travere’s website at ir.travere.com/events-and-presentations. Following the live webcast, an archived version of the call will be available for 30 days on the Company’s website.
About Civorebrutinib
Civorebrutinib (also known as EVER001) is a next-generation covalent reversible Bruton's tyrosine kinase (BTK) inhibitor in development globally for the treatment of renal diseases. BTK is an essential component of the B-cell receptor signaling pathways that regulate the survival, activation, proliferation, and differentiation of B lymphocytes. Targeting BTK with small molecule inhibitors has been demonstrated to be an effective treatment option for B-cell autoimmune diseases.
About Travere Therapeutics
At Travere Therapeutics, we are in rare for life. We are a biopharmaceutical company that comes together every day to help patients, families and caregivers of all backgrounds as they navigate life with a rare disease. On this path, we know the need for treatment options is urgent – that is why our global team works with the rare disease community to identify, develop and deliver life-changing therapies. In pursuit of this mission, we continuously seek to understand the diverse perspectives of rare patients and to courageously forge new paths to make a difference in their lives and provide hope – today and tomorrow. For more information, visit travere.com.
About Everest Medicines
Everest Medicines is a biopharmaceutical company focused on discovering, developing, manufacturing and commercializing innovative pharmaceutical products that address critical unmet medical needs for patients in global markets. The management team of Everest Medicines has deep expertise and an extensive track record both in China and with leading global pharmaceutical companies.
The Company’s therapeutic areas of focus include CKM (cardiovascular, kidney, and metabolic), autoimmune, ophthalmology and critical care. Everest Medicines has developed a fully integrated commercialization platform that combines omnichannel commercial capabilities with end-to-end product lifecycle management. Leveraging its proprietary mRNA platform, the Company is advancing its existing pipeline, including mRNA in vivo CAR-T and mRNA cancer vaccines, while selectively expanding into additional high-value therapeutic areas with blockbuster potential, and accelerating its global expansion. For more information, please visit the Company’s website: www.everestmedicines.com.
Forward Looking Statements
This press release contains “forward-looking statements” as that term is defined in the Private Securities Litigation Reform Act of 1995. Without limiting the foregoing, these statements are often identified by the words “on-track,” “positioned,” “look forward to,” “will,” “would,” “may,” “might,” “believes,” “anticipates,” “plans,” “expects,” “intends,” “potential,” or similar expressions. In addition, expressions of strategies, intentions or plans are also forward-looking statements. Such forward-looking statements include, but are not limited to, references to: statements regarding the Company's beliefs about the future potential of its pipeline and portfolio; statements regarding the Company's capabilities, competitive positioning, and strategic plans; statements and expectations regarding the potential of civorebrutinib to serve as a pipeline-in-a-product and to potentially become a best-in-class therapy across multiple immune-mediated kidney diseases, and its potential to provide differentiated efficacy, safety and convenience for the indications described herein; statements and expectations regarding the expected broad mechanistic applicability across diseases; statements and expectations regarding future treatment approaches and paradigms; statements and expectations regarding the clinical studies and data described herein; statements and expectations regarding potential future payments (including upfront, milestone and royalty payments) and, as applicable, the potential achievement and timing thereof; statements and expectations regarding the activities of the Company’s partners and collaborators; and statements related to the estimated sizes of patient populations. Such forward-looking statements are based on current expectations and involve inherent risks and uncertainties, including factors that could delay, divert or change any of them, and could cause actual outcomes and results to differ materially from current expectations. No forward-looking statement can be guaranteed. Among the factors that could cause actual results to differ materially from those indicated in the forward-looking statements are risks and uncertainties related to the license agreement with Everest, including the ability of the parties to obtain required regulatory approvals and satisfy other applicable conditions, and the ability of the Company to successfully advance the product through clinical trials toward potential future regulatory approval. The Company also faces risks and uncertainties related to its business and finances in general, the success of its commercial products, risks and uncertainties associated with its preclinical and clinical stage pipeline, risks and uncertainties associated with the regulatory review and approval process, risks and uncertainties associated with enrollment of
clinical trials for rare diseases, and risks that ongoing or planned clinical trials may not succeed or may be delayed for safety, regulatory or other reasons. Specifically, the Company faces risks associated with the commercial launch of FILSPARI in FSGS and the ongoing commercialization in IgAN, the timing and potential outcome of its and its partners’ clinical studies, market acceptance of its commercial products including efficacy, safety, price, reimbursement, and benefit over competing therapies, risks related to the challenges of manufacturing scale-up, risks associated with the successful development and execution of commercial strategies for such products, including FILSPARI, and risks and uncertainties related to the current administration, including but not limited to risks and uncertainties related to tariffs and the funding, staffing and prioritization of resources at government agencies including the FDA. The Company also faces the risk that it will be unable to raise additional funding that may be required to complete development of any or all of its product candidates, including as a result of macroeconomic conditions; risks relating to the Company’s dependence on contractors for clinical drug supply and commercial manufacturing; uncertainties relating to patent protection and exclusivity periods and intellectual property rights of third parties; risks associated with regulatory interactions; and risks and uncertainties relating to competitive products, including current and potential future generic competition with certain of the Company’s products, including potential ANDA filings or patent challenges, and technological changes that may limit demand for the Company’s products. The Company also faces additional risks associated with global and macroeconomic conditions, including health epidemics and pandemics, including risks related to potential disruptions to clinical trials, commercialization activity, supply chain, and manufacturing operations. You are cautioned not to place undue reliance on these forward-looking statements as there are important factors that could cause actual results to differ materially from those in forward-looking statements, many of which are beyond our control. The Company undertakes no obligation to publicly update any forward-looking statement, whether as a result of new information, future events, or otherwise. Investors are referred to the full discussion of risks and uncertainties, including under the heading “Risk Factors”, as included in the Company’s most recent Form 10-K, Form 10-Q and other filings with the Securities and Exchange Commission.
Contact Info
Media: 888-969-7879 | Investors: 888-969-7879 | ||||
Exhibit 31.1
CERTIFICATION OF CHIEF EXECUTIVE OFFICER
PURSUANT TO EXCHANGE ACT RULE 13a-14(a) OR 15d-14(a)
I, Eric Dube, certify that:
1. I have reviewed this Quarterly Report on Form 10-Q of Travere Therapeutics, Inc.;
2. Based on my knowledge, this report does not contain any untrue statement of a material fact or omit to state a material fact necessary to make the statements made, in light of the circumstances under which such statements were made, not misleading with respect to the period covered by this report;
3. Based on my knowledge, the financial statements and other financial information included in this report, fairly present in all material respects the financial condition, results of operations and cash flows of the registrant as of, and for, the periods presented in this report;
4. The registrant’s other certifying officer(s) and I are responsible for establishing and maintaining disclosure controls and procedures (as defined in Exchange Act Rules 13a-15(e) and 15d-15(e)) and internal control over financial reporting (as defined in Exchange Act Rules 13a-15(f) and 15d-15(f)) for the registrant and have:
(a) Designed such disclosure controls and procedures, or caused such disclosure controls and procedures to be designed under our supervision, to ensure that material information relating to the registrant, including its consolidated subsidiaries, is made known to us by others within those entities, particularly during the period in which this report is being prepared;
(b) Designed such internal control over financial reporting, or caused such internal control over financial reporting to be designed under our supervision, to provide reasonable assurance regarding the reliability of financial reporting and the preparation of financial statements for external purposes in accordance with generally accepted accounting principles;
(c) Evaluated the effectiveness of the registrant’s disclosure controls and procedures and presented in this report our conclusions about the effectiveness of the disclosure controls and procedures, as of the end of the period covered by this report based on such evaluation; and
(d) Disclosed in this report any change in the registrant’s internal control over financial reporting that occurred during the registrant’s most recent fiscal quarter (the registrant’s fourth fiscal quarter in the case of an annual report) that has materially affected, or is reasonably likely to materially affect, the registrant’s internal control over financial reporting; and
5. The registrant’s other certifying officer(s) and I have disclosed, based on our most recent evaluation of internal control over financial reporting, to the registrant’s auditors and the audit committee of the registrant’s board of directors (or persons performing the equivalent functions):
(a) All significant deficiencies and material weaknesses in the design or operation of internal control over financial reporting which are reasonably likely to adversely affect the registrant’s ability to record, process, summarize and report financial information; and
(b) Any fraud, whether or not material, that involves management or other employees who have a significant role in the registrant’s internal control over financial reporting.
| Date: August 4, 2026 | |||||||||||
| /s/ Eric Dube | |||||||||||
| Eric Dube | |||||||||||
| Chief Executive Officer | |||||||||||
| (Principal Executive Officer) | |||||||||||
Exhibit 31.2
CERTIFICATION OF CHIEF FINANCIAL OFFICER
PURSUANT TO EXCHANGE ACT RULE 13a-14(a) OR 15d-14(a)
I, Christopher Cline, certify that:
1. I have reviewed this Quarterly Report on Form 10-Q of Travere Therapeutics, Inc.;
2. Based on my knowledge, this report does not contain any untrue statement of a material fact or omit to state a material fact necessary to make the statements made, in light of the circumstances under which such statements were made, not misleading with respect to the period covered by this report;
3. Based on my knowledge, the financial statements and other financial information included in this report, fairly present in all material respects the financial condition, results of operations and cash flows of the registrant as of, and for, the periods presented in this report;
4. The registrant’s other certifying officer(s) and I are responsible for establishing and maintaining disclosure controls and procedures (as defined in Exchange Act Rules 13a-15(e) and 15d-15(e)) and internal control over financial reporting (as defined in Exchange Act Rules 13a-15(f) and 15d-15(f)) for the registrant and have:
(a) Designed such disclosure controls and procedures, or caused such disclosure controls and procedures to be designed under our supervision, to ensure that material information relating to the registrant, including its consolidated subsidiaries, is made known to us by others within those entities, particularly during the period in which this report is being prepared;
(b) Designed such internal control over financial reporting, or caused such internal control over financial reporting to be designed under our supervision, to provide reasonable assurance regarding the reliability of financial reporting and the preparation of financial statements for external purposes in accordance with generally accepted accounting principles;
(c) Evaluated the effectiveness of the registrant’s disclosure controls and procedures and presented in this report our conclusions about the effectiveness of the disclosure controls and procedures, as of the end of the period covered by this report based on such evaluation; and
(d) Disclosed in this report any change in the registrant’s internal control over financial reporting that occurred during the registrant’s most recent fiscal quarter (the registrant’s fourth fiscal quarter in the case of an annual report) that has materially affected, or is reasonably likely to materially affect, the registrant’s internal control over financial reporting; and
5. The registrant’s other certifying officer(s) and I have disclosed, based on our most recent evaluation of internal control over financial reporting, to the registrant’s auditors and the audit committee of the registrant’s board of directors (or persons performing the equivalent functions):
(a) All significant deficiencies and material weaknesses in the design or operation of internal control over financial reporting which are reasonably likely to adversely affect the registrant’s ability to record, process, summarize and report financial information; and
(b) Any fraud, whether or not material, that involves management or other employees who have a significant role in the registrant’s internal control over financial reporting.
| Date: August 4, 2026 | |||||||||||
| /s/ Christopher Cline | |||||||||||
| Christopher Cline | |||||||||||
| Chief Financial Officer | |||||||||||
| (Principle Financial Officer) | |||||||||||
Exhibit 32.1
CERTIFICATION OF
CHIEF EXECUTIVE OFFICER
PURSUANT TO 18 U.S.C. SECTION 1350
AS ADOPTED PURSUANT TO
SECTION 906 OF THE SARBANES-OXLEY ACT OF 2002
In connection with the accompanying Quarterly Report on Form 10-Q of Travere Therapeutics, Inc. (the “Company”), for the period ending June 30, 2026 (the “Report”), the undersigned officer of the Company hereby certifies pursuant to 18 U.S.C. Section 1350, as adopted pursuant to Section 906 of the Sarbanes-Oxley Act of 2002, that, to such officer’s knowledge:
1. The Report fully complies with the requirements of Section 13(a) or 15(d) of the Securities Exchange Act of 1934; and
2. The information contained in the Report, fairly presents, in all material respects, the financial condition and results of operations of the Company.
| Date: August 4, 2026 | |||||||||||
| /s/ Eric Dube | |||||||||||
| Eric Dube | |||||||||||
| Chief Executive Officer | |||||||||||
| (Principal Executive Officer) | |||||||||||
Exhibit 32.2
CERTIFICATION OF
CHIEF FINANCIAL OFFICER
PURSUANT TO 18 U.S.C. SECTION 1350
AS ADOPTED PURSUANT TO
SECTION 906 OF THE SARBANES-OXLEY ACT OF 2002
In connection with the accompanying Quarterly Report on Form 10-Q of Travere Therapeutics, Inc. (the “Company”), for the period ending June 30, 2026 (the “Report”), the undersigned officer of the Company hereby certifies pursuant to 18 U.S.C. Section 1350, as adopted pursuant to Section 906 of the Sarbanes-Oxley Act of 2002, that, to such officer’s knowledge:
1. The Report fully complies with the requirements of Section 13(a) or 15(d) of the Securities Exchange Act of 1934; and
2. The information contained in the Report, fairly presents, in all material respects, the financial condition and results of operations of the Company.
| Date: August 4, 2026 | |||||||||||
| /s/ Christopher Cline | |||||||||||
| Christopher Cline | |||||||||||
| Chief Financial Officer | |||||||||||
| (Principal Financial Officer) | |||||||||||